


INTRODUCTION Counterfeit and substandard drugs
Counterfeit and substandard drugs present unique and complex challenges to the fields of
forensic science and criminal justice. These goods seriously threaten public health and safety.
Analytical testing can be challenging because forensic analysis of counterfeit drugs typically
goes beyond more traditional pharmaceutical methods of drug analysis. Criminal justice and
law-enforcement organizations are being forced to evaluate current policies and practices so
successful investigation and adjudication in these types of cases is possible. The internet has
made it possible to easily access these drugs even in places like the United States that have a
well-controlled legitimate pharmaceutical supply chain. The problem is multidimensional and
incorporates matters of law, science, criminal justice, public health, and public policy. There is a
need to delineate the integral aspects of the counterfeit drug trade if an effective solution is to be
proposed.
Detection and identification of counterfeit drugs is a critical step in the process of
addressing the counterfeit-drug problem. The ability to individualize is the foundation of the
discipline of criminalistics and typically the goal of the forensic scientist. Classification is also
important. (De Forest, Gaensslen, & Lee, 1983) Scientific testing performed to achieve these
goals must use methods that work accurately and reliably. Analytical testing of counterfeit drugs
in the field is usually performed to differentiate an authentic from a non-authentic article and to
classify these goods. This capability has value and may help determine source or origin. In
situations where evidence will be used court, technologies and methods used must meet the
standards of admissibility of scientific evidence. Historically, results of tests performed using
field instruments were used in a presumptive manner (Fedchak, 2014) to support investigations
and for probable cause. More recently, the need to collect court-admissible data in the field is

desirable because it may save time and money in the long term. Another significant advantage of
the use of diagnostic field tests is the ability to deal with ‘‘dissipating evidence’’ such as gunshot
residue or explosive traces on the hands of suspects. (Almog, 2006) When analyzing counterfeit
drugs, additional considerations make the forensic testing of these goods in the field important.
This testing is unique and challenging.
When analyzing a drug in a pharmaceutical setting to confirm identity, infrared or mass
spectra may be collected from the sample and used for identification. An additional routine test
may then be used to quantify the amount of drug present or establish its distribution throughout
the final dosage form. Once the tablet is identified and quantified using standard test methods
that were developed during the drug-development process, analysis is usually complete. When
analyzing counterfeit drugs in a forensic setting, however, identification of the active
pharmaceutical ingredient (API) is likely only one of many steps in the analytical work flow.
Analysis of counterfeits may begin with the evaluation of packaging used. A comparison of
coatings on laminates used for both dosage form and bulk packaging may serve as a means for
differentiation. Physical comparisons of markings and other sample features may also be used
for this purpose. Once inside the packaging, physical and chemical characteristics of the dosage
form may then be evaluated. If the API is present, further testing is likely required. Counterfeits
may have the proper API in the wrong dosage or solid-state form. Solid-state form of the API is
important for a variety of reasons. An example is polymorphism where different polymorphs of
the same chemical may exhibit different bioavailability. The polymorphic form created during
manufacture may also have legal significance because one polymorphic form may be covered
under patent whereas others may not. In the case of different solid-state hydrates of the API, the
wrong hydration level may result in under dosing tablets when hydration levels are too high, or

over dosing tablets when hydration levels are too low. In addition dosages and solid-state form
differences, particle size or distribution throughout the dosage form may be different, or residual
solvents present may be different. These differences may be used as points of comparison
amongst samples. When evaluating counterfeits, the inactive ingredients may be different from
the authentic versions of the drug and need to be identified and characterized. Foreign particles
present in the counterfeit versions may serve as an additional feature for differentiation and
identification. In these cases, non-routine methods of analysis need to be employed.
The analytical testing of counterfeit drugs is also unique because it is typically the goal of
the forensic scientist to establish provenance when counterfeit drugs are presented for analysis.
No method or combination of methods has been shown to be universally applied to achieve this
goal. Attempts are usually made to link samples to each other using some chemical or physical
marker present in the sample. (Alabdalla, 2005; Lopatka & Vallat, 2011; Ortiz, et al., 2011)
These attempts have been extended to the sourcing of illicit drugs as well. Creating these links
between samples is important. Counterfeiters are frequently members of terrorist organizations
or organized crime groups. (United Nations Office on Drugs and Crime, 2014) Establishing
distribution chains and linking samples from different geographic locations may help bring
justice. This justice may be important criminally, for example, in cases when counterfeits cause
harm or death; or they may be important in civil cases where patent-infringement violations have
cost brand owners significant costs for which they seek compensation. A significant limitation of
all methods currently used to establish provenance is in the limited number of samples available
for analysis for any given study. (Pérez-Bernal, Amigo, Fernández-Torres, Bello, & Callejón-
Mochón, 2011) Testing of samples in the field and creation of a mechanism for lawenforcement
agencies from different locations to share this data would be very valuable, and would provide

the forensic scientist with access to larger numbers of samples. These shared data could be used
for comparisons, to evaluate trends, and perform other types of data review useful in establishing
provenance.
The counterfeit-drug problem is also complex because of both the patenting system and
laws governing patented drugs. In the United States, patents are granted through the United
Stated Patent and Trademark Office (USPTO) and are integrated within the development
lifecycle of pharmaceutical innovators and generic manufacturers. The United States patenting
system as applied to the pharmaceutical industry was completely redefined by The Drug Price
Competition and Restoration Act of 1984, also known as the Hatch-Waxman Act. This law was
designed to protect rights of innovators, while at the same time allowing for generic drugs to
come to the market as quickly as possible. It fundamentally changed the patenting system in the
United States and has had a significant impact on patenting of pharmaceutically relevant
compounds, drug development, the Food and Drug Administration (FDA) regulatory process,
pharmaceutical litigation, patent protection, and patent enforcement. It is important that the
patenting and drug development processes, and specifically the impact of the Hatch-Waxman Act
on these processes, is understood. The impact of counterfeit drugs on criminal justice and
forensic science are directly related to both patenting and drug development.
While it is important to understand the patenting processing and its integration with drug
development, understanding the value of patents is also important. Patents are granted because
the patented invention will provide a benefit to society. This is especially true in the
pharmaceutical industry. Drugs patented in the United States are developed and tested in
accordance with FDA guidelines with the intention of bringing the patented invention to the
market in the form of a new drug. Without these patented inventions, society loses and the future

of pharmaceutical development is challenged. Pharmaceutical innovation is an important part of
society. It helps cure and treat disease, and improves quality of life. New drugs will be needed
to remedy the health problems being created and perpetuated by the presence and proliferation of
counterfeit and substandard drugs in many regions of the world. In places like Africa and Asia,
anti-infective medicines containing sub-therapeutic amounts of API are commonly encountered.
These bogus drugs, whether counterfeit or substandard, increase the risk of the spread of
resistance of disease to available drugs. (Newton, et al., 2006) Subtherapeutic amounts of drug
promote resistance. This factor, coupled with other issues that indicate diseases like malaria are
impacted by environmental change (Siraj, et al., 2014), makes the counterfeit-drug problem even
more troubling.
The future of pharmaceutical innovation is in question. The drug-development life cycle
is long, risky, and expensive. It seems fair to question the future of pharmaceutical innovation
and the development of new medicines in a global environment where intellectual property (IP)
rights are not protected and enforced. These rights are an inherent aspect of the counterfeit-drug
problem. Therefore, it is important that the role patents and protection of patent rights play both
nationally and internationally are understood.
Counterfeit drugs present challenges to practitioners of law and justice. Identification and
prosecution of criminals responsible for the infiltration of counterfeit drugs into the market is
extremely challenging for a variety of reasons. For instance, determination of source or origin of
the drug is not easy, especially considering the global nature of this problem. In situations where
determination of source is possible, local laws and jurisdictional issues frequently hamper
prosecution. In addition, prosecution of counterfeiters is almost impossible in some regions of
the world where IP rights are meaningless. Even in places like the United States, enforcement of

IP protection for non-pharmaceutical products can be very difficult and require a level of
diligence on the part of the brand owner not usually required of victims of crimes. While some
relate these cases to being civil cases, IP rights are protected by law and breaking them is a
criminal activity. (Carruthers & Ariovich, 2004) For pharmaceutical products, FDA regulations
highlight the criminal aspect of this problem a little bit better. While brand owners may have an
unusual burden to achieve criminal justice, there is, at least, a regulatory, judicial, and legal
framework in the United States within which these issues may be addressed. Pharmaceutical
suppliers governed by FDA must adhere to and work in the United States legal and regulatory
systems. Within this jurisdiction, regulations require that suppliers adhere to certain guidelines
and meet minimum safety, efficacy, and other criteria in order to receive approval to sell
pharmaceutical drug products to the American consumer. If these guidelines are not met, laws
and statutes are in place to ensure that these drug products are not legally available for sale.
Interestingly, although historically the United States has experienced a tightly controlled
pharmaceutical supply chain, the FDA appears to be exercising more stringent control over
manufacturing sites located overseas. (Harris, 2014; Siddiqui & Chatterjee, 2014) Although
firms that legally supply drugs to the United States have always been governed by FDA,
inspections of these facilities are difficult due to geographical location. (Staton, 2012) In
February 2014, The New York Times reported that India, the second-largest exporter of overthe-
counter and prescription drugs to the United States, is coming under increased scrutiny by
American regulators for safety lapses, falsified drug test results, and selling fake medicines. This
is important to the American consumer because India's pharmaceutical industry supplies 40
percent of over-the-counter and generic prescription drugs consumed in the United States.

(Harris, 2014) In March 2014, the FDA banned imports from Indian generic manufacturer’s Sun
Pharmaceutical Industries Ltd's plant at Karkhadi in the western state of Gujarat, India. (Siddiqui
& Chatterjee, 2014) Although the exact reason for the ban was not cited, these recent events
indicate that the drug problem, specifically quality and counterfeits, is serious.
Just as importantly, while it is possible to stop sale within the United States in the
legitimate supply chain, it is very difficult to apply statutes and laws, and enforce these laws
when they are jurisdictionally unenforceable. The United States has no jurisdiction in many
regions of the world where counterfeit drugs are a serious problem. This situation is further
complicated by the fact that counterfeit drugs present significant public-health risks, especially in
less developed and third-world societies where counterfeit drugs are most prolific. In order to
address this issue, the role that trade agreements play in enforcement of the goods is important.
It is not always possible to apply local laws and enforcement in foreign lands where counterfeits
are produced for sale locally, and methods proposed to stop the proliferation of these goods,
including trade sanctions must be considered and reviewed.
CHAPTER 2. PROJECT SUMMARY
Overview
Counterfeit drugs are a problem for criminal justice. Theft of IP in the pharmaceutical
industry has caused sickness and death (Newton, et al., 2006; World Health Organziation, 2008;
Wondemagegnehu, 1999). It is necessary that the perpetrators of these crimes are brought to
justice. In some cases, threats to health and safety are short term; in others they are long term.
This research describes these threats in detail.

The problem is complex, and there are many important factors that need to be considered
to successfully address the problem. Public-health challenges, drug-development challenges,
legal-challenges, and financial considerations are all important aspects that need to be
understood. This research details these topics so appropriate solutions to the problem can be
proposed.
This project is also important to forensic science. It defines the challenges faced by the
field of forensic science and proposes methods that may be used to address these challenges.
This will enable forensic scientists to provide support to law enforcement and other members of
the legal system to successfully investigate and adjudicate these crimes.
This dissertation details and describes the integral aspects of the counterfeit-drug problem
related to forensic science and the criminal justice system. It is important to start, therefore, by
defining the term “counterfeit drug”. Depending upon the source, the geographic location, and
the context, this term may have different meanings. For the purpose of this dissertation, a
counterfeit drug is one which is deliberately and fraudulently mislabeled with respect to identity
and/or source. For instance, a counterfeit VIAGRA® tablet is designed to look as if the tablet
was manufactured by Pfizer, the brand owner. The color, size, shape, and tablet markings would
be such that they give the impression they are an authentic Pfizer VIAGRA tablet. Figure 1
(Hensley, 2013) shows the front and back of authentic and counterfeit VIAGRA tablets. The
tablets on the left (top and bottom) are counterfeit; the tablets on the right (top and bottom) are
authentic.

Figure 1. Counterfeit VIAGRA pills on the left (top and bottom)
and authentic VIAGRA on the right (top and bottom).
The problem of counterfeit drugs is also significantly impacted by substandard drugs.
Substandard drugs are considered in this dissertation. Sometimes with regard to drugs, the terms
“counterfeit drug” and “substandard drug” are mistakenly used interchangeably. Substandard
drugs include, but are not limited to, illegal imitations, expired drugs repackaged for sale, and
drugs that have not been stored properly and have may have lost efficacy. An illegal imitation is
a drug whose sale is in violation of government regulation (and is, therefore, illegal), but is not a
counterfeit because the drug is not purported to be the brand name of the patent holder (the drug
is “generic”). An example of an illegal imitation is an unregulated generic version of a drug. For
instance, tablets of sildenafil citrate (VIAGRA) sold within the United States, but that do not
represent themselves as being from Pfizer, the brand owner of VIAGRA, are illegal imitations.
Generic versions of VIAGRA are not legal for sale in the United States. They are “illegal”.
They are “imitations” because they are not purported to be VIAGRA, but rather they imitate
VIAGRA. Figure 2 shows an example of an illegal imitation. The tablet on the left is generic
VIAGRA that was purchased in the United States. The “DP” marking on the tablet is not likely
to be confused with the markings on the authentic VIAGRA tablet, but the product is intended to
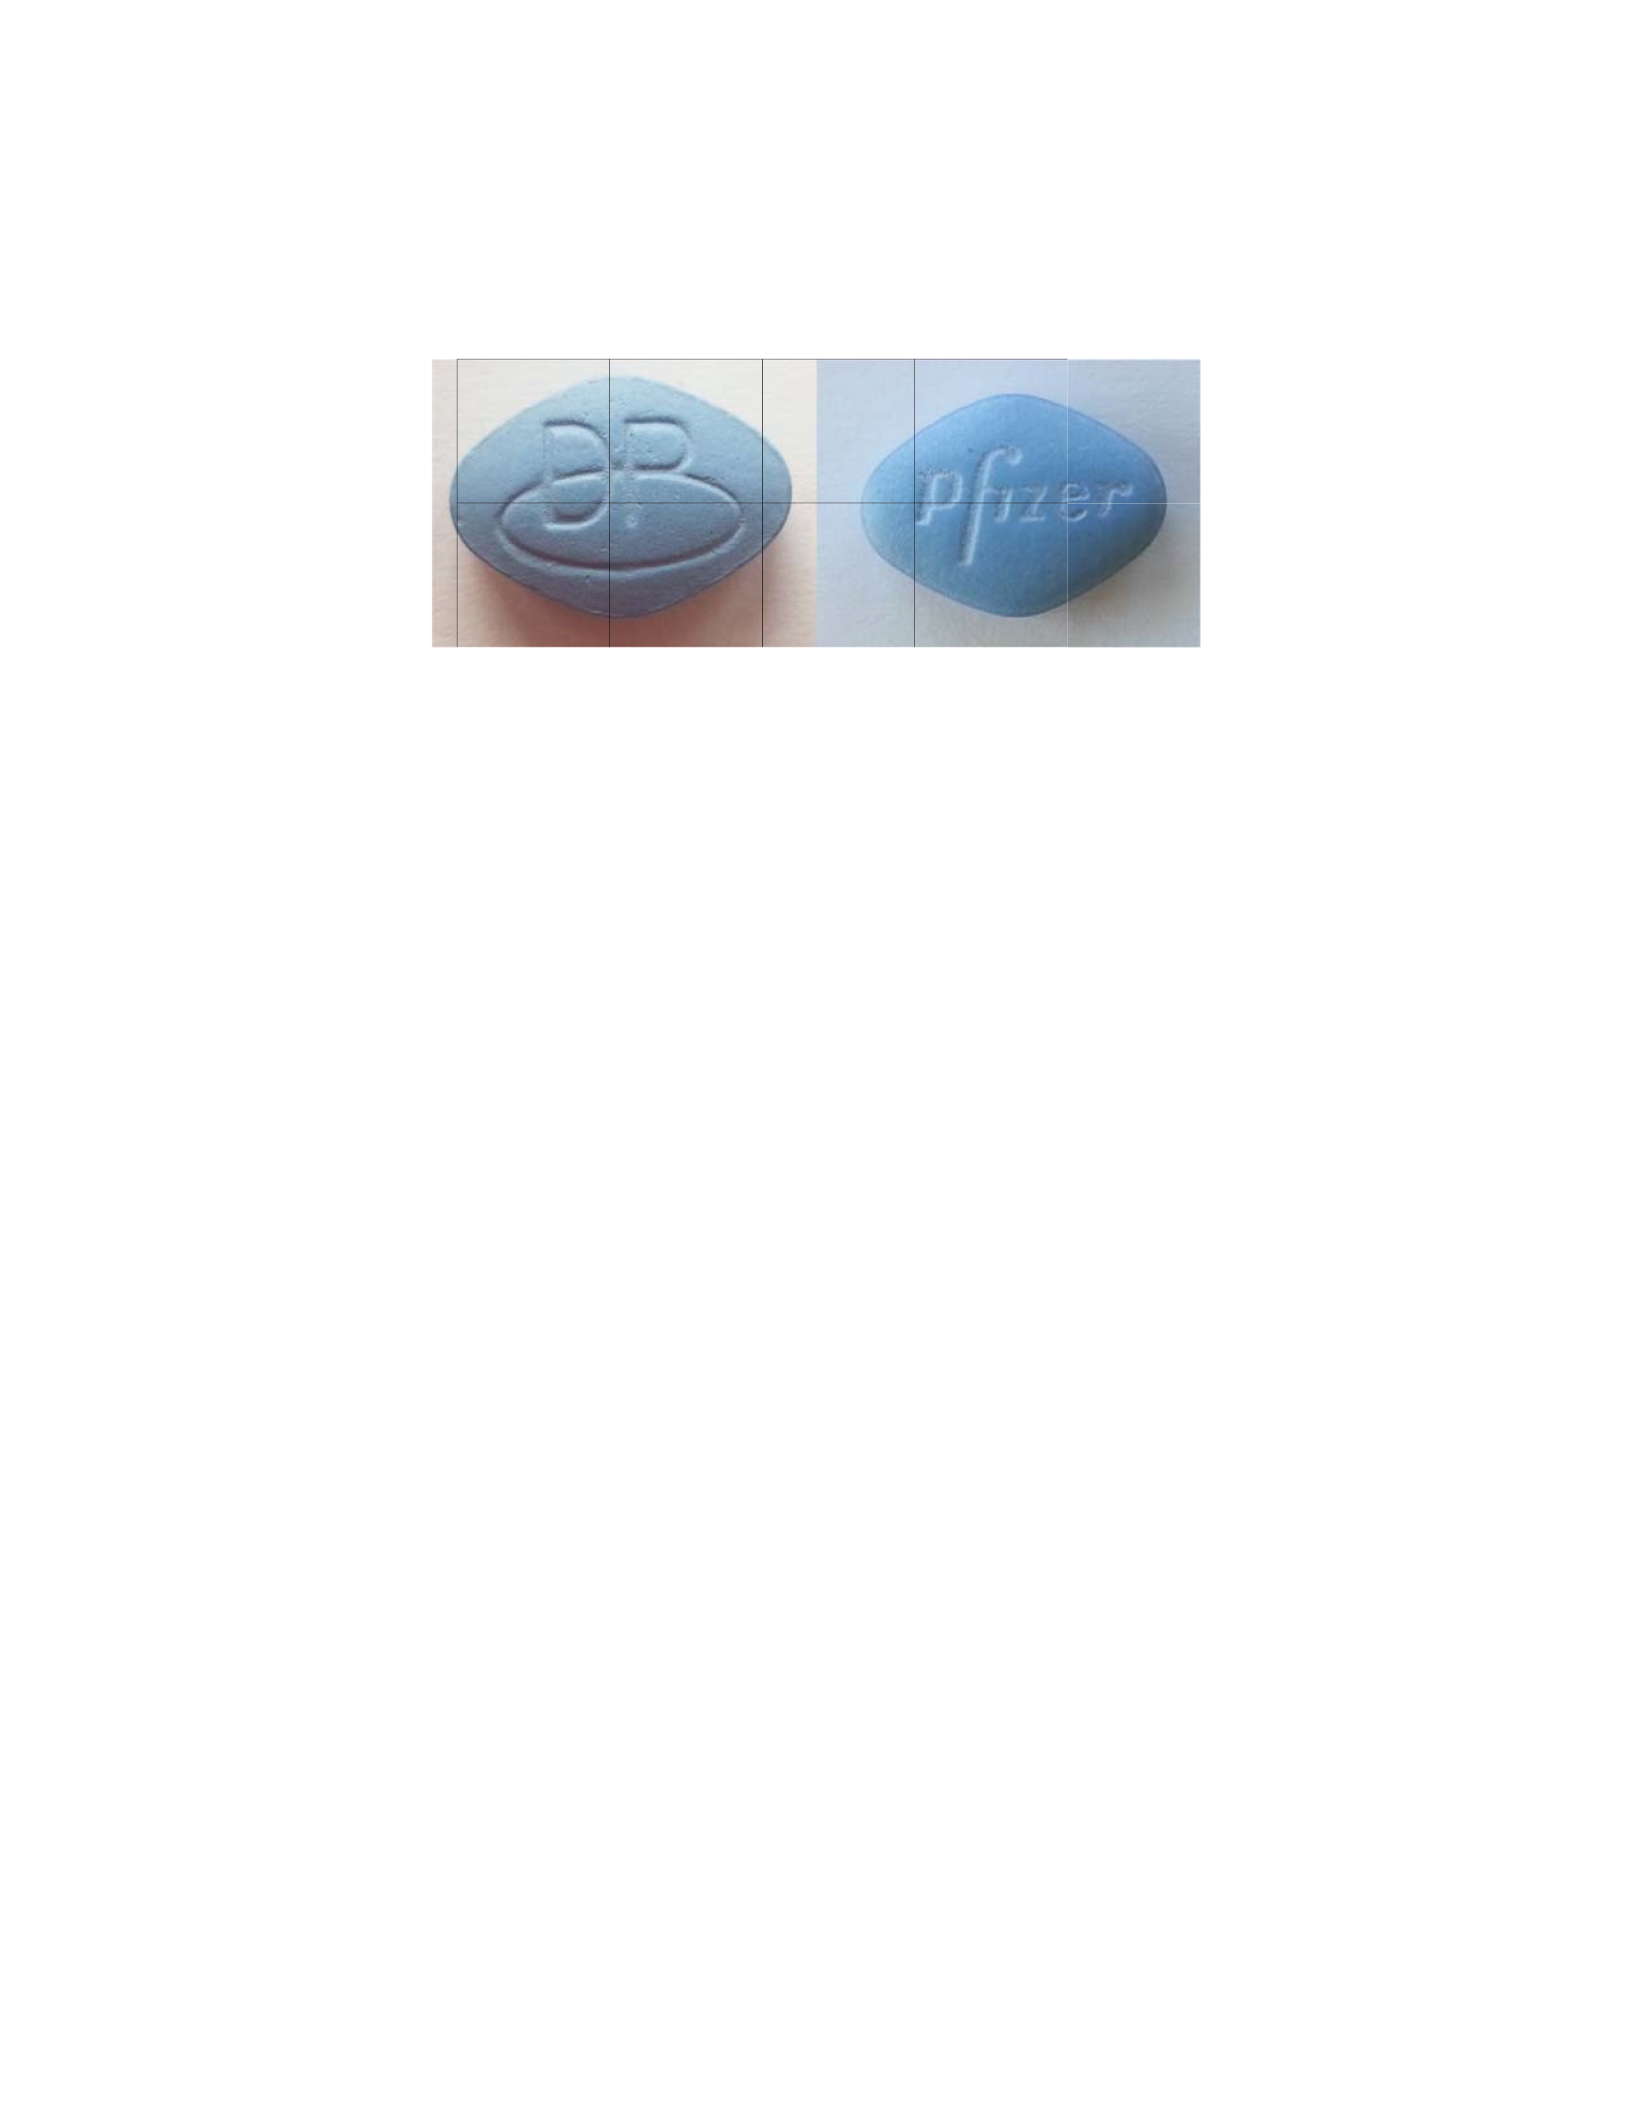
imitate the authentic VIAGRA product. It is extremely important to recognize that these
definitions may be convoluted and should always be defined in context.
Figure 2. The tablet on the left is an illegal imitation (substandard)
of the authentic VIAGRA tablet shown on the right.
Immediate and long-term threats to public health are the primary reason counterfeit drugs
are such an important issue. The impact of counterfeit drugs on public health is described in
chapter 3. Analytical challenges faced by the forensic scientist are described in chapter 4.
Chapter 5 describes the values and challenges of field-portable instrumentation, while chapter 6
details the samples and sampling matrix for this research. Chapters 7 through 10 describe
analytical testing performed on samples using field-portable instruments to aid in the successful
investigation and adjudication of counterfeiters. Dimensional measurements, infrared
spectroscopy, Raman spectroscopy, and gas chromatography-mass spectrometry (GC-MS) were
all used to determine the ability of each technology to identify and source counterfeits. Chapter
11 summarizes the analytical testing by describing a proposed method and workflow for the
analytical testing of non-authentic samples in the field using portable instruments. In addition,
the challenges of admissibility as evidence in United States courts of each of these technologies
for this intended use in the field is presented. Chapters 12 through 16 detail the aspects of the
counterfeit-drug trade that impact public health, and therefore, impact forensic science and
criminal justice. It is important to describe these roles and processes so that the problem is

understood and an appropriate solution can be proposed. Chapter 12 describes the
drugdevelopment process regulated by FDA. Chapters 13 and 14 describe patents and the
patenting process as related to the pharmaceutical industry, including a discussion of the Hatch-
Waxman Act and critical aspects relevant to IP protection and future of the pharmaceutical
innovation. Chapter 15 describes ways to enforce IP rights including enforceable laws in some
jurisdictions, and sanctions and trade agreements in others. Chapter 16 describes funding of
pharmaceuticals. Chapter 17 summarizes the long-term challenges counterfeit drugs have on
public health. This chapter will describe how counterfeit drugs will impact the future of
pharmaceutical innovation. Finally, the thesis is summarized in Chapter 18. The value and
contribution of this project to forensic science and criminal justice is described.
Research Questions
The research questions that were answered in this dissertation are as follows:
(1) To what extent do counterfeit and substandard drugs present a public-health
problem?
(2) Is it possible to develop an analytical scheme to identify counterfeit drugs
using field-portable instrumentation?
(3) How will failure to protect and enforce IP rights threaten the future of
pharmaceutical innovation?
CHAPTER 3. TO WHAT EXTENT DO COUNTERFEIT AND SUBSTANDARD DRUGS
PRESENT A PUBLIC-HEALTH PROBLEM?
Counterfeit and substandard drugs pose immediate and long-term dangers to public health
in both developed and developing societies. The immediate dangers are usually directly related

to the physical and chemical composition of the drug product. In some instances, poisoning
occurs from ingestion; in others, an incorrect dosage of API renders the drug ineffective or
harmful. These immediate dangers may even be fatal and will be described in this chapter.
There are at least two long-term dangers counterfeit and substandard drugs pose to public
health. The first is the development of resistance to drugs that results from the proliferation of
counterfeit and substandard drugs; the second is the threat to pharmaceutical innovation that
these goods present. The development of drug resistance will be covered within this chapter.
The threat to pharmaceutical innovation is more complicated, and understanding this threat
requires an understanding of matters related to criminal justice, regulatory affairs, public health,
law, and science. These individual topics will be covered throughout this dissertation, and then
the threat to pharmaceutical innovation will be covered in chapter 17.
Immediate Danger of Counterfeit and Substandard Drugs
The immediate dangers of the counterfeit and substandard drug problem are highlighted
in the recent actions of the FDA in India. In February 2014, The New York Times reported that
India, the second-largest exporter of over-the-counter and prescription drugs to the United States,
is coming under increased scrutiny by American regulators for safety lapses, falsified drug test
results, and selling fake medicines. This is important to the American consumer because India's
pharmaceutical industry supplies 40 percent of over-the-counter and generic prescription drugs
consumed in the United States. (Harris, 2014) In March 2014, the FDA followed up this report
by banning imports from Indian generic manufacturer’s Sun Pharmaceutical Industries Ltd's
plant at Karkhadi in the western state of Gujarat, India. (Siddiqui & Chatterjee, 2014)
These events are important for a few reasons. First, they show the importance of the issue
of drug safety to regulators in the United States. Larger numbers of drugs are coming from

overseas where regulators traditionally have been less strict about enforcing drug safety and
other issues that would be easier to enforce locally at the point of manufacturer. For example,
resources available to perform facility inspections overseas have been limited. (Food and Drug
Administration, 2010; Staton, 2012) It seems, though, that the threat these goods are posing to
the American consumer is becoming too great and action at this time is required. (Harris, 2014;
Siddiqui & Chatterjee, 2014; Staton, 2012) Second, it shows the amount of control the FDA has
to protect its regulated drug trade. Although these plants are not located within the United States,
they are required to follow FDA guidelines since they will be selling their product within the
United States. This is important because regulation, including enforcement, helps to bring safety,
but is expensive. Cost is a factor when evaluating the problem of counterfeit drugs. Third, India
supplies a large percentage of generic prescriptions to the United States market, but the amount
of innovative medications coming from this region of the world is significantly lower than
innovation in the United States and Europe. This is important because the cost of innovation is
significantly higher than the cost of manufacturing generics. This is a very important aspect with
regard to the future of pharmaceutical innovation. These details will be discussed in later
chapters.
While these recent events are important and show there is a need to address the
counterfeit-drug problem in the United States, there are no geographical limits to the immediate
dangers of counterfeit and substandard drugs. In fact, you are much more likely to encounter a
counterfeit drug outside of the United States than you are within its borders. The record of
public harm due to counterfeit and substandard drugs throughout the world is well established.
Some of these instances have been summarized below.

In 2004, an otherwise healthy 22-year-old Argentinean woman being treated for a mild
case of anemia died after the seventh injection of a ten-injection treatment with a toxic
counterfeit iron supplement. Subsequent analysis showed that instead of iron sorbitol, the API in
AstraZeneca’s legitimate Yectafer®, the counterfeit samples contained a different iron derivative
in a concentration three times as high as in the original product. This caused fatal liver failure.
Authorities managed to prosecute several members of the counterfeit drug’s distribution chain,
but those who actually produced the drug were never identified or caught. (Loewy, 2007) On a
larger scale, over 100 people in Panama died as a result of taking toxic counterfeit glycerin in
2006. These deaths were directly linked to a Chinese manufacturer that made and exported
diethylene glycol as 99.5% pure glycerin. This toxic solution of diethylene glycol was
unwittingly mixed into 260,000 bottles of cold medicine. (Bogdanich & Hooker, 2007)
Death due to the lack of the proper dosage of API in counterfeit and substandard drugs is
also documented. In February 2005, a 23-year-old man presented with a fever to a rural hospital
in eastern Burma where he was diagnosed as having uncomplicated hyperparasitaemic
falciparum malaria by microscopy (4.2% infected red blood cells). He was treated with oral
artesunate, labeled as made by Guilin Pharmaceutical (Guangxi, People’s Republic of China), 4
mg/kg once a day, the treatment of choice in this region. On the third night the young Burmese
man became unconscious and was transferred to another hospital where he was found to be in a
coma (Glasgow Coma Score = 3/15), with renal failure and a higher parasitaemia (5.5% infected
red blood cells). He was perfused with intravenous fluids, received an injection of intravenous
artesunate (2.4 mg/kg) and was transferred to a third hospital where he died within 12 hours of
arrival from cerebral malaria. The amount of artesumate in the tablet was determined to be 10
mg, rather than the purported dosage of 50 mg. (Newton, et al., 2006) These authors refer to this

death and others like it as manslaughter. They present as part of their evidence published
research showing that artemisinin derivatives were used in this region to treat malaria, and not
one of 600 patients prospectively studied with ≥4% parasitaemia had died.
These examples resulted in immediate harm to either the patient or the public in regions
outside of the jurisdiction of the United States. These counterfeit or substandard samples are
either poison or ineffective and lead to harm, disease, or death. There are many more examples
throughout the world, but it is important to note that in many cases, instances are not reported, or
the details of the event are not reported well for various reasons. (Cockburn, Newton, Agyarko,
Akunyili, & White, 2005)
In the United States, the situation is usually identified and reported more quickly than in
other regions of the world. Recent events related to counterfeit versions of the drug Avastin® in
the United States were reported and mitigated almost immediately. Other examples of
substandard or counterfeit drugs in the United States market are usually responded to with
similar expediency. (Pills for birth control seized as counterfeit, 1984; Wax, 1995)
One significant challenge to FDA regulators, though, is in management of drugs in the
United States market that are the product of a global supply chain. The recent events related to
more stringent control in India are evidence of this fact. (Siddiqui & Chatterjee, 2014) In
addition, the inability to effectively manage this supply chain resulted in substandard versions of
the drug heparin infiltrating the highly regulated pharmaceutical market in the United States.
The consequences were fatal.
In January 2008, the United States Centers for Disease Control and Prevention began a
nationwide investigation of severe adverse reactions that were first detected in a single
hemodialysis facility. Preliminary findings suggested that heparin was a possible cause of the

reactions. (Blossom, et al., 2008) Ultimately, a total of 785 adverse-reaction reports including 81
deaths in the United States were associated with the contamination of heparin. Intensive
investigations, inspections, testing, and chemical analysis related to Baxter’s heparin identified a
previously unknown contaminant, oversulfated chondroitin sulfate. Oversulfated chondroitin
sulfate is a modified form of chondroitin sulfate and was detected in samples of heparin crude
materials, heparin API’s, and finished heparin drug products.
Chondroitin sulfate is a naturally occurring in animal cartilage and is often used as a
dietary supplement and in supplements to treat arthritic joints. Oversulfated chondroitin sulfate,
however, doesn’t occur naturally and was created by chemical modification of chondroitin
sulfate. The production of crude material for the production of heparin starts with the isolation of
heparin from pig intestines. Many of those pigs come from rural farms in China, with the
intestines initially processed at unregulated small workshops. (Labadie, 2012) During this crisis,
it was found that the United States Pharmacopeia (USP) testing monograph for unfractionated
heparin sodium did not detect the presence of oversulfated chondroitin sulfate in heparin. New
tests and specifications have since been developed by the FDA and the USP and put in place to
not only detect the contaminant oversulfated chondroitin sulfate, but also to improve assurance of
quality and purity of the drug product. (Keire, et al., 2011)
Long-Term Danger of Counterfeit and Other Substandard Drugs
Drug Resistance
Counterfeit drugs also present long-term challenges to public health. They allow for the
development of resistance to current medications. Frequently, this type of situation occurs in
regions of the world where reports indicate that as much as, if not more than, half of prescription
drugs are suspected to be counterfeits. It has been reported that counterfeit artesunate (artesunate

is one of a number of drugs used for the treatment of malaria) comprises between 33% and 53%
of the supply of this drug in mainland Southeast Asia. (Senior, 2008) Anti-infective medicines
containing sub-therapeutic amounts of API, whether counterfeit or substandard, increase the risk
of spreading resistance to the drug. (Shakoor, Raylor, & Behrens, 1997) The recent discovery of
fake artesunate containing small quantities of API (Fernández, et al., 2006; Newton, Green,
Fernández, Day, & White, 2006) raises the likelihood that these counterfeits will facilitate the
selection and spread of Plasmodium falciparum parasites resistant to artemisinin derivatives.
(Newton, et al., 2006)
Unfortunately, poor-quality drugs are rarely mentioned as important in the evolution of
drug resistance. This is in part due to the fact that differentiating the relative roles of widespread
unregulated use of inadequate anti-infectives from poor drug quality is very difficult in these
geographical regions. (Newton, Green, Fernández, Day, & White, 2006) According to World
Health Organization (WHO) data, malaria infected around 219 million people in 2010, killing
around 660,000 of them - the vast majority in sub-Saharan Africa. Robust figures are hard to
establish for a disease that affects mainly poor communities in rural areas of developing
countries, and some global health experts say the annual malaria death toll could be double that.
(Kelland, 2014)
Aside from malaria, other diseases are reported to develop resistance due to
counterfeiting. The high prevalence of substandard chloramphenicol and cotrimoxazole in
Burma could well have contributed to the high frequency of typhoid antibiotic resistance. (Shwe,
Nyein, & Yi, 2002) Poor quality rifampicin and pyrazinamide (Wondemagegnehu, 1999) are
also likely to be fueling the spread of multidrug-resistant mycobacterium tuberculosis. Loss of

these medicines would likely lead to therapeutic failure. The need for the development of new
anti-infectives is indicated, when so few new ones are being produced. (Trouiller, et al., 2001)
This observation is especially problematic when considering results of a recent study of
global warming and the occurrence of malaria at higher elevations. These authors reported that
future global warming could lead to a significant increase in malaria cases in densely populated
regions of Africa and South America unless disease monitoring and control efforts are increased.
(Siraj, et al., 2014) In a study of the mosquito-borne disease that infects around 220 million
people a year, researchers from Britain and the United States found what they describe as the first
hard evidence that malaria creeps to higher elevations during warmer years and back down to
lower altitudes when temperatures cool. (Kelland, 2014) This in turn "suggests that with
progressive global warming, malaria will creep up the mountains and spread to new high-altitude
areas." (Neuhauser, 2014) People who live in these areas have no protective immunity because
they are not used to being exposed to malaria, so they will be particularly vulnerable to more
severe and fatal cases of infection. (Kelland, 2014)
For their study, Siraj and colleagues analyzed data from Ethiopia and Colombia, looking
at malaria case records from the Antioquia region of western Colombia from 1990 to 2005 and
from the Debre Zeit area of central Ethiopia from 1993 to 2005. By excluding other factors that
influence malaria case numbers, such as mosquito-control programs, resistance to anti-malarial
drugs and fluctuations in rainfall, they found that the median altitude of malaria cases shifted to
higher levels in warmer years and back to lower levels in cooler years. The authors purport that a
clear, unambiguous signal that can only be explained by temperature changes. They state this is
indisputable evidence of a climate effect. The researchers noted that their work was limited to

two countries on two continents, and suggested it should be replicated in more countries with
malaria in highland regions before more general trends are assumed. (Siraj, et al., 2014)
It is clear that both counterfeit and substandard drugs have a negative impact on public
health and, therefore, are a problem for both the forensic scientist and other practitioners within
the criminal justice system.

CHAPTER 4. FORENSIC SCIENCE AND COUNTERFEIT DRUGS
The Role of the Forensic Scientist in the Counterfeit-Drug Problem
The threat posed by counterfeit drugs to public health and safety was established in the
previous chapters. These goods are manufactured and distributed by individuals and criminal
organizations that need to be identified and brought to justice. (United Nations Office on Drugs
and Crime, 2014) Executing this capability relies upon the practice of good forensic science.
Counterfeit samples need to be identified using methods of analysis that are admissible in court.
Classification of these samples provides useful information that may be used in an investigative
manner to establish provenance and identify counterfeiting groups or organizations. It is
important that this capability be performed in the field so that decisions about a sample’s
authenticity may be made without delay. Frequently and for various reasons, counterfeit drugs
are sold in foreign countries distant from their manufacture site. (Dean, 2013) If a Customs
agent were able to rapidly and reliably establish authenticity, the import of these goods to the
target nation could be denied before infiltration of these illicit goods into the local supply chain
occurs.
Literature Review with Regard to Analytical Testing of Counterfeit Goods
A review of the scientific literature shows that a method to routinely identify counterfeit
drugs has not been described. In addition, although attempts were successful in some specific
instances, a method for the routine analysis of counterfeit goods or drugs to determine
provenance is yet to be established. Finally, there is a lack of discussion in the literature with
regard to the analysis of counterfeit goods in the field. Virtually all of the literature describing
counterfeits references lab-based methods. Most are used to detect, identify and quantify the
API. From this information, provenance determinations are attempted. The ingenuity, especially
for counterfeit drugs, is in the details of the instrumentation or methods used for sample
introduction rather than in the approach used to address the counterfeit-drug problem.
A search of the term “counterfeit” in the electronic databases of the Journal of Forensic
Sciences and Forensic Science International for the years 1983 through March 2014 returned 76
and 78 articles, respectively. In more recent years, the frequency of articles has increased,
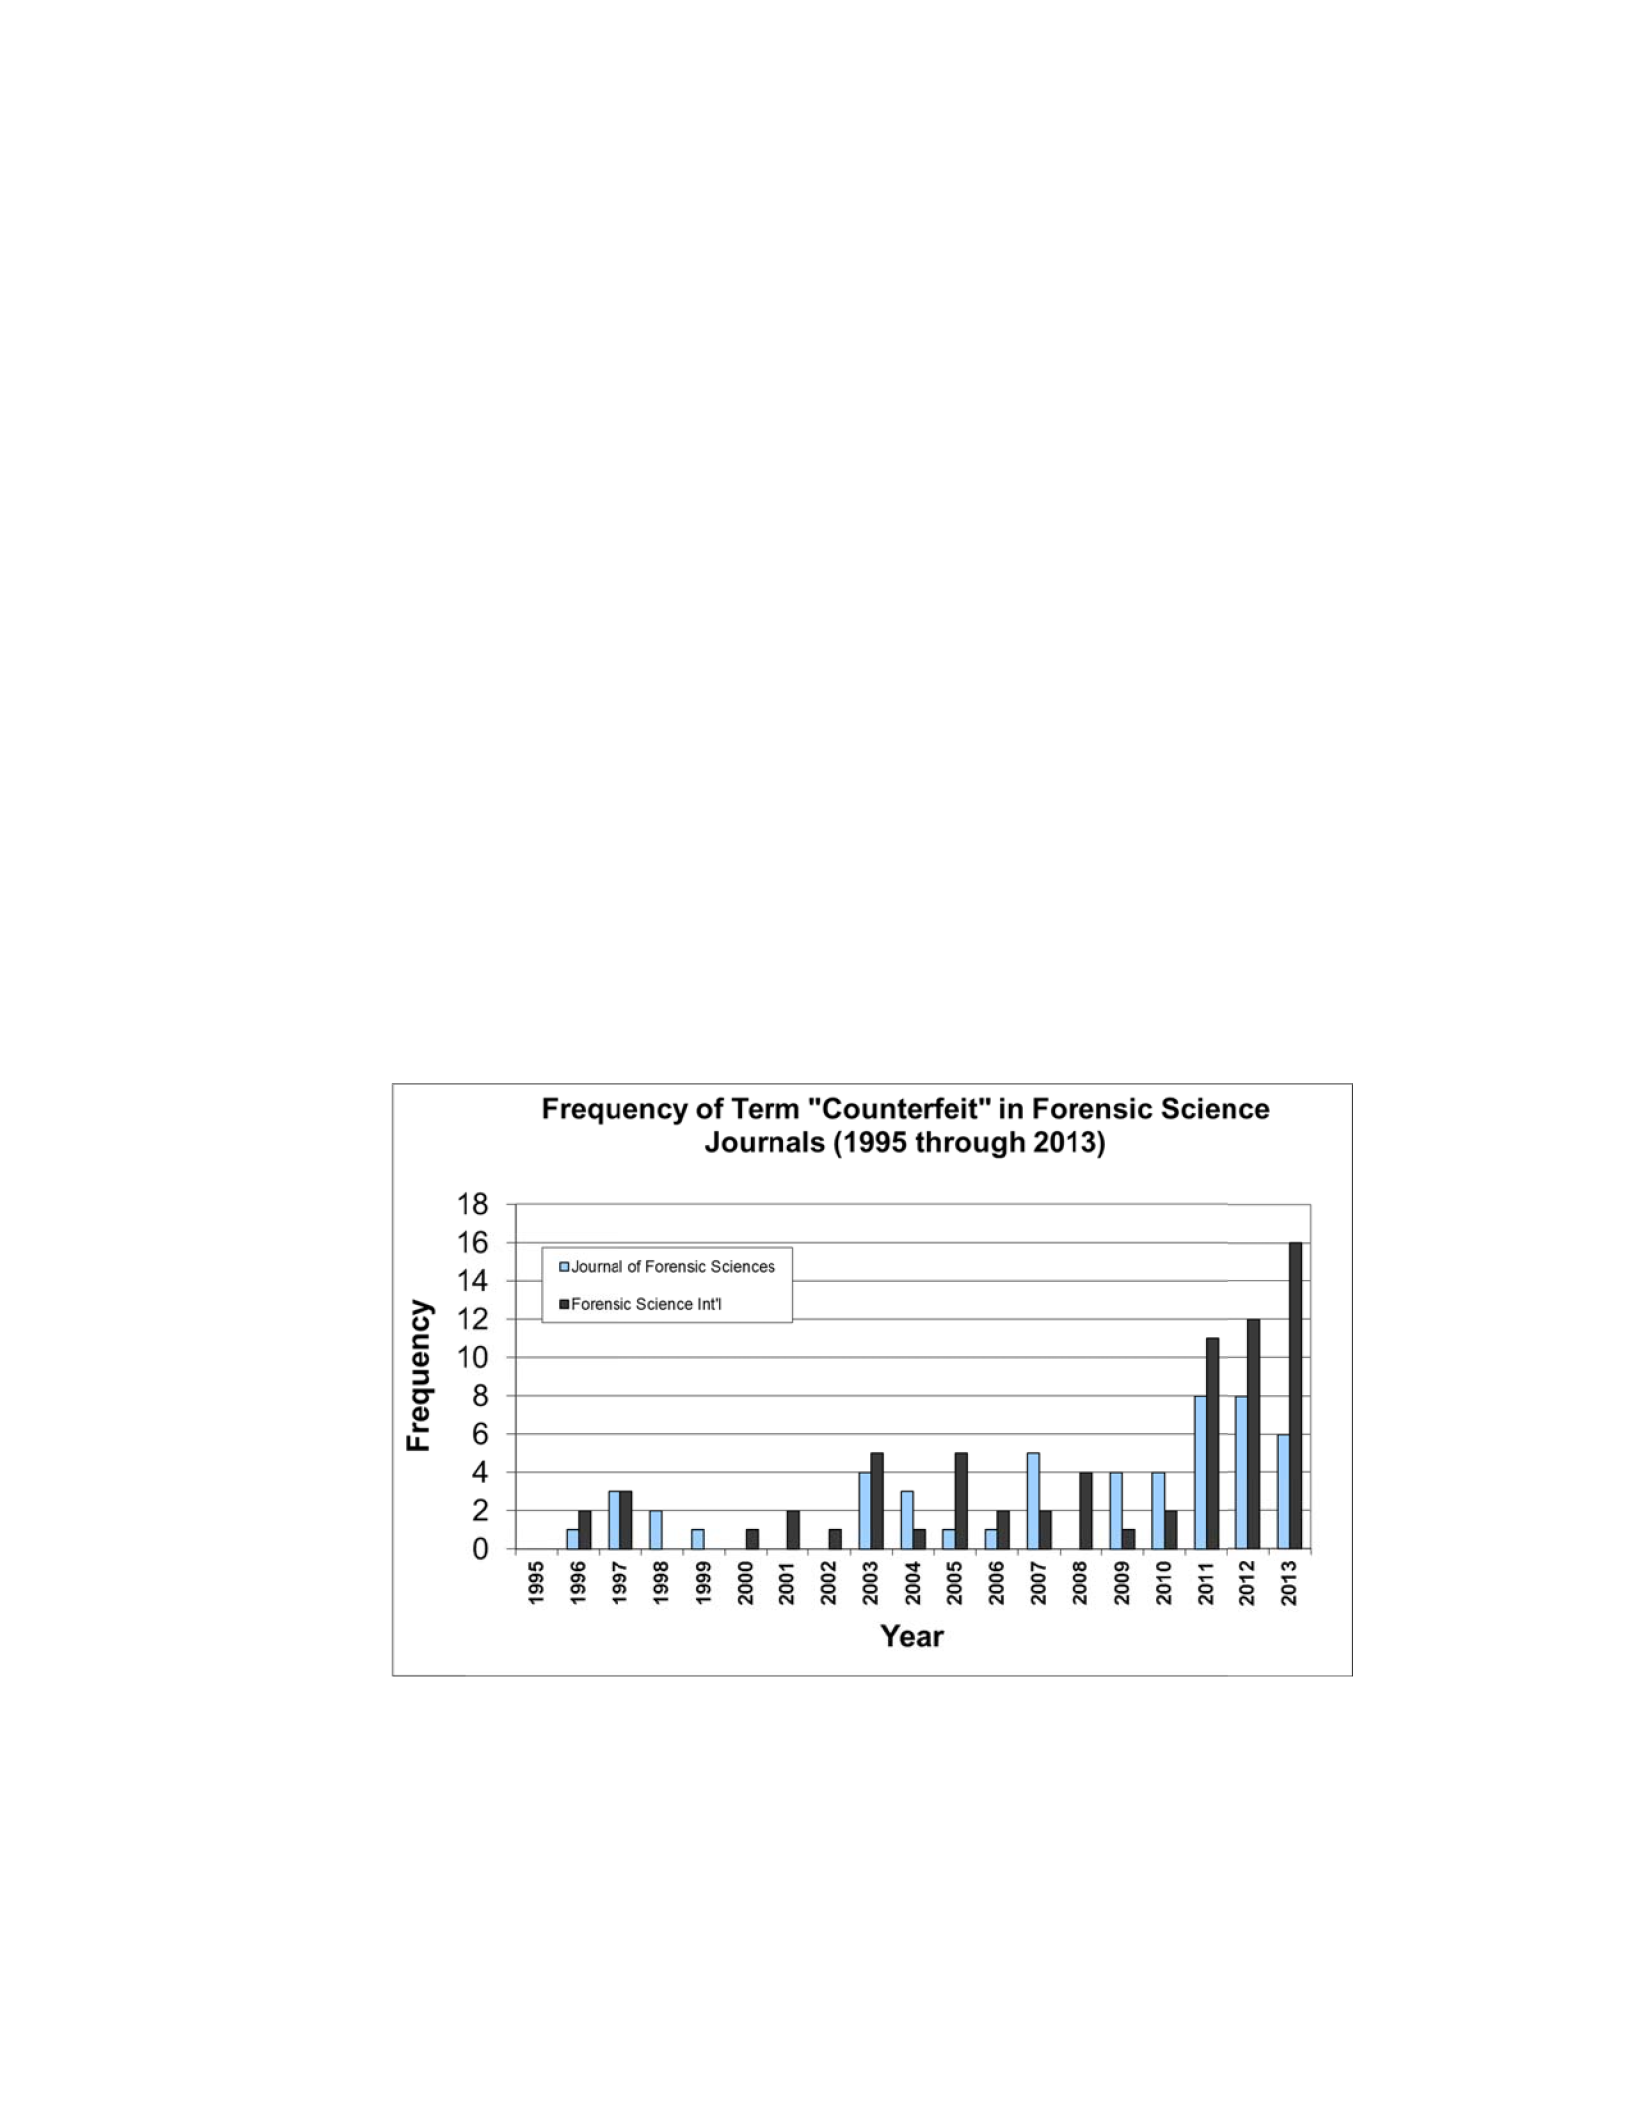
indicating this type of analysis is becoming more prevalent. Figure 3 shows the frequency of
occurrence of the term “counterfeit” between the years 1995 and 2013. This data shows an
increased interest by forensic scientists in the analysis of counterfeits as physical evidence in the
past few years. Interestingly, the type of evidence analyzed ranges from tobacco used to pass off
counterfeit cigarettes, to high-end designer drugs intended to circumvent both IP rights and
government regulation.
Figure 3. Frequency distribution of the term “counterfeit” in
forensic science journals.

Acampora, Ferranti, Malorni & Milone analyzed pigments in a counterfeit bank note
using direct-mixture-analysis mass spectrometry. Pigments identified in the note were the same
pigments contained within ink samples suspected to be used to produce the notes. The
conclusion drawn by the authors was that the bank note was consistent with being made using
inks from the suspect sample, but no attempts were made to individualize the inks to the bank
notes. (Acampora, Ferranti, Malorni, & Milone, 1991) Hida, Mitsui and Minami performed xray
fluorescence (xrf) and microscopy to analyze counterfeit coins. The counterfeit coins tested, just
like the authentic coins, were comprised primarily of copper and nickel. The amount of copper
and nickel present was higher in the counterfeit coins. In addition, the authors noted that
chromium, manganese, iron, nickel, and zinc profiles of the counterfeit coins were quantitatively
more variable than what was observed in the authentic coins. (Hida, Mitsui, & Minami, 1997)
Hida, Sato, Sugawara and Mitsui described analysis of counterfeit coins using xrf and x-ray
diffraction (xrd). Using statistical methods, these authors were able to cluster the coins into three
groups. One group included 250 counterfeit coins, the second group included 27 counterfeit
coins and the third group included the four authentic coins analyzed. (Hida, Sato, Sugawara, &
Mitsui, 2001) This same year, Hida and Mitsui described xrf testing of prepaid turnpike and
subway cards. They developed a multivariate statistical method to categorize 200 authentic
subway cards into at least four different groups. This same method was then applied to 20
counterfeit turnpike cards, as well as 12 authentic turnpike cards. Using this multivariate
statistical method, this group of 32 cards was divided into three groups. One was comprised of
the 20 counterfeit cards plus one authentic card; the other 11 authentic cards were divided into
two groups. (Hida & Mitsui, 2001) Bartle and Watling used laser ablation-inductively coupled
plasma-mass spectrometry (LA-ICP-MS) to analyze the isotope distribution profiles of porcelain.

From this work, they concluded that LA-ICP-MS can be used to identify and classify Oriental
porcelains and establish their origin of country. They furthered their conclusion by stating that
minor variations in the spectral fingerprints of porcelains from the same country can be attributed
to variations in production methods both over time and between individual potters. This
indicated to them a potential to further provenance Oriental porcelains to specific production
kilns. (Bartle & Watling, 2007) Sugawara used polarized infrared radiation to differentiate
counterfeit Japanese passports from authentic samples. When unpolarized radiation was used,
the authentic samples were indistinguishable from the counterfeits. When polarized radiation
was used, however, peak ratios were used for successful differentiation. (Sugawara, 2007)
Sugawara also successfully applied confocal-type laser microscopy to Japanese passports with
the goal of differentiating authentic samples from counterfeits. (Sugawara, Passport examination
by a confocal-type laser profile microscope, 2008) Suzuki used the acoustic characteristics of
500-yen coins to differentiate authentic coins from counterfeits. The author concluded that this
simple and rapid technique can not only be used to differentiate authentic from counterfeit coins,
but also that the method may be applied to the classification of counterfeit coins if databases of
coins were developed. (Suzuki, 2008) LaPorte and colleagues used microscopy to analyze the
printing defects in envelopes. These authors concluded that their study corroborates previous
assertions that assessments to determine whether two or more items may have a common origin
can be based upon the evaluation of printing defects. (LaPorte, Stephens, & Beuchel, 2010)
Pérez-Bernal and colleagues performed principal component analysis (PCA) on metallic
distribution data from the ash of tobacco from different brands and types of cigarettes. This data
indicates it may be possible to use this type of analysis to assess the brand of tobacco used for
manufacture of cigarettes. These authors state that a limitation of their study is in the number of

different types of tobacco analyzed, as well as in the number of different brands of cigarettes
analyzed. (Pérez-Bernal, Amigo, Fernández-Torres, Bello, & Callejón-Mochón, 2011) Romão et
al. used electrospray ambient ionization-mass spectrometry (EASI-MS) and showed this
technique may be successfully applied as a fast and non-destructive screening tool for
documents, specifically for Brazilian vehicle registration. (Romão, et al., 2012) Brzezinksi and
Craft summarized analysis performed on seized samples of counterfeit toothpaste. These
samples were analyzed to determine the number of microorganisms present. Although the
authors showed the samples contained high numbers of microorganisms, they were not able to
assign all samples to either a manufacturing site or method of manufacture. (Brzezinski & Craft,
2012) Cao, Gao, Fan, and Yan presented an automatic and efficient detection algorithm for copy-
move forgery for digital counterfeits. (Cao, Gao, Fan, & Yang, 2012) Nam and colleagues used
liquid chromatography-mass spectrometry-mass spectrometry (LC-MS-MS) to detect
corticosteroids in cosmetic creams. The method allowed for the analysis of large numbers of
samples in a short period of time. As a result of this work, illegal steroids were detected in four
different cosmetic products and resulted in these products being withdrawn from local markets.
(Nam, Keun, Lee, & Lee, 2012) Božičević and colleagues used micro Raman spectroscopy to
identify a common origin of toner-printed counterfeit banknotes. Their results show that
microRaman spectroscopy can be successfully applied as a method for the analysis of color toner
printed counterfeits, such as banknotes and documents, in some instances. (Božičević, Gajović,
& Zjakić, 2012). Baechler and colleagues applied chemometric methods to data from visual
markers to classify false identity documents. Their method generated low error rates and was
concluded to be able to link documents to a common source or to differentiate them. (Baechler, et
al., 2013)

With regard to the analysis of counterfeit drugs, most of the literature relevant to the field
of forensic science describes methods and techniques for the identification and/or quantitative
analysis of the API and other ingredients present in the counterfeit drug. Alabdalla discussed the
analysis of counterfeit samples of Captagon®, a drug product containing the API fenethylline,
using GC-MS. This drug is commonly abused in the country of Jordan and throughout the
Middle East. The author concluded that the comparison of chromatographic patterns is useful in
establishing links between samples under investigation. In addition, the author concluded that
profound similarities of chromatographic profiles support the assumption that samples could
have a common origin, or at least have been produced by the same manufacturer. (Alabdalla,
2005) Baer, Gurny, and Margot used near-infrared (NIR) analysis to evaluate cellulose and
lactose. Although they state it may be possible to use this type of analysis to source ecstasy
tablets, their data is preliminary and the intra- and inter-variability of compression batches using
this method is unknown. (Baer, Gurny, & Margot, 2007) Soltaninejad and colleagues discussed
the analysis of counterfeit buprenorphine using GC-MS and liquid chromatography (LC) with a
diode array detector. These authors showed that the counterfeited samples analyzed were derived
from illicit heroin. Due to the therapeutic and metabolic differences between buprenorphine and
heroin derivatives, the authors concluded that the use of counterfeit buprenorphine leads opiate
abusers to serious health risks. (Soltaninejad, Faryadi, Akhgari, & Bahmanabadi, 2007) Venhuis,
Blok-Tip and de Kaste used highperformance liquid chromatography (HPLC) with a diode-array
detector fitted with a mass spectrometer (MS) detector to analyze three different types of seized
natural aphrodisiacs. All three types of samples contained analogs of prescribed erectile
dysfunction drugs. Although there was nothing unusual about the analysis, this study is
interesting because these drugs were likely designed to circumvent patents by using unpatented

analogs. In addition, they are being used as natural supplements and, therefore, were also likely
designed to circumvent regulatory requirements. (Venhuis, Blok-Tip, & de Kaste, 2008) Been et
al. used NIR and Raman to propose a strategy for the classification of counterfeit medicine.
(Been, Roggo, Degardin, Esseiva, & Margot, 2011) Kauppila and colleagues purport the use of
desorption atmospheric pressure photoionization-mass spectrometry (DAPPI-MS) as a screening
tool for drug samples. A primary advantage of the DAPPI-MS technique compared with GC-MS
is speed. Their research showed the DAPPI-MS technique could be used to identify controlled
substances within drug samples within seconds. (Kauppila, et al., 2011) Lopatka and Vallat
evaluated surface granularity of tablets to differentiate different samples from each other. They
purport that this type of analysis contains valuable information with regard to production batch
identity. (Lopatka
& Vallat, 2011) Samms et al. used direct-analysis-in-real-time – time-of-flight – mass
spectrometry (DART-TOF-MS) to analyze street samples of Xanax®. The authors propose this
as a new method to identify API’s. (Samms, Jiang, Dixon, Houck, & Mozayani, 2011) Jung et al.
propose a method for quick and reliable identification of counterfeit VIAGRA® and Cialis®
using image processing and statistical analysis. Although this method seems useful, one of the
concerns about the technique is that there is potential for a large number of false positives if
tablet discoloration occurs. (Jung, Ortiz, Limberger, & Mayorga, 2012) Logan and colleagues
used a variety of techniques including thin-layer chromatography, GC-MS, HPLC, and liquid
chromatography – time-of-flight – mass spectrometry (LC-TOF-MS) to characterize a variety of
different types of samples including counterfeits. It is important to note that no counterfeit
samples analyzed as part of this study contained any active ingredient. (Logan, Reinhold, Xu, &
Diamond, 2012) Park and Ayn summarized quantitative analysis of sildenafil and tadalafil using

high-performance liquid chromatography-ultraviolet spectroscopy (HPLC-UV) analysis
performed on counterfeit drugs seized in Korea. One hundred five samples were analyzed.
More than half contained more than the prescription dose of sildenafil and tadalafil. (Park &
Ahn, 2012) Patterson et al. developed and described an electrospray ionization-liquid
chromatography-mass spectrometry (ESI-LC-MS) analytical method used to differentiate
sildenafil from its vardenafil analog. This work is valuable because these compounds are
difficult to differentiate using more common MS methods due to similarity in the fragmentation
patterns of the two compounds when analyzed using a quadrupole MS. (Patterson, Mabe,
Mitchell, & Cory, 2012) Ortiz et al. used attenuated total reflection (ATR) Fourier-transform
infrared spectroscopy (FT-IR) with PCA analysis to classify seized samples of Cialis and
VIAGRA. The PCA applied to ATR-FTIR data allowed grouping samples according to their
different chemical profiles, distinguishing successfully between authentic and counterfeits
samples. In addition, PCA scores inserted counterfeit drugs from different seizures in the same
cluster, suggesting a common illicit source for these medicines. (Ortiz, et al., 2013) Mariotti and
colleagues evaluated the trends in counterfeit amphetamine-type simulants after prohibition of
these goods in Brazil. These authors concluded that visual inspection and the physical
characteristics like average mass were useful in indicating forgery. However, they stated that this
analysis is limited and alone these measurements were not reliable to distinguish between
authentic and counterfeit samples. For some samples they recommended additional analysis
using GC-MS. For others they recommended other methods including PCA of ATR-FT-IR data.
(de Cássia Mariotti, Ortiz, Sousa, Fröehlich, & Limberger, 2013) Ortiz and colleagues used
chemometrics applied to UPLC-MS to analyze counterfeit samples of drugs used to treat erectile
dysfunction. (Ortiz, et al., 2013) Anzanello et al. proposed a framework for selecting the

analytical techniques providing the most conclusive data for categorizing seized drugs into
authentic and unauthentic classes. With regard to VIAGRA and Cialis, these authors proposed
the use of data from UPLC–MS, physical dimensions, and ATR FT-IR. (Anzanello, Ortiz,
Limberger, & Mariotti, 2014)
This literature review shows that there is a need for an analytical scheme to rapidly and
reliably identify counterfeit drugs and establish provenance using methods of analysis that work
well. Infrared, Raman, and gas chromatograpy-mass spectrometry (GC-MS) are methods that
are routinely used to physically and chemically characterize drugs and drug products.
Investigation of these methods for this use is proposed. Ideally, this scheme would be applied
using field-portable instruments in order to rapidly and reliably identify these goods at their port
of entry. The scope of this dissertation includes an attempt to establish such an analytical scheme
using field-portable instrumentation. Ultimately, the data collected in the field should be used to
create a database of counterfeit drugs analyzed in order to provide forensic scientists the data
they need to be able to effectively establish sample provenance.
CHAPTER 5. TESTING IN THE FIELD: PORTABLE ANALYTICAL
INSTRUMENTATION
Design, Performance, and Implementation of Field-Portable Instruments
It seems appropriate to start this chapter by describing the current state of field-portable
instrumentation. It is important that detection and identification of counterfeit drugs is
performed in the field using methods of analysis that are rapid and reliable. At Customs check
points, the ability to detect and identify these goods quickly and accurately will prevent them
from entering the local supply chain. It is not always practical or possible to delay entry of cargo

to wait on results of lab testing. In fact, border agents typically will only have a few minutes to
make a go/no-go decision at checkpoints under conditions of high stress. (Pomfret, 2006) If
detection and identification can occur in the field, law-enforcement organizations may take
immediate action to prevent the infiltration of these goods into the market. It may also help to
identify counterfeiters, establish provenance, and appropriately adjudicate these cases.
Significant developments have resulted in the availability of smaller, lighter, and faster
instruments. (Carrabba, Spencer, Rich, & Rauh, 1990; Lammert, Rockwood, Wang, & Lee, 2006;
Overton, 2010; Reffner & Martoglio, 1995) In many cases, technology advances have led to the
availability of handheld measuring devices and spectrometers such as infrared and Raman
systems. (Smiths Detection, 2014; Wasatch Photonics, 2014) In the case of GC-MS, portable
instruments are available that weigh less than 35 pounds. (Smiths Detection, 2014) These
handheld and portable instruments are smaller and lighter, but yet generate data that is
qualitatively comparable to data from laboratory instruments. There are, however, differences in
design, performance criteria and implementation between portable and lab-based instruments.
With regard to design, field portable instruments need to be small, light, rugged, and easy
to use. These systems may be hand carried to the site where analysis will be performed. Large
and heavy instruments are not practical or acceptable for this application. They are expected to
experience significant wear and tear during transit and at the analysis scene. Frequently,
specifications for drop testing and other ruggedization criteria are as critical as the specifications
for the signal-to-noise ratio of the spectrometer. Due to the sometimes-precarious environments
in which these instruments are expected to perform, they must have an easy-to-use software
interface which generates results-driven answers rather than spectral displays requiring
significant user interpretation. Automatic library searching using algorithms tailored to optimize

results in the field are a significant aspect of these instrument designs. In addition, sampling and
other accessories are sometimes required, but are not desirable. The more complex a
fieldportable instrument becomes, the less likely it will become a part of the routine on-scene
work flow. There is also a need to minimize impractical consumables. The more gear that needs
to be carried to make an instrument perform, the less likely the instrument will be successful in
the field.
Performance criteria of field-portable instrumentation, as previously stated, typically
include specifications for analytical performance as well as for size, weight, and ruggedization.
Specifications for analytical performance such as signal-to-noise ratio, sensitivity, and levels of
detection and quantitation will be less stringent than they are for lab-based instrumentation of
equivalent technologies. However, instruments will be specified to perform under much more
extreme environmental conditions than lab-based equivalents. For instance, specifications may
be included for performance in cold and hot temperatures, in rainy environments, or even
desertlike conditions. Field-portable instruments are expected to be drop resistant and
specifications allow for post-analysis decontamination. This is a significant factor for
instruments intended to be used to detect chemical warfare agents and other toxic chemicals that
may be harmful to future users of the instrument.
Ease of implementation of field-portable instruments is also a significant consideration.
Training requirements should be minimal. Frequently, the goal is to have non-scientists use
field-portable instruments. This translates to a need to minimize modes of operation, simplify
user interfaces, and generate results-driven answers. Electronic libraries and the search
algorithms used to search these databases of spectra are a critical factor when considering the
potential success of a field-portable instrument. This is because users in the field do not usually

have the time available at the scene, or the practical experiences required to process and interpret
complicated data files. It is common for field-portable instruments to use library-search
algorithms designed to identify mixtures rather than use an algorithm like correlation coefficient
which assumes pure samples. The ability of the user to separate a mixture sample in the field
prior to analysis are sometimes limited, so designing the library-search algorithms to compensate
for this issue during data processing adds value to these systems. In many ways, field portable
instruments are only as useful out in the field as the success of their library-search algorithms,
and the size and quality of the library searched.
Technologies Evaluated in this Study
As part of this research, analysis in the field was performed using portable infrared,
Raman and GC-MS spectrometers.
Attenuated Total Reflection Infrared Spectroscopy
Most field-portable mid-infrared systems that are used for rapid analysis in the field
perform ATR measurements. This method of infrared analysis is useful in field applications
because it requires little-to-no sample preparation. In addition, ATR spectra are very
reproducible, which is also useful for field applications.
Attenuated total reflection occurs when a sample is brought into contact with an
internalreflection element (IRE) that has a higher refractive index than the sample. The IRE is
also referred to as an ATR crystal. Typical IREs are ZnSe, diamond, and germanium. ZnSe and
diamond have refractive indices of 2.4; germanium has a refractive index of 4.0. If radiation is
brought through the IRE at an angle greater than the critical angle, the beam will be totally
internally reflected.
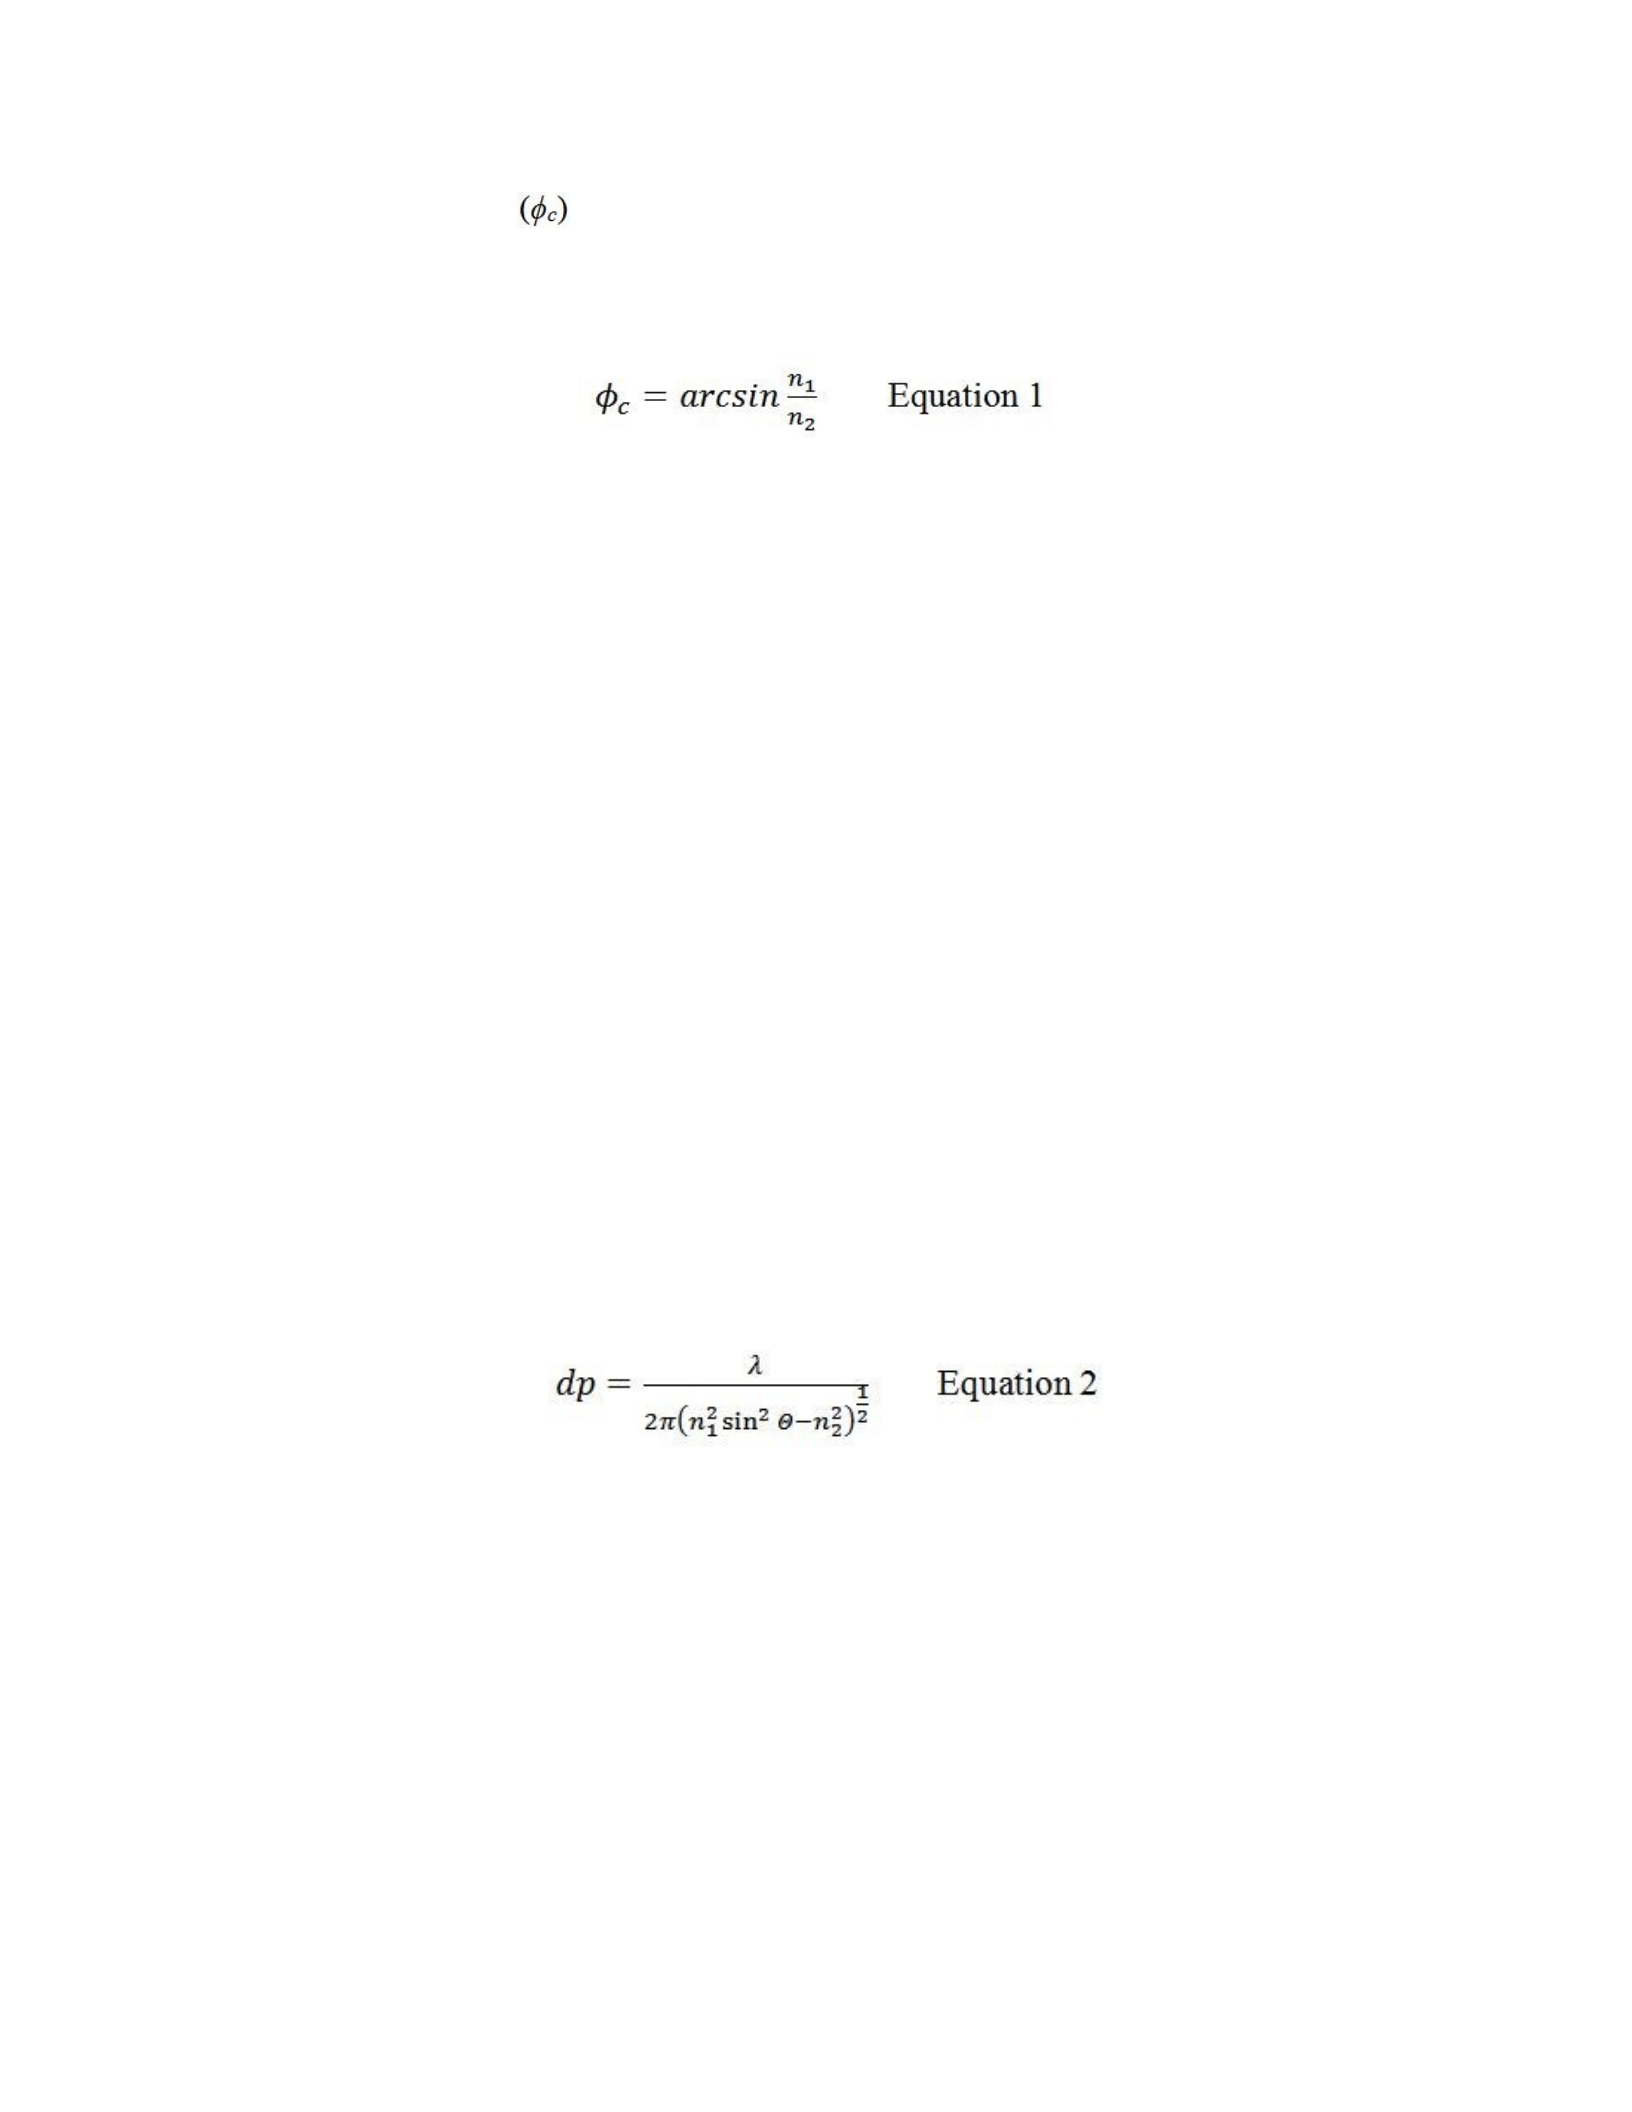
The critical angle between two media with different indices of refraction (n1 and n2)
is defined by the angle whose sin is n1/n2:
For ATR spectroscopy, n1<n2. A critical angle exists only for radiation traveling through a
higher-index medium into a lower-index medium. At angles greater than the critical angle, the
incident radiation is completely reflected, but there is an electromagnetic field that extends
beyond the IRE. This field’s strength decreases as the distance from the IRE surface increases;
it is referred to as an evanescent wave. It follows that if an absorbing material is brought into
contact with the IRE, the evanescent wave will be absorbed at wavelengths where the material
has an absorption band, and the amount of energy reflected back through the IRE will be
attenuated.
The distance the evanescent wave extends past the crystal surface and into any sample in
contact with the surface can be defined in terms of the depth of penetration. The depth of
penetration (dp) is defined as
where λ is the wavelength of radiant energy, n1 is the index of refraction of the IRE, n2 the index
of refraction of the sample, and Θ the angle of incident radiation on the interface. This depth of
penetration may essentially be considered the sampling depth. (Reffner & Martoglio, 1995)
Assuming contact between the IRE and the sample is made, the sampling depth is consistent
from measurement to measurement. For field analysis, this is very useful because it makes it
easier to collect reproducible spectra which can be easily compared with library spectra for
comparison and identification. It is important to recognize, though, that transmission spectra are
not wavelength dependent. Therefore, comparisons between ATR and transmission spectra
should allow for expected differences.
As part of this study, tablet cores were analyzed using a field-portable FT-IR ATR
spectrometer with a diamond IRE. In the system used, the diamond sampling area is 1.3 mm x
0.8 mm. Less than 1 mg of sample was pressed against the diamond for analysis. The optical
engine is designed with a resistively heated wire source, a fiber optic laser diode reference,
cubecorner reflectors, and a thermo-electrically cooled deuterated L-alanine doped triglycine
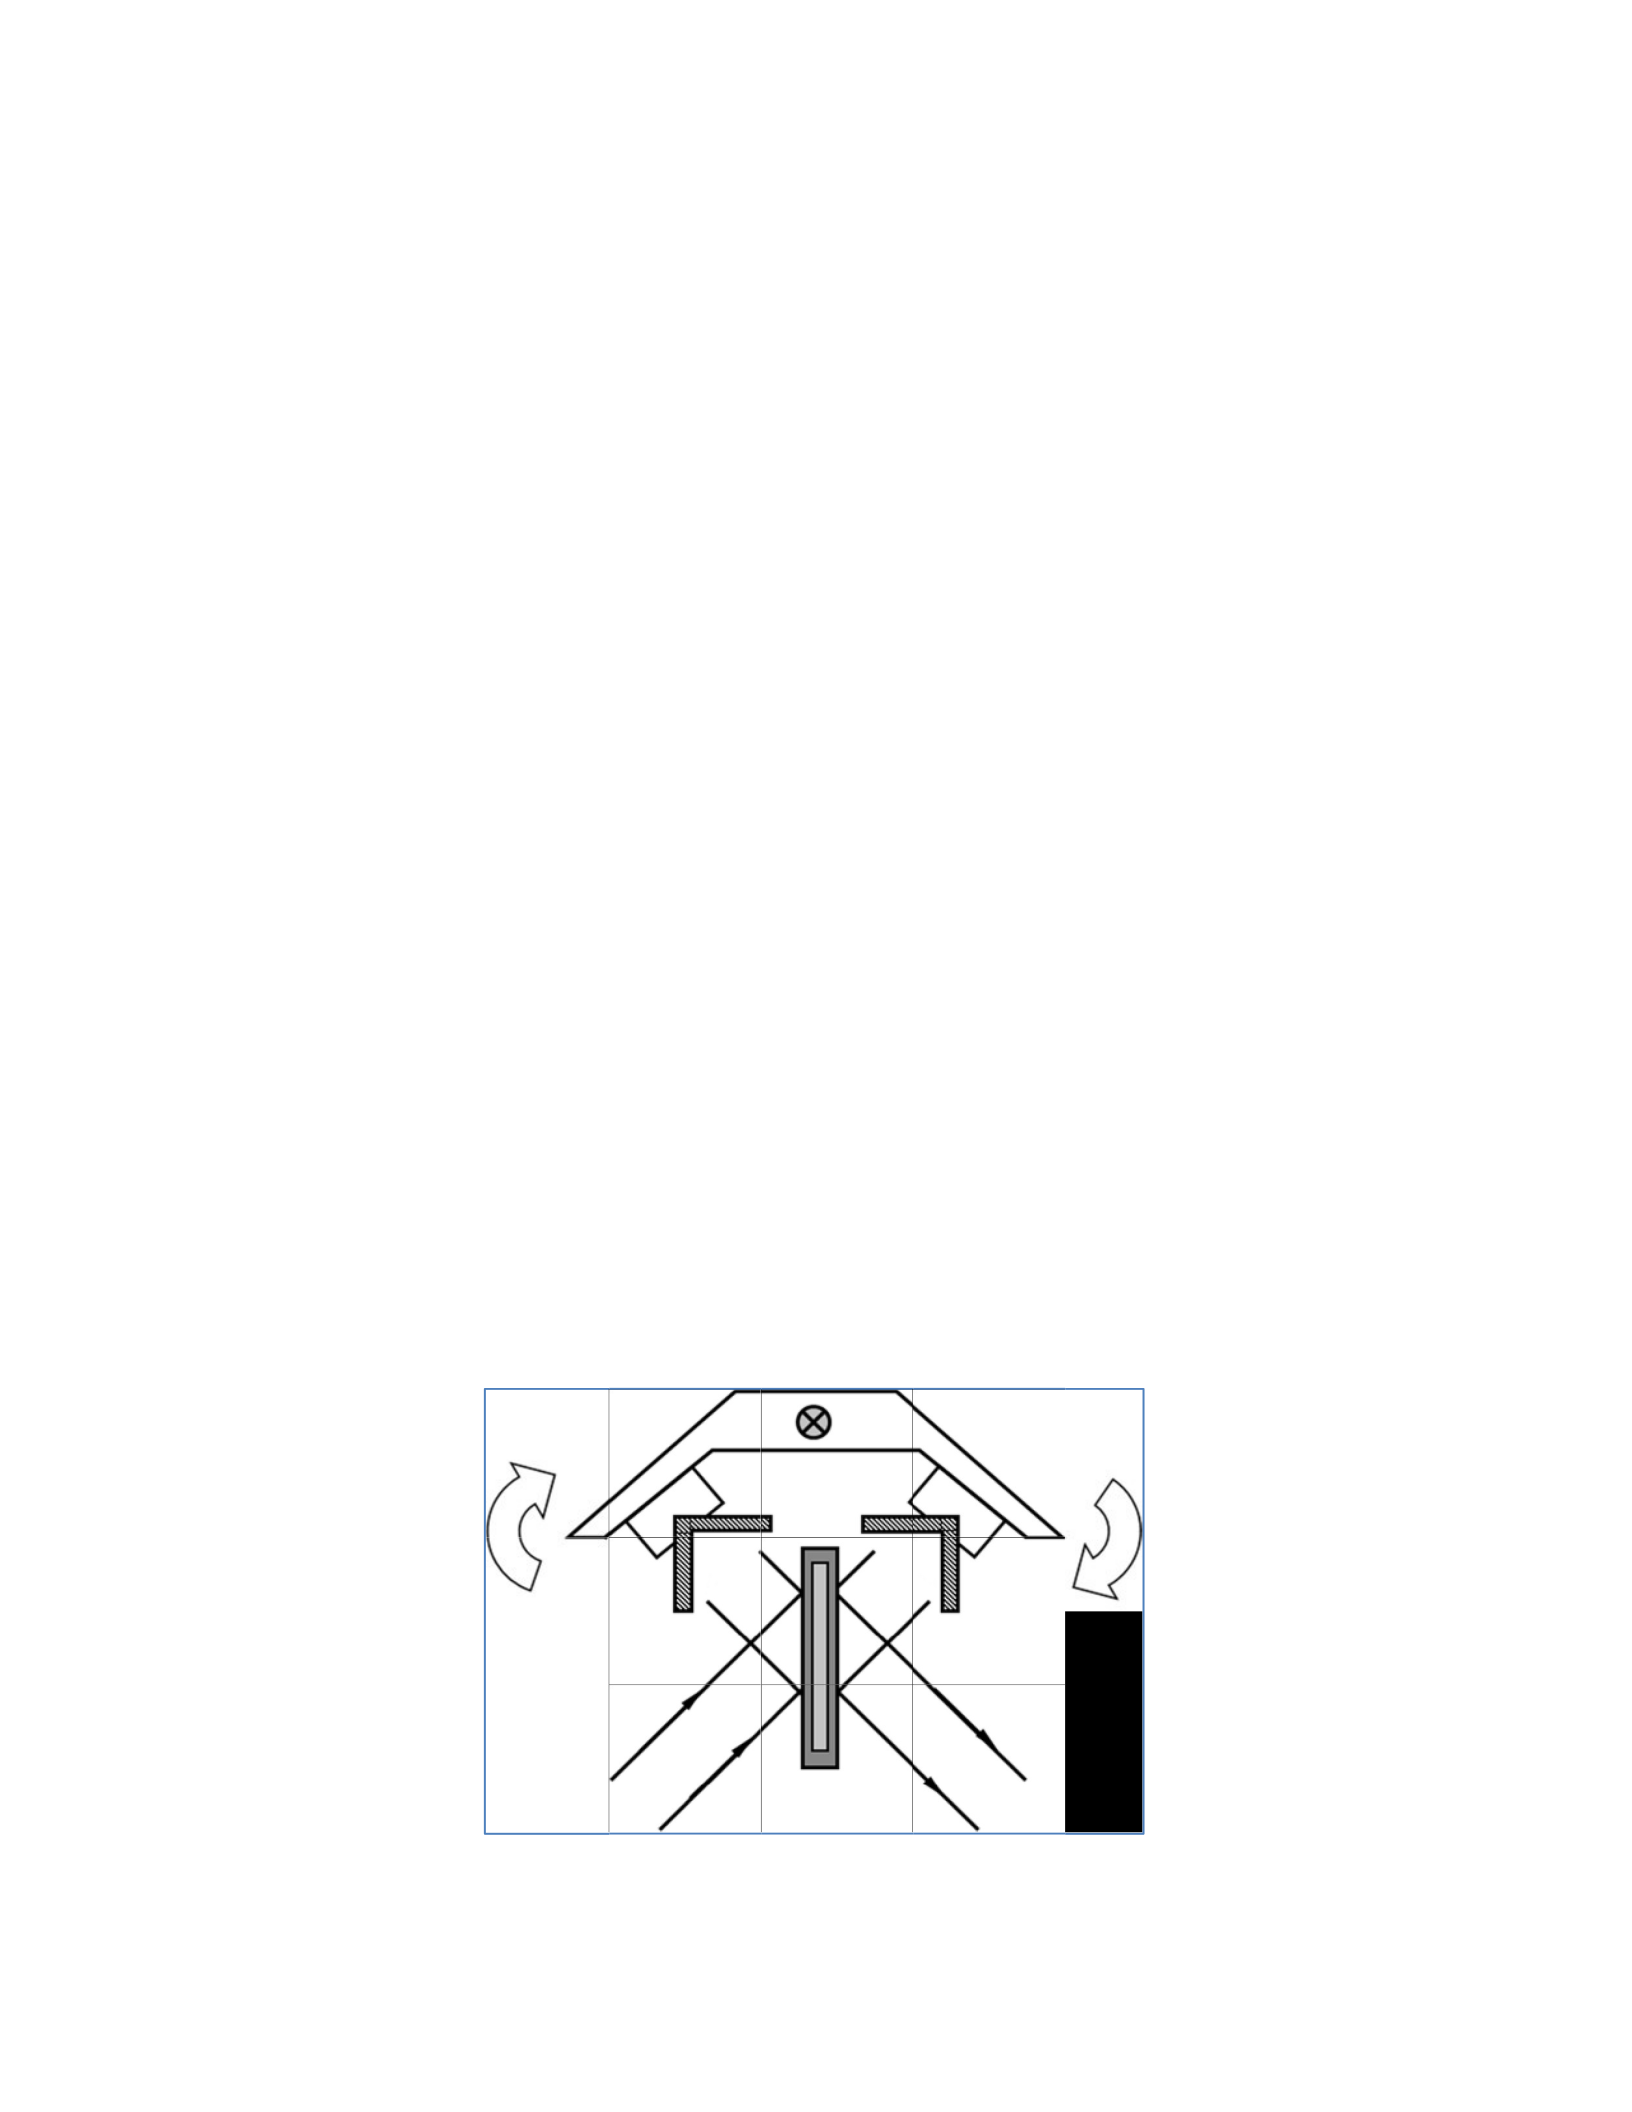
sulfate (DLaTGS) detector. Cube-corner reflectors are mounted on two arms of a double
pendulum modulator driven by an electromagnetic actuator. Interferograms are collected by
sweeping the reflectors over the required optical path difference. A schematic of this
interferometer is shown in figure 4. Both reflectors move synchronously so each reflector travels
half the linear distance required in a traditional Michelson interferometer. This optical engine is
advantageous in fieldportable systems because of its tilt compensation, reduced motion, inherent
self-alignment and disturbance rejection. (Arnó, et al., 2013) Although the system includes
automatic librarysearching capabilities, no library-searching functions were used in this study.

Figure 4. Schematic of corner-cube interferometer. Photo
courtesty of Mike Frunzi and Dustin Levy, Smiths Detection,
Danbury, CT.
Raman Spectroscopy
The Raman spectrometer used for these experiments uses a volume phase holographic
(VPH) grating to optimize signal. The use of holographic filters in Raman spectrometers is one
of the primary reasons Raman spectrometers were capable of becoming field portable.
The first reported use of holographic in Raman spectrometers was by Carrabba et al. in
1990. The filter was used for Rayleigh-line rejection in Raman spectroscopy. (Carrabba,
Spencer, Rich, & Rauh, 1990) The use of these filters was a significant advance because the
ability to reject the elastic scattering meant that any size monochromator could be used to collect
Raman spectra. Depending on the type of filter produced, and how it is mounted, lines as close
as 30 cm-1 from the laser line can be observed. (Adar, Delhaye, & DaSilvia, 2003).
When a beam of electromagnetic radiation impinges upon on a particle that is small with
respect to the wavelength of the radiation, the electrons of the particle are in an intense,
alternating field caused by the electric and magnetic components of the radiation. During this
interaction, the electrons of the particle oscillate with the same frequency of the incident
radiation and thereby produce electromagnetic radiation of the same frequency as the incident
radiation, but emanating from the particle in all directions. This appears to be scattered radiation
and is referred to as Rayleigh scatter. If, however, the polarizability of the particle, usually a
molecule, changes rather than remains constant, then the intensity of the scattered radiation
varies accordingly. Polarizability is related to the ease of separation of charges in an external
electrical field. If one or more normal modes of vibration of a molecule involve changes in the
polarizability, then the scattered radiation contains this vibrational frequency superimposed upon
the frequency of the incident radiation. This is the Raman effect. For a vibration to be active in
the Raman effect, the polarizability of the molecule must change during the vibration. Most
collisions of the incident photons with the sample molecules are elastic, or Rayleigh. Raman
scatter is significantly weaker than Rayleigh scattering. (Willard, Merritt Jr., Dean, & Settle Jr.,
1988) The use of lasers to generate sufficient Raman signal, coupled with holographic filters to
reject Rayleigh line scatter enabled significant advances in Raman instrumentation and the field
portability of these instruments.
Classical electromagnetic theory predicts the Raman effect, although a quantum
mechanical treatment is needed for detailed explanation. According to classical theory, the
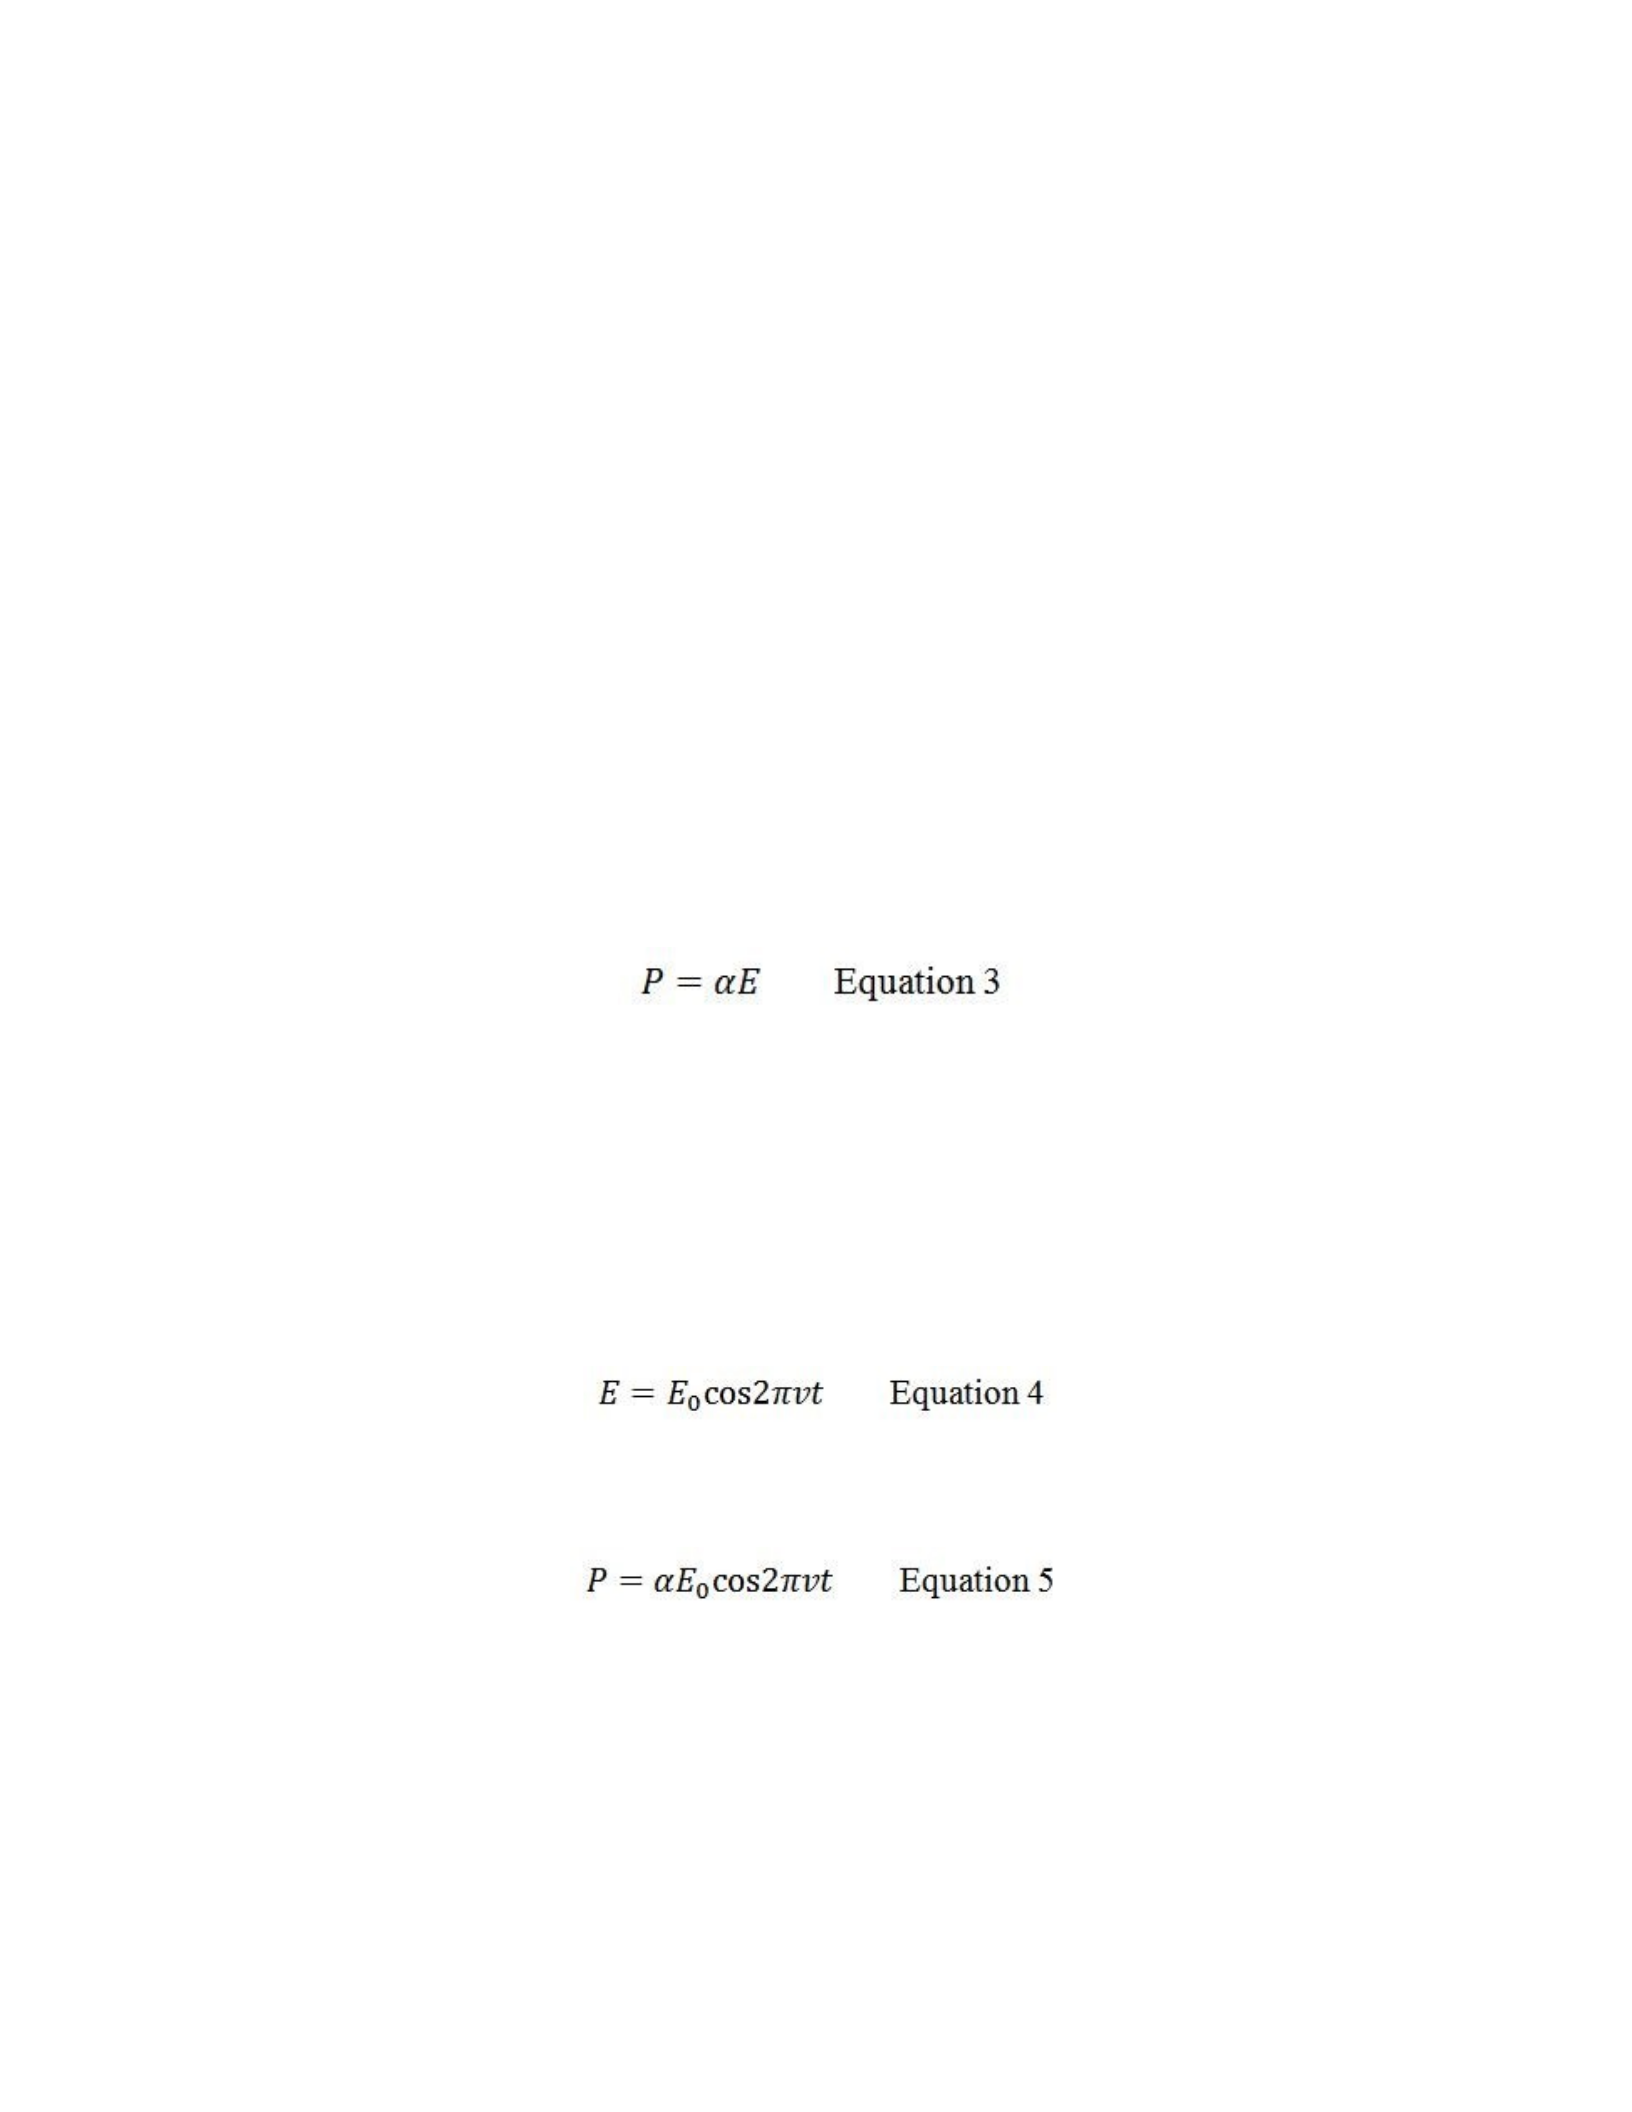
polarization, P, expressed as dipole moment per unit volume, is given equation 3
where E is the magnitude of the electric vector of the electromagnetic field that acts on the
molecule and α, the polarizability, is the proportionality constant. Since the magnitude of the
electric vector of the electromagnetic field varies with time, t, in a sinusoidal manner, E is
calculated in accordance with equation 4
and the polarization becomes
The polarizability consists of two parts: α0, the polarizability when the atoms of a molecule are
in their equilibrium positions, and a second term that is the sum of the polarizabilities of the
molecule due to the various rotational and vibrational motions. Each term of this second part
varies with the frequency associated with the particular rotation or vibration. Thus,
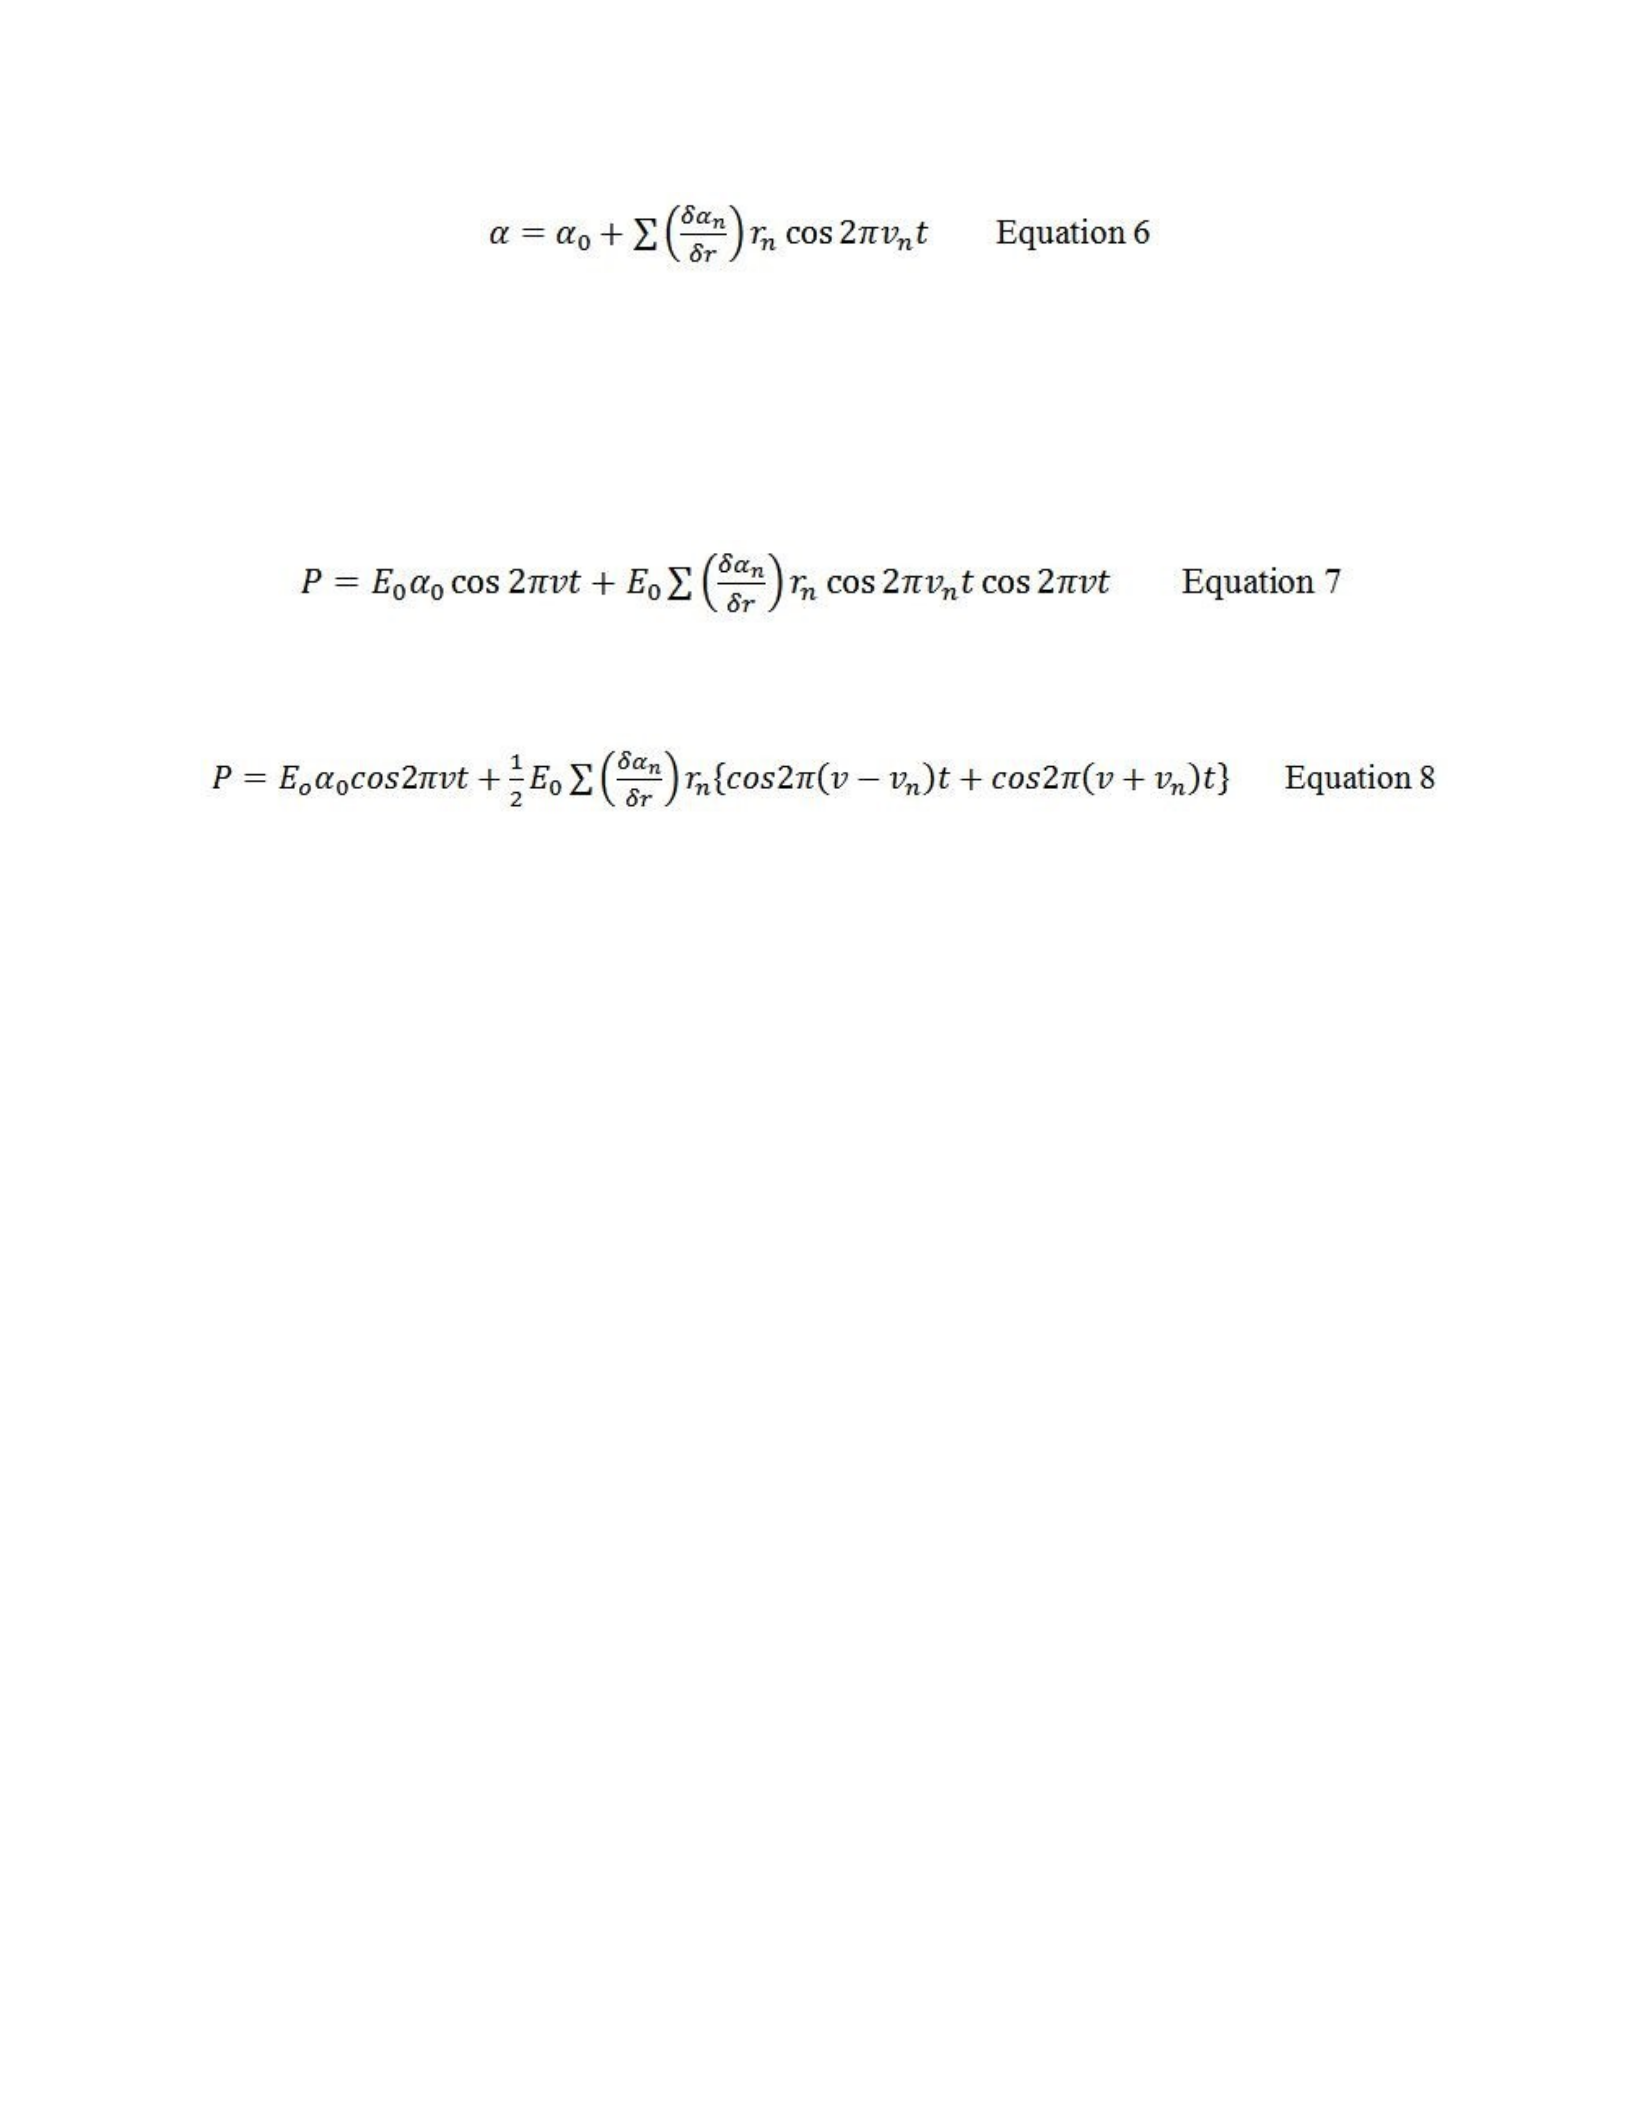
where αn is the polarizability associated with the nth rotational or vibrational mode and the rn is
the maximum displacement of the involved atoms. Combining equations 5 and 6, the
polarization becomes
which may be expressed
The first term has the frequency of the incident radiation and is the Rayleigh scattering. The
second term is the basis for the Raman effect and represents both the Stokes (v - vn) and
antiStokes (v + vn) Raman bands. As previously mentioned, most collisions of the incident
photons with the sample molecules result in Rayleigh scatter. Only a small portion of the excited
molecules (10-6 or less) may undergo changes in polarizability during one or more of the normal
vibration modes, i.e., the Raman effect. Therefore, holographic filters and their ability to reject
Rayleigh-line scatter are a significant advancement.
For their experiment, Carrabba et al. used holographic Bragg diffraction (HBD) filters
designed for laser-line rejection at 514.5 nm. The technique for fabricating the HBD filters is
based upon recording of a hologram in a dichromate gelatin/polymer graft emulsion
approximately 20-μm thick between transparent plates. During the recording process, a
standing-wave pattern generates as set of interference fringes which are recorded in the
holographic emulsion as successive planes of high and low refractive index. The planes are
separated by λ/2, where λ is the wavelength of the recording light in the medium. When used as
a filter, only those wavelengths fulfilling the Bragg condition will be diffracted, while other
wavelengths will be transmitted. In this way, the filter diffracts at the same wavelength at which
it was recorded, if recording is carried out at normal incidence. The holographic images exhibit
nearly ideal physical properties. (Carrabba, Spencer, Rich, & Rauh, 1990)
The Raman spectrometer used in this research employs a built-in 785-nm laser with f#/1.3
numerical aperture (NA) and volume phase holographic (VPH) grating. The detector used was a
back-thinned silicon array CCD with low-noise electronics. There are some advantages to this
portable system. The built-in laser with f#/1.3 NA enables high throughput with minimal loss of
efficiency. The VPH grating allows for high diffraction efficiency, minimizes polarization
effects, and is optimized over a broad range of wavelengths (although there may be a tradeoff
between maximizing the bandwidth and maximizing the peak efficiency). (Baldry, Bland-
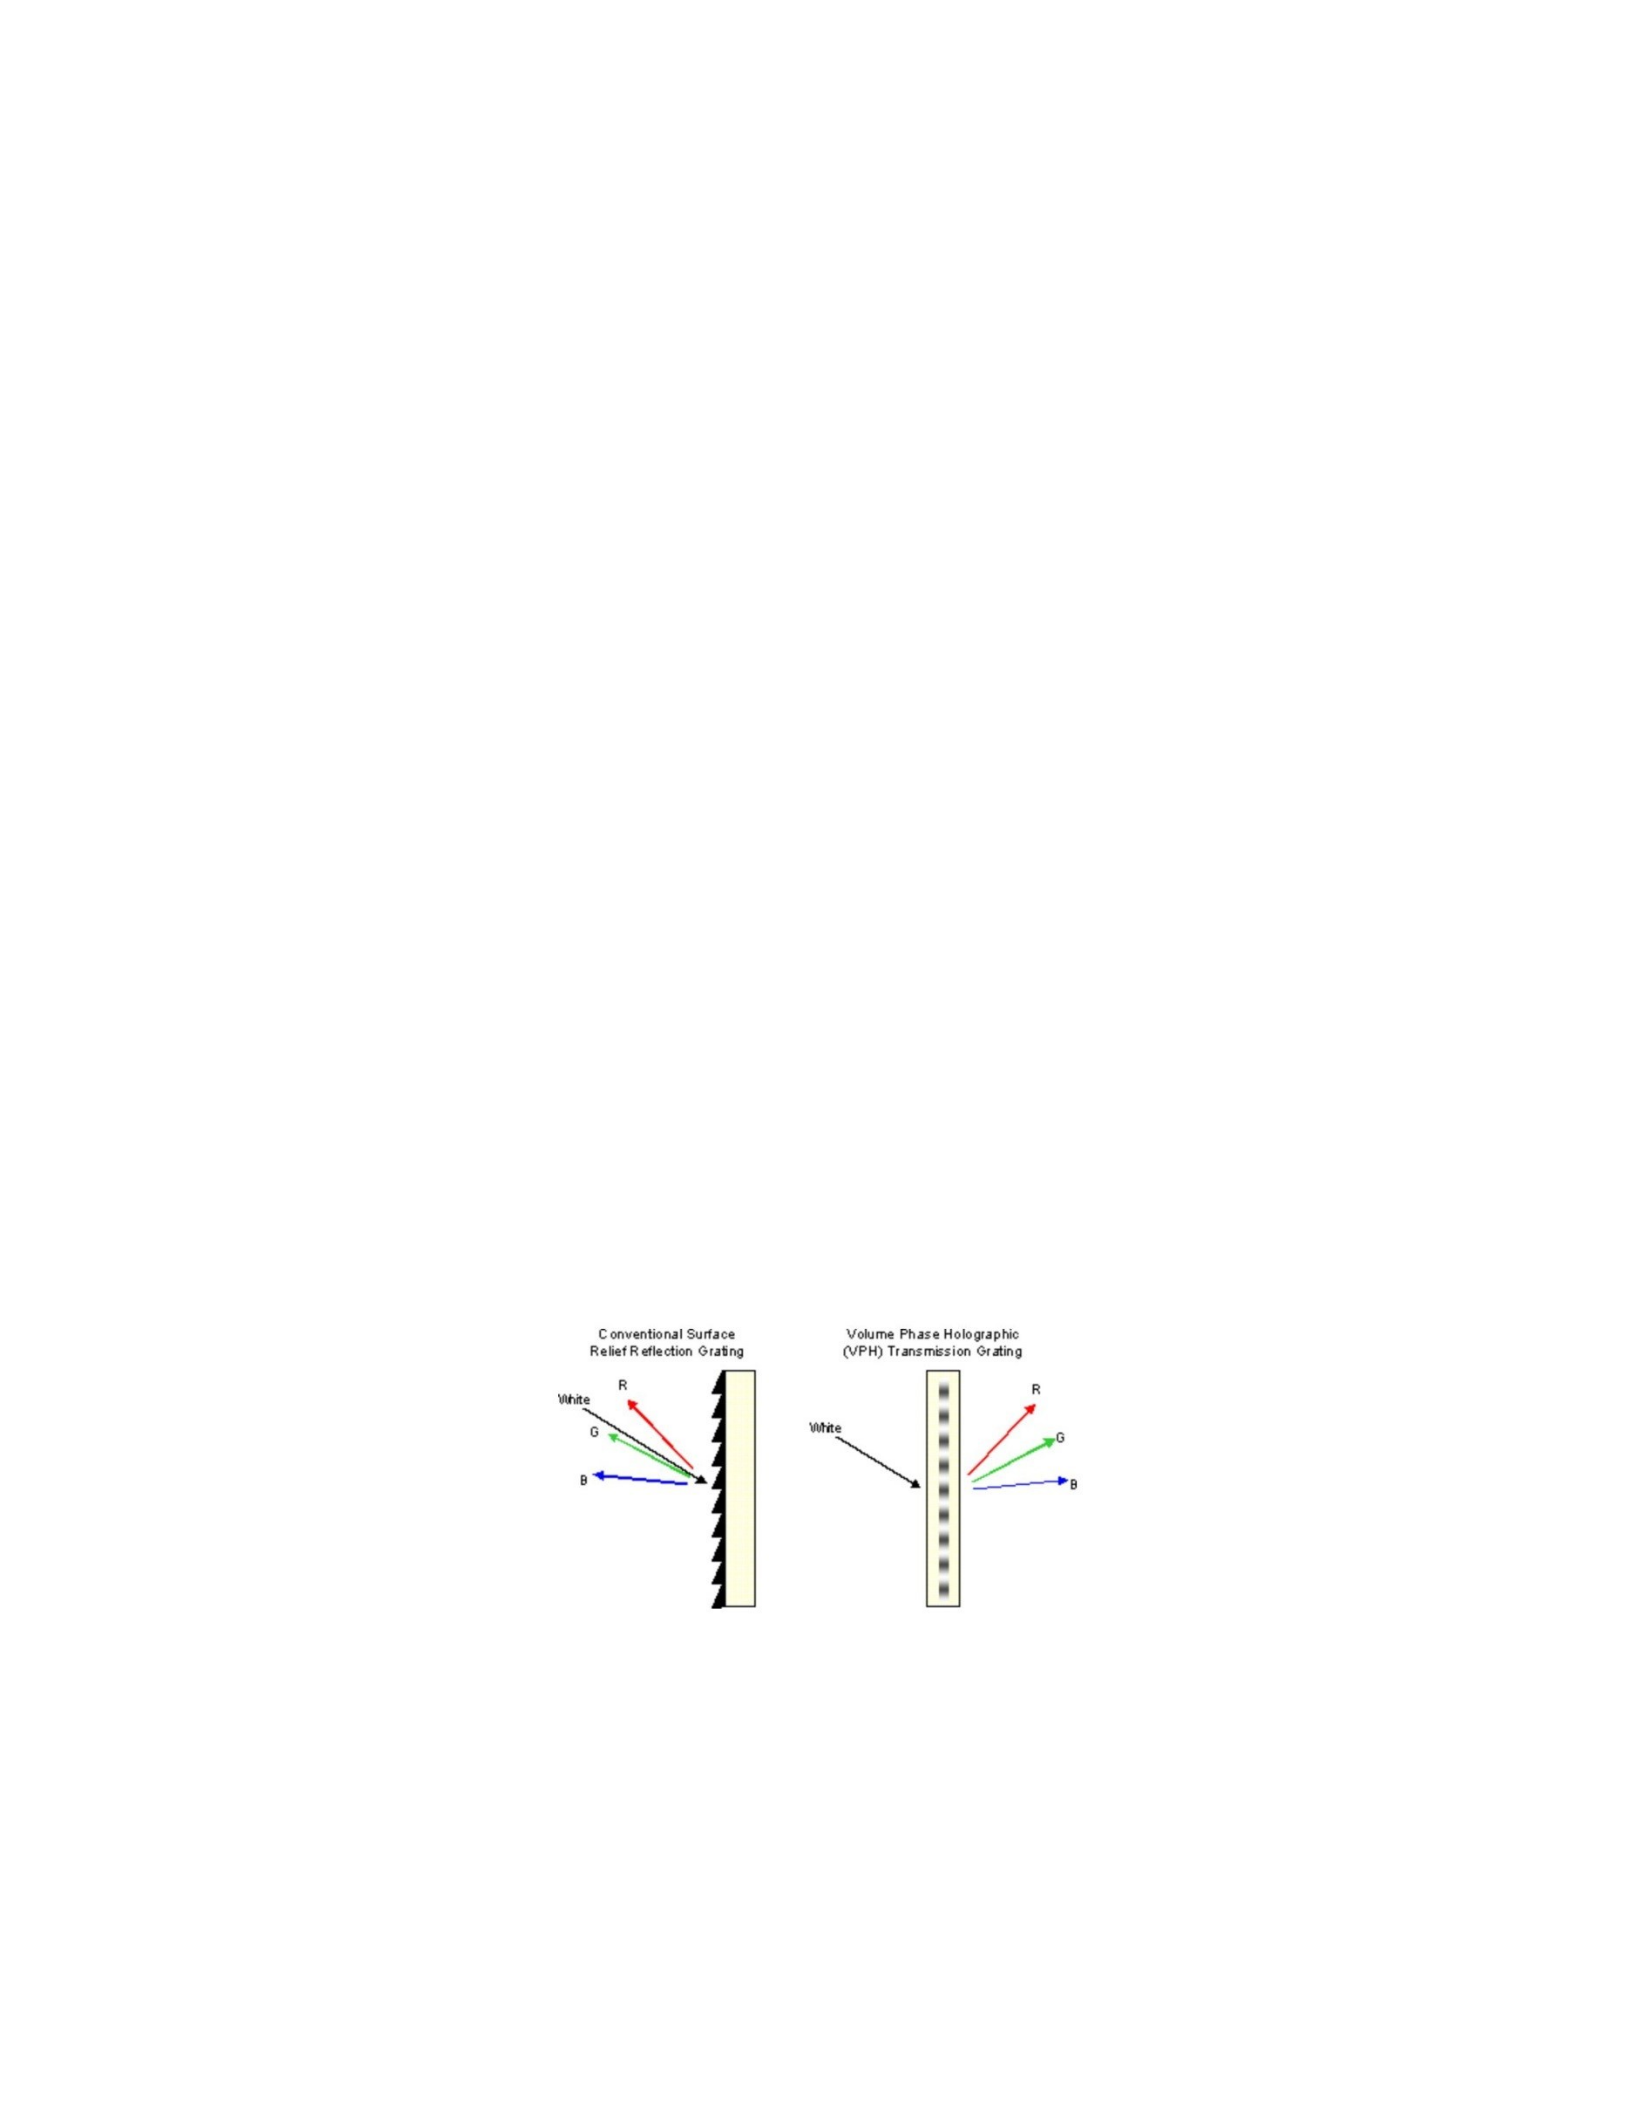
Hawthorn, & Robertson, 2004) Rather than having surface structure as in classical gratings,
VPH gratings diffract light by refractive index modulations within a thin layer of material
sandwiched between two glass substrates. Figure 5 shows a schematic (Kaiser Optical Systems,
Inc., 2014) of a conventional grating compared with a VPH grating.
Figure 5. Schematic of diffraction from conventional and VPH
gratings.
The intensity of the refractive index modulation and the depth of the grating layer are
critical parameters in the performance of the grating. Light is diffracted at angles corresponding

to the classical grating equation as a function of the incident angle and the frequency of the index
modulation at the surface of the grating. The diffraction efficiency, however, is a strong function
of the relationship between the angle of incidence and angle of diffraction with respect to the
fringes formed by the refractive-index modulations within the volume of the grating. If these
relationships satisfy the Bragg condition, which also depends on the depth of the grating volume
and on the intensity of the grating fringes, then high peak diffraction efficiencies, approaching
100%, are possible. (Barden, 1998)
As part of this study, tablet cores were crushed in a mortar and pestle and approximately
25 mg was transferred to a glass vial. The sample was analyzed using the Raman spectrometer
through the glass to evaluate chemical composition. The resulting Raman spectrum may also be
sensitive to solid-form differences. Although approximately 25 mg of sample is present in the
vial, the analysis spot size was approximately 50 μm for each spectrum collected.
Gas Chromtography-Mass Spectrometry
As part of this study, tablets were analyzed using a field-portable GC-MS to evaluate
organic volatiles and solvents remaining in the tablet after production (residual solvents). The
portable system used employs a low-thermal-mass (LTM) resistively heated capillary column
directly linked with a miniaturized toroidal ion-trap MS. This design is advantageous in
fielddeployed settings for a few reasons. The LTM GC column enables fast separation and
thermal recovery (approximately five minutes between the start of two consecutive injections).
This rate of thermal recovery is achieved because only the capillary column of the GC is heated
during the analysis. The miniaturized toroidal ion-trap MS minimizes power and vacuum
requirements, making the technology amenable for field analysis.

Ion-trap MS systems, by virtue of their simplicity, are ideal candidates for
miniaturization. They are inherently small and have only a few ion optic elements that do not
require highly precise alignment compared with other types of mass analyzers. The operating
pressure for ion traps is higher than for other forms of MS, allowing for less stringent pumping
requirements. Furthermore, since the radio frequency (rf) trapping voltage is inversely
proportional to the square of the analyzer radial dimension, a modest decrease in analyzer size
results in a large reduction in operating voltage and, hence, lower power requirements.
(Lammert, Rockwood, Wang, & Lee, 2006) A primary challenge for ion traps, especially when
maximizing reduction in size, is the management of ion-ion repulsions in the trap. The
toroidalion-trap design increases trapping volume and, therefore, minimizes these ion-ion
repulsions.
The toroidal ion trap can be viewed as a conventional three-dimensional ion trap cross
section that has been rotated on an edge through space. Figure 6 shows the geometry of the
conventional ion trap with axis of rotation shown in red (left), and the toroidal ion trap with axis
of rotation shown in blue (right). As can be seen, the result of this edge rotation is a trapping
field that is in the shape of a torus. This toroidal rf ion-trap design maintains a given trapping
field radius while increasing the ion storage volume of the ion trap. (Lammert, Plass, Thompson,
& Wise, 2001) This capability is significant.
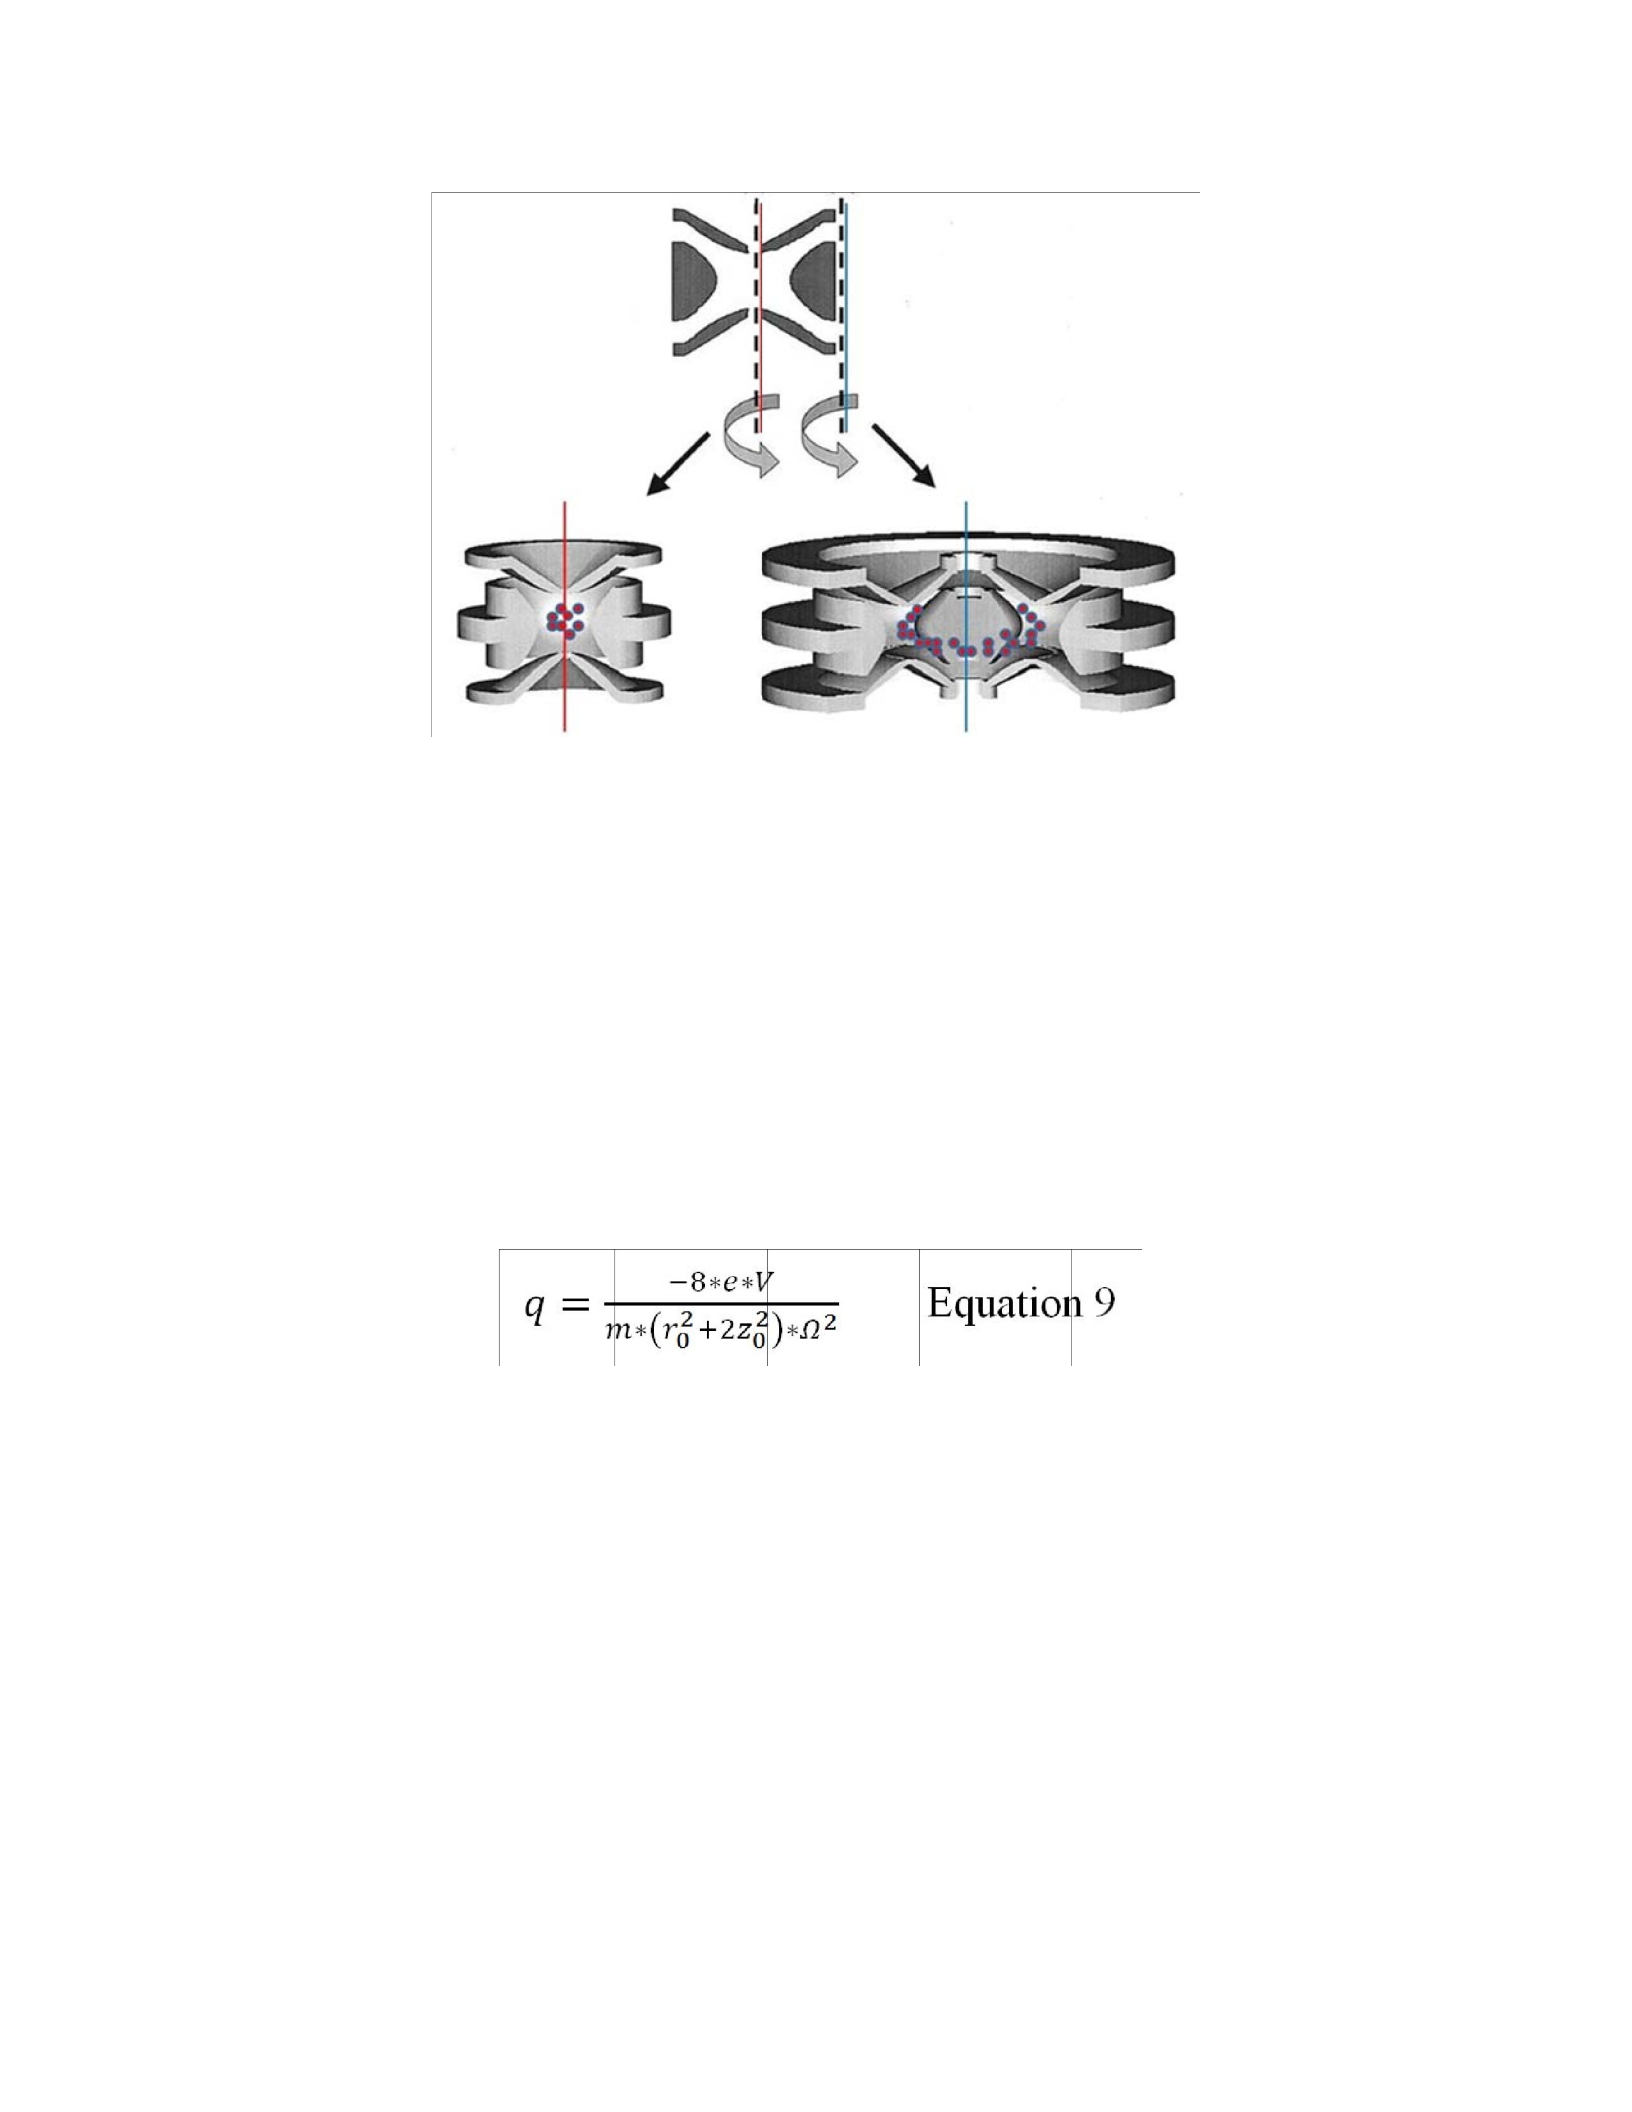
Figure 6. Cross section of conventional ion trap (left) and toroidal
ion trap (right). The axis of rotation is shown as a colored line for
each trap design
Equation 9 governs mass stability in a quadrupole ion-trap MS, where q is one of the two
Mathieau stability parameters, r0 is the radial dimension, V is the operating voltage, z0 is the axial
dimension, m is the ion mass, and Ω is the rf frequency. (Lammert, Rockwood, Wang, & Lee,
2006)
The ability to maintain the same radius, while increasing the trapping volume, gives the
advantages of minimizing ion-ion interactions while working at lower voltage and, therefore,
power requirements. The ability to operate at higher pressures, as well as the reduction in power
requirements are the primary advances that enabled field portability.
In this study, tablets were gently crushed and then heated in a vial for headspace analysis.
Gases and volatiles in the headspace were sampled using solid-phase microextraction (SPME).
Samples were thermally desorbed from the SPME upon injection into the GC-MS system. Fast

separation (less than three minutes) was performed on the GC. Analytes eluted directly from the
column into the ionization chamber of the toroidal ion-trap MS. An rf trapping field was applied,
and resonance ejection was achieved by applying a voltage of known frequency to the filament
end cap and increasing the amplitude of the rf trapping field such that the frequency of the ion
becomes the same as that applied to the end cap (mass-selective instability scan). Dynamic-
ionization optimized the number of ions in the trap for each analyte. Dynamic ionization control
is a feature that adjusts the filament current in real time to optimize the number of ions being
generated. This prevents space charging and other problems that result from overload of the ion
trap. The resulting scanned mass range was from 42 to 500 m/z.
Chemometric Methods
When collecting large amounts of chemical data, it can be useful to apply mathematical
methods to extract useful chemical information. As part of this research, the chemometric
methods of PCA and canonical variate analysis (CVA) were applied to both the infrared and
Raman data after autoscaling to investigate the classification and discrimination power of each of
these methods. The deconvolution data, specifically the peak positions and areas, also were
analyzed using hold-one-out cross validation (HOO-CV) PCA-CVA to determine an estimated
error rate for the classification of sildenafil citrate tablets using each of these methods.
As previously stated, data was autoscaled prior to PCA analysis. Autoscaling is the
combination of mean centering and variance scaling. Mean centering is performed by
calculating the average data vector of all n rows in a data set and subtracting it point by point
from each vector in the data set. Graphically, mean centering corresponds to a shift in the origin
of the plot. Variance scaling is the process of normalizing each column so that the sum of
squares equals one. Variance scaling is performed in order to give equal weight or importance to
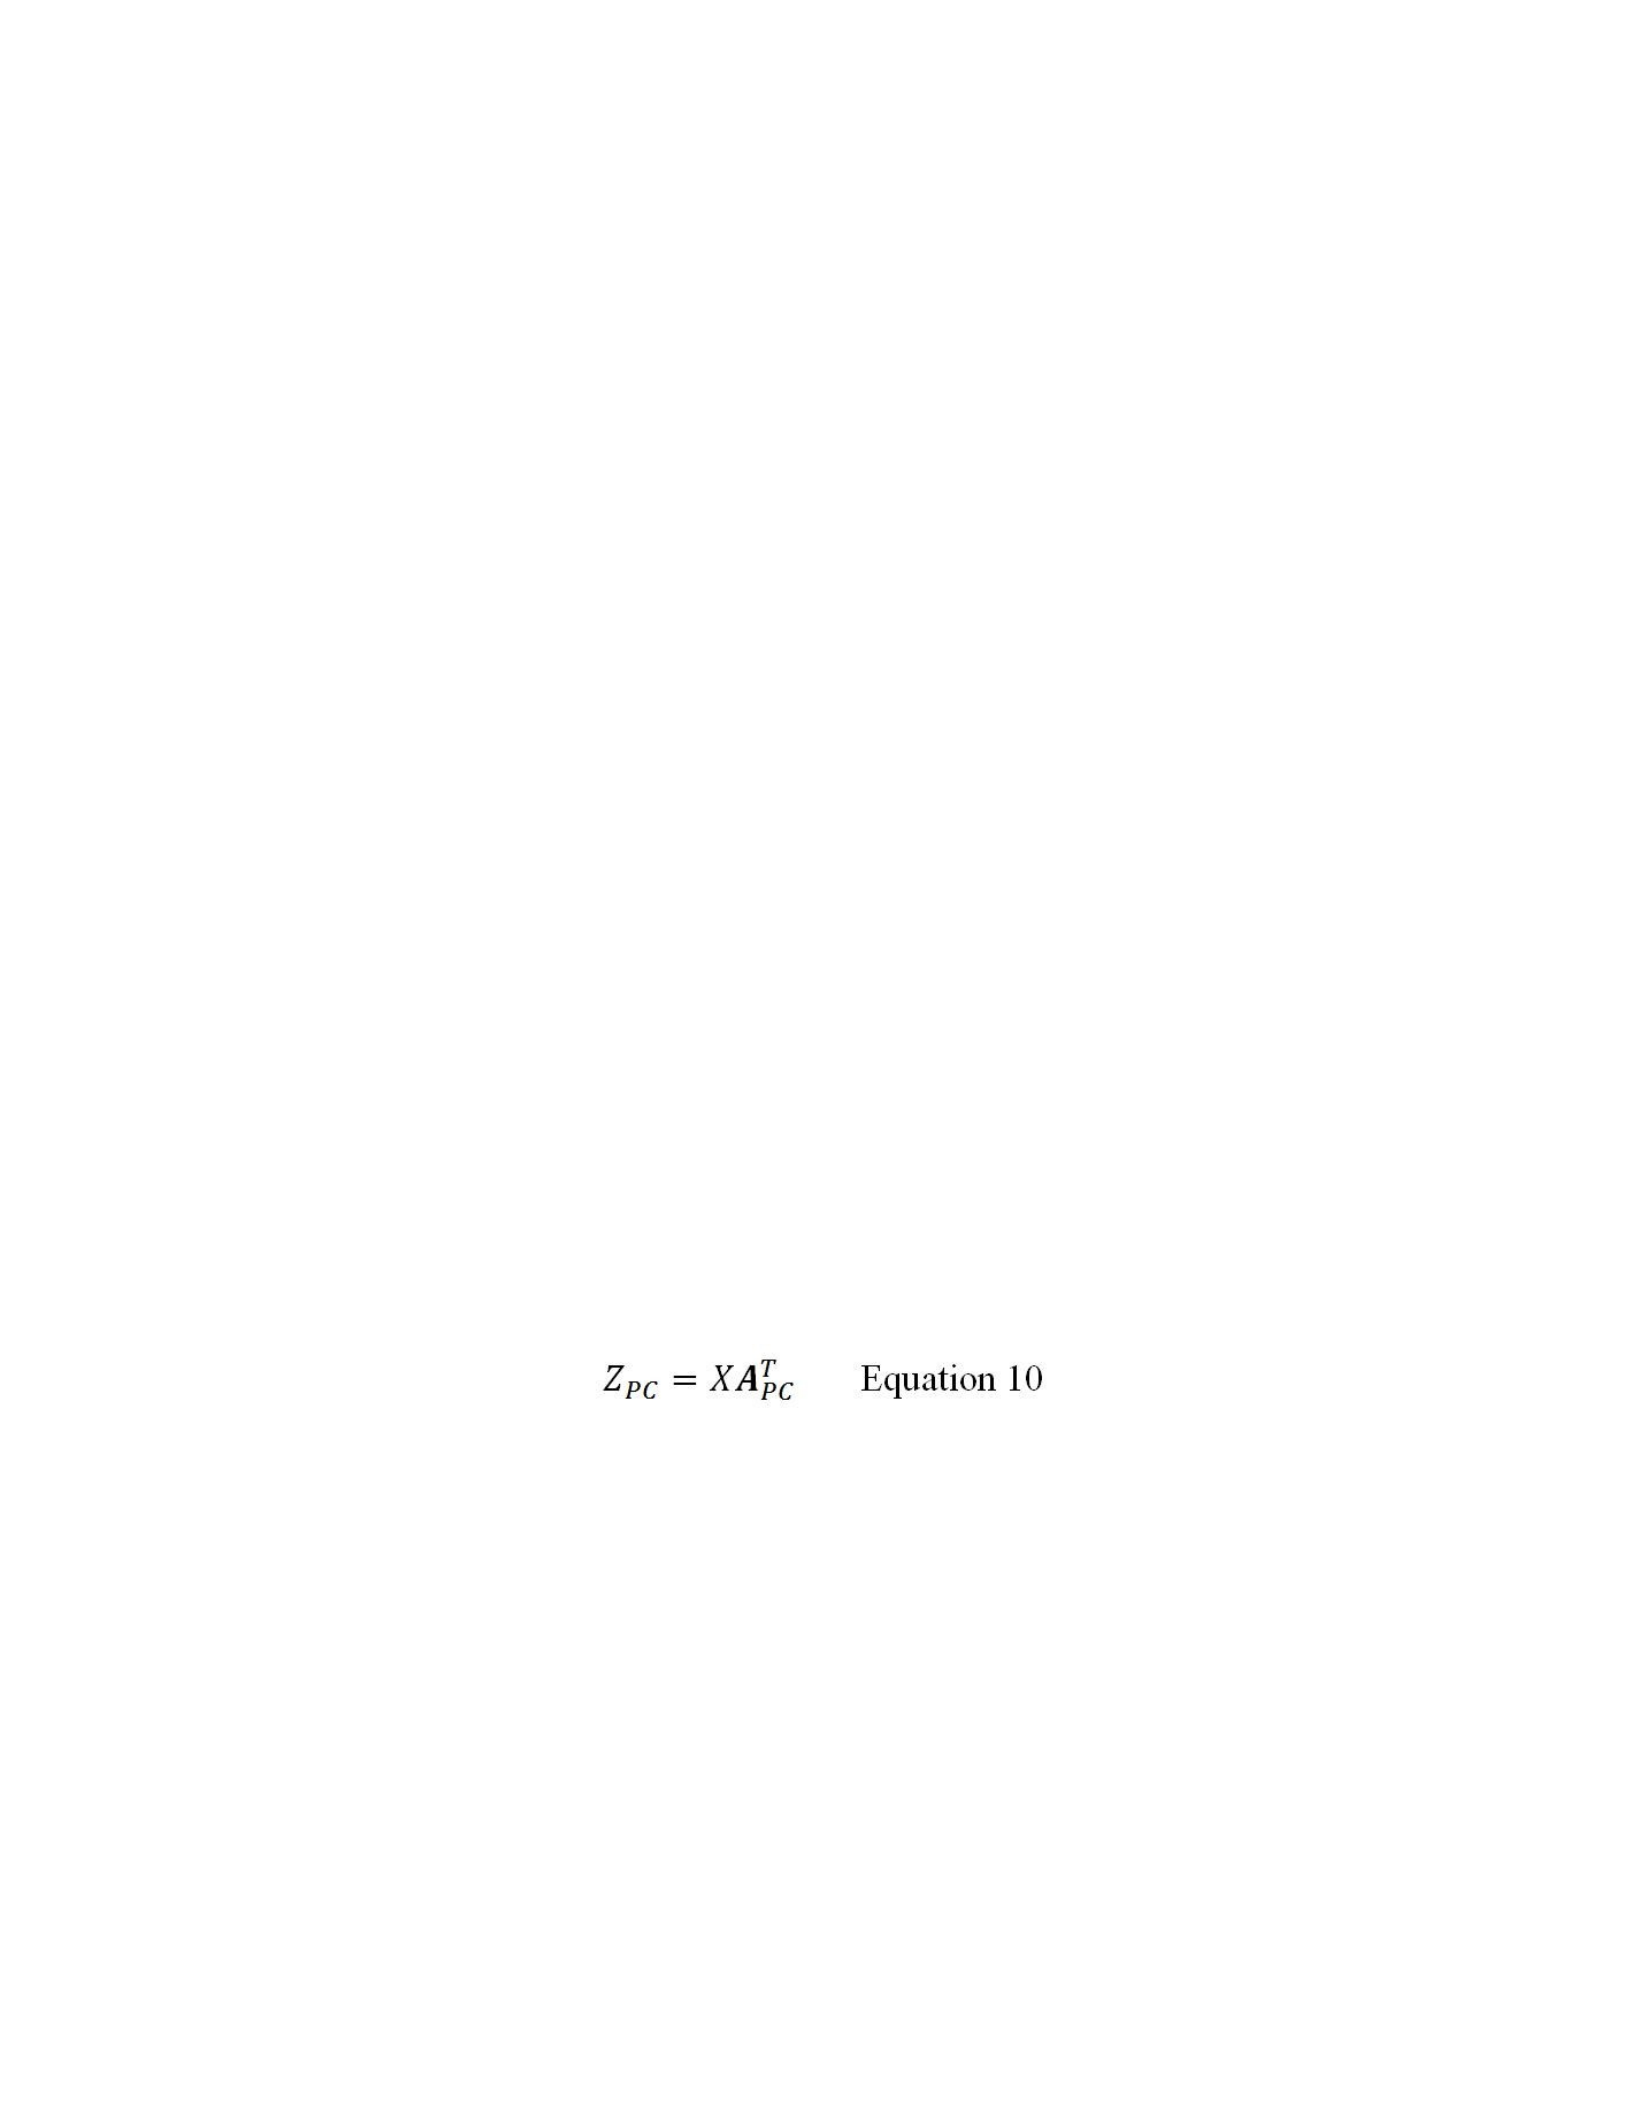
all variables (or wavenumbers) in the measured data set, and must be completed after mean
centering. (Varmuza & Filmozer, 2009)
Principal Component Analysis
PCA is a chemometric method that reduces the dimensionality of data based upon the
variance in the data. This allows the user to remove the large amounts of data that contain the
least amount of variance. As part of this research, the number of starting dimensions was equal
to the number of wavenumber data points for each spectrum. For the infrared data, the number
of original dimensions was 1425; for the Raman data, the number of dimensions was 938. Of
course this is a highly redundant representation and should be reduced in dimension. (Petraco,
2014)
The first PC, i.e., first reduced dimension, is the linear combination of the original
variables with maximum variance. The second PC is the linear combination that exhibits the
second greatest amount of variance, but is orthogonal to the first PC. Each successive PC
exhibits less variance than the one before it. The linear combination of the original variables is
used to form a set of derived variables, i.e., PC’s. The matrix form of these derived variables is
where the superscript T represents the transpose of APC. The new data set Z is derived from the
original variables, but sorted in order of decreasing variance so PC’s containing minimal
information can be removed. This allows for the reduction in the number of derived variables
used without the loss of a significant amount of information from the data. (Varmuza &
Filmozer, 2009)
The matrix APC is computed by diagonalizing the p x p maximum likelihood covariance
matrix (S) of X
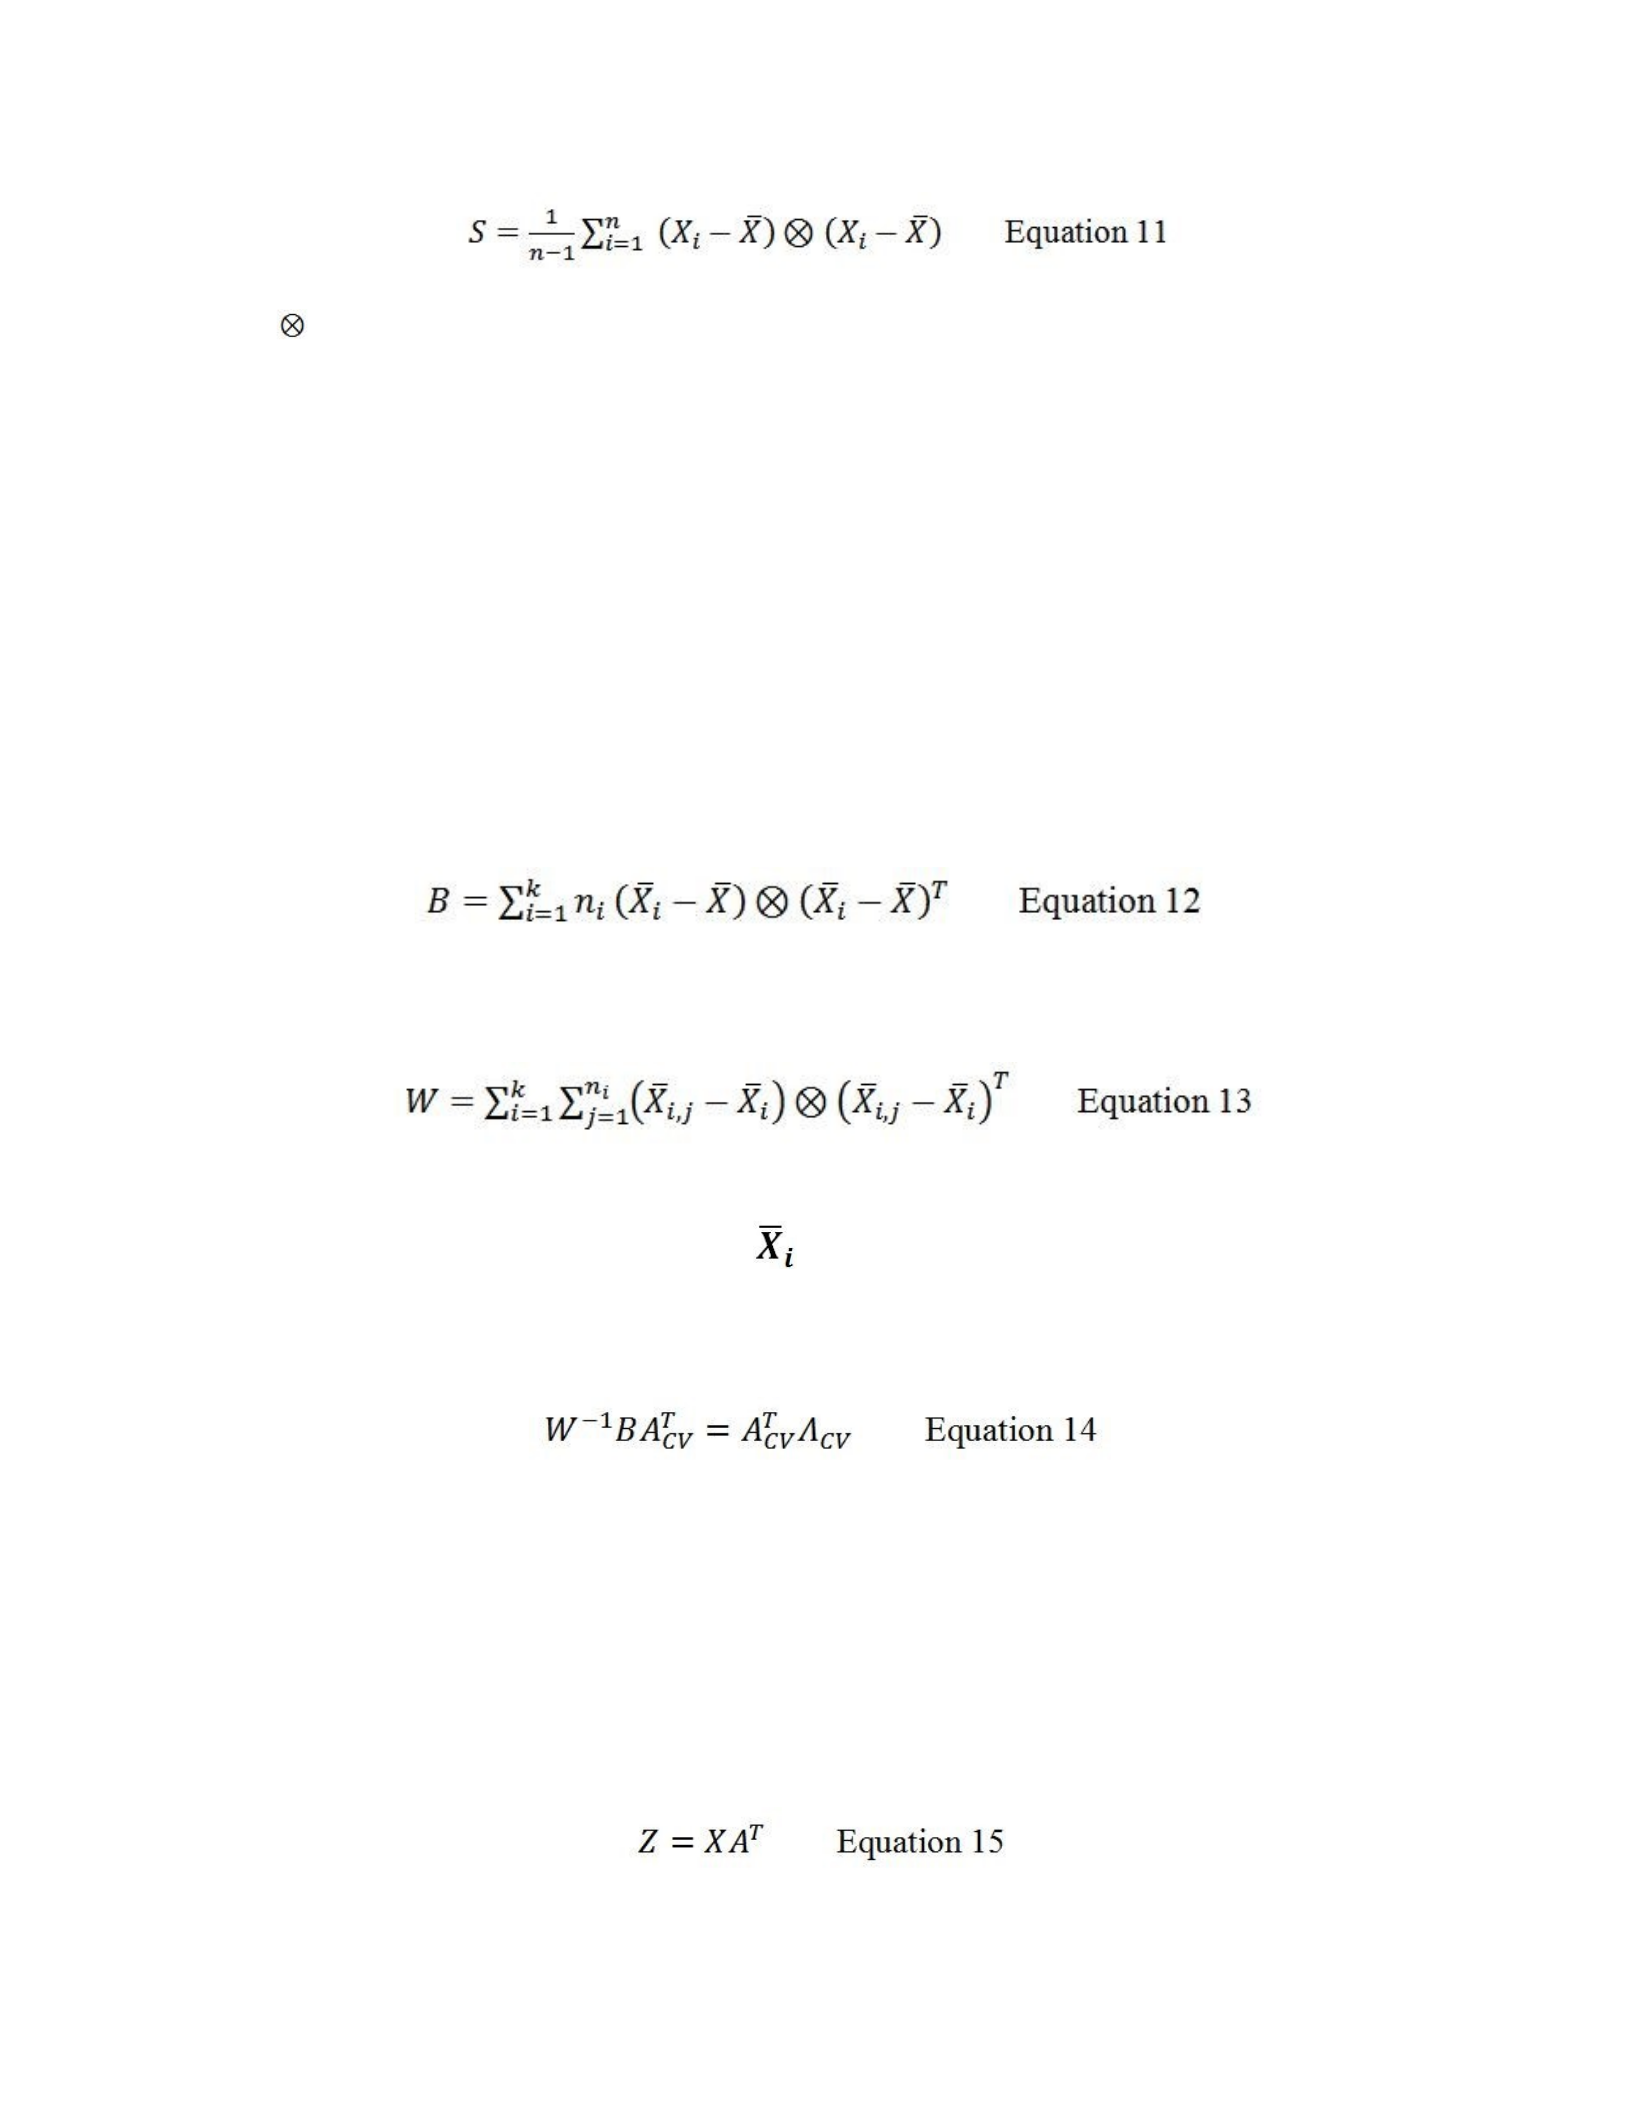
where is the Kronecker product of vectors.
Canonical Variate Analysis
For this research, CVA was performed on the derived data computed from PCA. CVA
uses a covariance matrix, W-1B, to find the direction of best group separation. The analysis is
based on the relationship between-group (B) variance and within-group (W) variance.
The CVs, ACV, and their eigenvalues, ΛCV, are computed by diagonalizing the covariance
matrix (W-1B), where
and
The non-singular W is inverted using a standard inversion method. Xi,j represents the jth
absorbance response in the ith spectrum and is the average of all of the spectra in the ith
sample. There are ni spectra in sample i. The eigenproblem for CVA is
and is not symmetric and, therefore, its eigenvectors are not necessarily orthogonal. Thus as
opposed to PCA, the CV’s are often not perpendicular. As a consequence, covariance in the data
causes the CVA model to fail, which is why spectral data must first undergo PCA. The last step
in CVA is the transformation of the data on the basis of the retained CV’s as
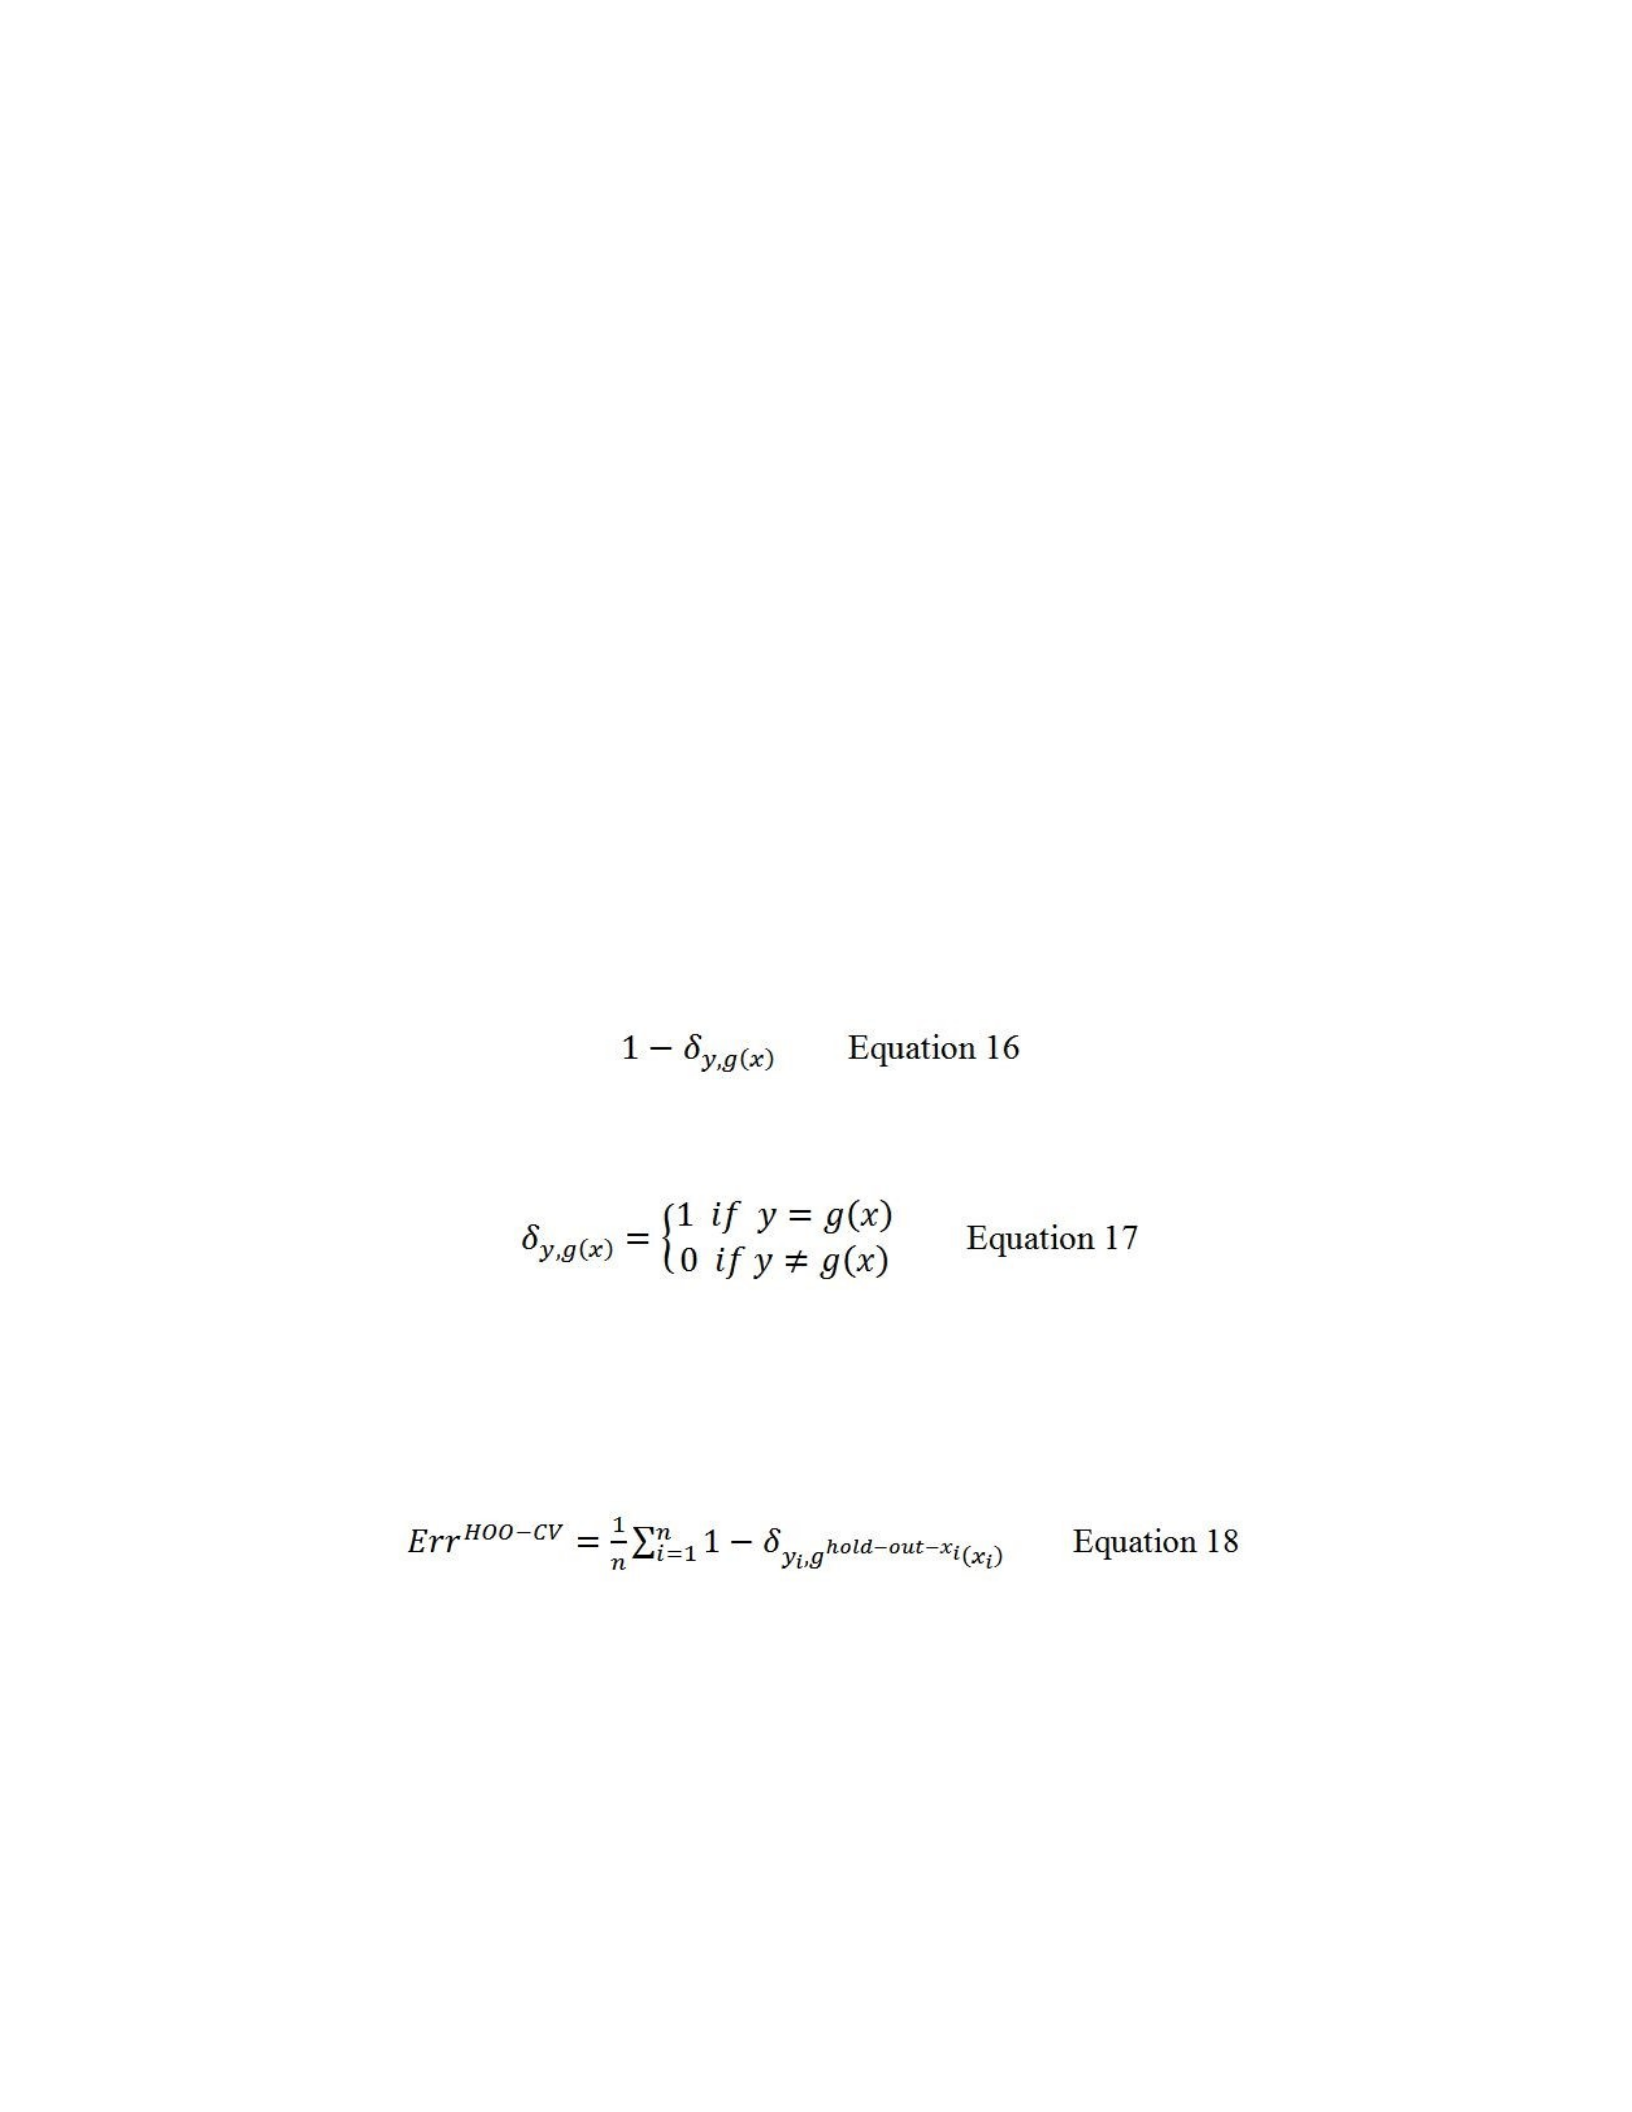
Hold-One-Out Cross Validation
HOO-CV is a method for estimating an algorithm’s error rate (called the estimated error
rate) for a population of spectra using data from which it was not trained. HOO-CV was used as
part of this research to assess the estimated error rates of the mathematical modes created using
PCA-CVA. HOO-CV is a useful method to estimate error rates when large data sets are not
available for testing. (Effron & Tibshirani, 1993) HOO-CV computes the decision rules using all
but one of the spectra in the data set. The HOO-CV estimated error rate is calculated by first
determining whether the decision rule was correct on a held-out spectrum, x, with true identity, y,
where ghold-out-x(x) denotes the identity of x assigned by the “hold-one-out” decision rules. Error
or misclassifications are assigned a 1 and correct classifications a 0, which is symbolically
written as:
where the Kronecker delta denotes 1 if y = g(x) and 0 otherwise.
The HOO-CV procedure is repeated for each spectrum in the data set and the results are averaged
to calculate an estimated error rate shown in equation 18.

CHAPTER 6. SAMPLES AND SAMPLING MATRIX
Sample Procurement
Testing performed as part of this research was used to develop an analytical scheme to
rapidly and reliably detect counterfeit drugs in the field. Attempts were made to procure
“known” counterfeit drugs for testing. Many pharmaceutical companies that manufacture drugs
known to be routinely counterfeited were contacted and requests for samples were made.
However, no suppliers contacted were willing to provide test samples. For this reason, samples
were ordered online from websites offering to sell prescription medication direct to a consumer
located within the United States without a prescription. The drug ordered was sildenafil citrate
100-mg tablets. No prescription was provided at the time of any online purchase, so it was
expected that the samples ordered online would be non-authentic versions of Pfizer’s VIAGRA
100-mg tablets. This is because prescription medications (like VIAGRA) are not allowed to be
legally sold within the United States without a prescription. There are many reported reasons
this drug is a target of counterfeiters in the United States and around the world, and reports with
regard to the online purchase of VIAGRA indicate most are fake. (Fiore, 2012) For comparison,
sixteen tablets expected to be authentic were purchased in the United States through
FDAgoverned pharmacies.
VIAGRA is not a drug commonly tested in crime labs in the United States. This is in
spite of the fact that there are many reports of availability of non-authentic versions, and any
non-authentic version of this drug sold in the United States is illegal. (35 U.S.C. § 271; Federal
Food, Drug, and Cosmetic Act, 1938) The reason non-authentic VIAGRA samples are illegal is
because they violate FDA regulations, and IP law. Reasons for this discrepancy are variable and
include availability of resources within law enforcement, jurisdiction issues, and other
enforcement challenges. (Dean, 2013; Faucon & Whalen, 2012; Sen, 2012) However, it is a
good drug of choice for inclusion in this testing for a few reasons. First, VIAGRA is currently
under patent in the United States. (Ellis & Terret, 2002) Therefore, any non-authentic sample
purchased in the United States, by definition, is either an illegal counterfeit or an illegal imitation
version of the drug. Second, samples ordered online were purchased from “pharmacies” that
appeared to be looking for repeat purchasers. At the time of purchase, each website requested
contact emails for marketing purposes. They all also asked for permission to contact the
purchaser to send a reminder when a “refill” was due. It was expected, therefore, that tablets
received would likely contain at least some therapeutic amount of API. This was valuable from
an analytical perspective because it required that methods used as part of this scheme be able to
distinguish samples that were chemically similar to each other. It would be significantly easier to
identify counterfeits or imitations if no API or a different API were present in the final dosage
form.
Sample Summary
Websites from which tablets purchased online were ordered were found using various
search terms and search engines. Purchases from any website were only made once, i.e., no
repeat purchases were made from the same website by this researcher. Table 1 is a summary of
all samples included in this testing from online purchases and purchases made from
FDAgoverned pharmacies in the United States.
C of Postmarking on External
S ID P
I1 India
I2 India
I3 India
I4 India
I5 India
S7 India
S8 China
SING1
Singapore (although blister packaging
identifies country of manufacture as India)
SING2
Singapore (although blister packaging
identifies country of manufacture as India)
S13 FDA-governed pharmacy in US
S14 FDA-governed pharmacy in US
S15 FDA-governed pharmacy in US
S16 FDA-governed pharmacy in US
Table 1. Summary of samples used for analytical testing.
Sampling Workflow
Analysis of each sample followed a step-by-step process that included photographic
documentation, dimensional analysis, and physical/chemical characterization. The details of
each method performed will be described in chapters 7 through 10, but the general testing
process is summarized in figure 7.
Figure 7. Analytical workflow for all test samples.
Sampling Matrix
Although attempts were made to analyze at least seven tablets from every sample, this
was not always possible because in some cases seven tablets were not available for testing.
Attempts were also made to analyze all seven tablets from each sample for GC-MS analysis.
This was also not always possible because for some samples, a GC-MS system was not available
at the time of analysis of that sample, or instrumental problems occurred that prevented analysis.
Due to the fact that the entire tablet for GC-MS analysis was heated to generate a headspace
appropriate for sampling, repeat testing at a different time was not an option if instrumental
problems occurred.
Clarification of sampling is also necessary for replicate spectra collected from the same
tablet. For some Raman spectra collected, there was a file corruption that occurred during data
transport from the instrument. Although data for the sample was collected, in some cases, the
third replicate analysis performed could not be included due to this file corruption.
To clarify the exact testing performed, a sample matrix showing all testing performed is
shown in Tables 2 through 4.
Summary of All Measurements Performed
tablet length measurements 210
tablet width measurements 210
tablet depth measurements 210
tablet weight measurements 210
infrared spectral measurements 231
Raman spectral measurements 177
GC-MS measurements 207
total number of measurements (not 1455
including photographs)
Table 2. Summary of measurements performed on all samples.
sample ID
sample
classification Photographs tablet length tablet width tablet depth tablet weights
S13 T1 Authentic Yes 3 3 3 3
S13 T2 Authentic Yes 3 3 3 3
S13 T3 Authentic Yes 3 3 3 3
S13 T4 Authentic Yes 3 3 3 3
S14 T1 Authentic Yes 3 3 3 3
S14 T2 Authentic Yes 3 3 3 3
S14 T3 Authentic Yes 3 3 3 3
S14 T4 Authentic Yes 3 3 3 3
S15 T1 Authentic Yes 3 3 3 3
S15 T2 Authentic Yes 3 3 3 3
S15 T3 Authentic Yes 3 3 3 3
S15 T4 Authentic Yes 3 3 3 3
S16 T1 Authentic Yes 3 3 3 3
S16 T2 Authentic Yes 3 3 3 3
S16 T3 Authentic Yes 3 3 3 3
S16 T4 Authentic No 0 0 0 0
I1 T1 Illegal Imitation Yes 3 3 3 3
I1 T2 Illegal Imitation Yes 3 3 3 3
I1 T3 Illegal Imitation Yes 3 3 3 3
I1 T4 Illegal Imitation Yes 3 3 3 3
I1 T5 Illegal Imitation Yes 3 3 3 3
I1 T6 Illegal Imitation Yes 3 3 3 3
I1 T7 Illegal Imitation Yes 3 3 3 3
I2 T1 Illegal Imitation Yes 3 3 3 3
I2 T2 Illegal Imitation Yes 3 3 3 3
I2 T3 Illegal Imitation Yes 3 3 3 3
I2 T4 Illegal Imitation Yes 3 3 3 3
I2 T5 Illegal Imitation Yes 3 3 3 3
I2 T6 Illegal Imitation Yes 3 3 3 3
I2 T7 Illegal Imitation Yes 3 3 3 3
I3 T1 Illegal Imitation Yes 3 3 3 3
I3 T2 Illegal Imitation Yes 3 3 3 3
I3 T3 Illegal Imitation Yes 3 3 3 3
I3 T4 Illegal Imitation Yes 3 3 3 3
I3 T5 Illegal Imitation Yes 3 3 3 3
I3 T6 Illegal Imitation Yes 3 3 3 3
I3 T7 Illegal Imitation Yes 3 3 3 3
I4 T1 Illegal Imitation Yes 3 3 3 3
I4 T2 Illegal Imitation Yes 3 3 3 3
I4 T3 Illegal Imitation Yes 3 3 3 3
I4 T4 Illegal Imitation Yes 3 3 3 3
I4 T5 Illegal Imitation Yes 3 3 3 3
I4 T6 Illegal Imitation Yes 3 3 3 3
I4 T7 Illegal Imitation Yes 3 3 3 3
I5 T1 Illegal Imitation Yes 3 3 3 3
I5 T2 Illegal Imitation Yes 3 3 3 3
I5 T3 Illegal Imitation Yes 3 3 3 3
I5 T4 Illegal Imitation Yes 3 3 3 3
I5 T5 Illegal Imitation Yes 3 3 3 3
I5 T6 Illegal Imitation Yes 3 3 3 3
I5 T7 Illegal Imitation Yes 3 3 3 3
S7 T1 Illegal Imitation Yes 0 0 0 0
S7 T2 Illegal Imitation Yes 3 3 3 3
S7 T3 Illegal Imitation Yes 3 3 3 3
S7 T4 Illegal Imitation Yes 0 0 0 0
S7 T5 Illegal Imitation Yes 0 0 0 0
S7 T6 Illegal Imitation Yes 0 0 0 0
S7 T7 Illegal Imitation Yes 3 3 3 3
S7 T8 Illegal Imitation Yes 0 0 0 0
S8 T1 Counterfeit Yes 0 0 0 0
S8 T2 Counterfeit Yes 0 0 0 0
S8 T3 Counterfeit Yes 0 0 0 0
S8 T4 Counterfeit Yes 3 3 3 3
S8 T5 Counterfeit Yes 3 3 3 3
S8 T6 Counterfeit Yes 0 0 0 0
S8 T7 Counterfeit Yes 0 0 0 0
S8 T8 Counterfeit Yes 3 3 3 3
SING1 T1 Illegal Imitation Yes 3 3 3 3
SING1 T2 Illegal Imitation Yes 3 3 3 3
SING1 T3 Illegal Imitation Yes 3 3 3 3
SING1 T4 Illegal Imitation Yes 3 3 3 3
SING1 T5 Illegal Imitation Yes 3 3 3 3
SING1 T6 Illegal Imitation Yes 3 3 3 3
SING1 T7 Illegal Imitation Yes 3 3 3 3
SING2 T1 Illegal Imitation Yes 3 3 3 3
SING2 T2 Illegal Imitation Yes 3 3 3 3
SING2 T3 Illegal Imitation Yes 3 3 3 3
SING2 T4 Illegal Imitation Yes 3 3 3 3
SING2 T5 Illegal Imitation Yes 3 3 3 3
SING2 T6 Illegal Imitation Yes 3 3 3 3
SING2 T7 Illegal Imitation Yes 3 3 3 3
Total Number of
Measurements
210 210 210 210
Table 3. Summary of photography and dimensional measurements performed
on each tablet.
sample ID infrared spectra core Raman spectra core
GC MS at 60C standard ‐
fiber GC MS at 60C carboxen fiber‐
GC MS at 110C standard ‐
fiber GC MS at 110C carboxen fiber‐
S13 T1 3 0 1 1 1 1
S13 T2 3 0 1 1 1 1
S13 T3 0 0 0 0 0 0
S13 T4 3 0 1 1 1 1
S14 T1 3 0 0 0 0 0
S14 T2 3 0 1 1 1 1
S14 T3 3 0 0 0 0 0
S14 T4 3 0 0 0 0 0
S15 T1 3 3 1 1 1 1
S15 T2 3 3 1 1 1 1
S15 T3 3 3 1 1 1 1
S15 T4 3 3 1 1 1 1
S16 T1 3 3 1 1 1 1
S16 T2 3 3 1 1 1 1
S16 T3 3 3 1 1 1 1
S16 T4 0 0 1 1 1 1
I1 T1 3 2 1 0 1 1
I1 T2 3 3 1 0 1 1
I1 T3 3 2 1 0 1 1
I1 T4 3 3 1 0 1 0
I1 T5 3 2 1 0 1 0
I1 T6 3 3 1 0 1 0
I1 T7 3 3 1 0 1 0
I2 T1 3 3 0 1 1 0
I2 T2 3 3 0 1 1 0
I2 T3 3 2 1 0 1 0
I2 T4 3 3 0 1 1 0
I2 T5 3 3 1 0 1 0
I2 T6 3 3 1 0 1 0
I2 T7 3 2 1 0 1 0
I3 T1 3 3 1 1 1 1
I3 T2 3 2 1 1 1 1
I3 T3 3 3 1 1 1 1
I3 T4 3 3 1 1 1 1
I3 T5 3 3 1 1 1 1
I3 T6 3 3 1 1 1 1
I3 T7 3 3 1 1 1 1
I4 T1 3 3 1 1 1 1
I4 T2 3 3 1 1 1 1
I4 T3 3 2 1 1 1 1
I4 T4 3 3 1 1 1 1
I4 T5 3 3 1 1 1 1
I4 T6 3 3 1 1 1 1
I4 T7 3 3 1 1 1 1
I5 T1 3 3 1 1 1 1
I5 T2 3 3 1 1 1 1
I5 T3 3 3 1 1 1 1
I5 T4 3 3 1 1 1 1
I5 T5 3 3 1 1 1 1
I5 T6 3 3 1 1 1 1
I5 T7 3 3 1 1 1 1
S7 T1 3 0 0 0 0 0
S7 T2 3 3 1 0 1 0
S7 T3 3 3 0 0 0 0
S7 T4 3 0 1 1 1 1
S7 T5 3 0 1 1 1 1
S7 T6 0 0 0 0 0 0
S7 T7 3 2 0 0 0 0
S7 T8 3 0 0 0 0 0
S8 T1 3 0 0 1 1 0
S8 T2 0 0 0 0 0 0
S8 T3 3 0 0 0 0 0
S8 T4 3 2 0 0 0 0
S8 T5 3 3 0 0 0 0
S8 T6 3 0 1 1 1 1
S8 T7 3 0 0 0 0 0
S8 T8 3 3 0 1 1 0
SING1 T1 3 3 0 0 0 0
SING1 T2 3 3 1 0 1 0
SING1 T3 3 3 1 0 1 0
SING1 T4 3 3 0 1 1 0
SING1 T5 3 3 0 0 0 0
SING1 T6 3 3 0 1 1 0
SING1 T7 3 3 0 0 0 0
SING2 T1 3 3 1 1 1 1
SING2 T2 3 3 1 1 1 1
SING2 T3 3 3 1 1 1 1
SING2 T4 3 3 1 1 1 1
SING2 T5 3 3 1 1 1 1
SING2 T6 3 3 1 1 1 1
SING2 T7 3 3 1 1 1 1
Total Number of
Measurements
231 177 57 50 64 46
Table 4. Summary of infrared, Raman and GC-MS measurements
performed on each tablet.
CHAPTER 7. DIMENSIONAL ANALYSIS AS A TOOL FOR IDENTIFICATION AND
DIFFERENTIATION OF AUTHENTIC DRUGS

Introduction
The manufacture of pharmaceutical tablets is a complicated process. Raw materials are
blended together using a variety of solvents, usually at elevated temperatures. These materials
are ultimately dried and compressed into tablets of specified physical dimensions.
Manufacturing equipment such as the die’s used to impress product-specific labeling into the
tablet also change the physical features of the tablet. For coated tablets, coating materials and the
process used to perform the coating may also impart upon the final dosage form characteristics
that are specific to the manufacturer. Analysis and comparison of these dimensional
measurements and other physical and chemical properties between authentic and non-authentic
samples may sometimes result in the ability to quickly and easily differentiate authentic from
non-authentic samples. (Jung, Ortiz, Limberger, & Mayorga, 2012) This data may be further
used to classify counterfeit drugs providing investigative leads useful in establishing source or
origin. They are also amongst the easiest of analyses to perform in the field.
Tablet weight is an important measurement because it is difficult, unless samples are
manufactured identically using the same types and amounts of ingredients, to generate a
reproducible tablet weight. An analytical balance will easily detect these differences, especially
in cases where tablets might otherwise appear grossly to be the same. Physical dimensions such
as length, width, depth, and overall tablet shape may also be different if different manufacturing
equipment is used to press the final product. A caliper may be used to measure these properties.
Tablet color and coating details will also depend upon both the raw materials used, as well as the
process used to coat the tablet. Aside from color and chemical composition, coatings may
appear and behave differently. Photographs may be used to document these differences. In this
study, photographs of tablets were taken and dimensional measurements were performed to

determine whether or not it was possible to differentiate between authentic from non-authentic
samples.
Experimental Details
Each tablet was weighed and photographed. The tablet’s length, width, and depth were
measured using a digital caliper. The average weight, length, width, and depth were calculated
for all tablets of a given sample. Results were compared for different tablets of the same sample,
as well as for different samples.
Photography
Photographs were taken using a Samsung Galaxy phone with camera, model SCH-1535,
Android version 4.1.2. Tablet images were collected from both sides of each tablet at three
different distances. Images captured from a distance of 60.4 mm were selected for inclusion in
this summary. Magnification was set to 4.0 times, but autofocus was used.
Tablet Dimensions
Measurements of tablet length, width, and depth were taken for each tablet using a
Ferance Construction Co. digital caliper: For each dimension, three measurements were taken
for each tablet. The “AVERAGE” function of Microsoft Excel was used to calculate the tablet’s
dimension value from the three measured values (Microsoft Excel 2010 running on Windows 7
Enterprise, Service Pak 1, 64-bit Operating System). The average dimension and standard
deviation for each sample was then calculated from the determined values for each tablet of the
sample using the “AVERAGE” and “STDEV” functions within Microsoft Excel. From these
values, the %RSD was calculated for each sample using Microsoft Excel with the formula
“=(STDEV/AVERAGE)*100”.

Tablet Weights
Tablet weights were measured using two different balances. The balance used was based
upon availability at the time of analysis. Each balance had a readability as low as 0.1 mg. Both
balances were under service contracts for calibration at all times measurements were made. For
each tablet, sample weight was measured three times. The balance was tared between
measurements. The “AVERAGE” function of Microsoft Excel was applied to the three values to
determine tablet weight (Microsoft Excel 2010 running on Windows 7 Enterprise, Service Pak 1,
64-bit Operating System). The average weight and standard deviation of the sample were then
calculated from the weights all tablets of the sample using the “AVERAGE” and “STDEV”
functions within Microsoft Excel. The %RSD was using Microsoft Excel with the formula
“=(STDEV/AVERAGE)*100”.
Results and Discussion
It was possible to differentiate the authentic samples from all non-authentic samples
based upon visual observation. For eight of the nine non-authentic samples, the tablets could
very easily be distinguished from the authentic samples with the unaided eye. These eight
nonauthentic samples exhibited obvious differences in color, tool-mark impressions, and even
shape, from the authentic samples. Weight measurements were also frequently different from
each other. The ninth sample, sample S8, was counterfeit and was more similar in general
appearance than the other eight samples to the authentic samples. These tablets were still
differentiable, however, by visual observation, when compared with authentic VIAGRA. Just
like for authentic 100-mg VIAGRA tablets, the tablets of sample S8 were coated, light-blue
tablets. They were labeled “Pfizer” on one side and “VGR 100” on the reverse side. Figure 8
shows the front of a tablet from sample S8 alongside a tablet of an authentic sample. Figure 9
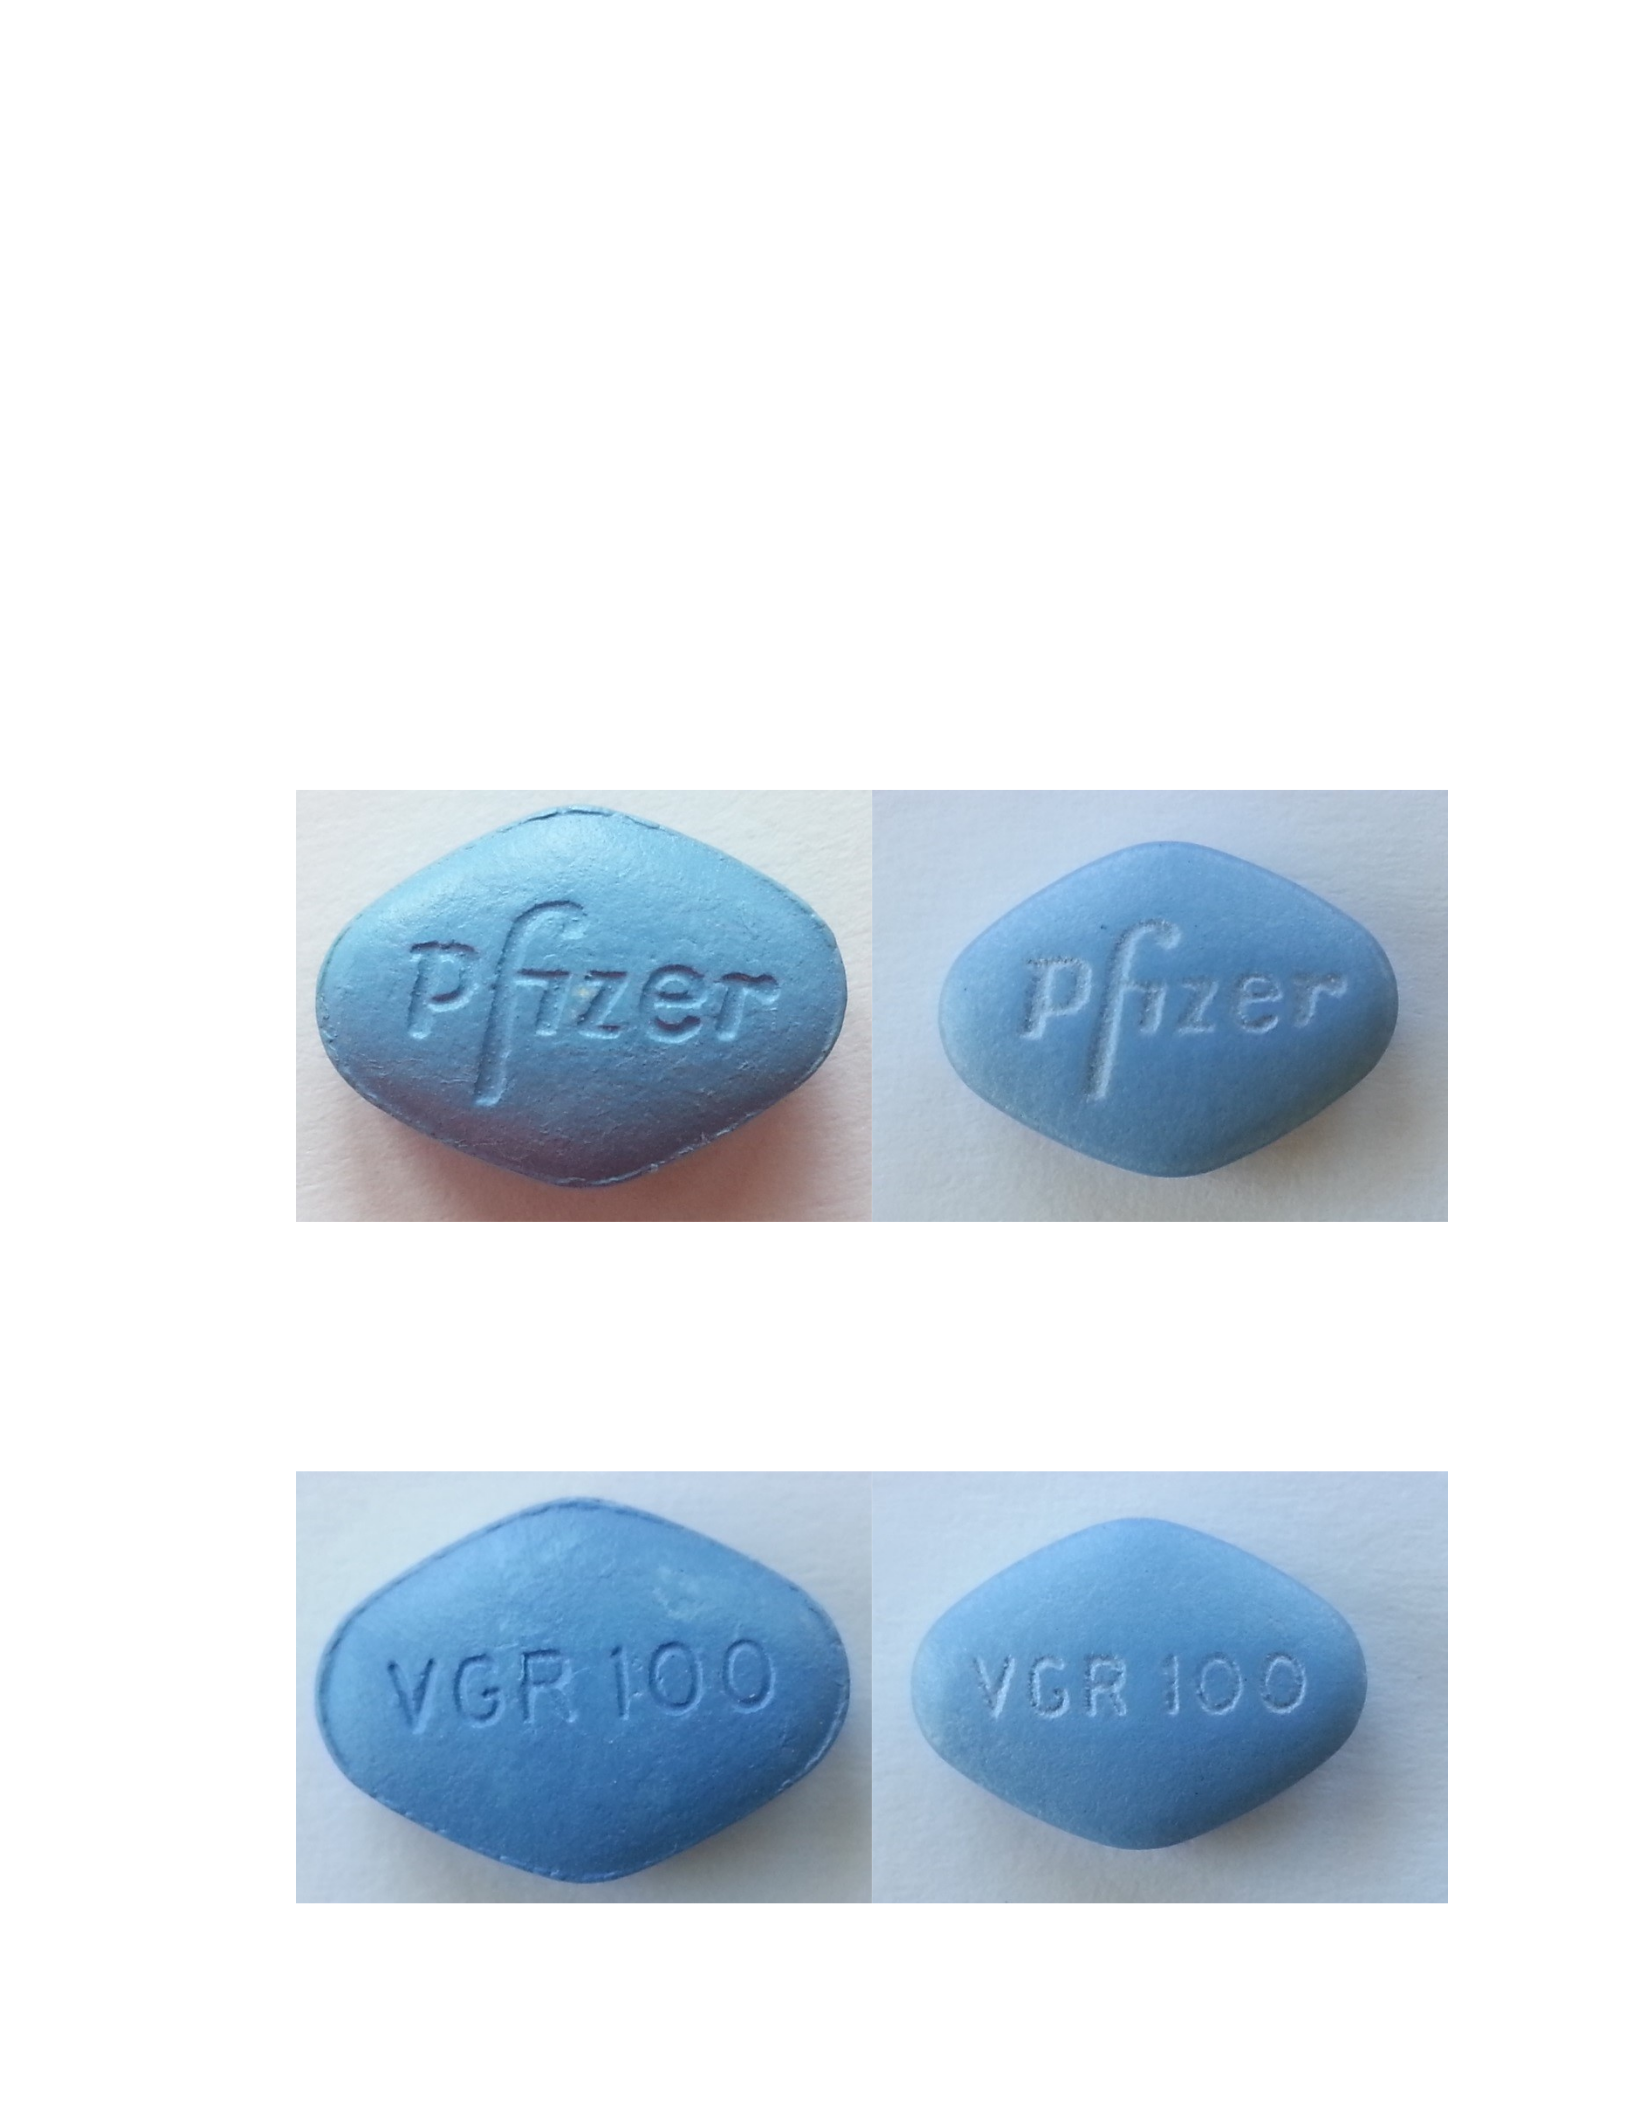
shows the back side of both tablets. Although the counterfeit sample is intended to look like an
authentic VIAGRA, the appearance of this tablet is different from that of authentic VIAGRA
when a side-by side comparison of the two samples is made. Aside from size and color, tool-
mark impressions appeared different between tablets of the authentic samples and the tablets of
sample S8. The benefit of having an authentic sample for comparison made this differentiation
based upon visual observation alone quite easy. If no authentic for comparison was available, it
would have been more difficult to identify this tablet as a counterfeit without measuring tablet
size and/or weight.
Figure 8. Front side of a counterfeit VIAGRA tablet (left)
alongside an authentic VIAGRA tablet (right). Both tablets are
displayed on the same scale.
Figure 9. Back side of a counterfeit VIAGRA tablet (left)
alongside an authentic VIAGRA tablet (right). Both tablets are
displayed on the same scale.
Table 5 summarizes the dimensional analysis for all the samples analyzed. The tablets of
sample S8 are slightly larger than the authentic samples in length and width, and weighed
approximately 100 mg more than did the authentic samples. Standard deviations were also
calculated for each measurement of each tablet and are shown in this table. The relative standard
deviation (RSD) amongst different tablets of the same samples was always less than 2% for each
sample. This shows that for a specific sample, all measured values were consistent from tablet to
tablet.
Sample
Number
Length
(mm) St Dev %RSD
Width
(mm) St Dev %RSD
Depth
(mm) St Dev %RSD
Weight
(mg) St Dev %RSD
I1 14.39 0.01521 0.11 10.42 0.01894 0.18 5.57 0.0969 1.74 609.1 7.01 1.15
I2 14.63 0.02085 0.14 10.48 0.01755 0.17 5.36 0.0398 0.74 557.9 5.37 0.96
I3 11.62 0.01357 0.12 11.85 0.02460 0.21 5.35 0.0581 1.09 528.1 4.63 0.88
I4 14.43 0.02175 0.15 10.46 0.00970 0.09 5.66 0.0251 0.44 602.4 7.67 1.27
I5 14.52 0.01538 0.11 10.50 0.00720 0.07 5.10 0.0316 0.62 642.3 4.64 0.72
S13
(authentic)
14.77 0.02722 0.18 10.65 0.07120 0.67 5.77 0.0160 0.28 633.0 10.59 1.67
S14
(authentic)
14.70 0.04787 0.33 10.57 0.02722 0.26 5.79 0.0397 0.69 633.8 9.36 1.48
S15
(authentic)
14.75 0.02331 0.16 10.62 0.00694 0.07 5.77 0.0073
9
0.13 618.7 12.08 1.95
S16
(authentic)
14.71 0.01644 0.11 10.61 0.01262 0.12 5.78 0.0083
9
0.15 615.7 5.36 0.87
S7 14.77 0.00839 0.06 10.54 0.01836 0.17 4.82 0.0735 1.52 502.3 7.31 1.45
S8 14.92 0.00839 0.06 10.78 0.01333 0.12 5.68 0.0301 0.53 733.6 5.15 0.70
SING1 11.69 0.01638 0.14 11.92 0.01835 0.15 5.47 0.0507 0.93 541.1 4.49 0.83
SING2 11.72 0.02193 0.19 11.95 0.03173 0.27 5.46 0.0305 0.56 533.2 5.97 1.12
Table 5. Summary of dimensional measurements of samples
analyzed.
Although it was possible to distinguish the non-authentic samples from the authentic
samples, analysis of variance (ANOVA) was applied to the data to determine whether the
variance could be used to differentiate non-authentic samples that were very similar in
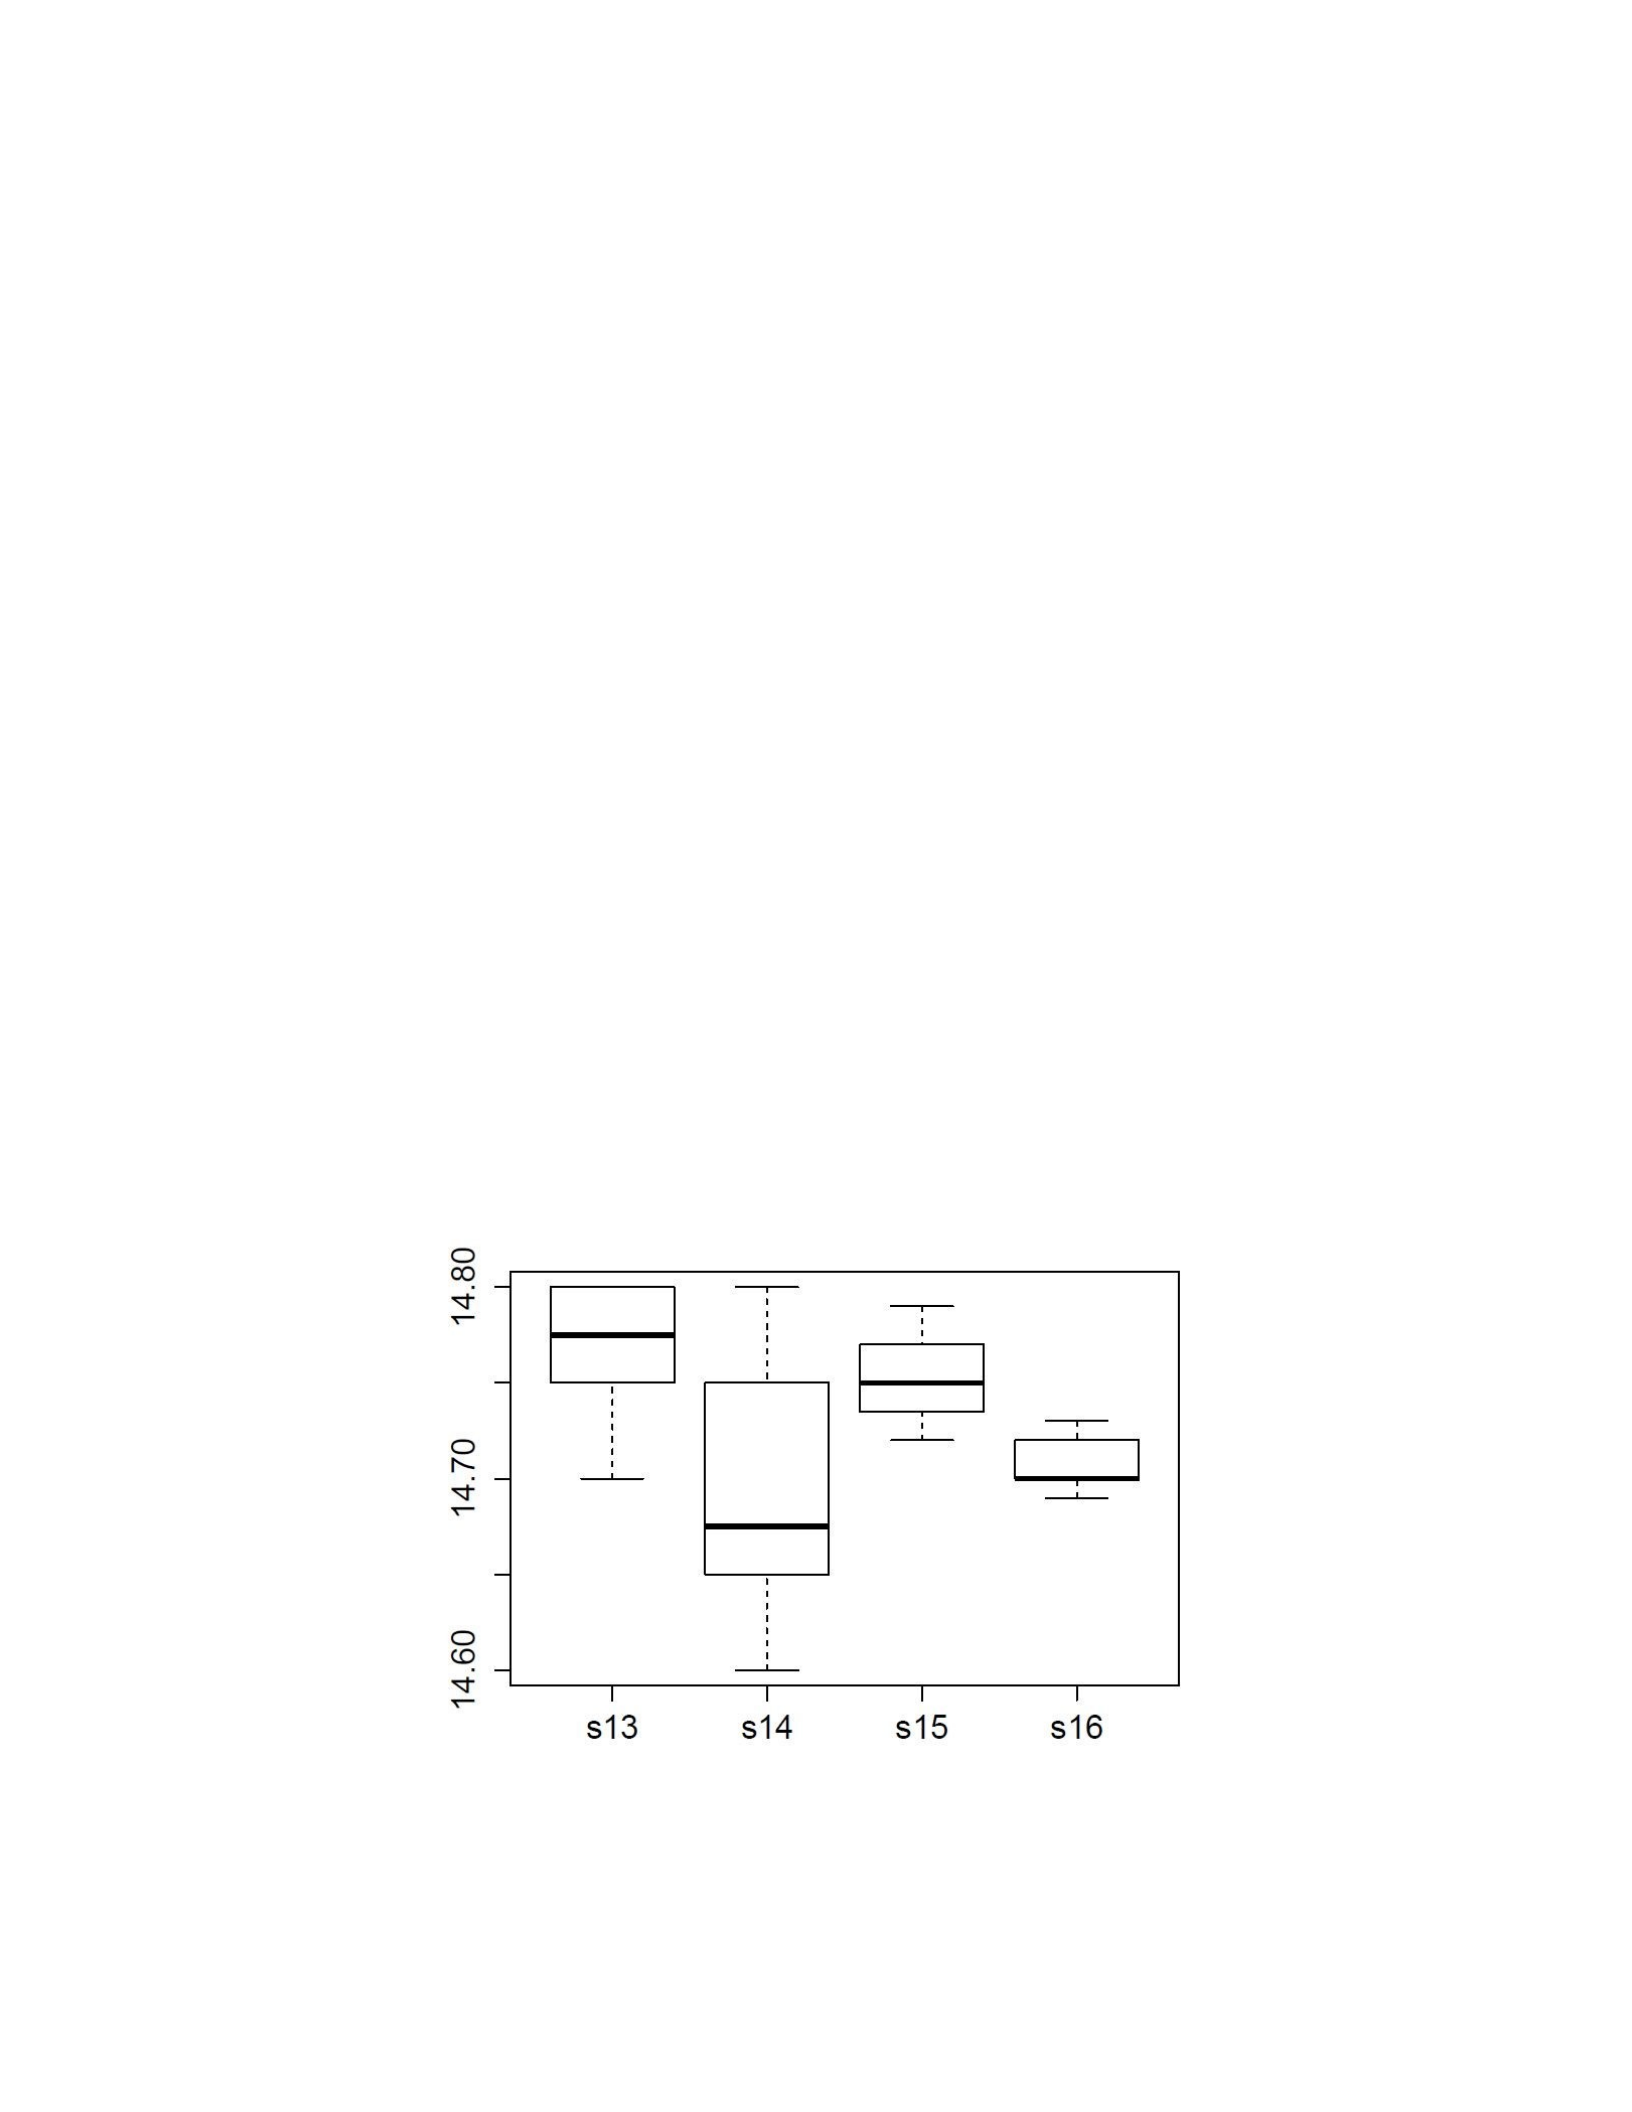
appearance to each other. For example, samples I1 and I4 were very similar in dimensional
measurements and physical appearance to each other. Using the physical measurements of
length, width, depth, and weight, t-tests indicate that there is a statistically significant difference
between I1 and I4 at the 5% level of significance. However, using the physical measurements of
length, width, depth, and weight, ANOVA analysis of the four authentic samples indicates there
is also a statistically significant difference even amongst these authentic samples. This variance
in these four authentic samples indicates t-tests and ANOVA analysis of the
physicalmeasurements data collected from these samples alone is not an appropriate method of
discrimination.
The box plots generated from ANOVA analysis of the length measurements of authentic
samples is shown in figure 10. The box plots generated from the t-test analysis of the length
measurements of I1 and I4 re shown in figure 11. The box plots highlight the median value for
each sample. Although only the results from the length measurements are shown, the same
general amount of variance was observed for the measurements of width, depth, and weight.
Figure 10. Box plots generated from length measurements of the
authentic samples.
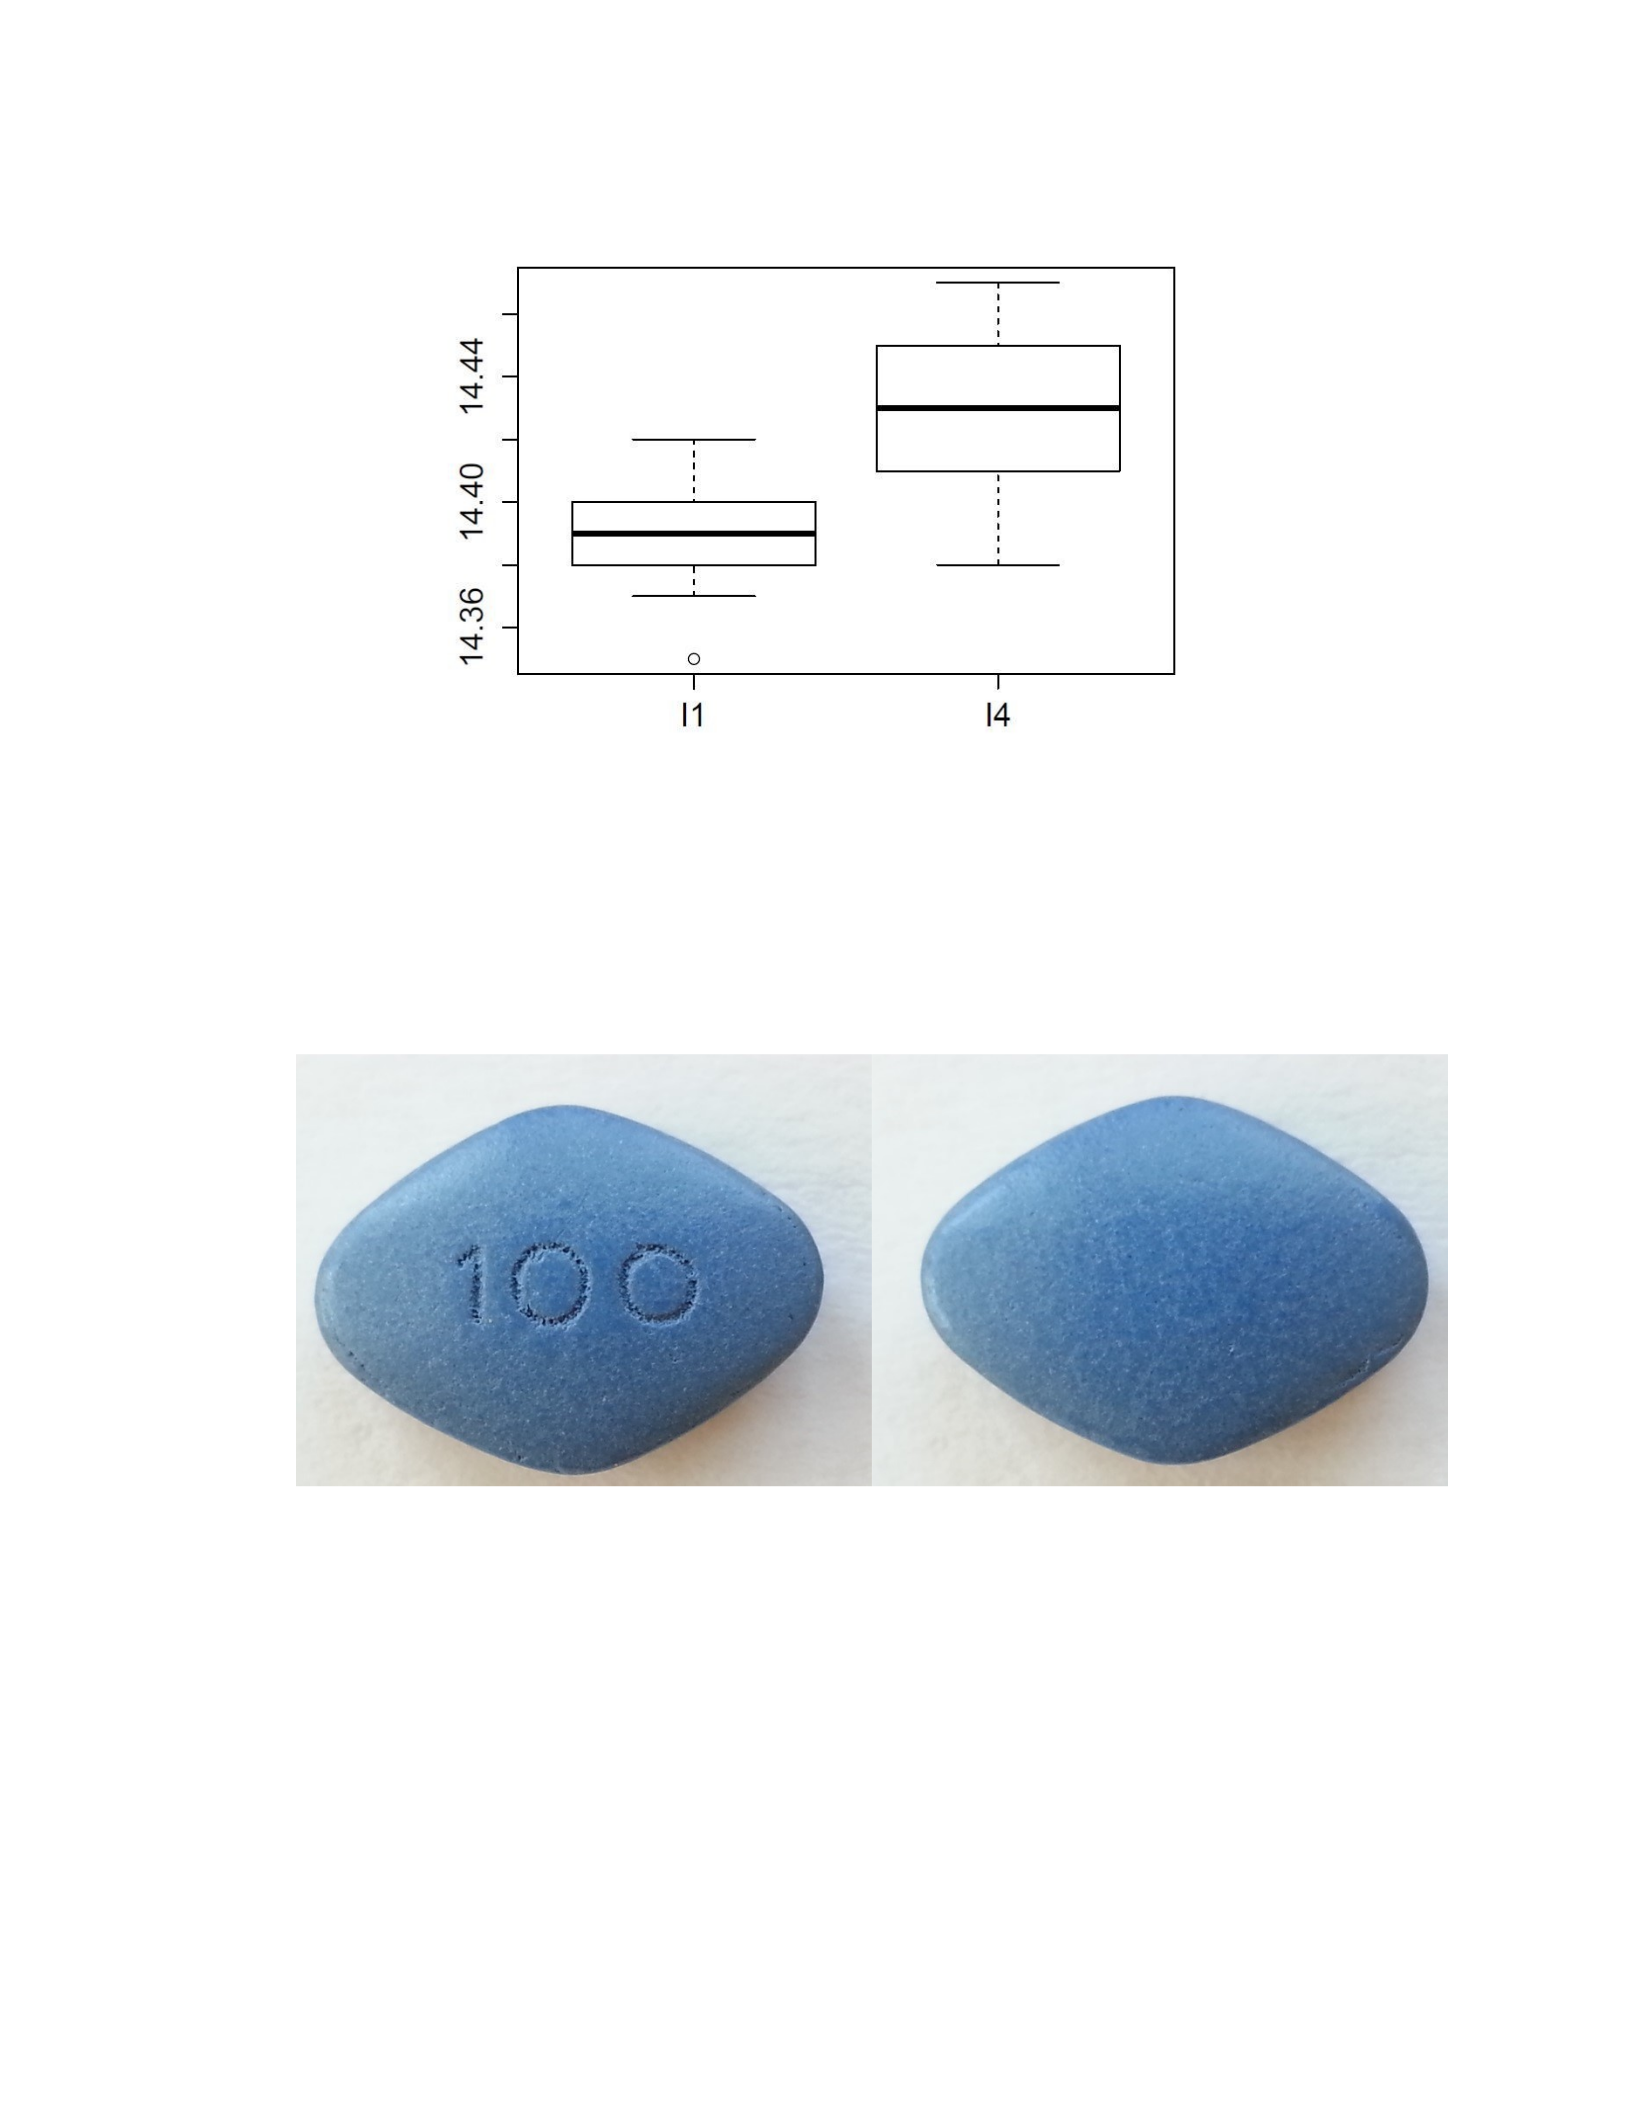
Figure 11. Box plots generated from length measurements of
samples I1 and I4.
Figures 12 through 19 show the front and back sides of tablets of each of the other eight
non-authentic samples.
Figure 12. Images of the front and back of a tablet of I1.
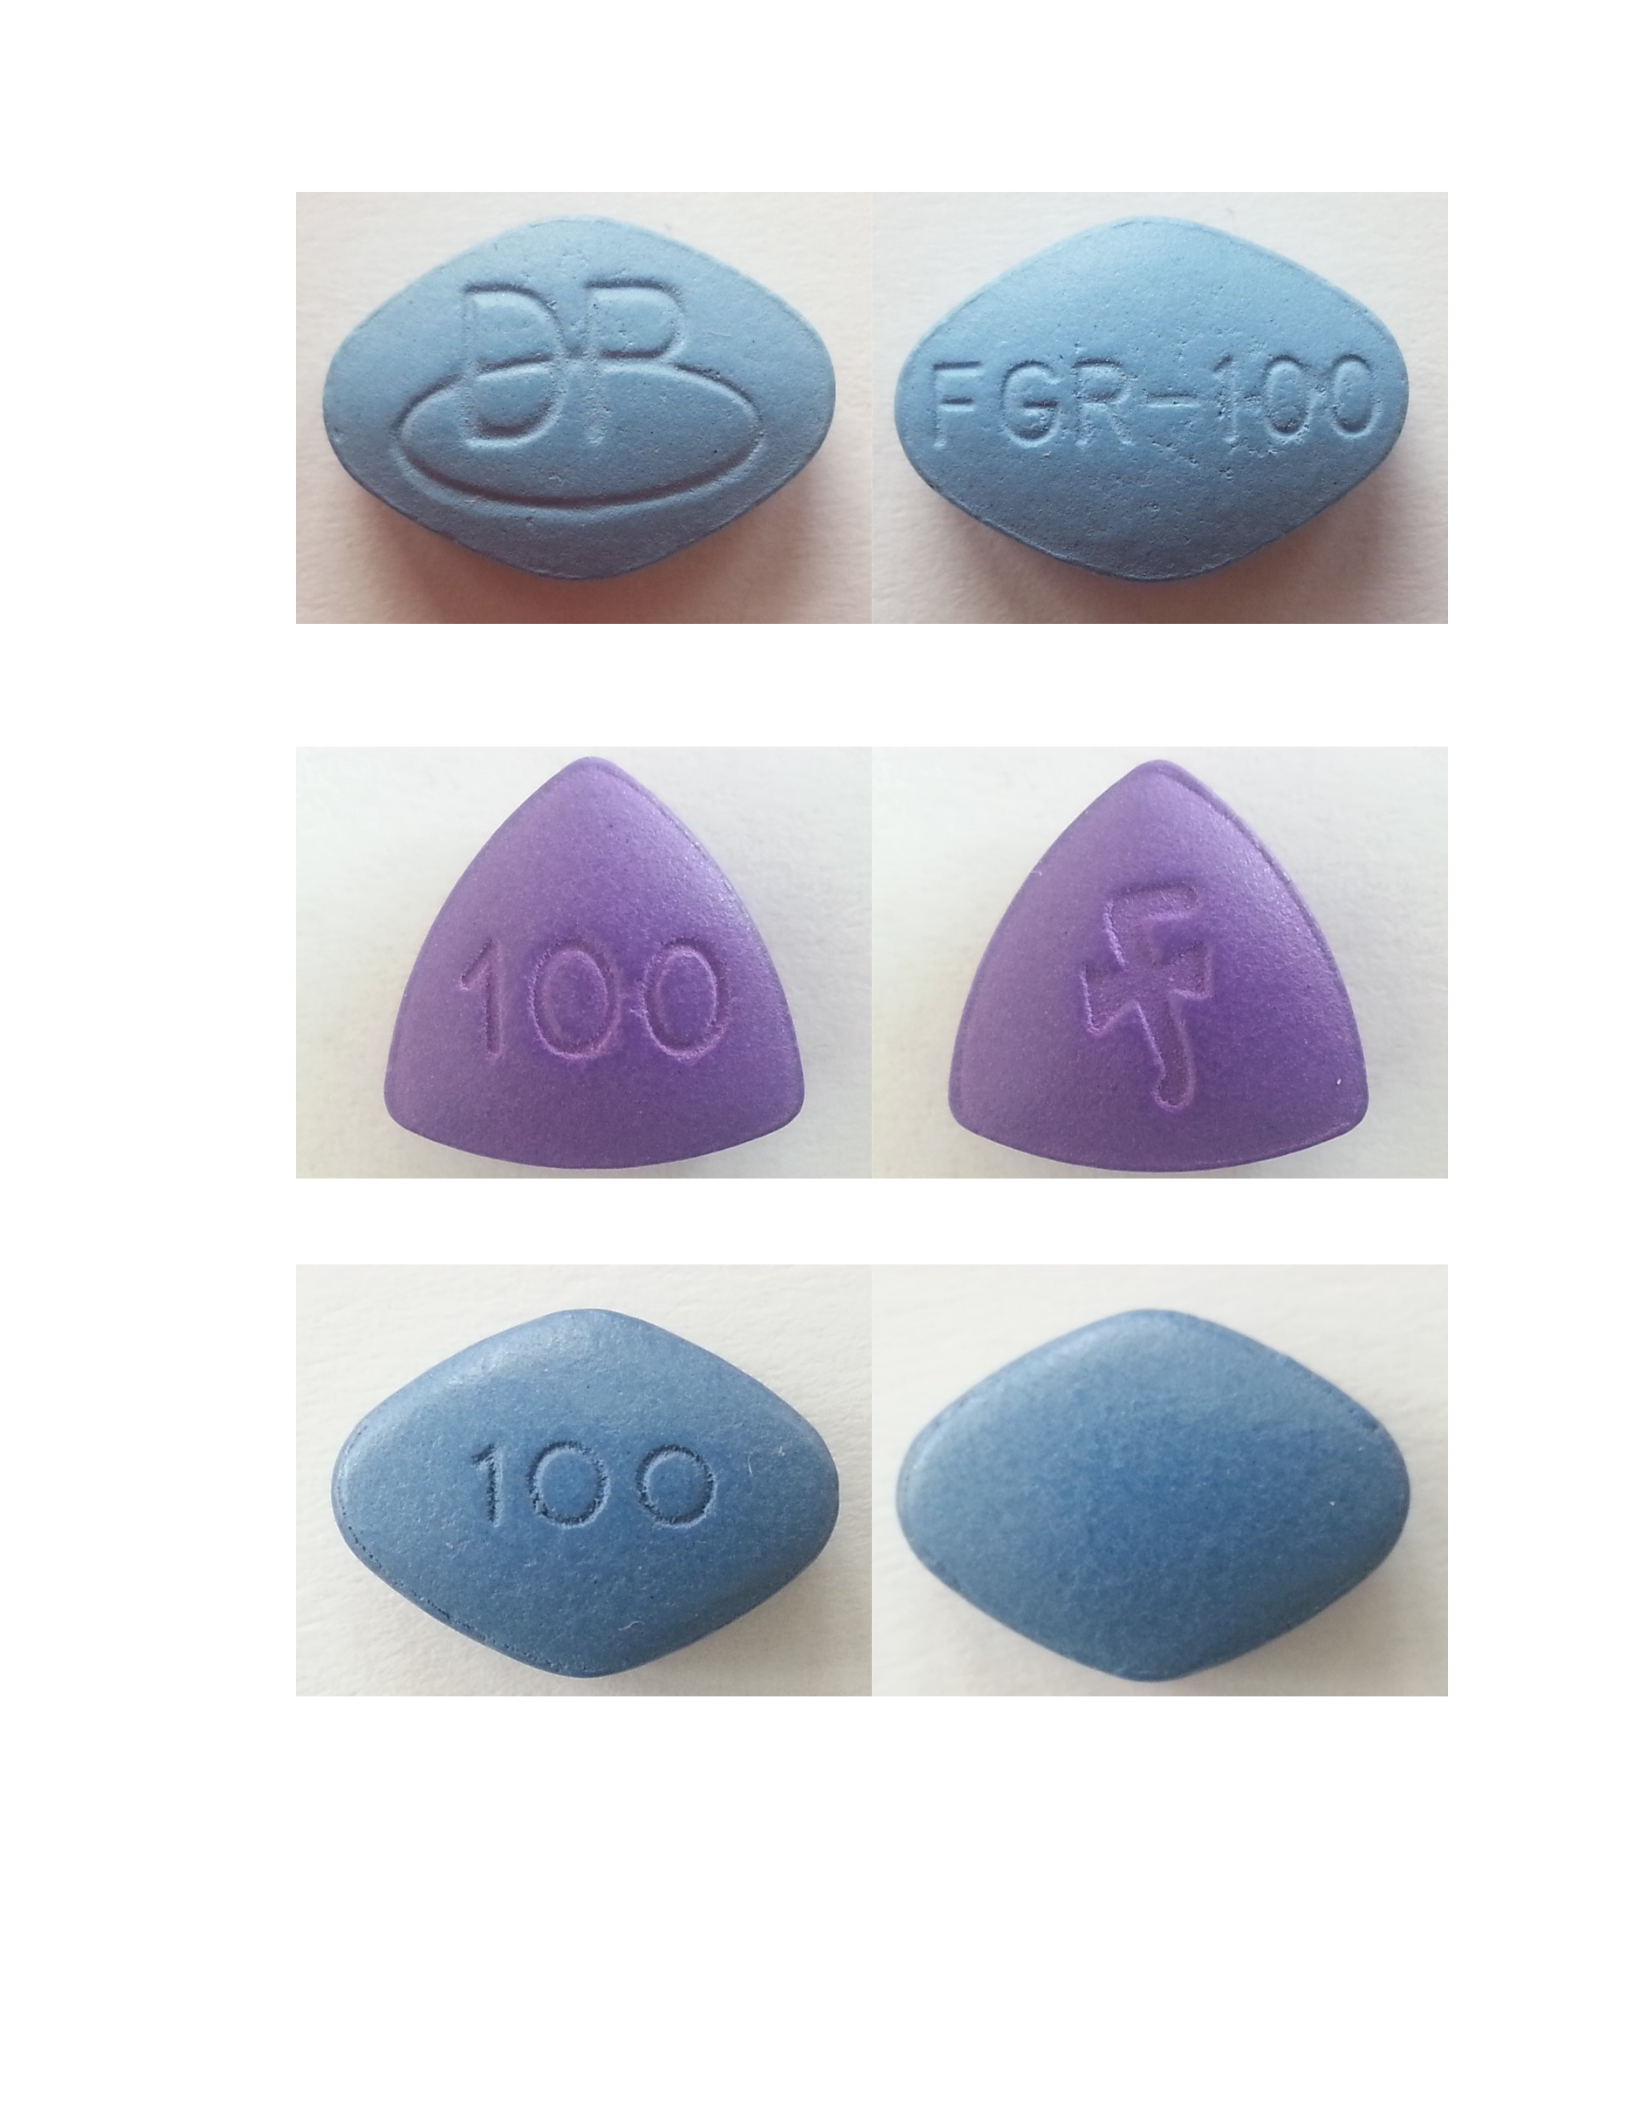
Figure 13. Images of the front and back of a tablet of I2.
Figure 14. Images of the front and back of a tablet of I3.
Figure 15. Images of the front and back of a tablet from I4.
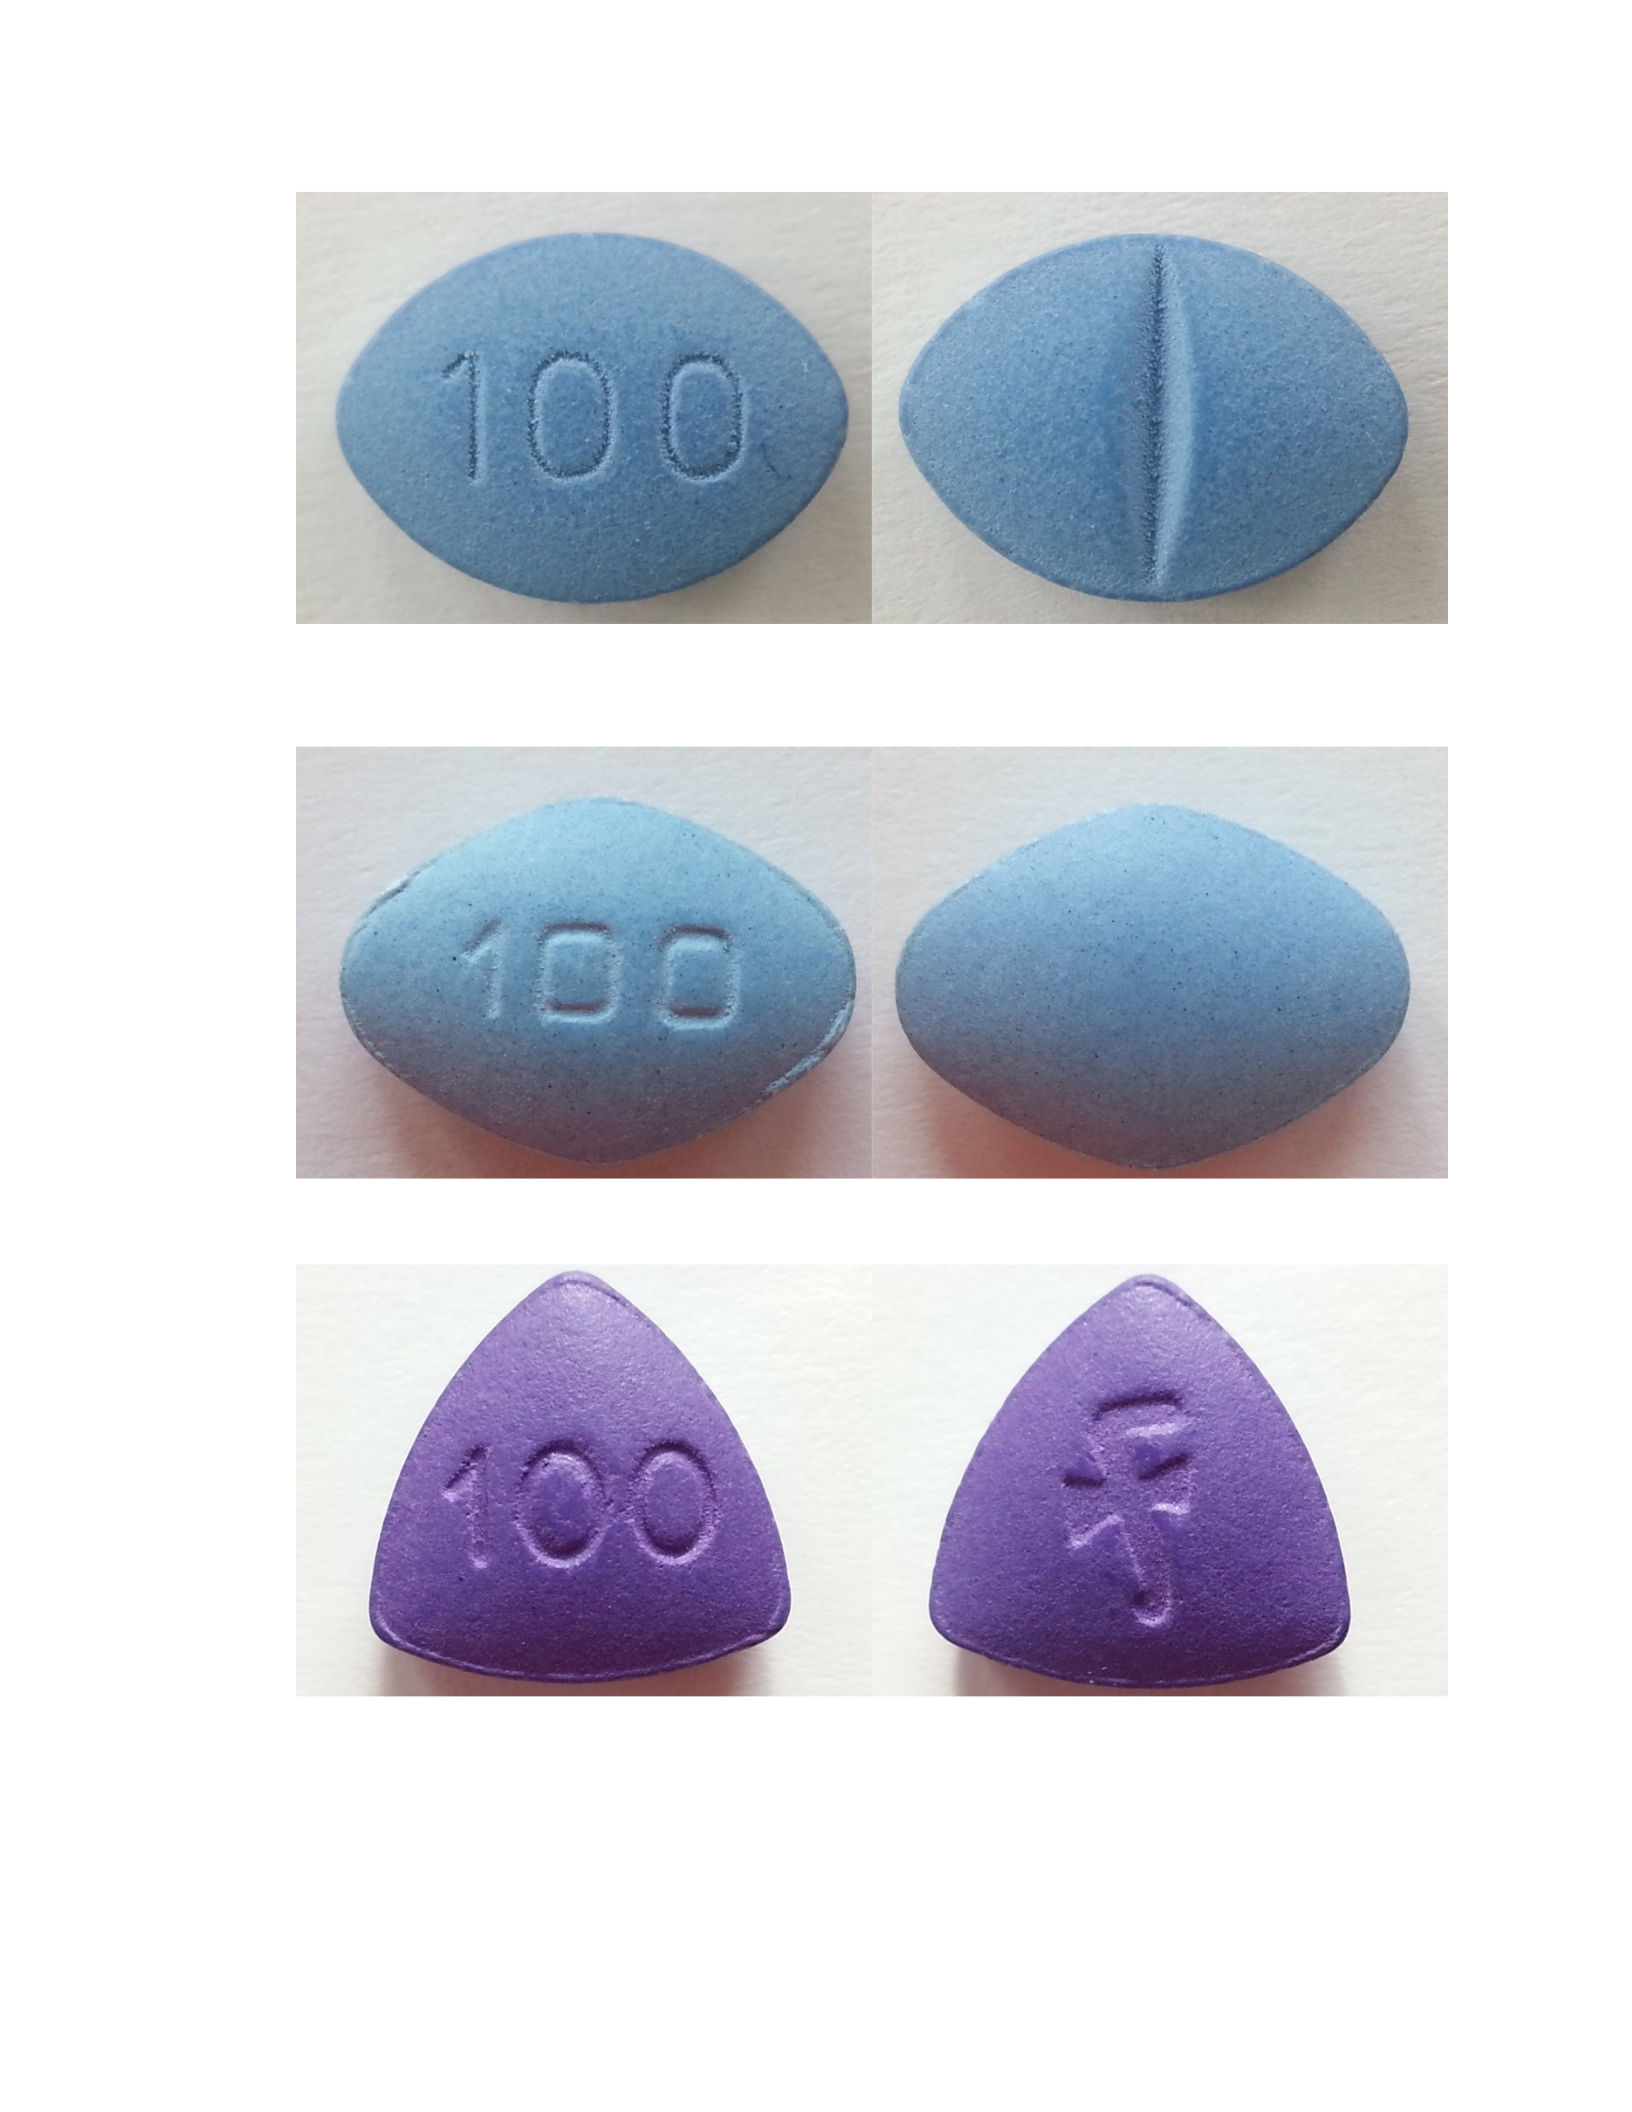
Figure 16. Images of the front and back of a tablet from I5.
Figure 17. Images of the front and back of a tablet from S7.
Figure 18. Images of the front and back of a tablet from SING1.
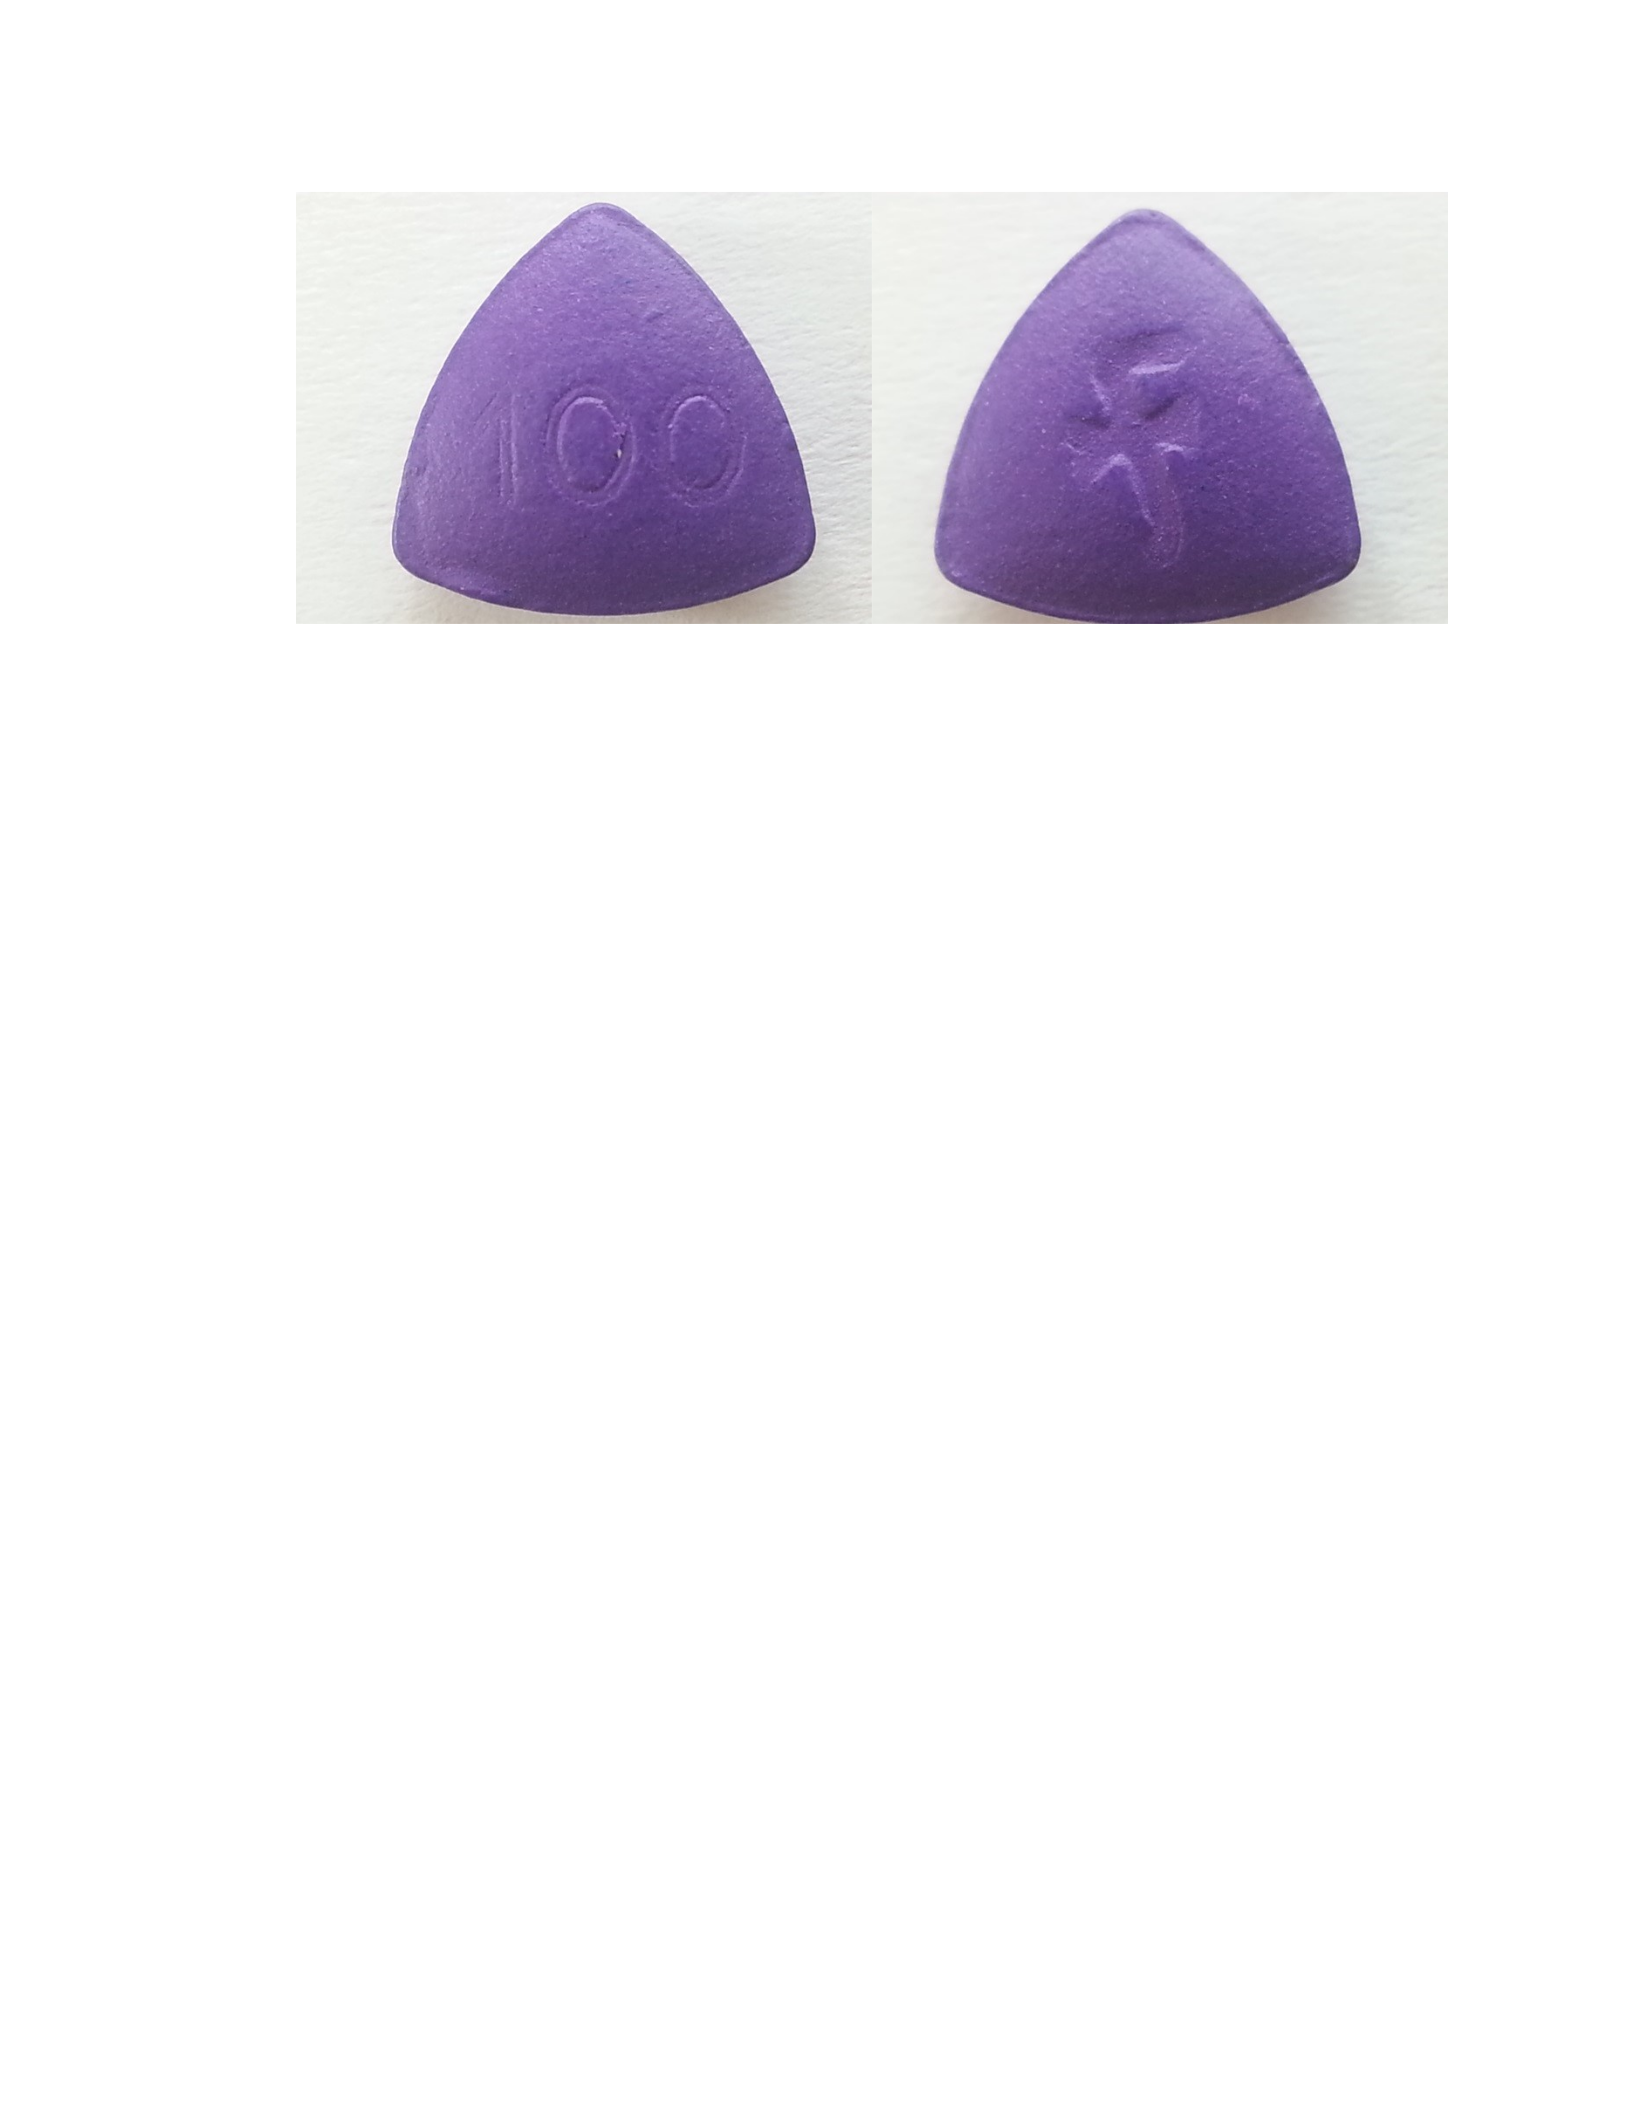
Figure 19. Images of the front and back of a tablet from SING2.
Summary and Conclusion
This research shows that detection and identification of all of the non-authentic samples
was possible based upon visual observation. In addition, tablet weights and other dimensional
measurements could be used to differentiate the non-authentic samples from the authentic
versions. These measurements were consistent for different tablets of the same sample. For all
tablets within a sample, the percent RSD for all measurements of length, width, depth, and
weight were less than 0.34, 0.68, 1.75 and 1.96, respectively. T-tests and ANOVA analysis of the
physical-measurements data collected from these samples alone is not an appropriate method of
discrimination. The within-group variance of these measurements is equal to or greater than
the between-group variance.

CHAPTER 8. INVESTIGATION AND EVALUATION OF CHEMICAL AND PHYSICAL
PROPERTIES OF FINAL DOSAGE FORMS USING INFRARED SPECTROSCOPY
Introduction
Pharmaceutical tablets are typically comprised of active and inactive ingredients blended
together using specific manufacturing processes. Specific manufacturing processes are used
because different processes may alter these ingredients, causing changes in both physical and
chemical form. An example of a change in physical form may be the conversion from one
solidstate form of the drug to another; an example of a change in chemical form may be the
formation of degradation products. Both the active (and inactive) ingredients may undergo
changes in solid-state form. An example of this type of transition is the conversion of a drug
substance from a crystalline form to an amorphous form.
Drugs may also sorb water or solvent to form hydrates or solvates. Some water is sorbed
interstitially. This water does not become part of the crystal structure of the drug or incorporate
itself in the drug chemically. Water may also be sorbed resulting in differences in crystal
structure between two hydrated forms of a drug. In these crystalline hydrates, water is part of the
drugs crystal lattice. These hydrates and solvates may or may not desorb during the final
manufacturing steps.
Ingredients may also physically or chemically react with each other during tablet
manufacture or storage. An example of a physical interaction is the formation of a eutectic. An
example of a chemical interaction would be the formation of a degradation product. These
interactions may result in tablets that exhibit differences in therapeutic effects, storage stability,
and physical or chemical composition.

When manufacturing processes are well controlled and adhere to regulatory guidelines as
is typical for an authentic product sold in the United States, the physical and chemical properties
of the final dosage form is well controlled and reproducible. The ability to detect and identify
the physical and chemical form of the final dosage form in a reliable manner is useful, providing
a compositional fingerprint that is representative of the authentic product and may be used to
differentiate authentic from non-authentic tablets. This fingerprint may also be used to classify
counterfeit drugs, providing investigative leads useful in establishing source or origin.
Infrared spectroscopy is used to evaluate the chemical and physical form of tablets and
other drug products. Data can be evaluated many different ways when applied to the analytical
challenges associated with counterfeit drugs. In this study, infrared spectra were compared to
differentiate authentic from non-authentic samples. The reproducibility of spectra collected from
different tablets of a sample was also evaluated. Spectra were then evaluated to establish
differences in the relative concentration of the API. Finally, chemometrics analysis, specifically
PCA-CVA with HOO-CV, was performed to classify samples and determine an estimated error
rate for this classification. Spectra used in this comparison were collected from the core matrix
of the tablet.
Experimental Details
Instrumentation
Infrared spectra were collected using a HazMatID™ Elite infrared spectrometer (Smiths
Detection, Danbury, CT). Performance of the spectrometer was tested at the start and end of
every work day. The system always passed performance-test criteria. The performance test
method used was the method recommended by the instrument manufacturer. This test requires
that the performance standard meets a minimum library match factor when compared with a
reference standard spectrum.
The HazMatID Elite is specifically intended for field use. Some of the specifications for
this system, especially those intended to support portability and ruggedization of the instrument,
are shown in Table 6. An image of the system is shown in figure 20.
HazMatID Elite Specifications for Portability and Ruggedization
Compact size 10 5/8 in x 5 5/8 in x 3 1/8 in)
Weight 2.29 kg
Operating Environmental
Temperature
-20°C to 50°C
Operating Environmental Humidity 0-100%
Decontamination IP-67 rated and sealed for decontamination by
immersion
User interface 4.3 inches internally bonded LCD color display for high
visibility in direct sunlight conditions; individually lit
keypad and instructional graphics guide users through
the operation of the device
Power Rechargeable lithium-ion battery for 4 hours of
operation;
compatible with disposable 123A battery
Repeater RF repeater for extending the wireless data transfer
range
Colors Olive drab or yellow
Wireless Embedded RF modem for 1 km line-of-sight data
transfer and command control; FIPS 140-2
compliant encryption; GPS included
Table 6. HazMatID Elite specifications for portability and
ruggedization of importance to this study.
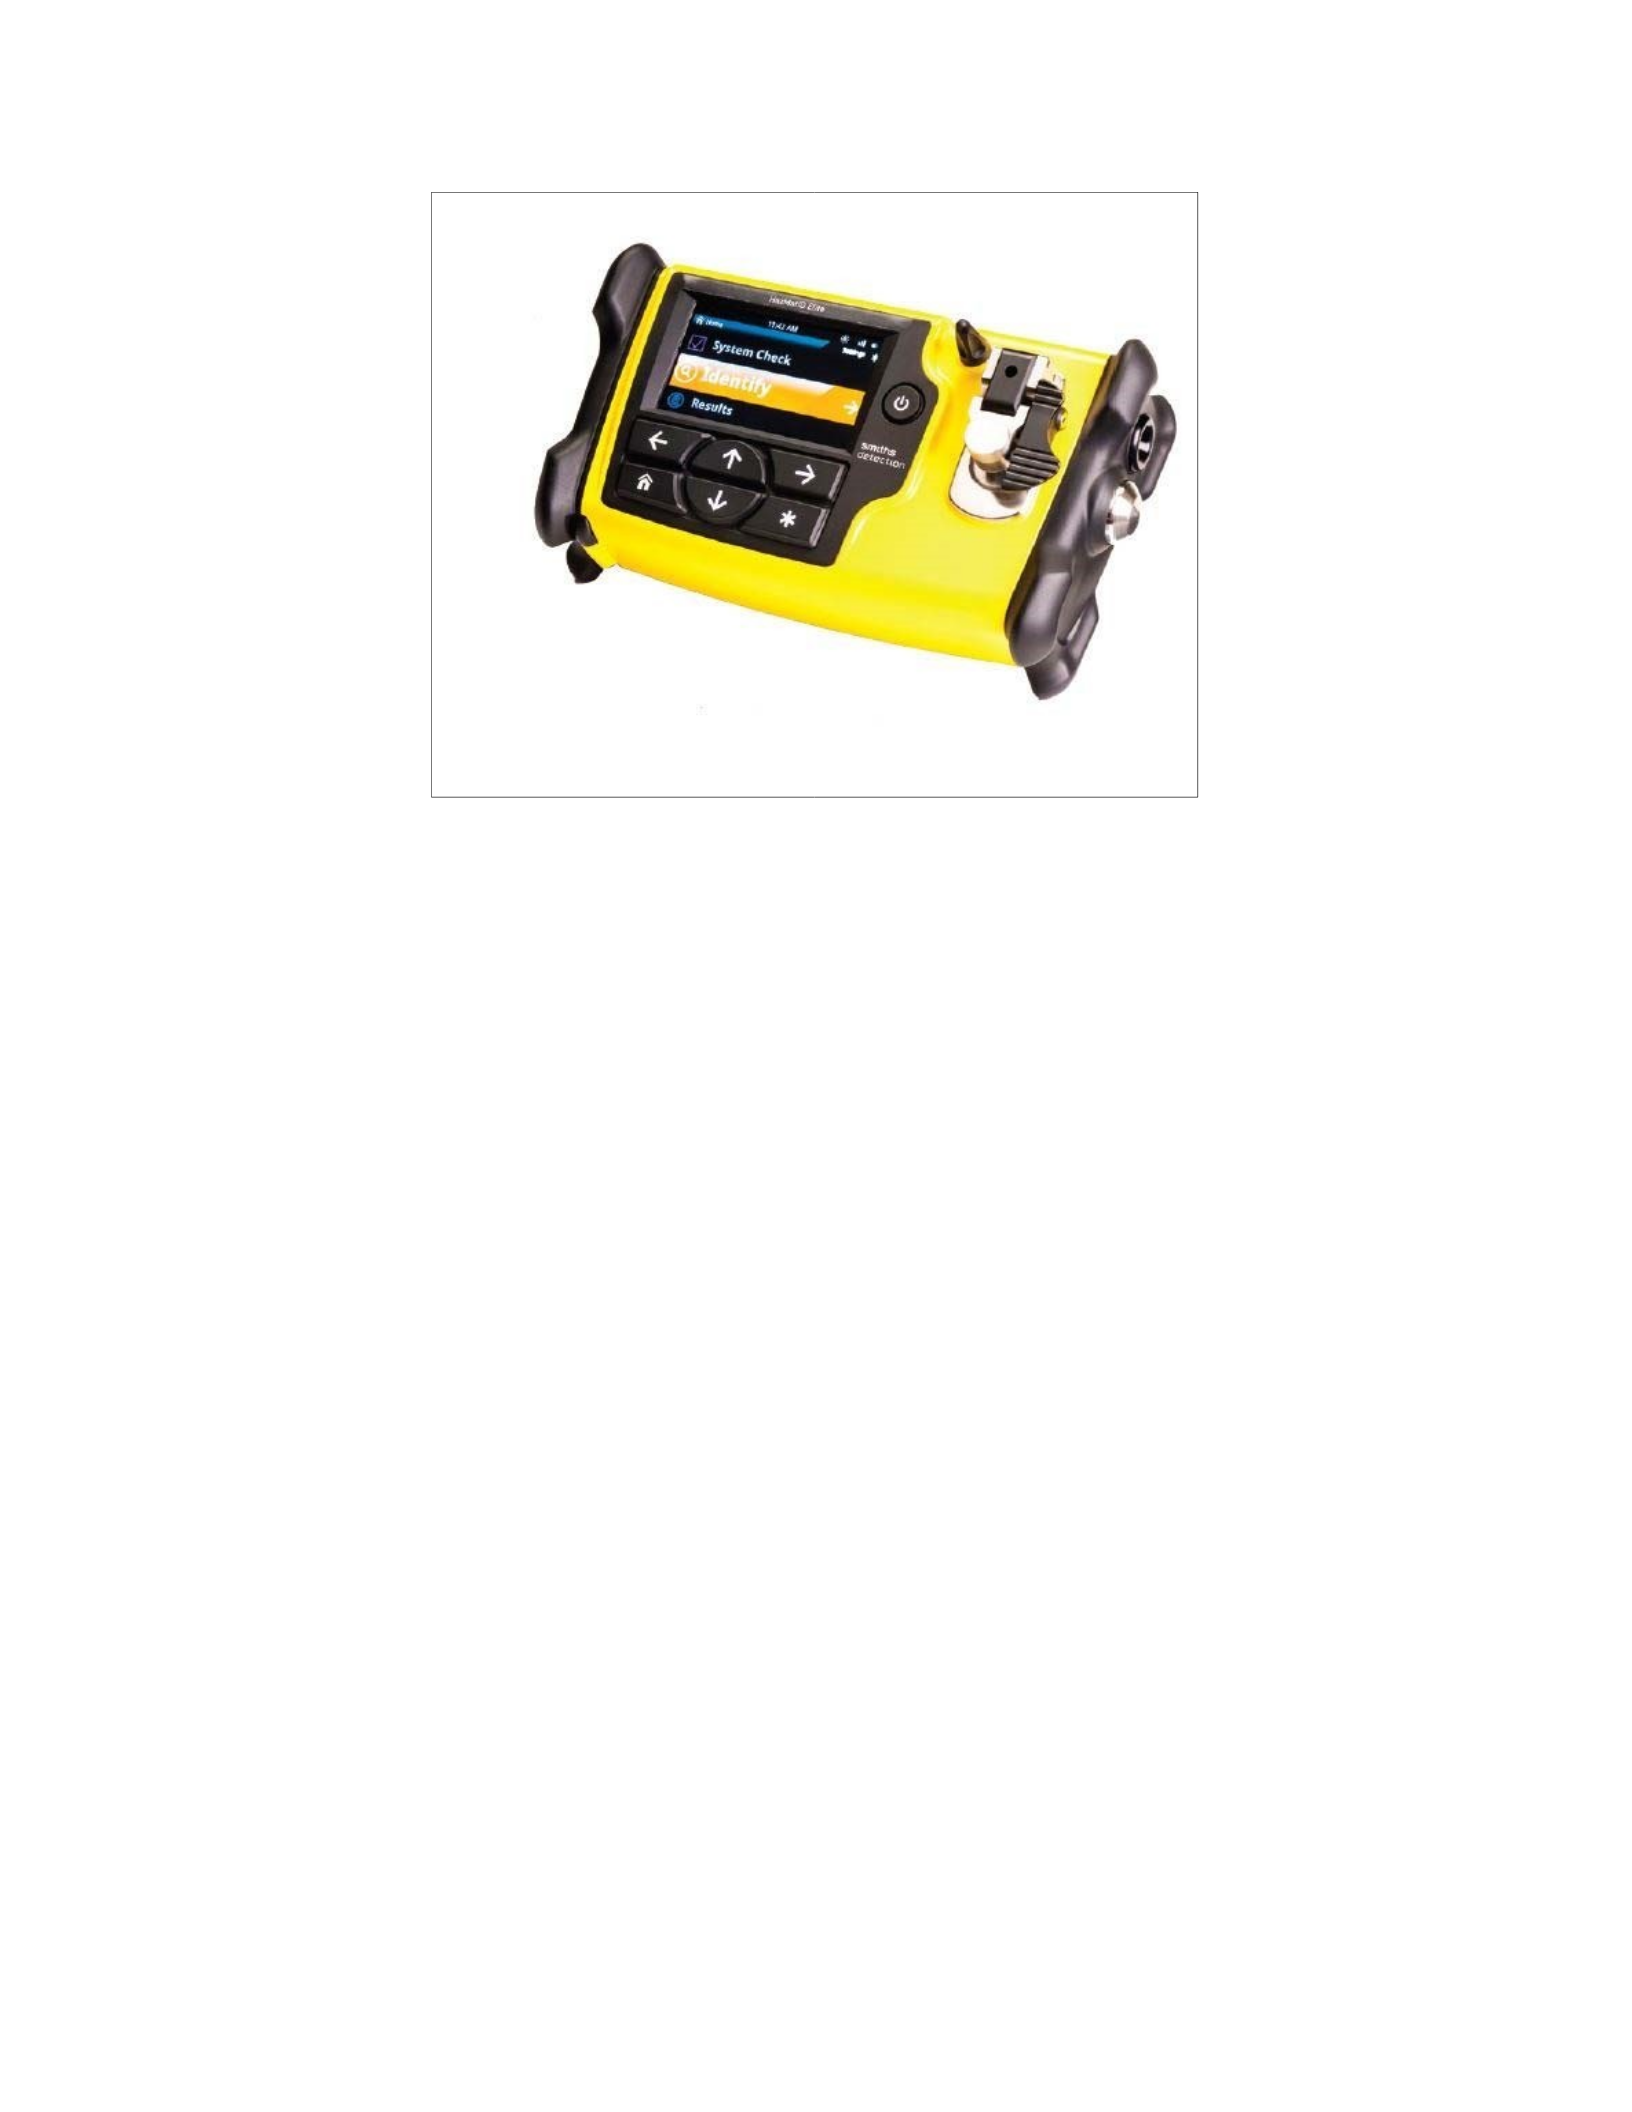
Figure 20. Photograph of HazMatID Elite diamond ATR FT-IR
spectrometer. Courtesy of Smiths Detection, Danbury, CT.
Infrared Method
Prior to each sample analysis, the diamond IRE was cleaned using a low-lint tissue and, if
necessary, with an appropriate solvent. A background spectrum was collected using the default
method for the instrument set for 8-cm-1 resolution and 32 co-added scans. Each tablet was
gently crushed and a small amount of the tablet’s core was pressed against the diamond area until
the instrument screen displayed in green. In order for this green readout to appear, the
absorbance signal level for the spectrum’s largest absorption band must reach a value of at least
0.11. A sample spectrum was then collected. The infrared spectrum was automatically
calculated and displayed on the screen of the instrument. For each tablet, three infrared spectra
from different samples of each tablet’s core were collected.

Data Analysis and Chemometrics
All spectra were then exported from the HazMatID Elite via a USB storage hub and
transferred to a personal computer. Each spectrum was opened using the Grams Suite 9.0
software (Thermo Fisher Scientific, Philadelphia, PA). Using Grams, a linear multipoint baseline
correction was performed and corrected to account for diamond uncompensation in the spectral
region between 2350 cm-1 and 1850 cm-1. Using the Grams Add-On Excel Exchange function,
spectra were exported to Microsoft Excel (Microsoft Excel 2010 running on Windows 7
Enterprise, Service Pak 1, 64-bit Operating System). Within Microsoft Excel, each spectrum was
normalized to an area under the curve equal to one. The normalized spectrum was then imported
to Grams using the Grams Add-On Excel Exchange.
The chemometric methods of PCA and CVA were applied to investigate the classification
and discrimination power of infrared spectroscopy. Prior to chemometric analysis, the spectral
region between 2350 cm-1and 1850 cm-1 was deleted from all spectra. Non-authentic samples
were sorted into seven groups based upon physical appearance, dimensions, and receipt date;
authentic samples were categorized into a single group. The deconvolution data, specifically the
peak positions and areas, also were analyzed using HOO-CV PCA-CVA to determine an
estimated error rate for the classification of sildenafil citrate tablets.
Results and Discussion
It was possible to differentiate the authentic samples from all non-authentic samples using
this method. In most cases, differences in spectra were small but reproducible. Using PCA-
CVA, 3D-scores plots were created and visual separation of samples was observed. This
observation indicates that not only can infrared spectroscopy be used to differentiate authentic
from non-authentic samples, but also that this method may be used to sort non-authentic samples
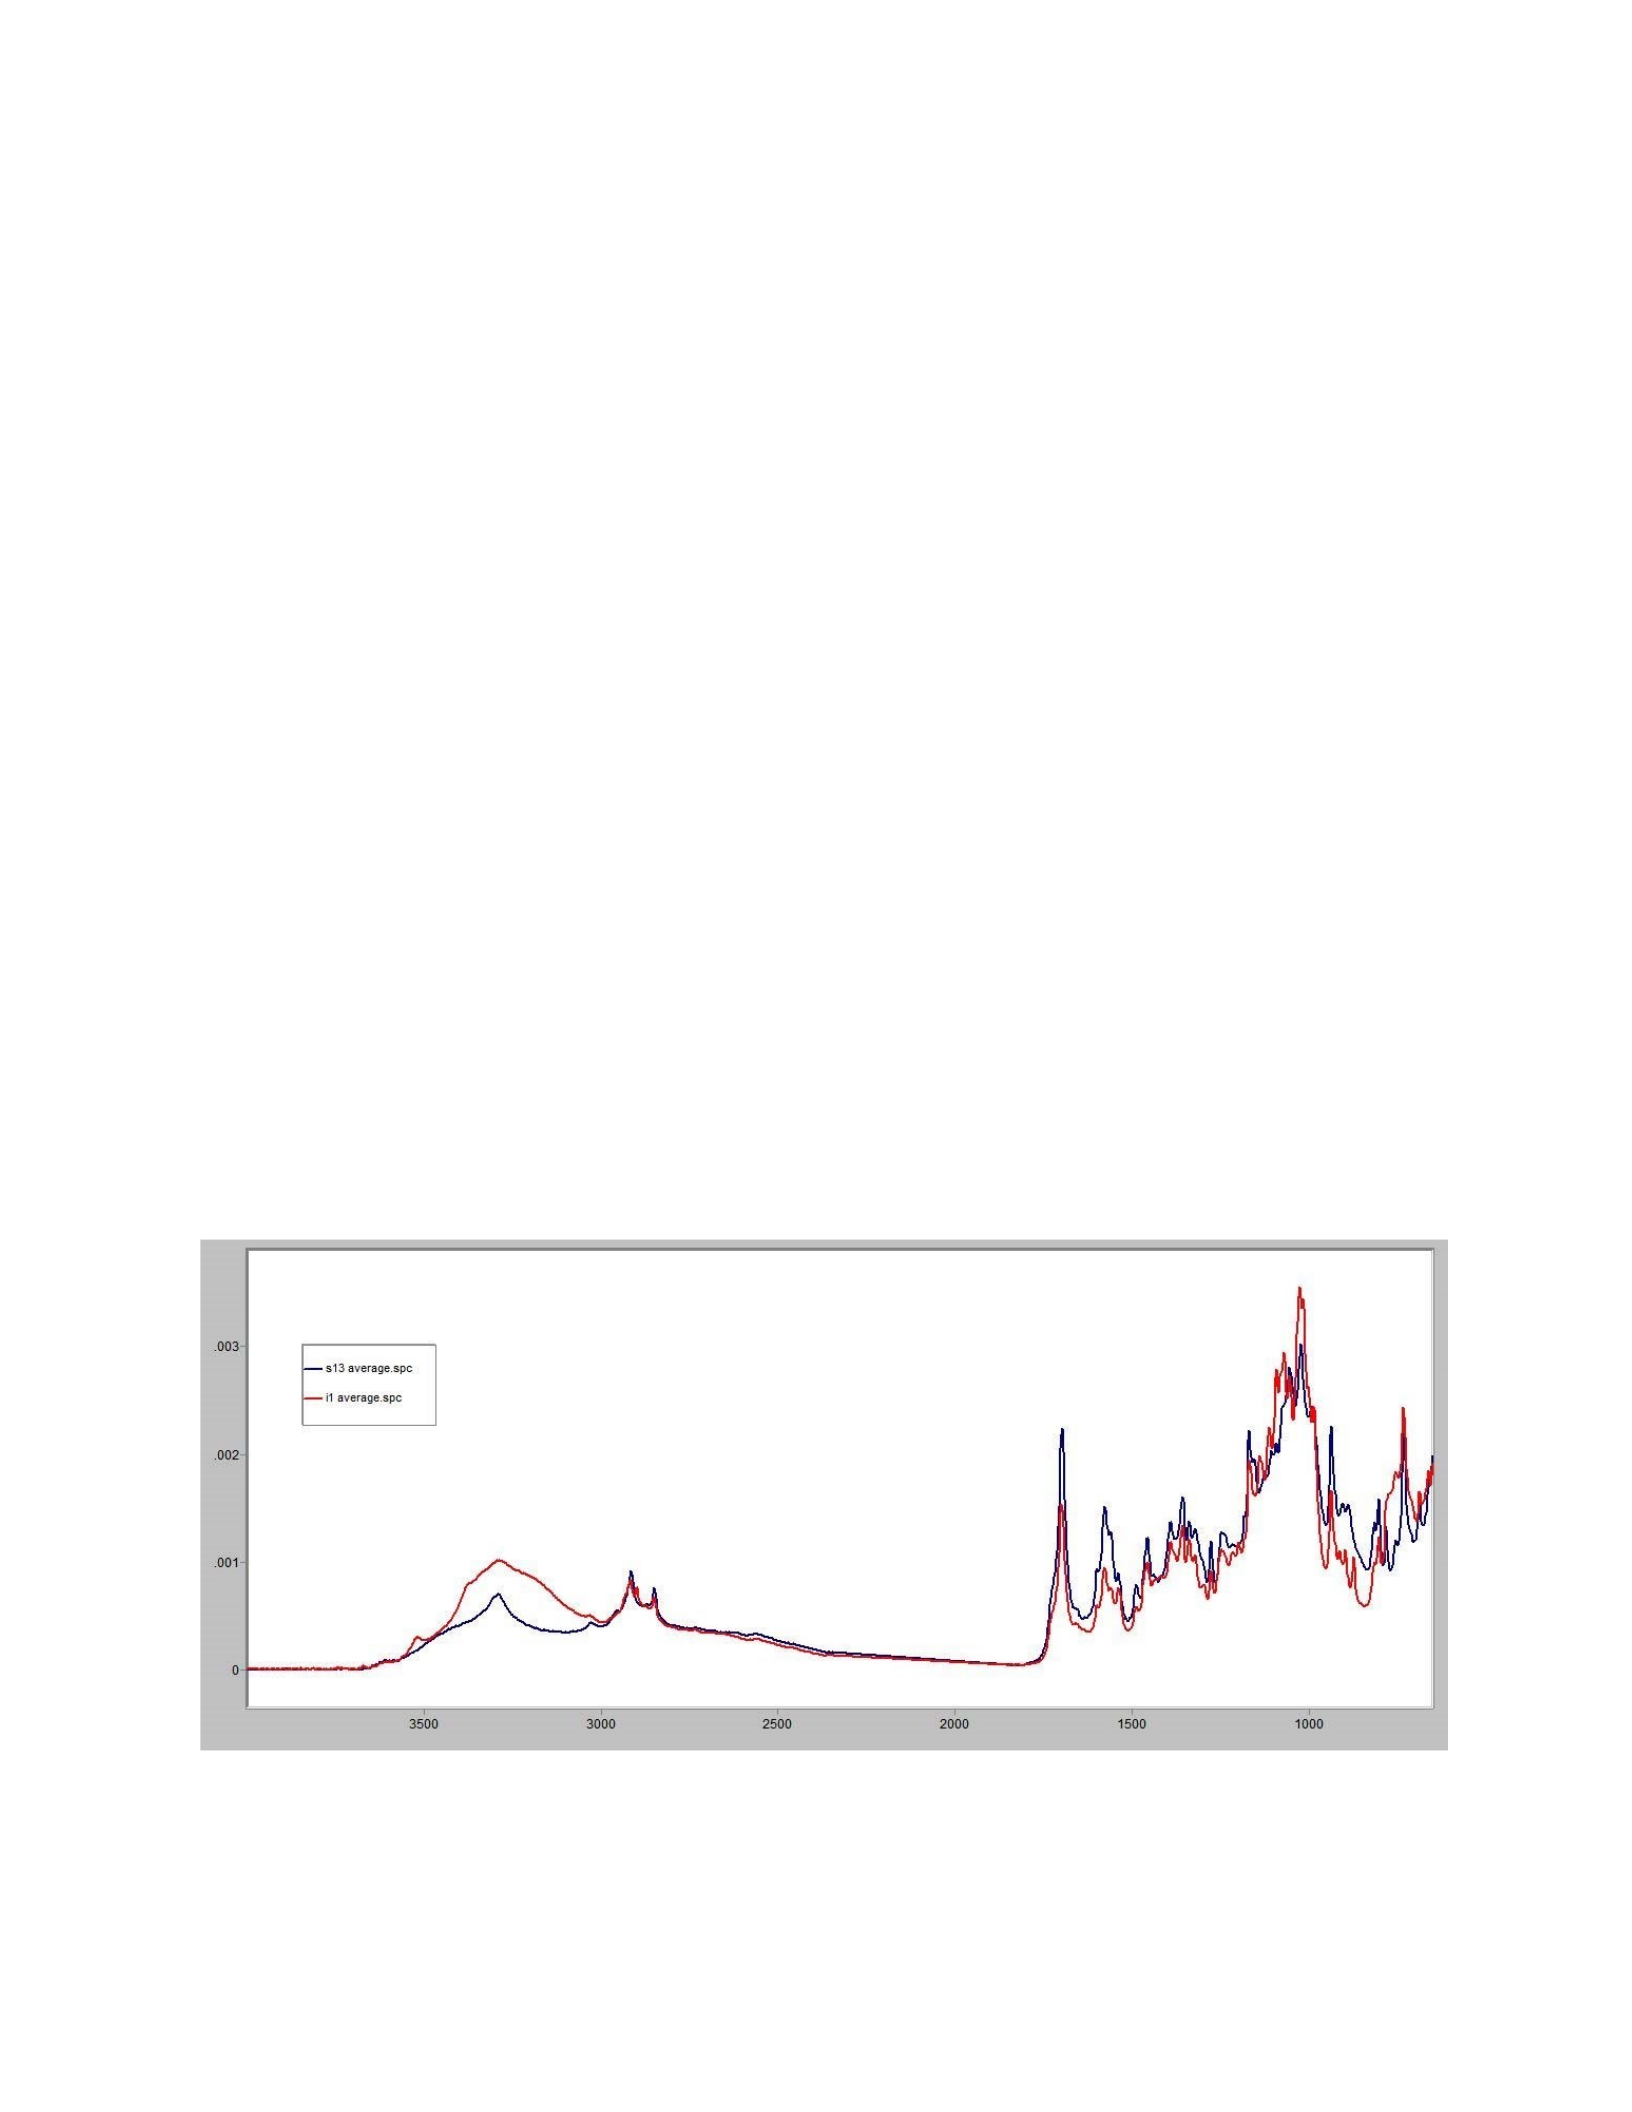
and ultimately lead to the establishment of provenance. Using HOO-CV, an estimated error rate
of 2.16% was calculated using 12 principal component’s (PC’s). The error was due to the
misclassification of samples I1 and I4. A review of the infrared spectra shows that this
misclassification is likely due to differing amounts of water in the samples. Otherwise, the
spectra were indistinguishable. It is likely based on all other data collected that these two lots
share a common origin.
Figures 21 through 29 show the average infrared spectrum of an authentic sample
compared with the average spectrum from each of the nine non-authentic samples. Note that
with the exception of sample S8, differences in the fingerprint region of the spectra of the
nonauthentic samples were small indicating that the chemical composition of the non-authentic
samples was similar, but not identical to, the chemical composition of the authentic samples.
Therefore, it was possible to differentiate all non-authentic samples from the authentic ones.
Sample S8 contained an additional broad absorption band of high intensity that is likely due to
the presence of calcium sulfate in the sample.
Figure 21. Infrared spectrum of authentic sample shown in blue
compared with non-authentic I1 shown in red (spectra are area
normalized).
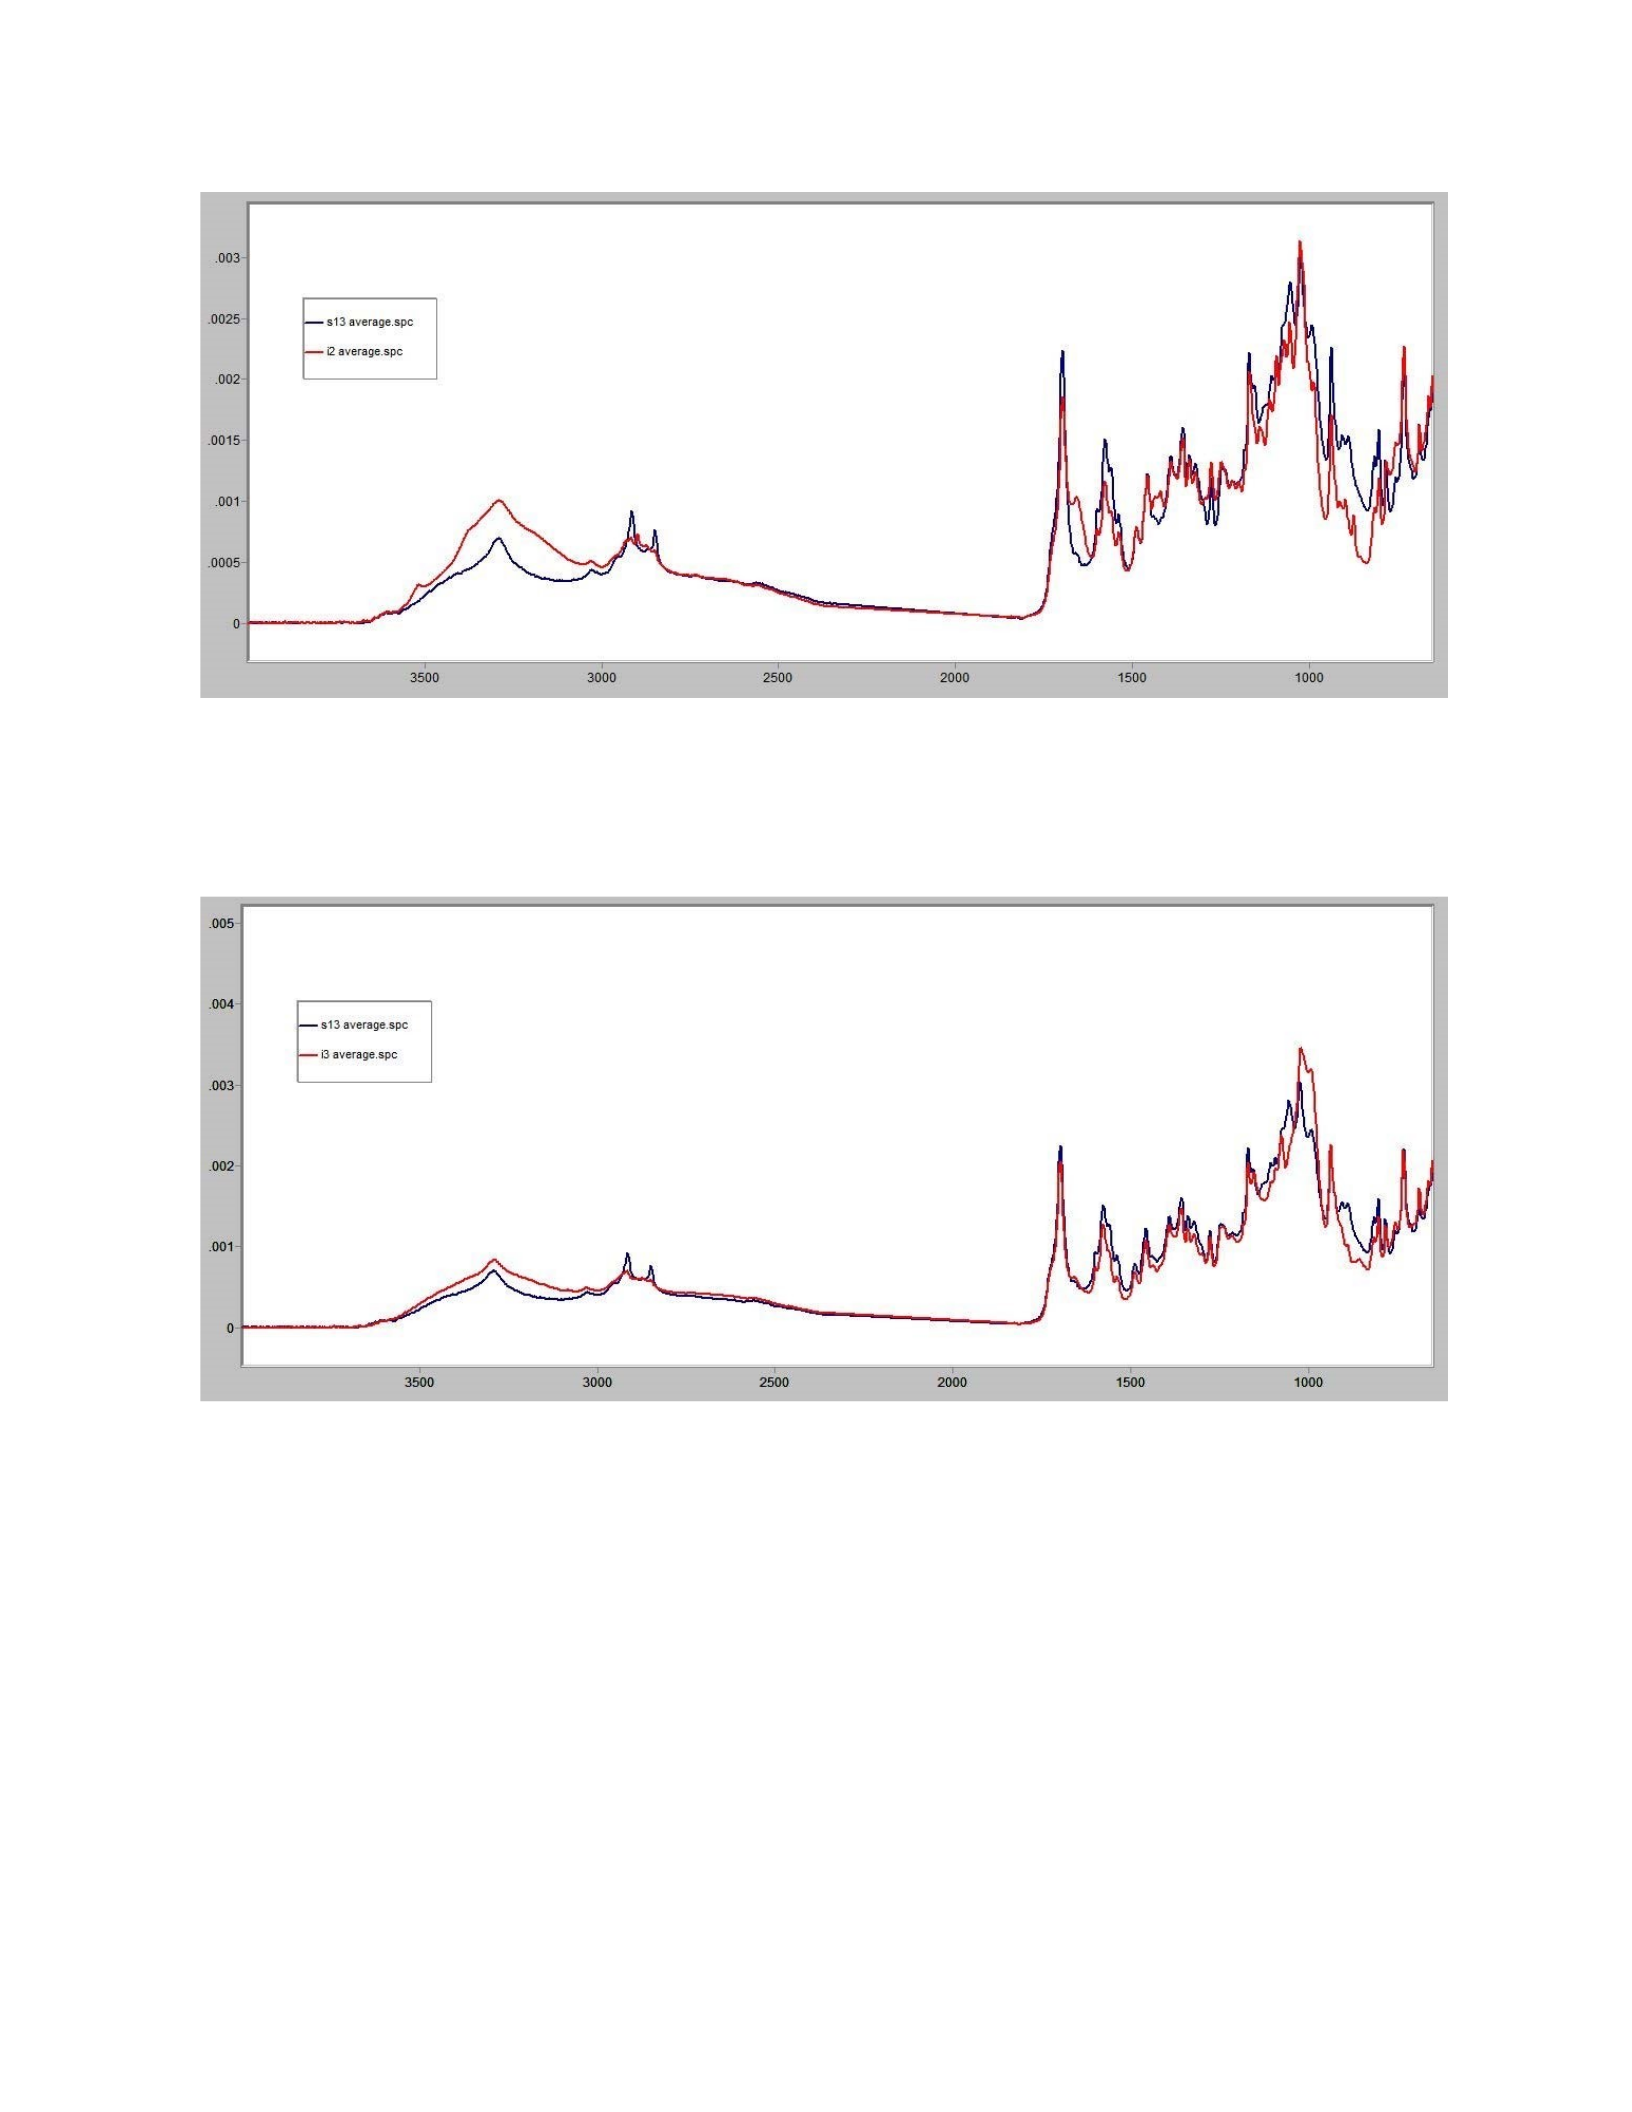
Figure 22. Infrared spectrum of authentic sample shown in blue
compared with non-authentic I2 shown in red (spectra are area
normalized).
Figure 23. Infrared spectrum of authentic sample shown in blue
compared with non-authentic I3 shown in red (spectra are area
normalized).
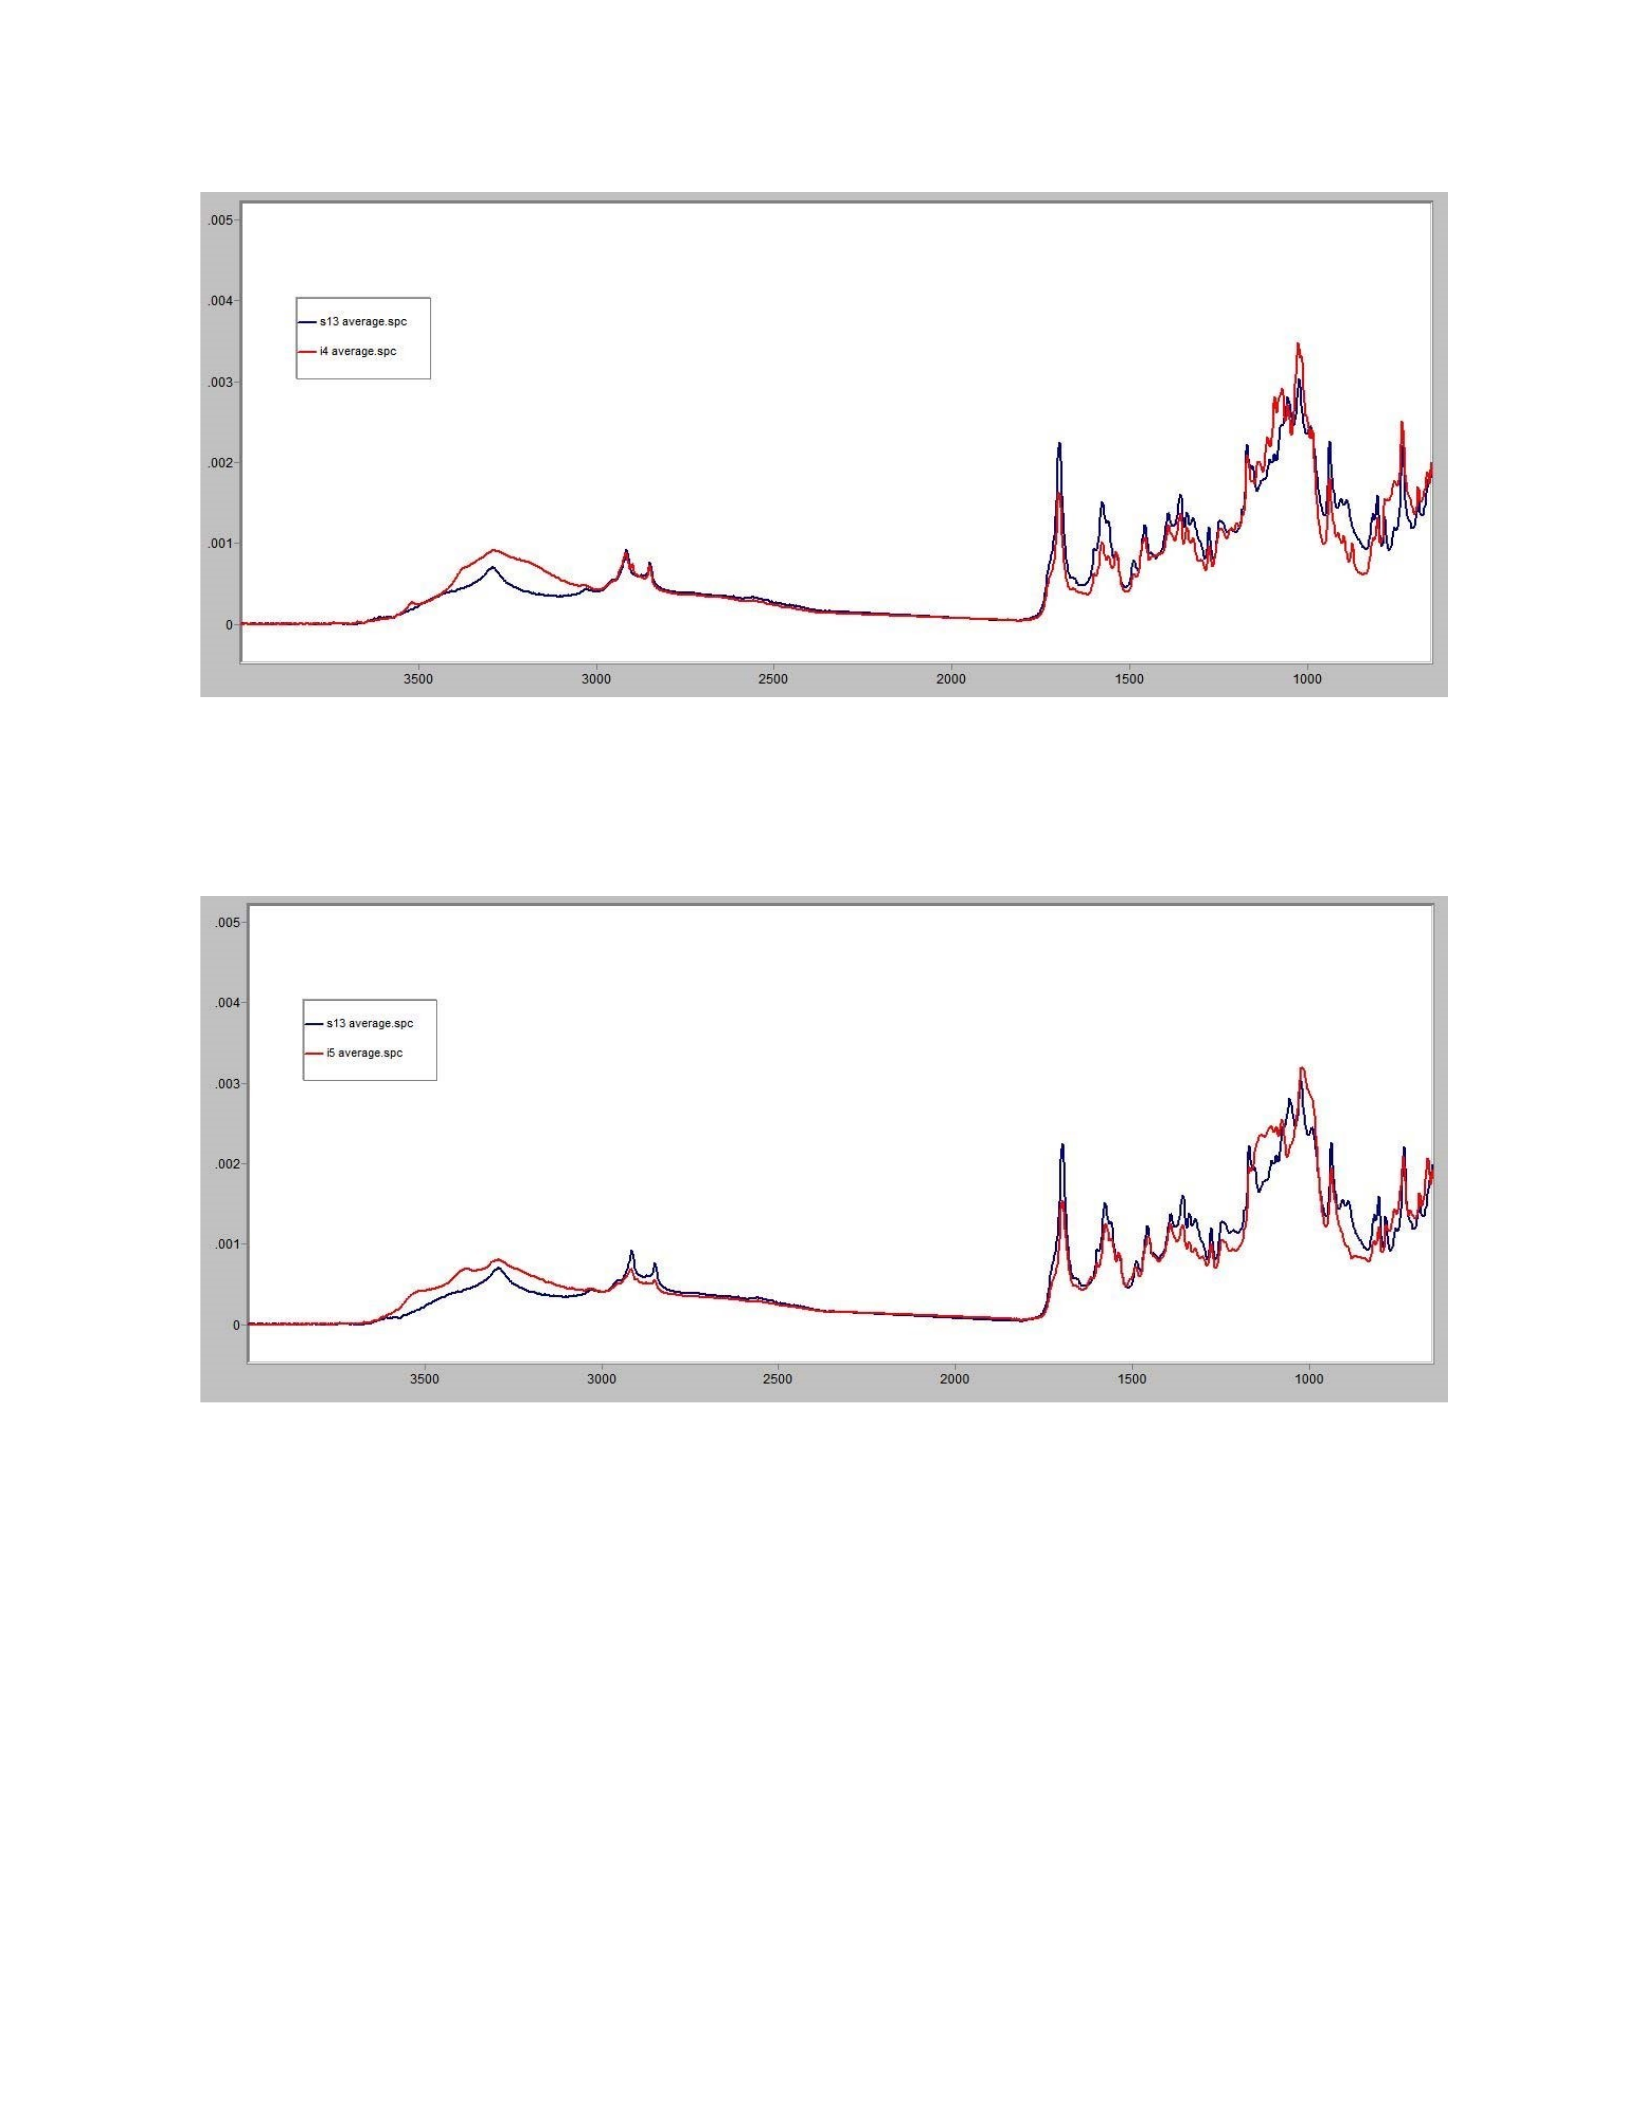
Figure 24. Infrared spectrum of authentic sample shown in blue
compared with non-authentic I4 shown in red (spectra are area
normalized).
Figure 25. Infrared spectrum of authentic sample shown in blue
compared with non-authentic I5 shown in red (spectra are area
normalized).
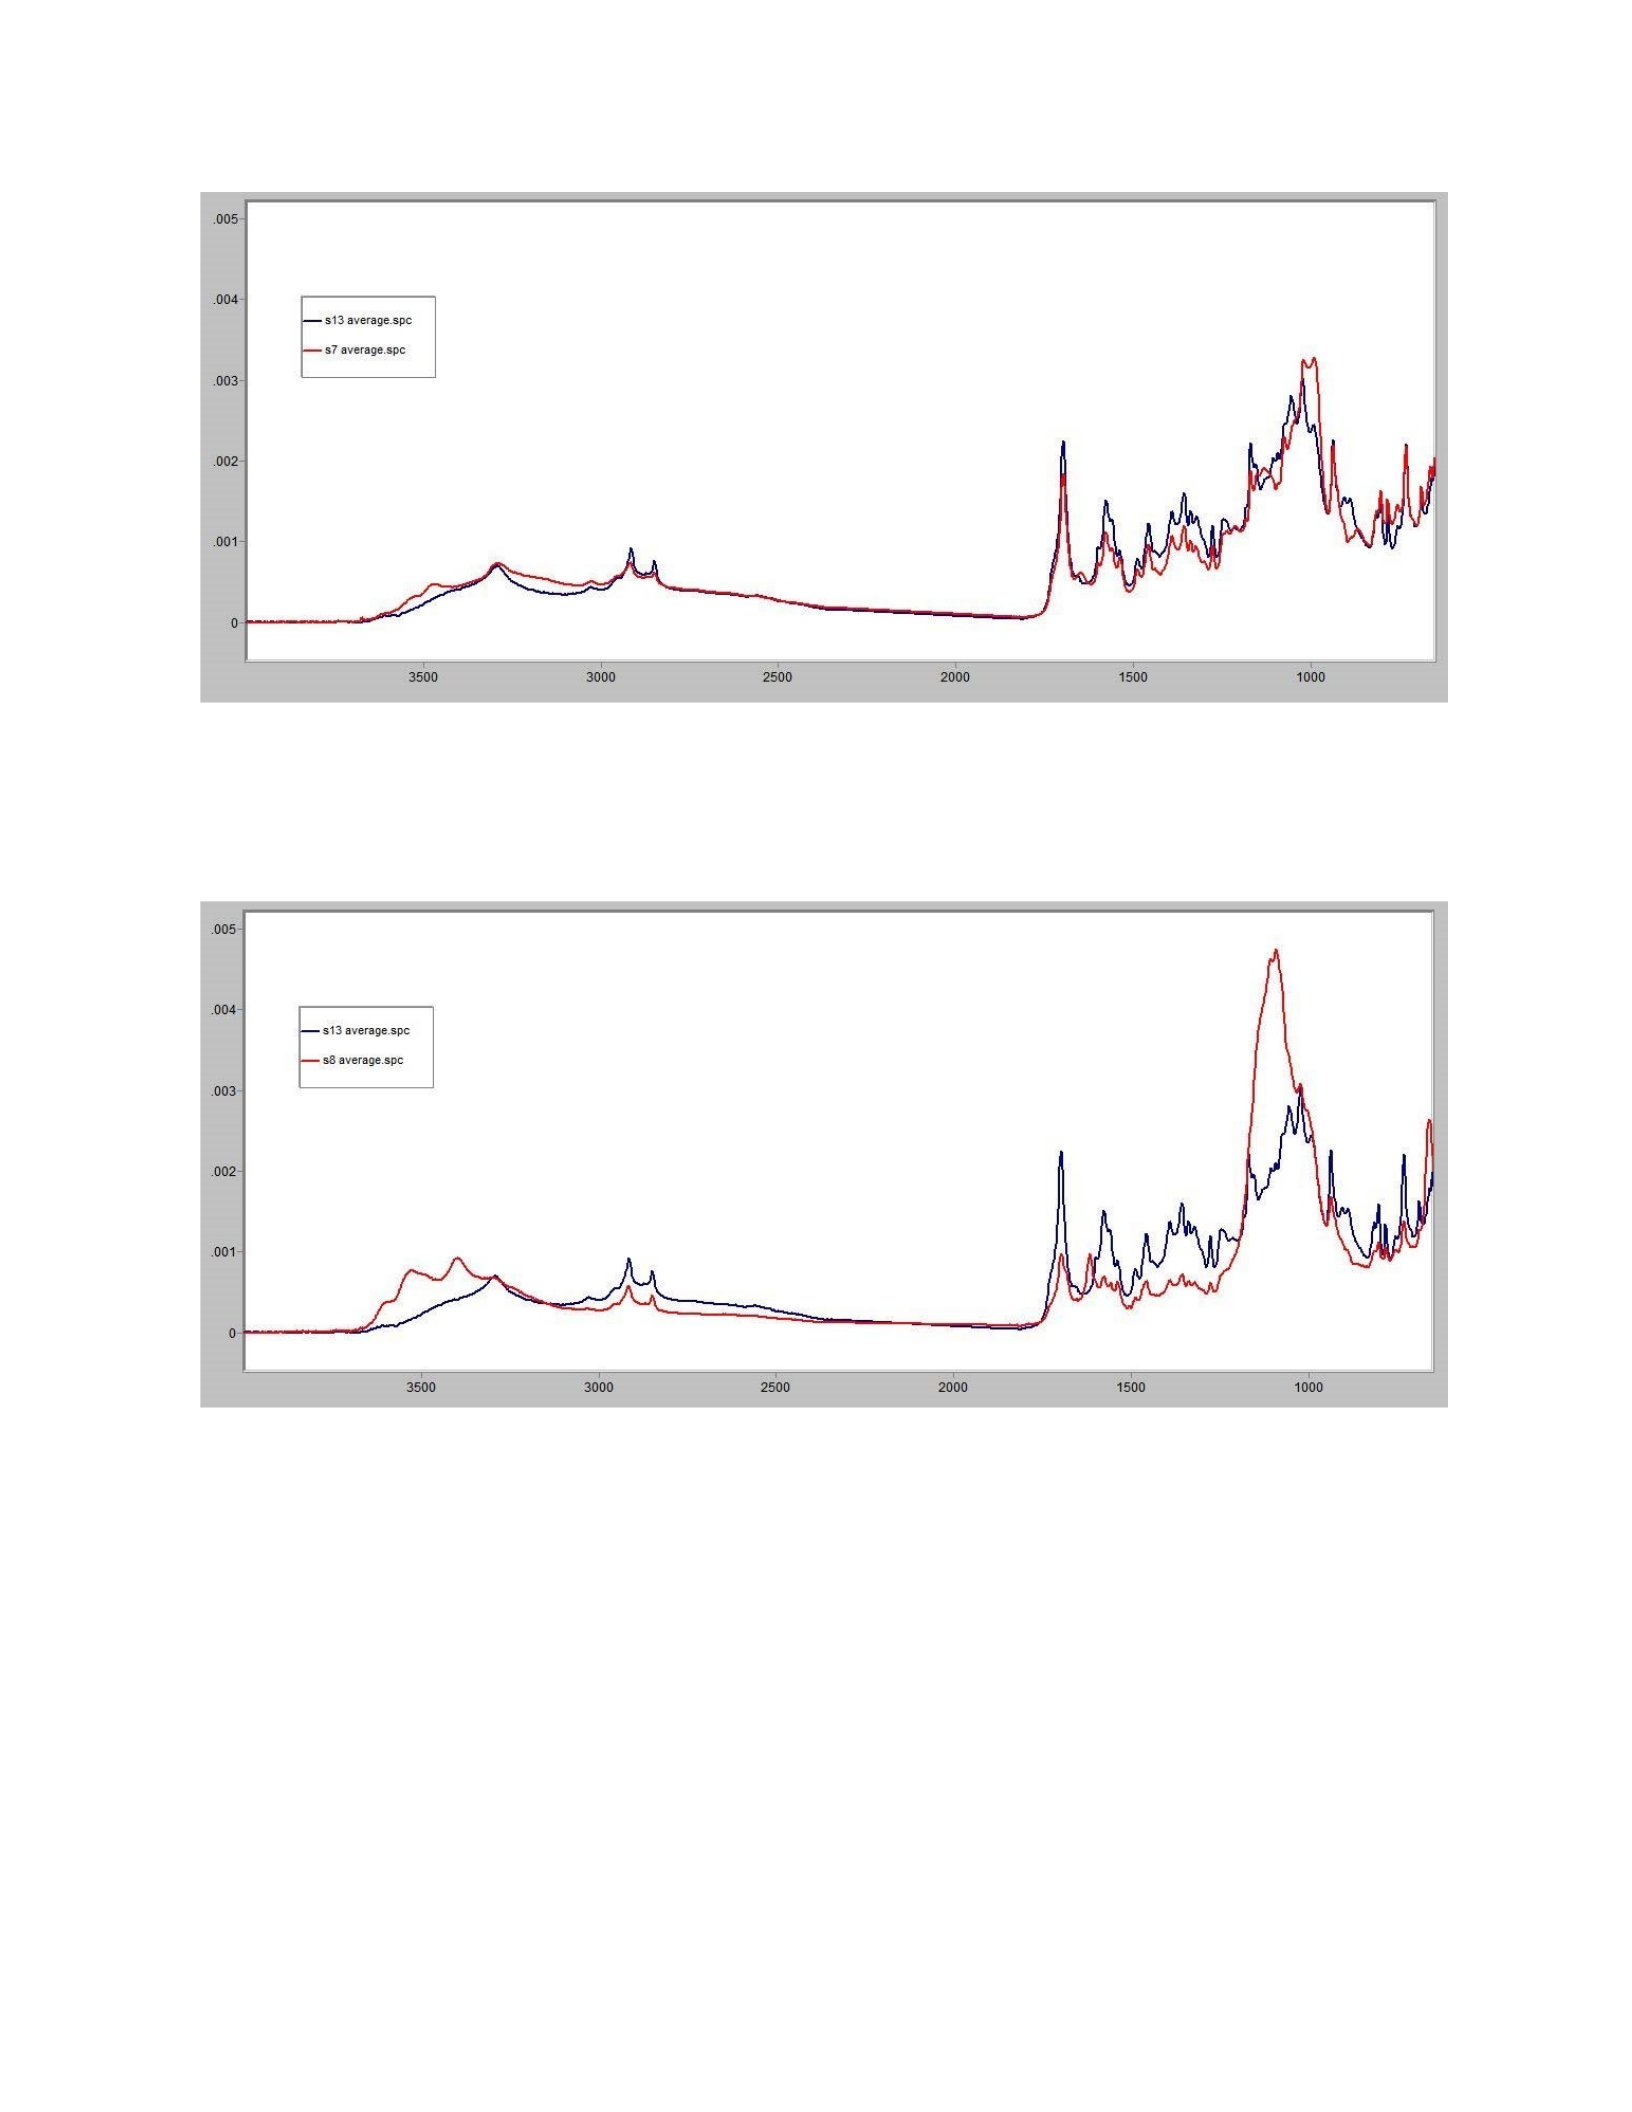
Figure 26. Infrared spectrum of authentic sample compared shown
in blue with non-authentic S7 shown in red (spectra are area
normalized).
Figure 27. Infrared spectrum of authentic sample shown in blue
compared with non-authentic S8 shown in red (spectra are area
normalized).
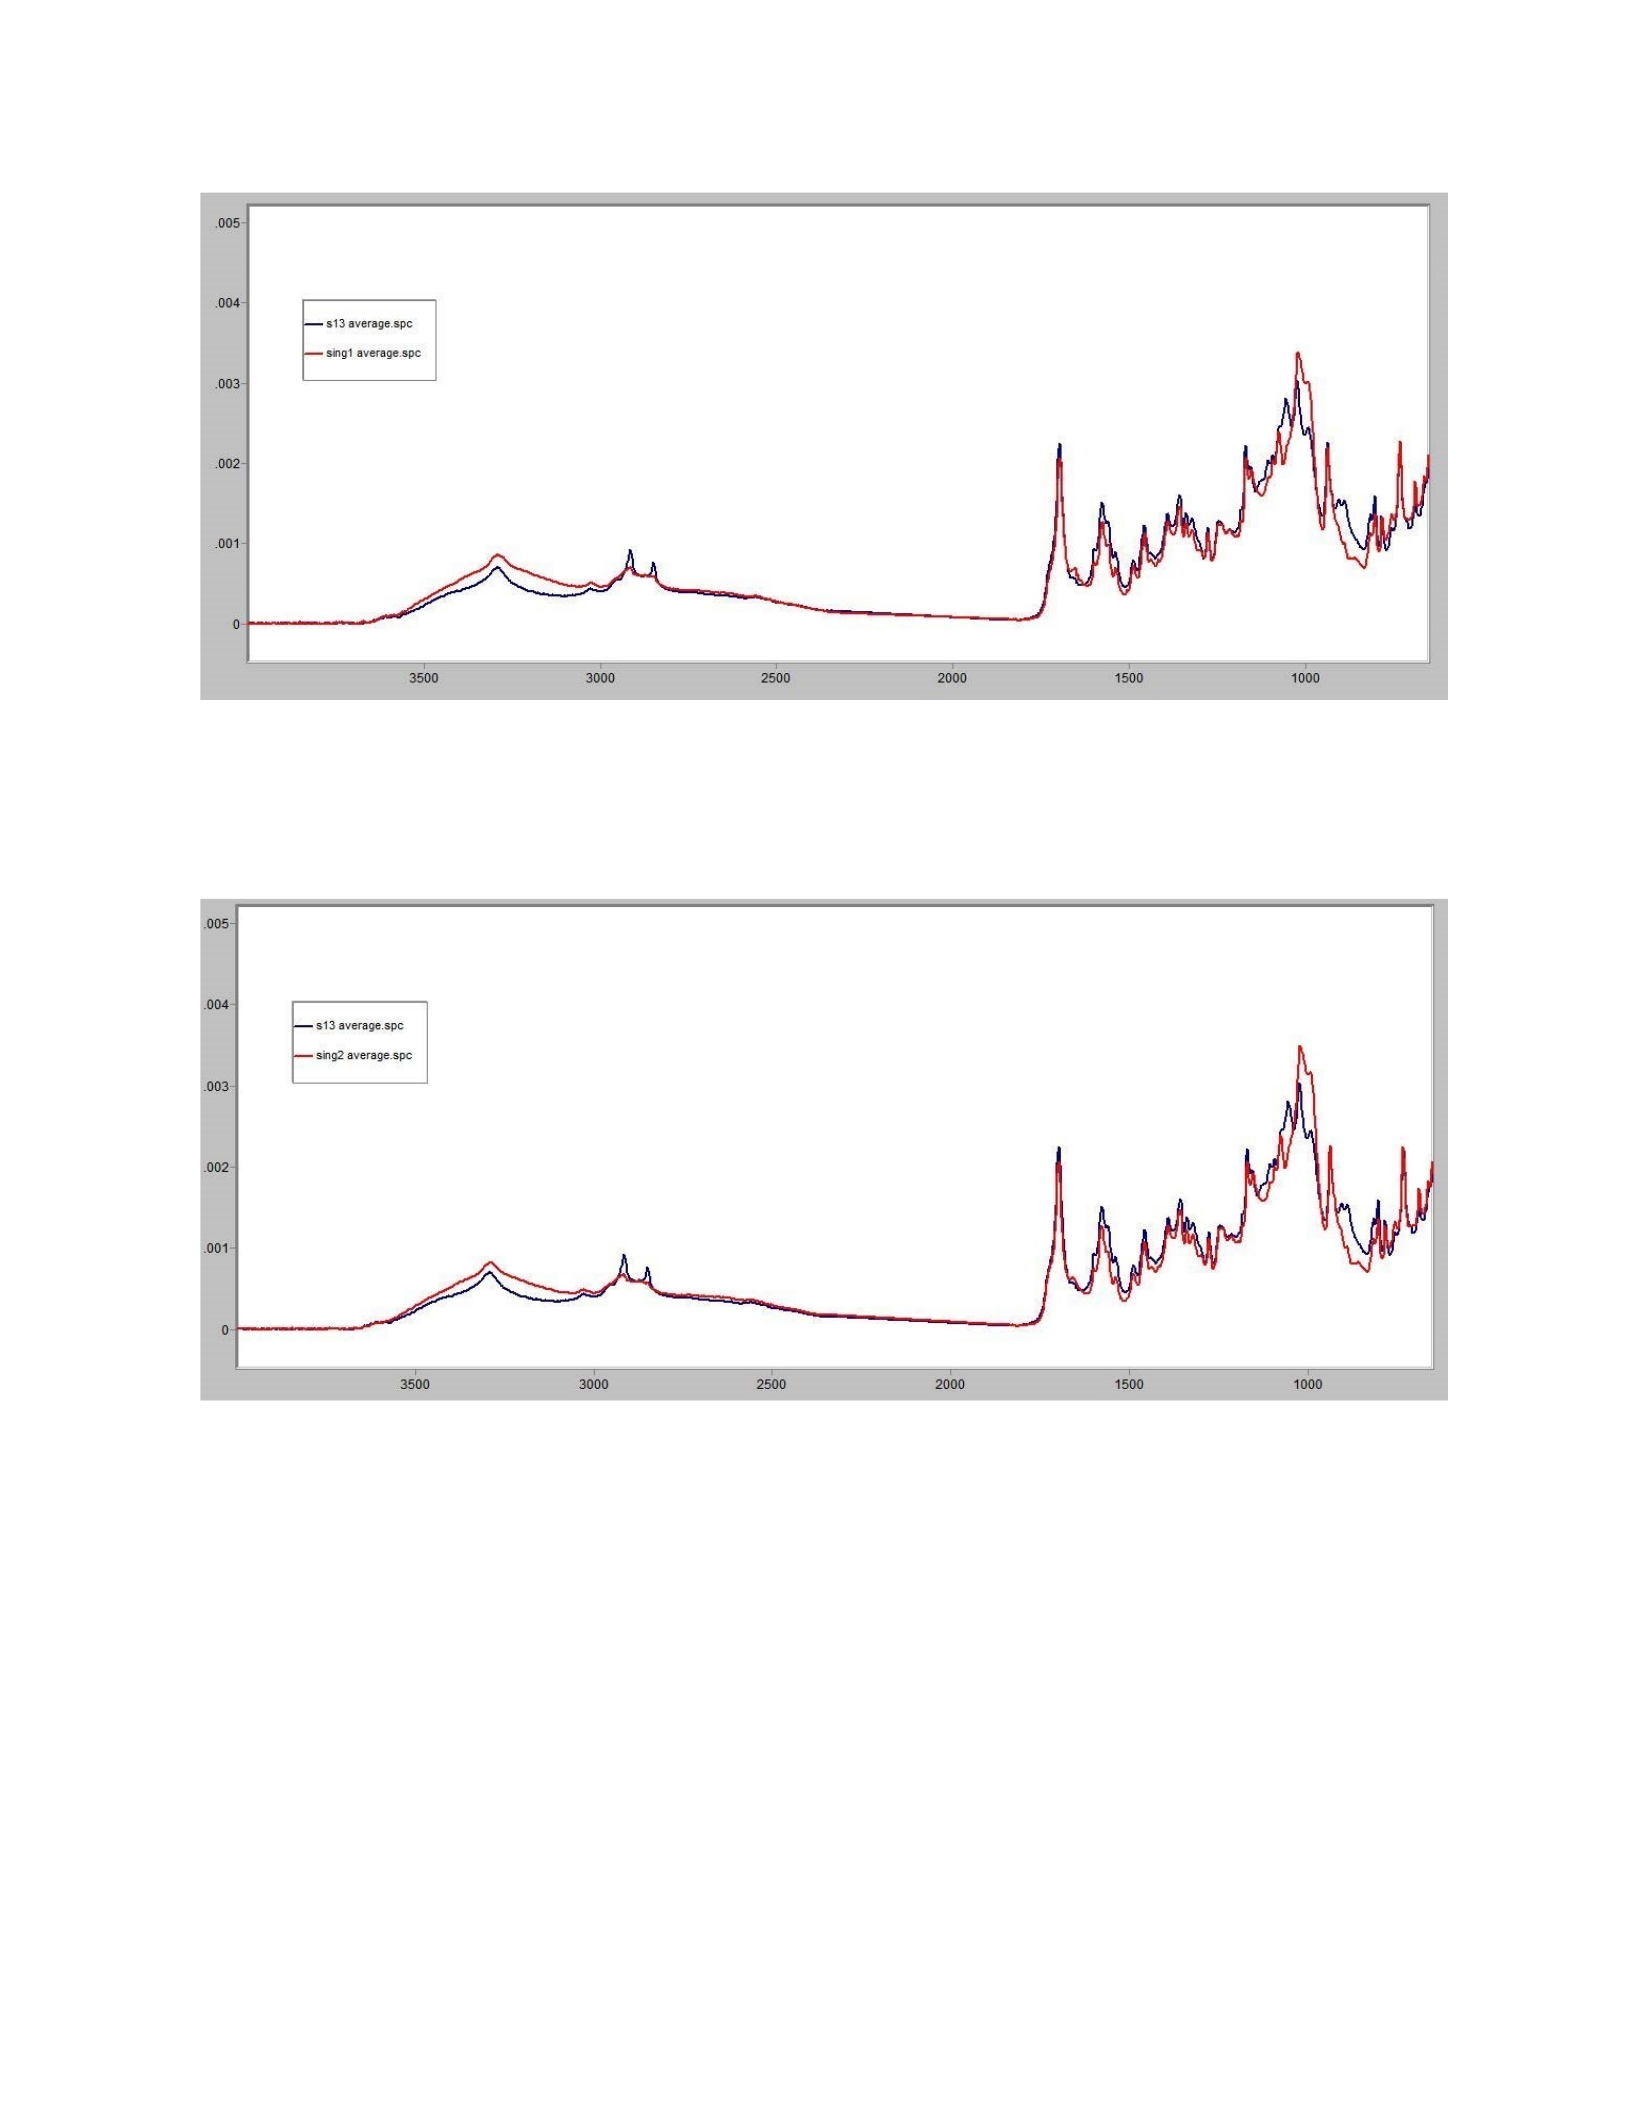
Figure 28. Infrared spectrum of authentic sample shown in blue
compared with non-authentic SING1 shown in red (spectra are
area normalized).
Figure 29. Infrared spectrum of authentic sample shown in blue
compared with non-authentic SING2 shown in red (spectra are
area normalized).
Although only the average spectrum for each sample is shown in the preceding nine
figures, all of the spectra collected from each tablet and from the same sample were consistent
with each other. There was some very small variance in the intensities of absorption bands
within a sample, but there wasn’t a single instance where two spectra from the same sample
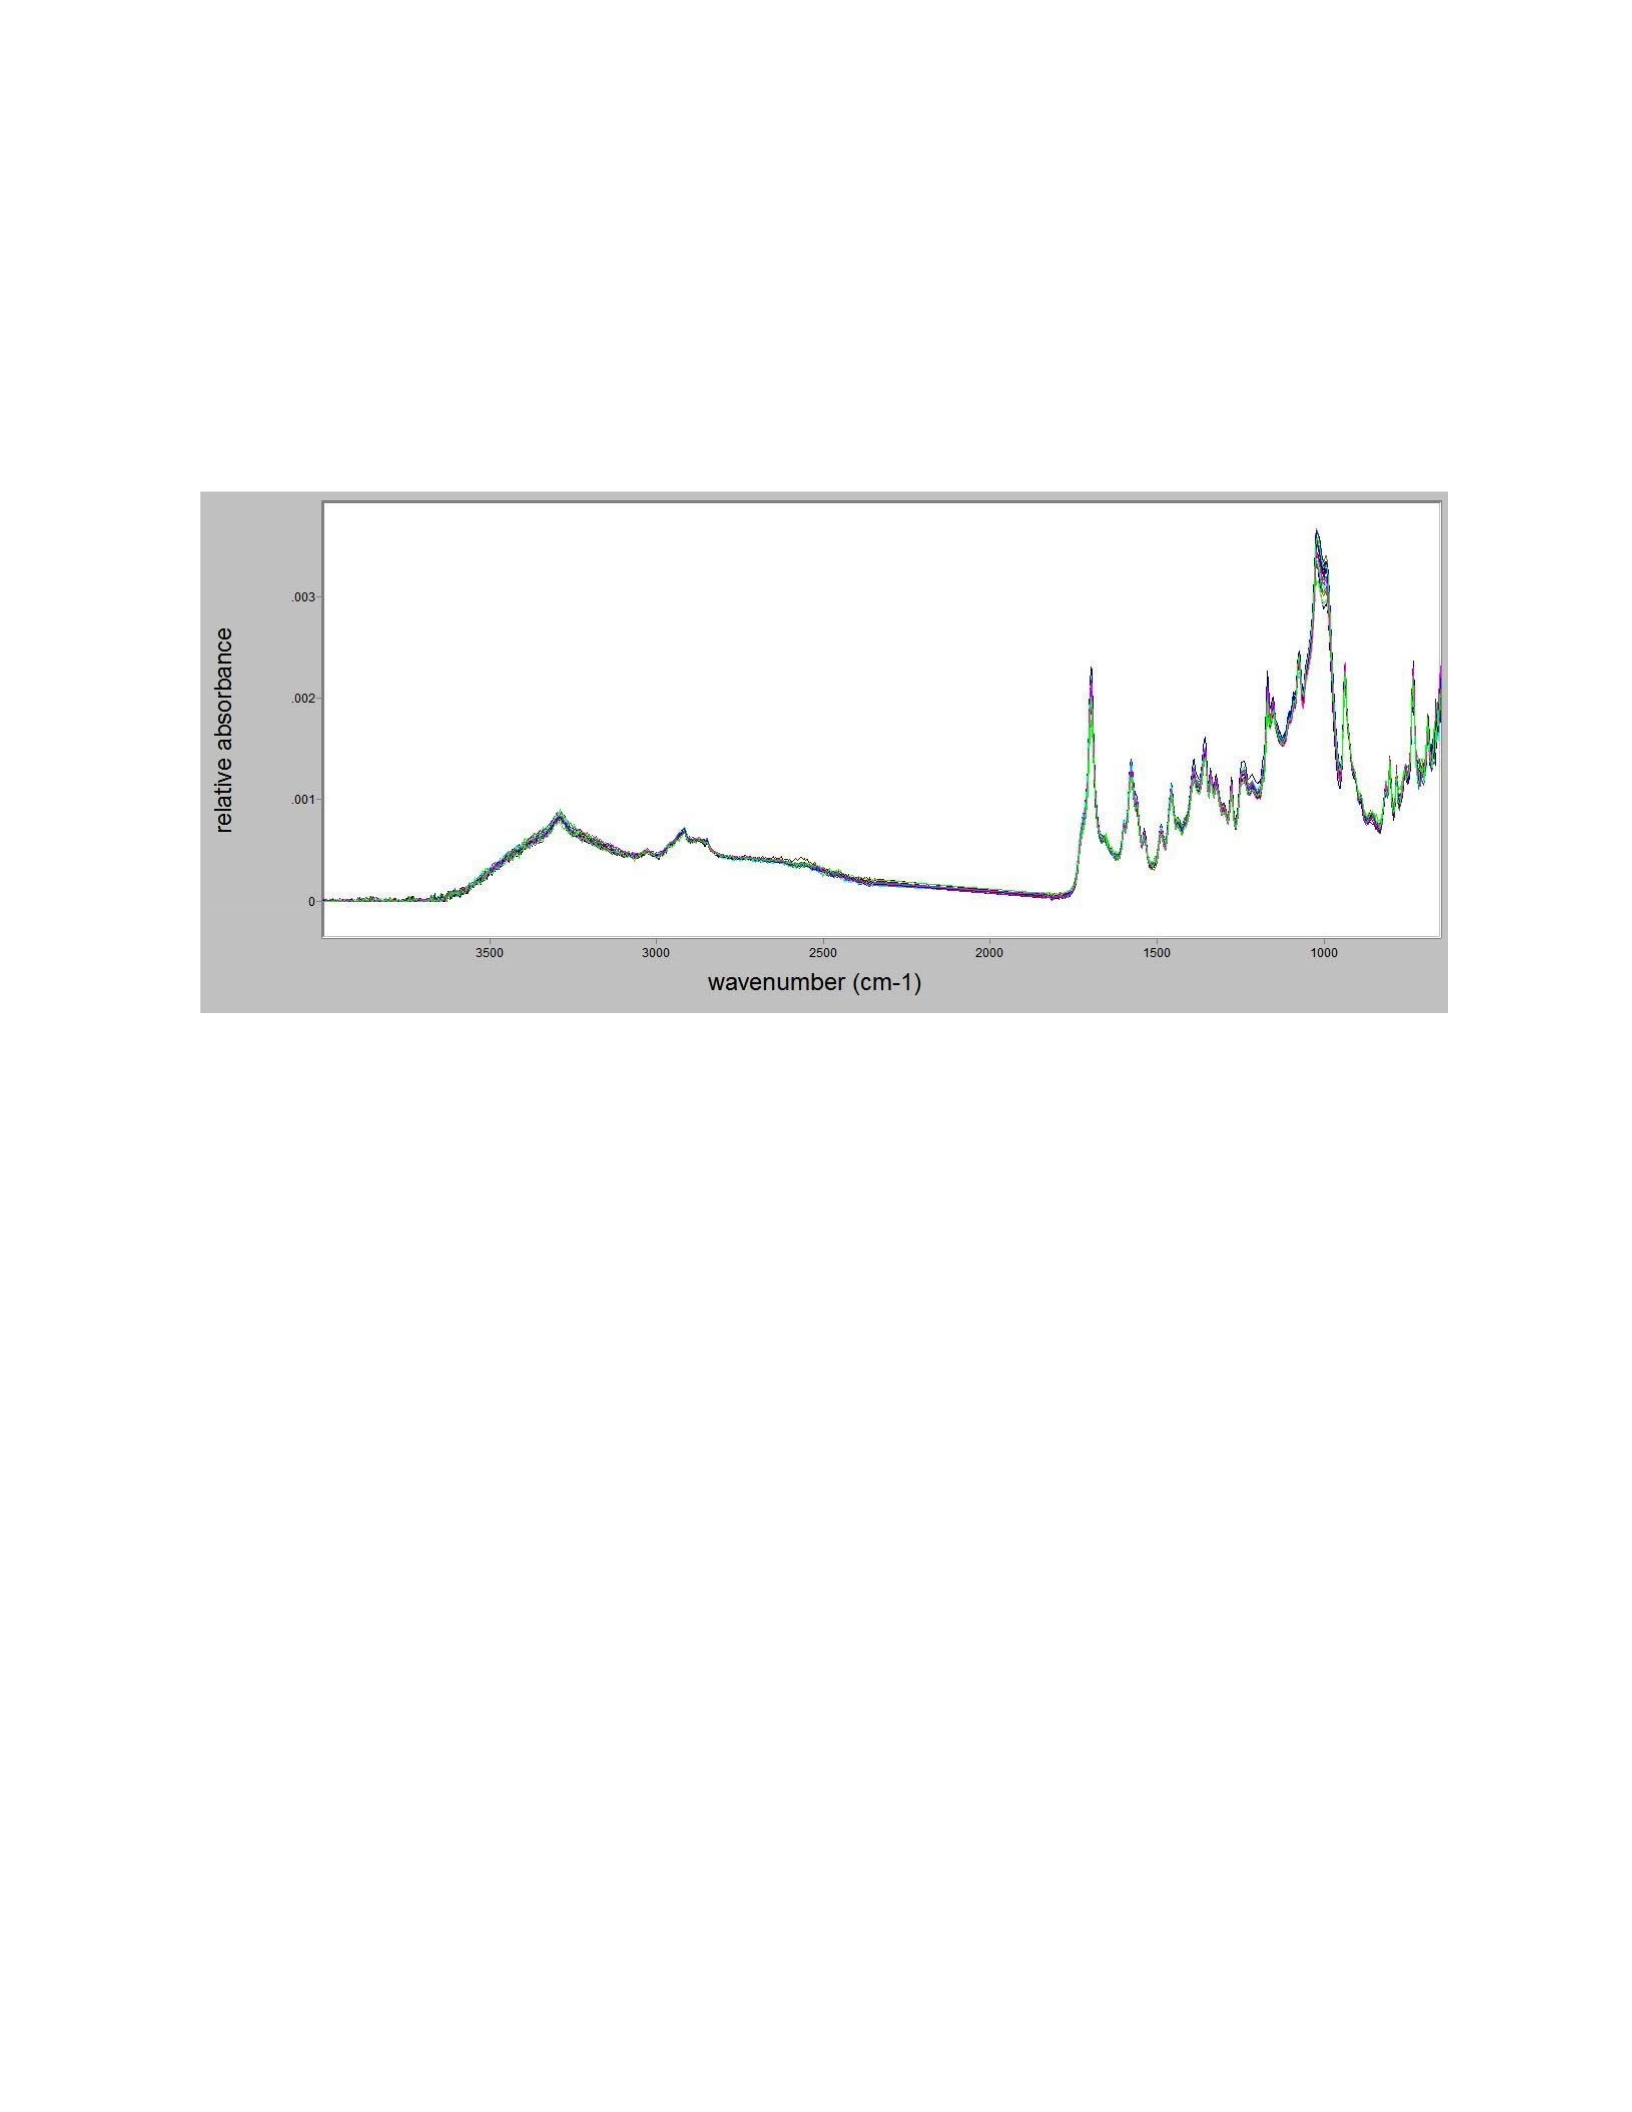
revealed different absorption bands by visual inspection performed by this researcher. Figure 30
shows the 21 spectra collected from the tablet core of the tablets of sample I3. The variance from
spectrum to spectrum shown in these figure were typical of the variance observed in spectra
collected from all tablets of a single sample.
Figure 30. Twenty-one spectra collected from seven tablets of
sample I3 (spectra are area normalized).
Figures 31 and 32 show the standard deviation of the absorbance values of each sample
plotted as a function of wavenumber. Figure 31 shows the data in the fingerprint region of the
spectrum and Figure 32 shows the data in the hydrogen-bonding region of the spectrum. The
standard deviation of the absorbance is variable across the spectrum. Different samples are more
variable than others in different regions of the spectrum. This indicates that variance is directly
related to the physical and chemical compositions of the sample, i.e., absorption bands in some
samples are more variable than they are in others. This data also indicates that the amount of
variance is not necessarily indicative of whether or not a sample is authentic. Although authentic
sample (S13) shows some of the least variance across the spectrum, it has the greatest amount of
variance of all samples at 2915 cm-1 and 2850 cm -1. This observation is consistent with the
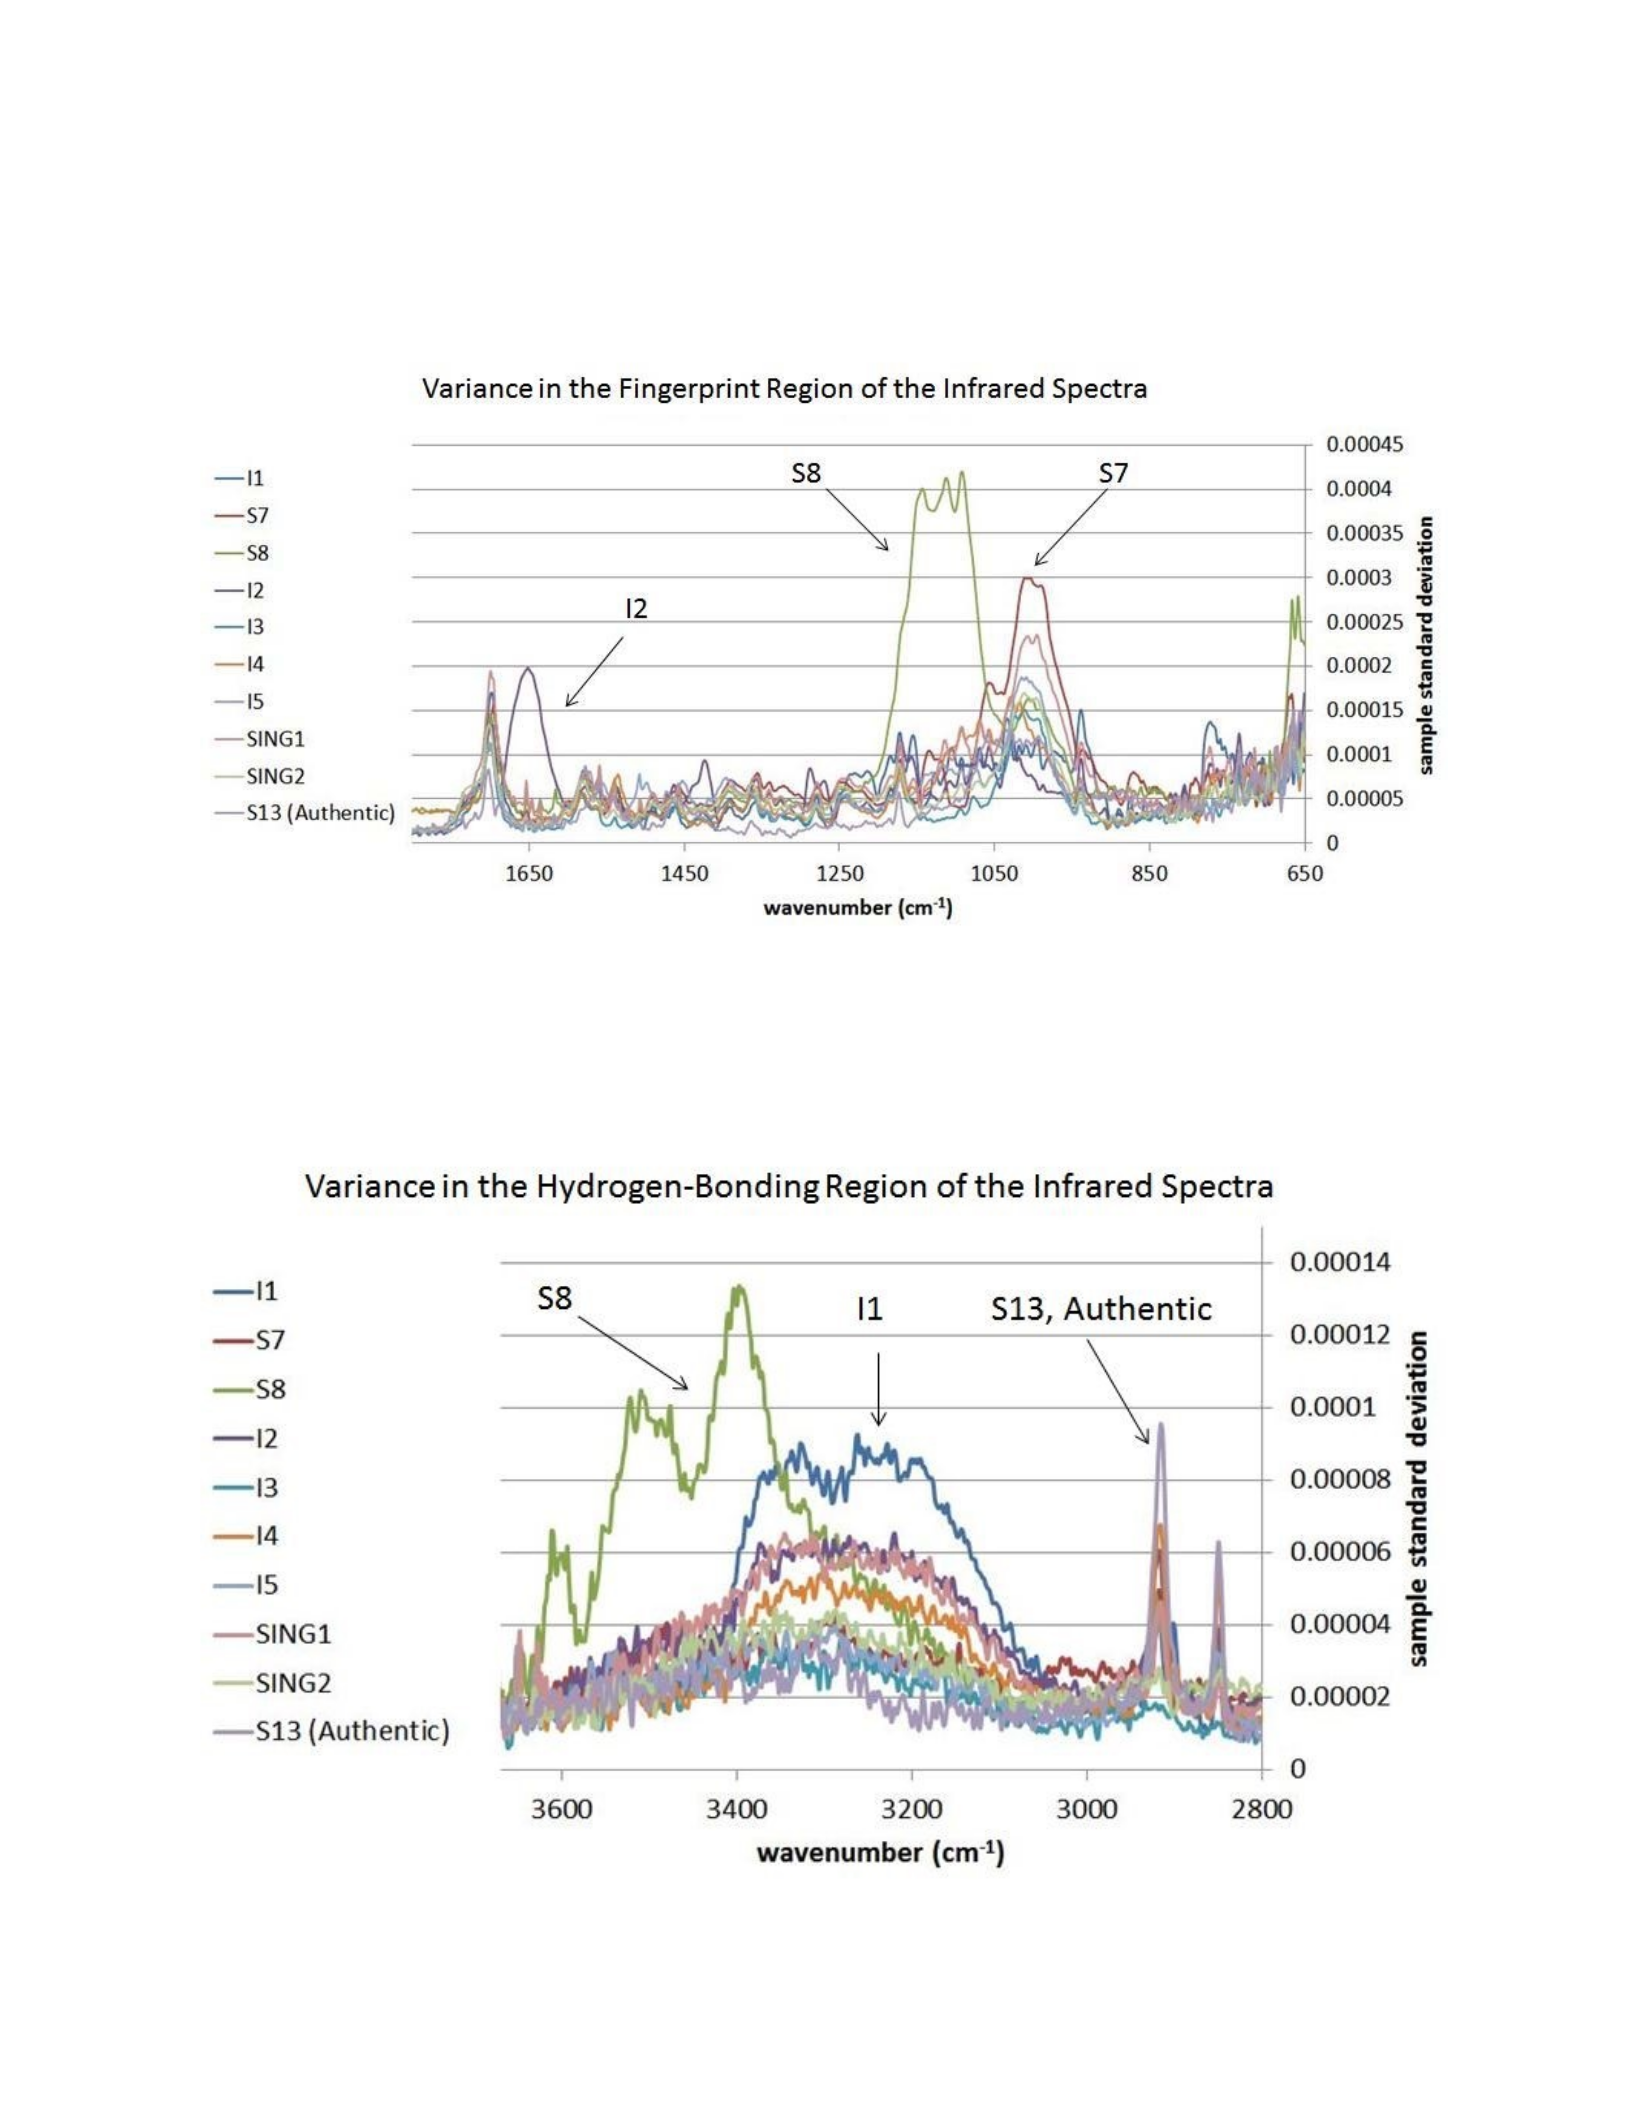
hypothesis that this sample, although manufactured in a well-controlled manner, has sorbed water
which causes variance in these two absorption bands.
Figure 31. Variance expressed as the standard deviation at each
wavenumber in the fingerprint region of the spectrum.
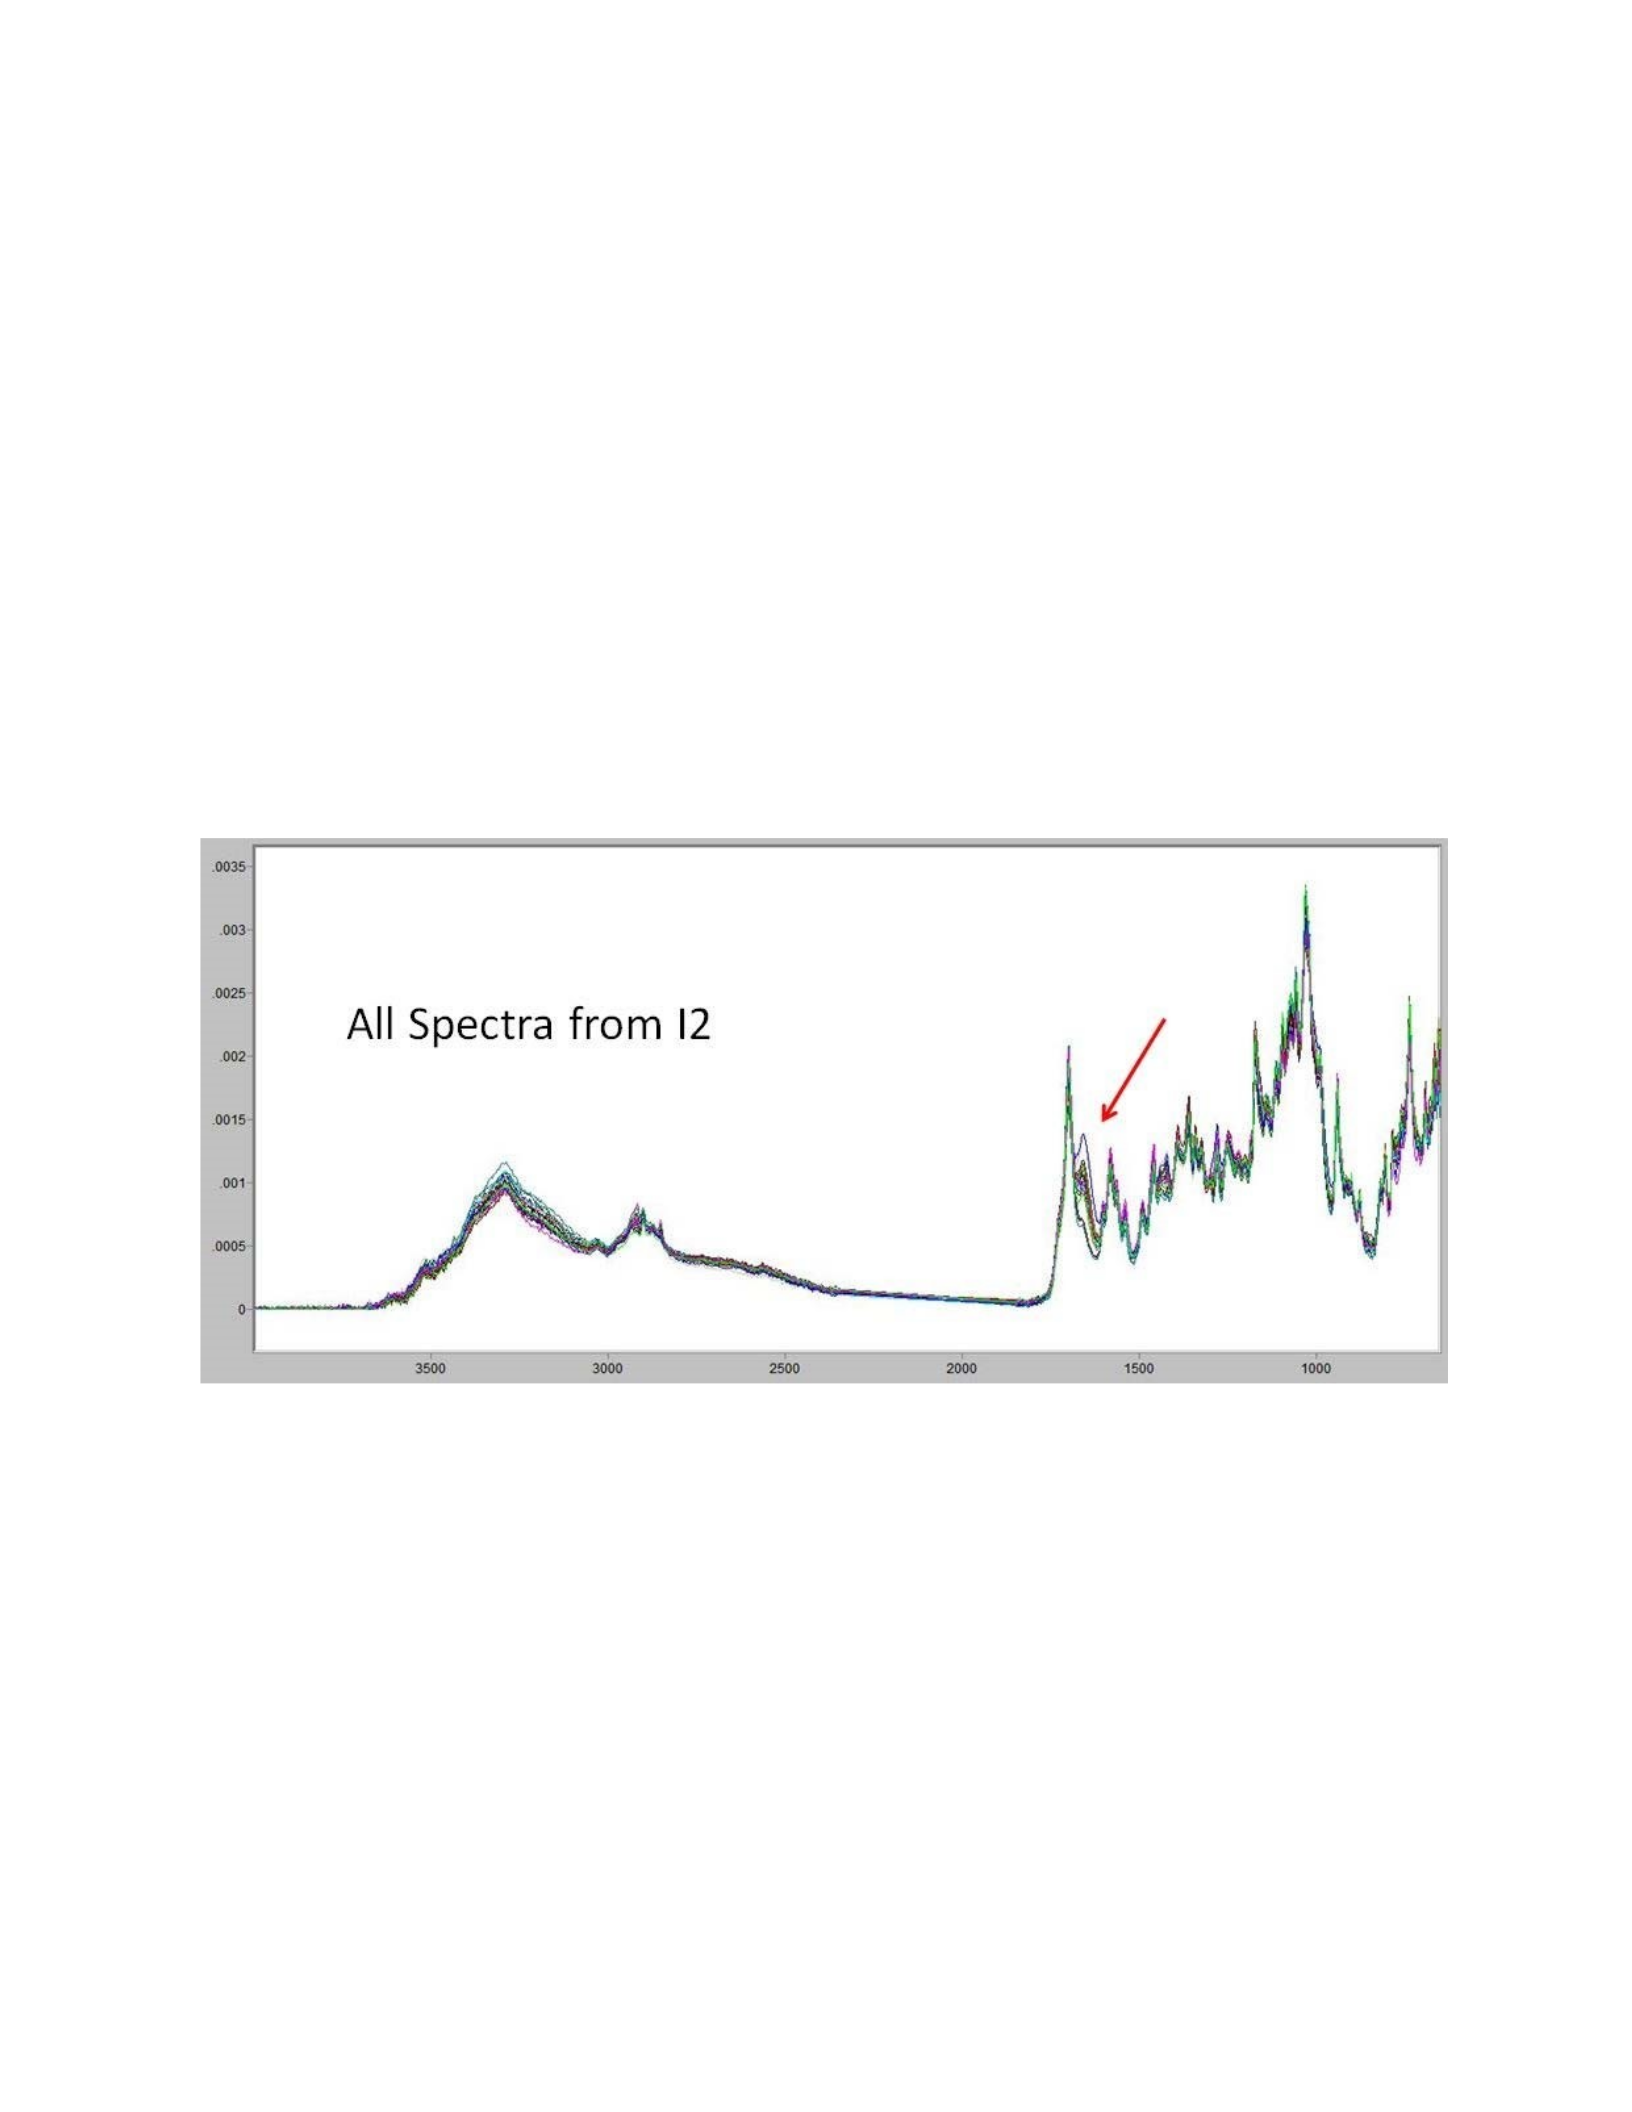
Figure 32. Variance expressed as the standard deviation at each
wavenumber in the hydrogen-bonding region of the spectrum.
Sample I2 showed the greatest amount of variance of all samples at 1655 cm-1. Figure 33
shows an overlay of all of the spectra collected from sample I2. Note that in this sample, the
greatest amount of variance is evident in the absorption band at 1655 cm-1. Figure 34 the same
overlay expanded to see the absorption band at 1655 cm-1 more clearly. In this instance, the
cause of this spectral variance is unknown. Although the variance in the authentic samples can
be explained by the presence of different amounts of water, interpretation of variance in the
infrared spectra in this and some of the other samples was not as straightforward.
Figure 33. All infrared spectra collected from non-authentic
sample I2 (spectra are area normalized).
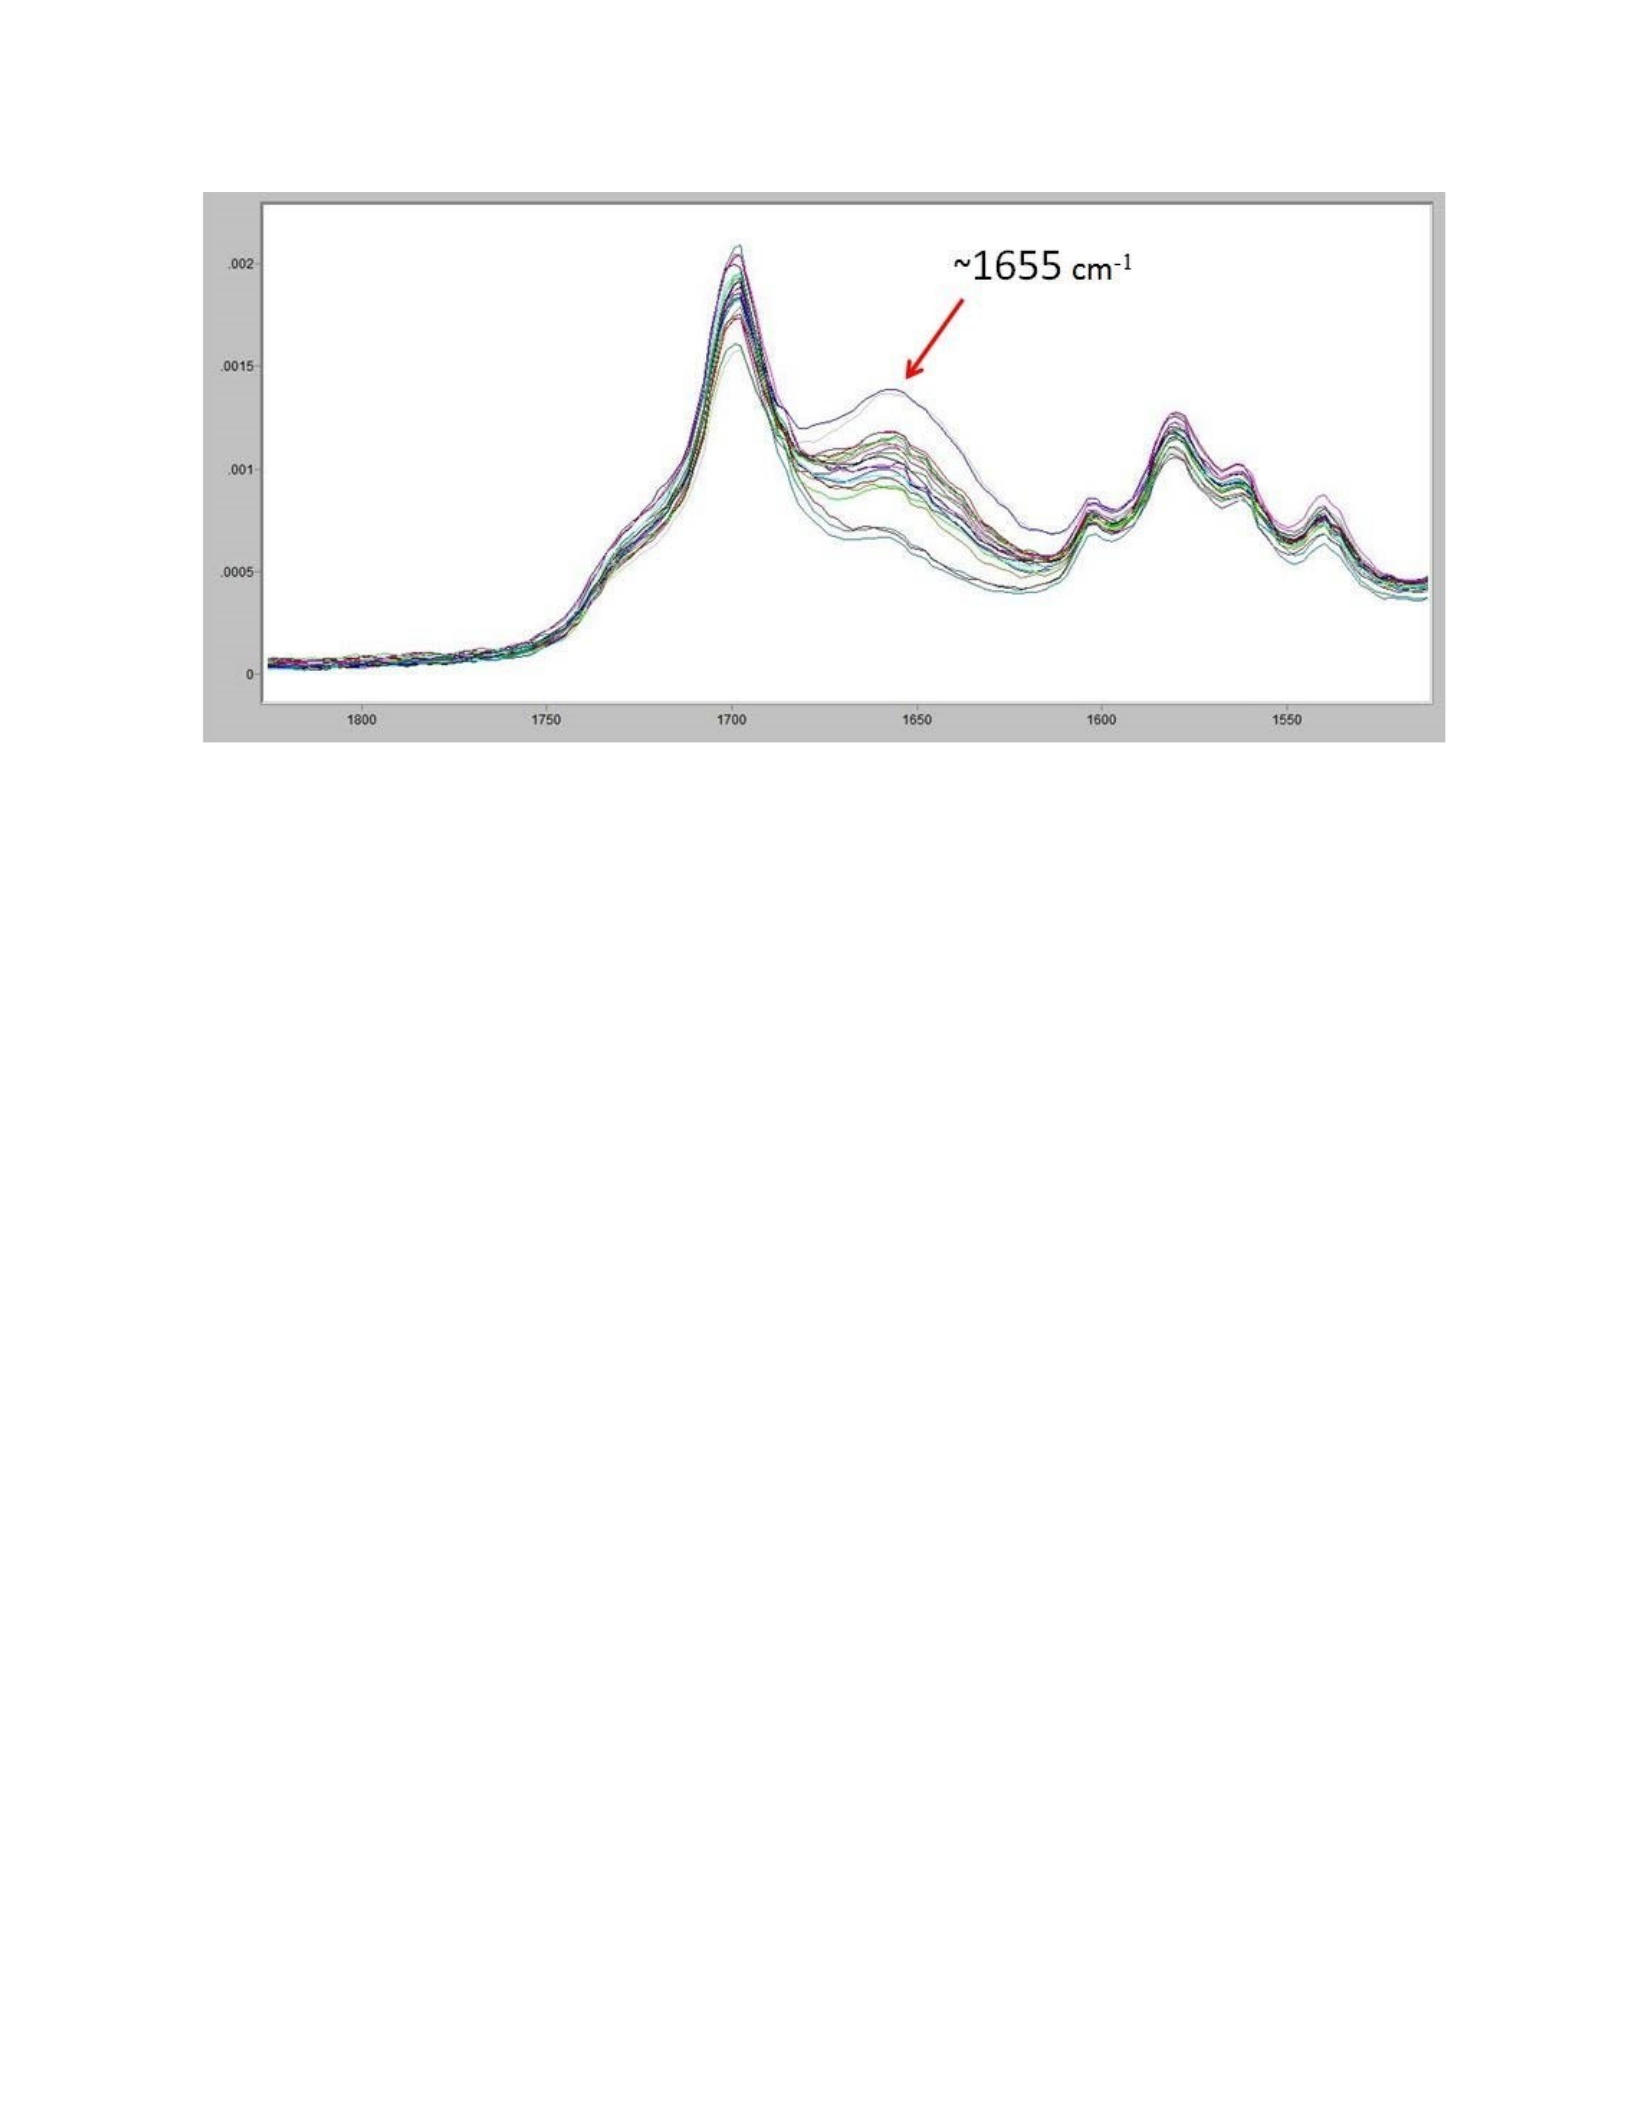
Figure 34. All infrared spectra collected from non-authentic
sample I2 at 1655 cm-1 (spectra are area normalized)
From this infrared data, it was also possible to show that eight of the nine non-authentic
samples contained less than 100 mg of sildenafil citrate, in spite of the fact that all were
purported to contain 100 mg of this API. This type of analysis using infrared spectroscopy was
possible for these samples because tablet weights for these eight samples were measured and
were lower than or approximately equal to what they were for the authentic VIAGRA.
Therefore, the percent by weight of sildenafil citrate in these samples should be equal to or
higher than for the authentic VIAGRA. Since the spectra are area normalized and the infrared
spectra were very similar, absorption bands due only to sildenafil citrate could be reviewed to
look for relative concentrations of sildenafil citrate. In all eight of these samples, the absorption
band due to sildenafil citrate at 1697 cm-1 was lower in intensity than it was for the authentic
VIAGRA. Therefore, the non-authentic samples could not contain 100 mg of sildenafil citrate
(as purported). This band intensity should be the same or higher than it is for these tablets if they
did, in fact, contain 100 mg of sildenafil citrate. It was not possible to determine whether or not
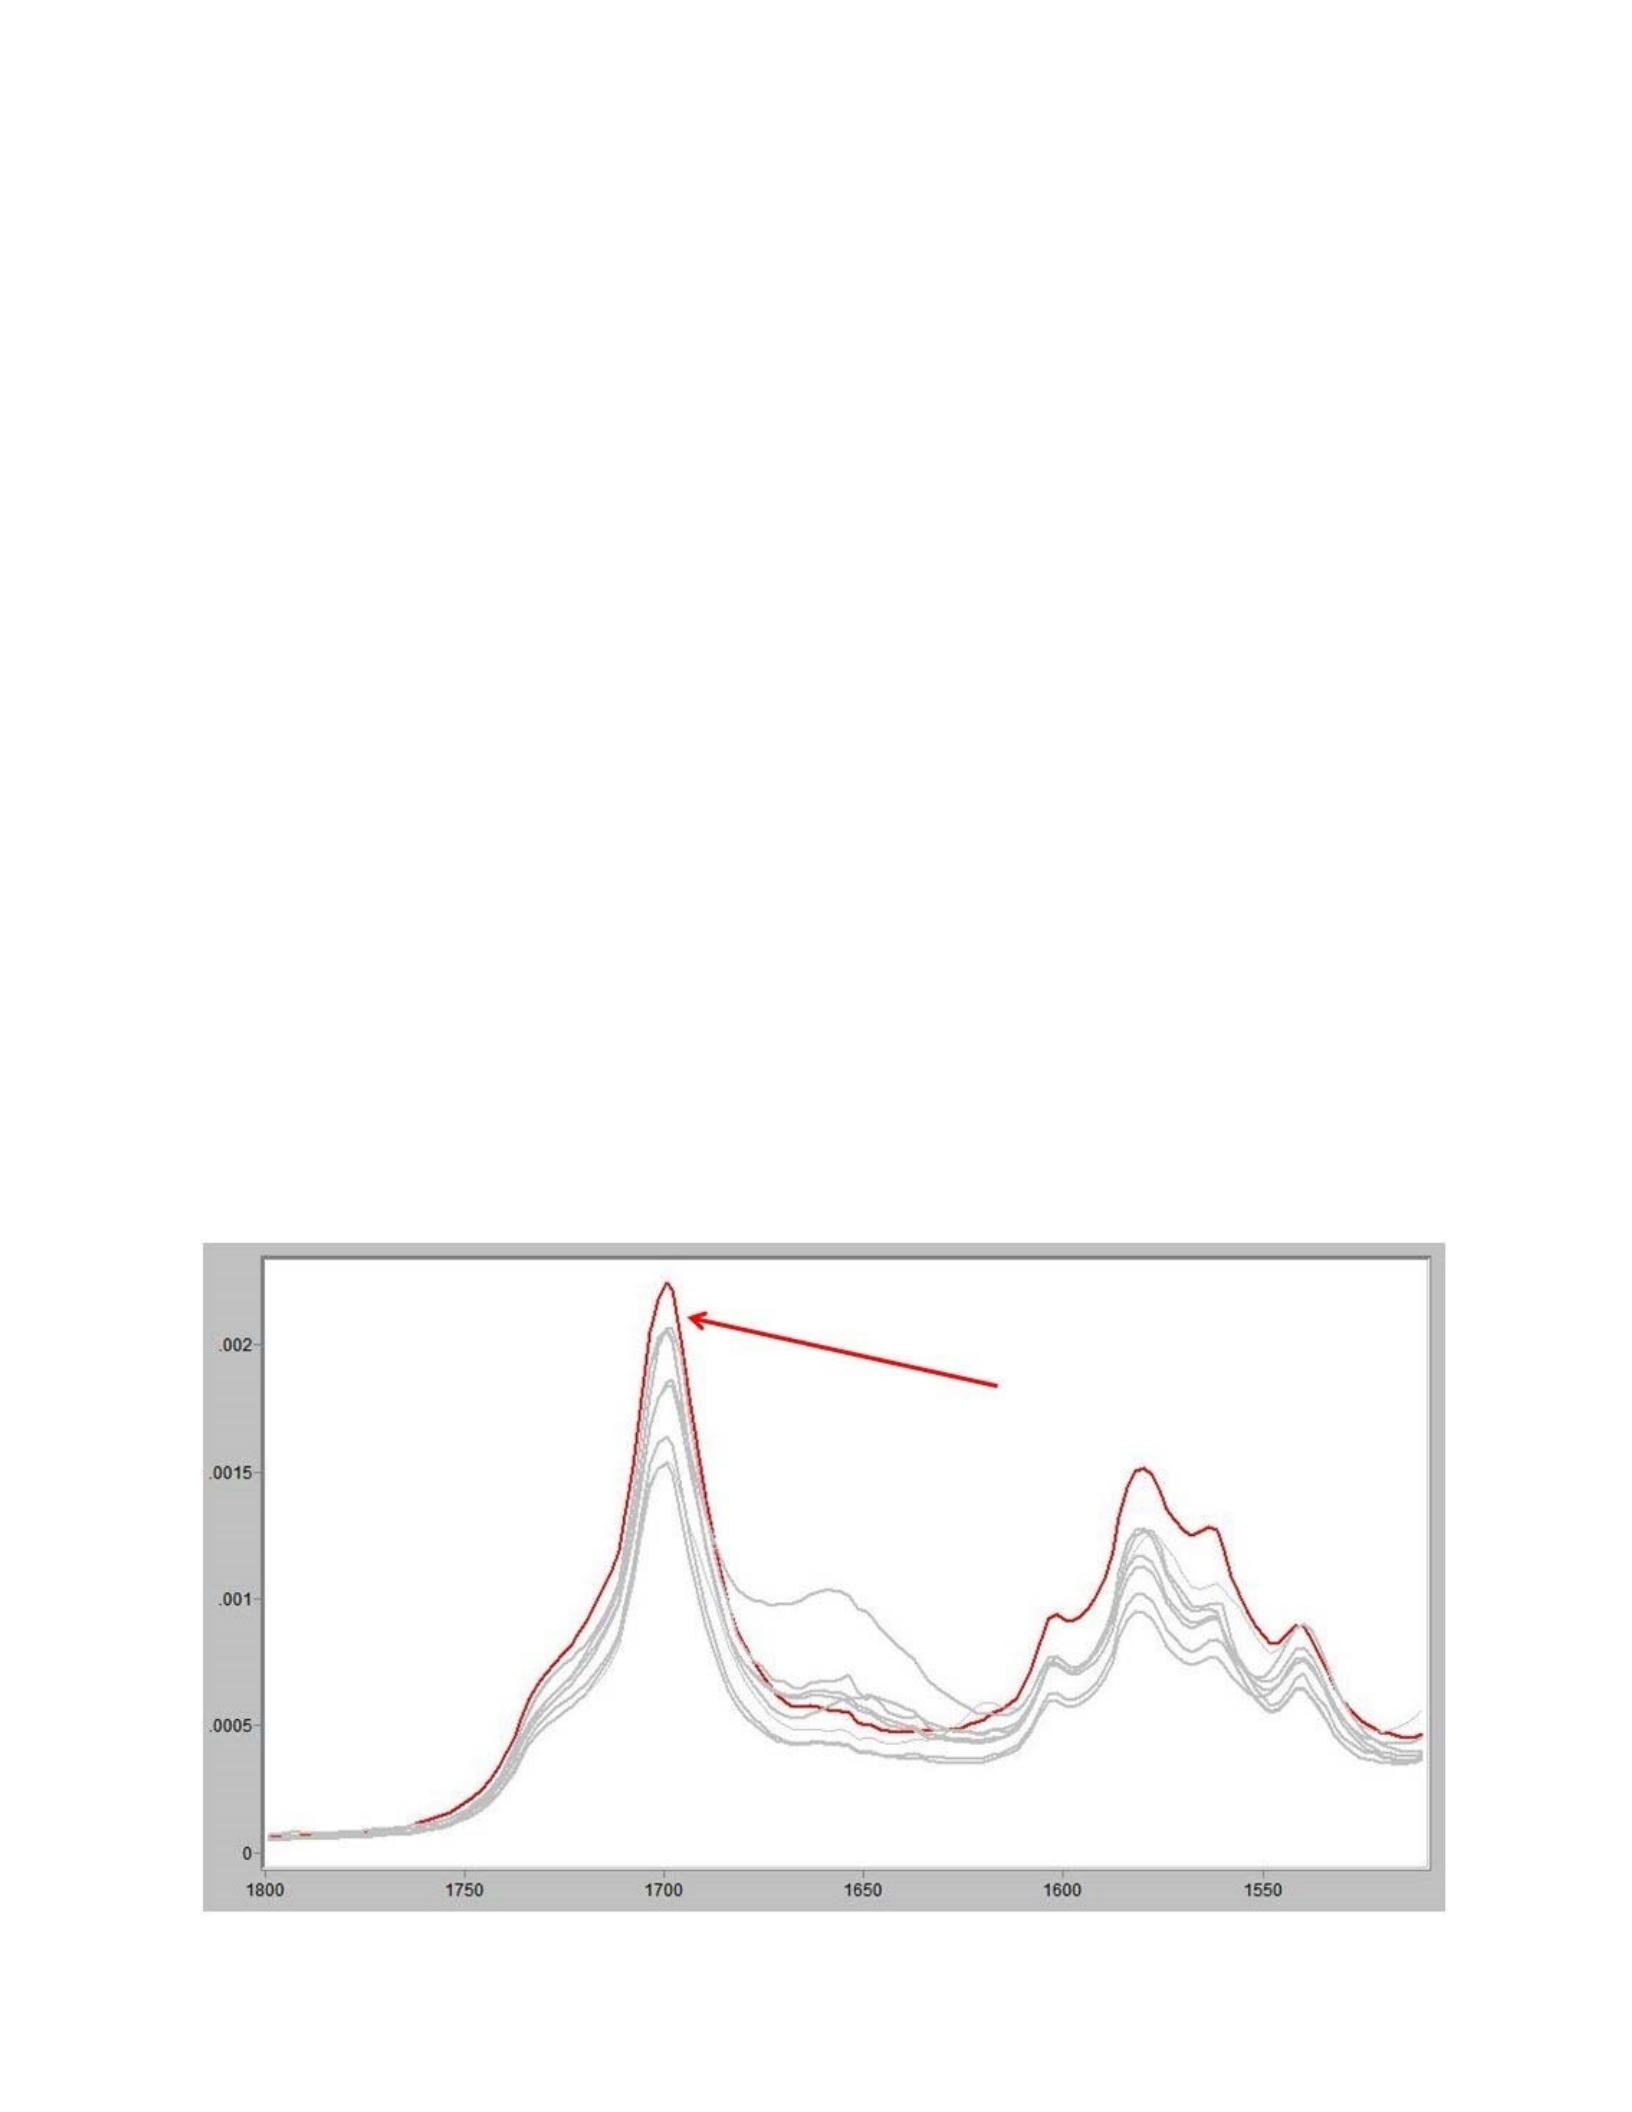
the ninth non-authentic sample contained less than 100 mg of sildenafil citrate because the tablet
weight for this sample was significantly higher than for authentic samples and, therefore, the
absorption band intensity for area normalized spectra should be lower. In addition, the infrared
spectrum for this sample was quite different than it was for the authentic sample. This is also a
problem when performing this type of analysis because intensities become distorted during
normalization due to differences in absorption-band intensities due to the presence of different
chemical functional groups. Aside from this relative evaluation, no attempts were made to
determine quantitatively the amount of sildenafil citrate present in these samples using infrared
spectroscopy or any other technology.
Figure 35 shows an overlay of the spectrum at this absorbance at 1697 cm-1 for the
authentic and non-authentic samples. The spectrum of the authentic sample is shown in red and
all other spectra shown are of the non-authentic samples (except S8). For all samples except for
sample S8, this indicates that less than the purported amount of 100 mg of sildenafil citrate is
present in the sample.
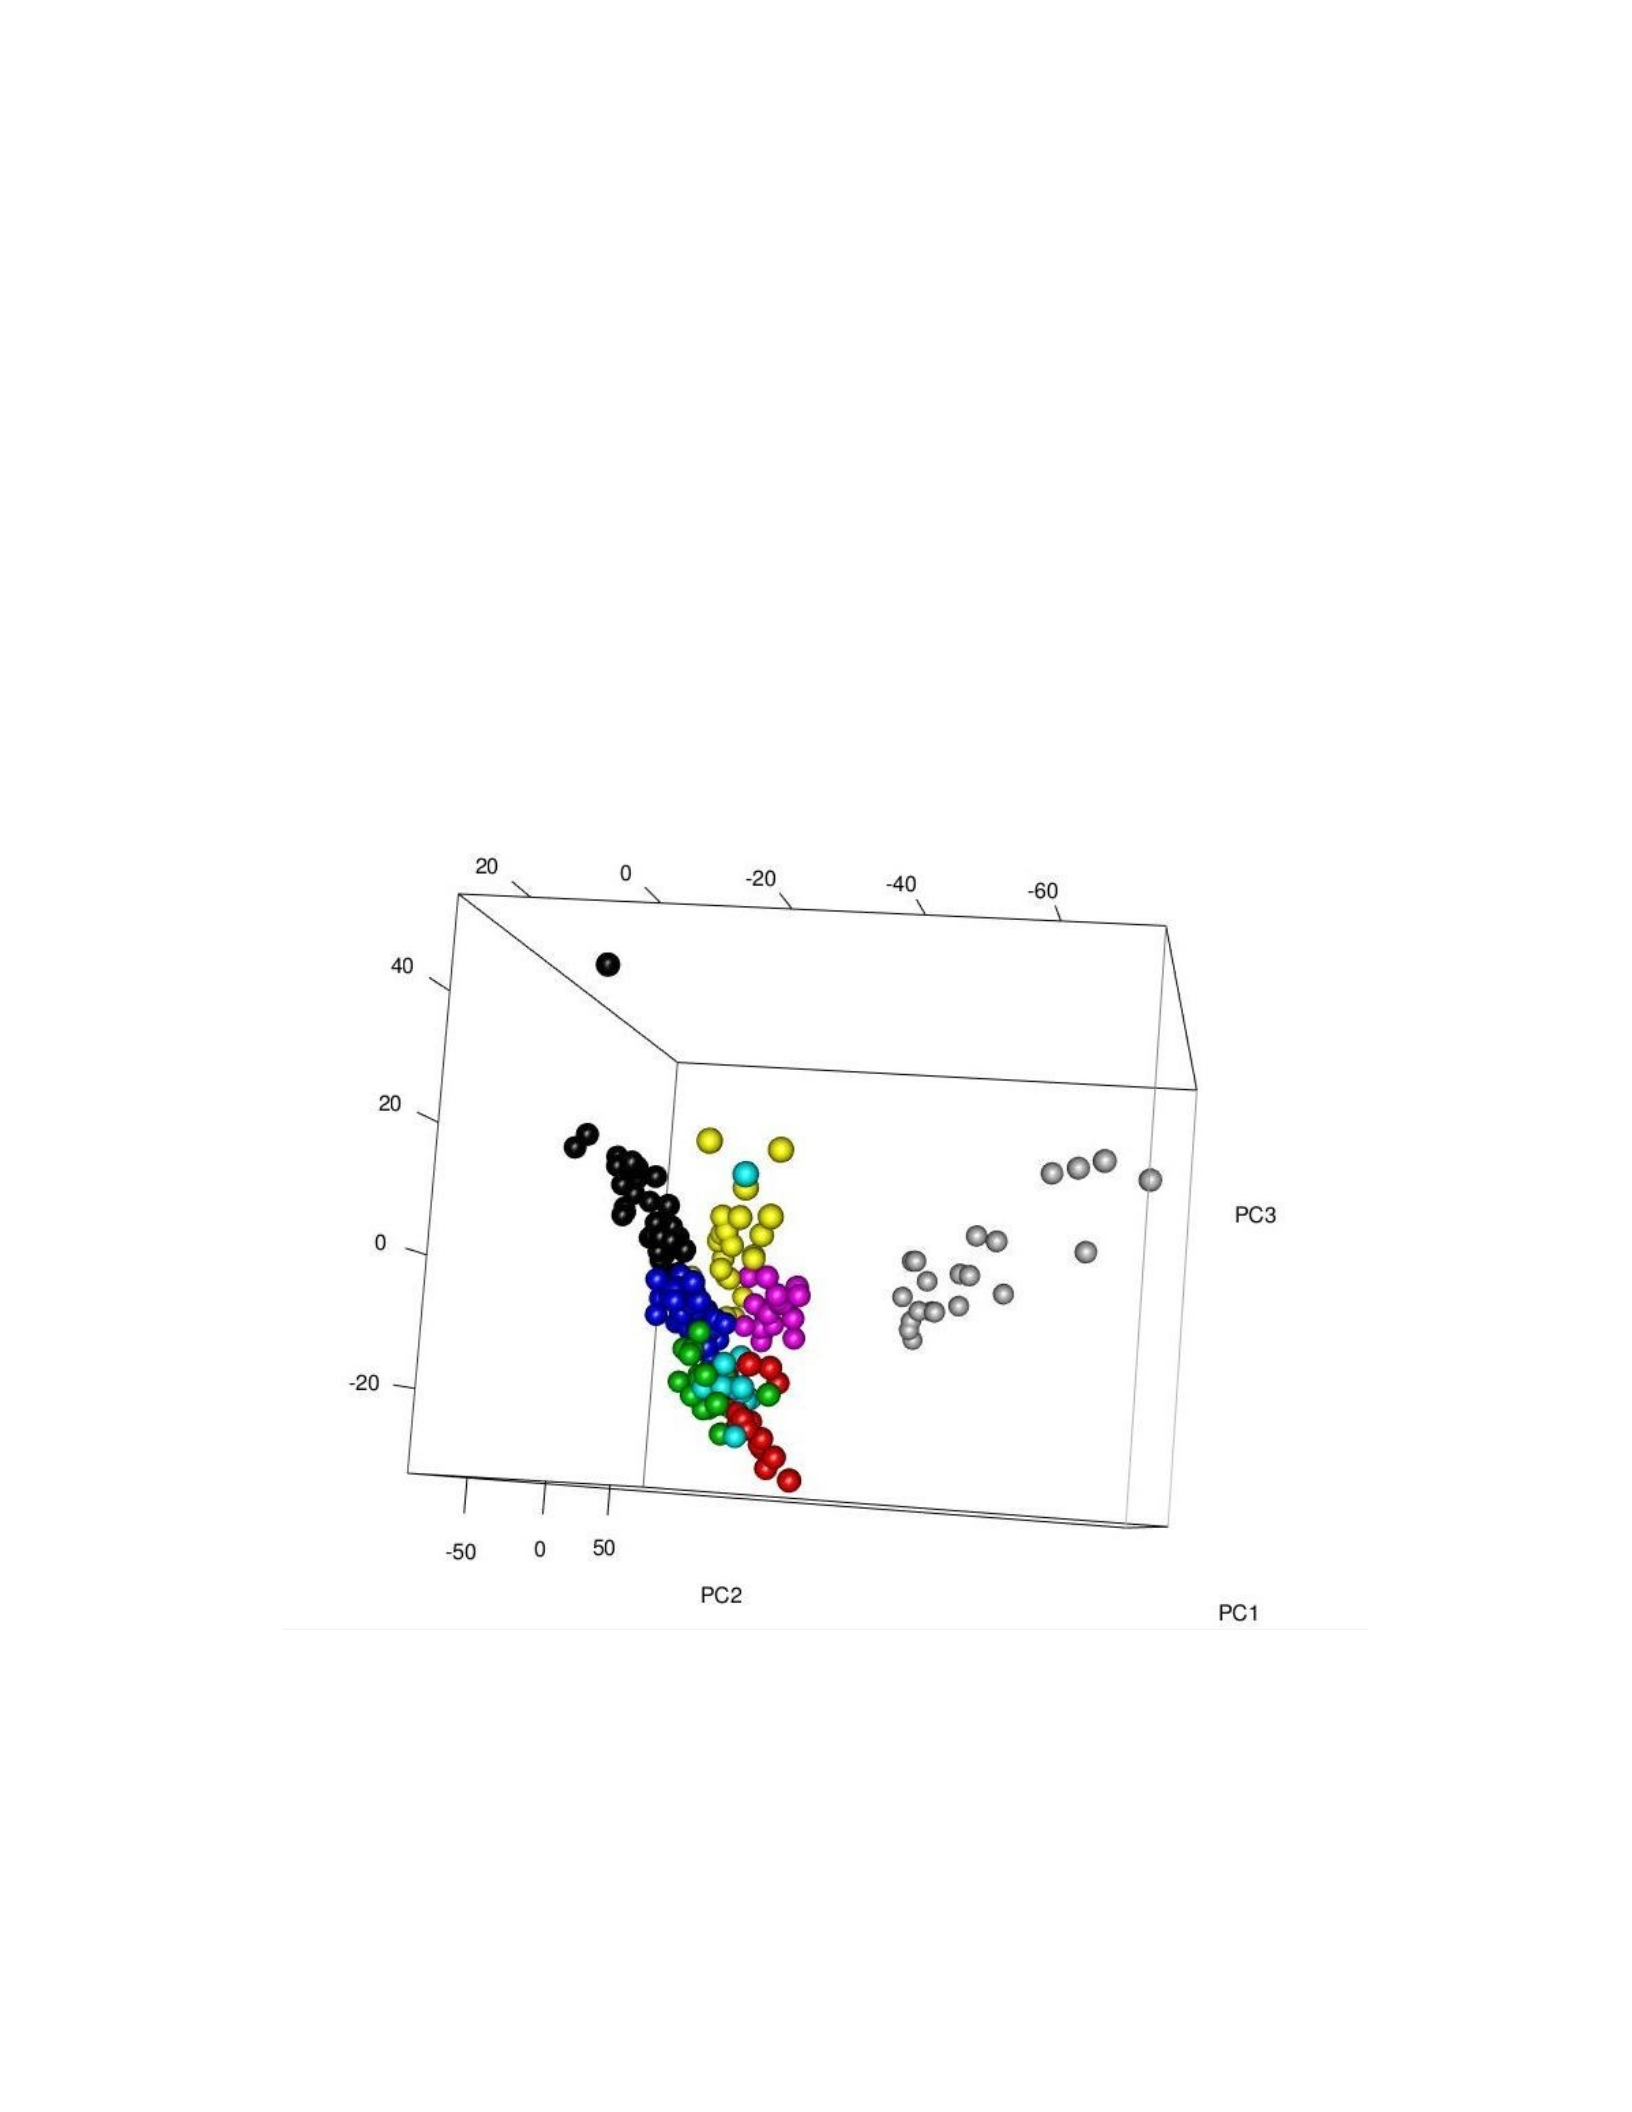
Figure 35. Spectra of authentic (red) and non-authentic samples
(except S8) showing the absorption band present due primarily to
the API sildenafil citrate (spectra are area normalized).
PCA-CVA showed that it was possible to classify samples based upon their infrared
spectrum into eight groups (seven non-authentic and one authentic). HOO-CV was used to
determine an estimated error rate of 2.16% using 12 PC’s. Figure 36 shows the 3D-scores plot.
In this figure, each grouping is colored differently and clustering based upon group assignment in
three dimensions is evident. Figure 37 shows the estimated error rates calculated as a function of
number of PC’s.
Figure 36. 3D-scores plot of infrared data (autoscaled).
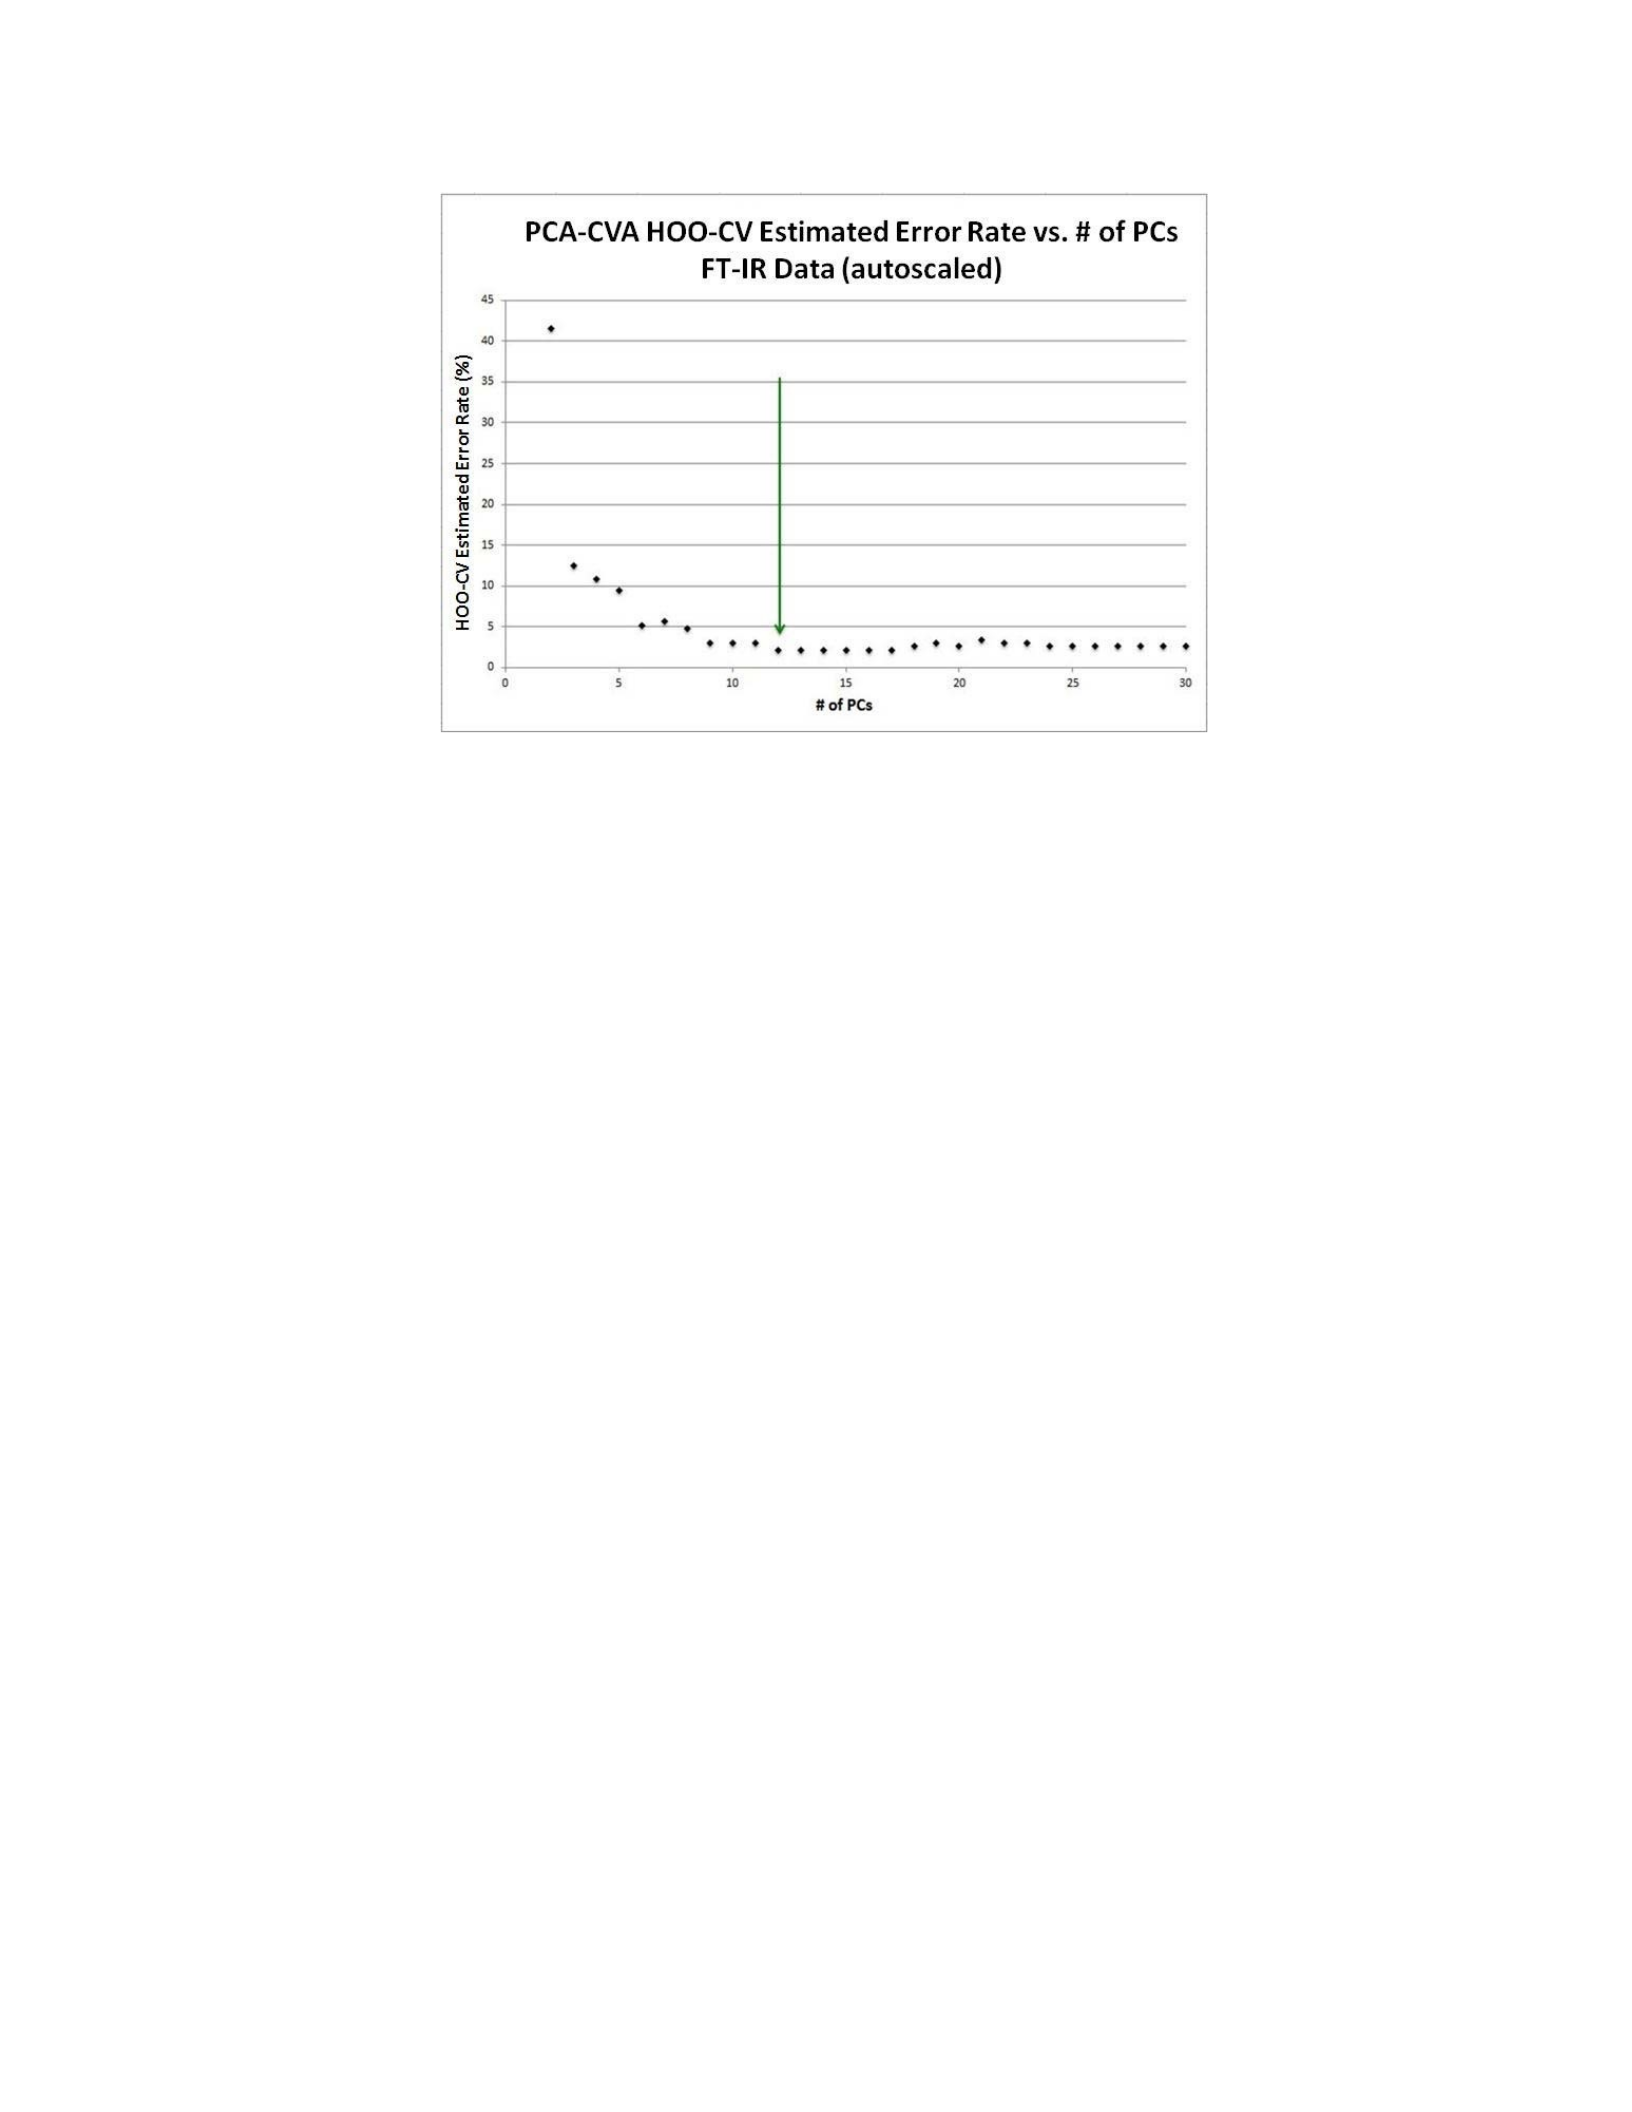
Figure 37. Estimated error rate displayed as a function of the
number of PC’s. An estimated error rate of 2.16% was observed at
12 PC’s.
The HOO-CV classification table is shown in Table 7. This data shows that four of the
five misclassifications occurred between groups 2 (sample I1) and 5 (sample I4). The spectra for
these two samples are shown in Figure 38. It is clear from these two spectra that the primary
difference between these samples as observed in the infrared spectra is in the water content of the
samples. The average spectrum of sample I4 contains less water than the average spectrum of
sample I1. This is an important observation because infrared analysis is sensitive to water, and
drug tablets are known to sorb different amounts of water based upon many factors. For this
reason, it is important that when grouping samples together, this fact be considered. In this study,
the region of the spectrum sensitive to water was not removed prior to chemometrics analysis
because water in pharmaceutical tablets may be present due to amorphous API. Even small
amounts of amorphous content can cause the sample to sorb more water than a crystalline form
of the drug. (Sacchetti, 2013) Amorphous content may be directly related to manufacturing
process (Hancock & Zografi, 1997) and may provide insight as to the source of the tablets.
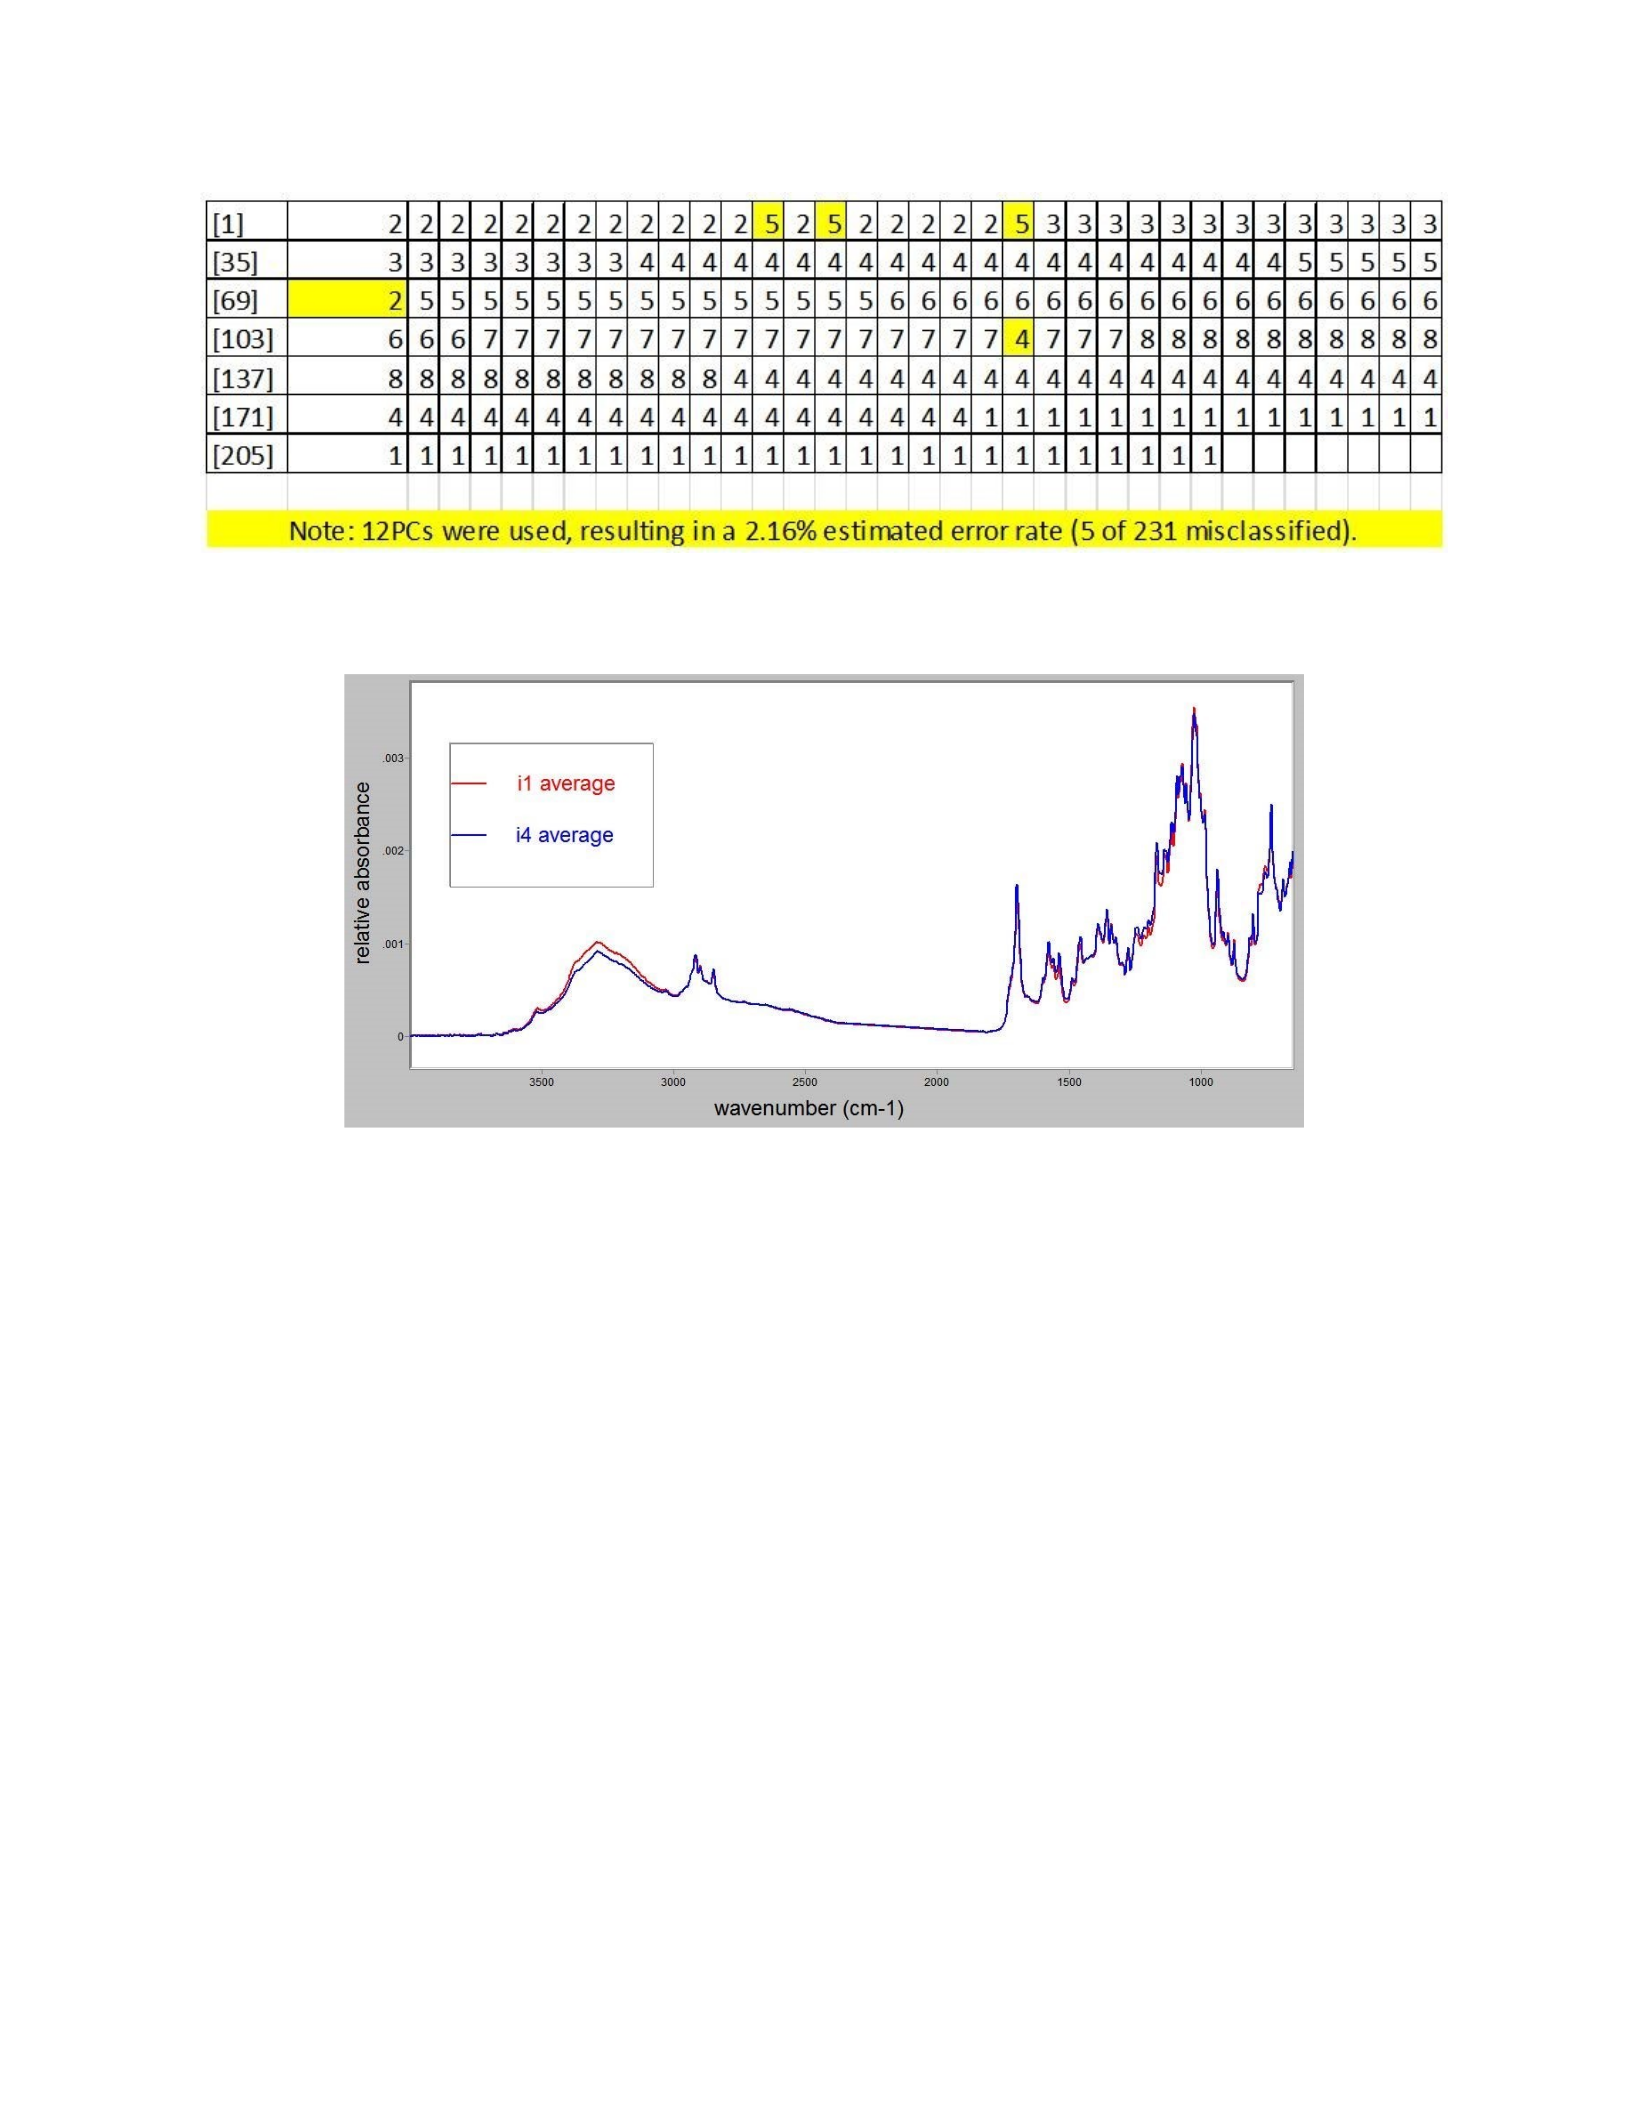
Table 7. HOO-CV estimated error-rate test results for PCA-CVA
analysis of infrared spectra.
Figure 38. Average infrared spectra of samples I1and I4 (spectra
area normalized).
Summary and Conclusion
This research shows that detection and identification of chemical and physical differences
in drug tablets using field-portable infrared spectroscopy provides a chemical signature or profile
that may be used to authenticate product. PCA-CVA HOO-CV applied to the infrared data
showed that infrared may also be used to differentiate non-authentic samples from each other.
This is important because it indicates that infrared analysis performed in the field may be used to
establish provenance. A visual comparison of the infrared spectra was used to differentiate
nonauthentic from authentic samples. Infrared spectra were reproducible within different tablets

of the same sample and may be used as a chemical fingerprint of a specific tablet and sample.
These spectra may be used to classify samples of unknown origin and provide investigators
insight into the sample’s manufacturing process and history.
CHAPTER 9. INVESTIGATION AND EVALUATION OF CHEMICAL AND PHYSICAL
PROPERTIES OF FINAL DOSAGE FORMS USING RAMAN SPECTROSCOPY
Introduction
The rationale for using Raman spectroscopy to evaluate counterfeit drugs is the same as
that for infrared spectroscopy (see chapter 8). The reason it is useful as a complement to infrared
spectroscopy is because these two methods have different selection rules. In order for a
vibrational mode to be active in the infrared region, the vibration must cause a change in the
permanent dipole moment of the molecule. The dipole moment is the product of the charge of
the dipole and the charge separation difference. On the other hand, for a vibration to be active in
the Raman effect, the polarizability of the molecule must change during the vibration.
Polarizability is the value of the induced dipole moment divided by the strength of the field that
causes the induced dipole moment. In other words, the electron cloud of the molecule must be
more readily deformed in one extreme of the vibration than in the other. (Willard, Merritt Jr.,
Dean, & Settle Jr., 1988) Due to these differences in selection rules, some samples are more
amenable to infrared analysis and some are more amenable to Raman analysis. For instance,
samples in an aqueous dispersant may have limited success when analyzed using infrared
spectroscopy. This is due to the strong infrared absorption of water which may overwhelm the
spectrum and mask the presence of other components in the sample. When analyzed using
Raman spectroscopy, water does not generate a strong Raman spectrum and will not usually
interfere with the Raman measurement of other components in the sample. On the other hand,

samples will frequently exhibit a strong fluorescence signal across the Raman spectrum which
may interfere with spectral interpretation and comparison. Raman spectroscopy also may cause
problems when analyzing colored (and some non-colored) samples because the sample may
burn. This is because the sample will absorb the light at the laser wavelength. (Arnó, Frunzi,
Kittredge, & Sparano, 2014) This energy is dissipated as heat due to non-radiative emissions.
(Frunzi, 2014) In these instances, infrared analysis may be more appropriate. Lasers are
required for Raman systems in order to generate a sufficient Raman scatter for detection.
Raman has a significant advantage in some applications, though. Raman signal may be
directly collected from samples through some transparent glass vials and plastic bags. This
capability has likely contributed to its success in field-portable applications. For these reasons,
infrared and Raman spectroscopy are considered complimentary techniques. When used
together, they can provide information that may enable a more complete understanding of the
chemical and physical properties of the samples. (Otieno-Alego & Speers, 2011)
Data generated from Raman analysis can be evaluated many different ways when applied
to the analytical challenges associated with counterfeit drugs. In this study, Raman spectra were
compared to differentiate authentic from non-authentic samples. Reproducibility of spectra
collected from a single tablet, as well as amongst different tablets of the same sample was also
evaluated. Samples were classified and the discrimination potential of Raman as a tool for the
analysis of counterfeit drugs was evaluated using PCA-CVA HOO-CV.
Experimental Details
Instrumentation
Raman spectra of the core matrices of tablets were collected using a 785L Raman
spectrometer (Wasatch Photonics, Durham, NC). Performance of the spectrometer was tested at
the start and end of every work day by analyzing cyclohexane and verifying peak position. At all
times, the system passed this check. The 785L is a small portable instrument. However, it is not
specified for ruggedization. It was used as part of this study primarily due to its size, portability
and performance characteristics. Some of the specifications for this system that are important
reasons for its use in this study are shown in Table 8. Figure 39 is an image of the model of
Raman instrument used.
Raman 785L Specifications of Importance to this Study
Compact size 5 in x 6.5 in x 2 in housing
Weight 1.2 kg
Laser type 785-nm single-mode laser, diode stabilized with volume Bragg grating
Diffraction grating HD 1624 LP/mm volume phase grating
Instrument design Lens-based spectrograph
f/# 1.3
Interface USB 2.0
Integration time 1 ms to 2 s
Resolution 10 cm-1 with 50-μm slit
Sample spot size <50 μm
Table 8. Raman 785L specifications of importance to this study.
Figure 39. Photograph of Wasatch Raman 785L spectrometer.
(http://wasatchphotonics.com/systems/vph-spectrometers/
ramanflourescence/stroker-785l/)
Raman Method
Approximately 25 mg of gently ground tablet core were transferred to a small, clear, glass
vial. The vial was placed in the vial sample chamber of the spectrometer and its position was
adjusted slightly to maximize signal intensity for each measurement. A black drape was placed
over the sampling chamber during scanning to minimize stray light. Ten scans were collected
with a resolution of 10 cm-1 and integration time of 1000 milliseconds for each scan. The
instrument’s Dash software was used to calculate an average of these ten scans. The laser was
then turned off and ten dark scans were collected using the same integration time. The system
software was used to calculate the average of these ten scans. The data from the spectrometer
including each of the individual scans and the average for both the sample spectra and the dark-
scan spectra were then transferred in a single worksheet to Microsoft Excel. Using Microsoft
Excel, the dark spectrum average value at each frequency was subtracted from the sample

spectrum average value at each frequency. The subtraction values along with their respective
frequencies were copied into a new worksheet in Microsoft Excel.
Data Analysis and Chemometrics
The “Excel Exchange” function of Thermo Grams was used to convert this worksheet
data into an *.spc file. Each *.spc file created was opened in Thermo Grams and then baseline
corrected using a quintic multipoint baseline correction. For each tablet, all spectra were
averaged using the array basic “average” function in Grams. For each sample, all spectra from
each tablet of the sample were averaged using the array basic “average” function in Grams. The
chemometric methods of PCA and CVA were applied to investigate the classification and
discrimination power of Raman spectroscopy. Non-authentic samples were sorted into seven
groups based upon physical appearance, dimensions, and receipt date; authentic samples were
categorized into a single group. The deconvolution data, specifically the peak positions and
areas, also were analyzed using HOO-CV PCA-CVA to determine an estimated error rate for the
classification of sildenafil citrate tablets.
Results and Discussion
It was possible to differentiate the authentic samples from all non-authentic samples using
Raman spectroscopy. In most cases, differences in spectra were small but reproducible. Using
PCA-CVA, 3D-scores plots were created and visual separation of samples was not evident in
these plots. Although separation was not evident in three dimensions for the Raman data, the
estimated error rate calculated using HOO-CV was lower than it was for the infrared data and
calculated to be 0.56% using 14 PC’s. This lower estimated error rate indicates that Raman
analysis may be just as useful as infrared analysis to establish provenance. Not only can Raman
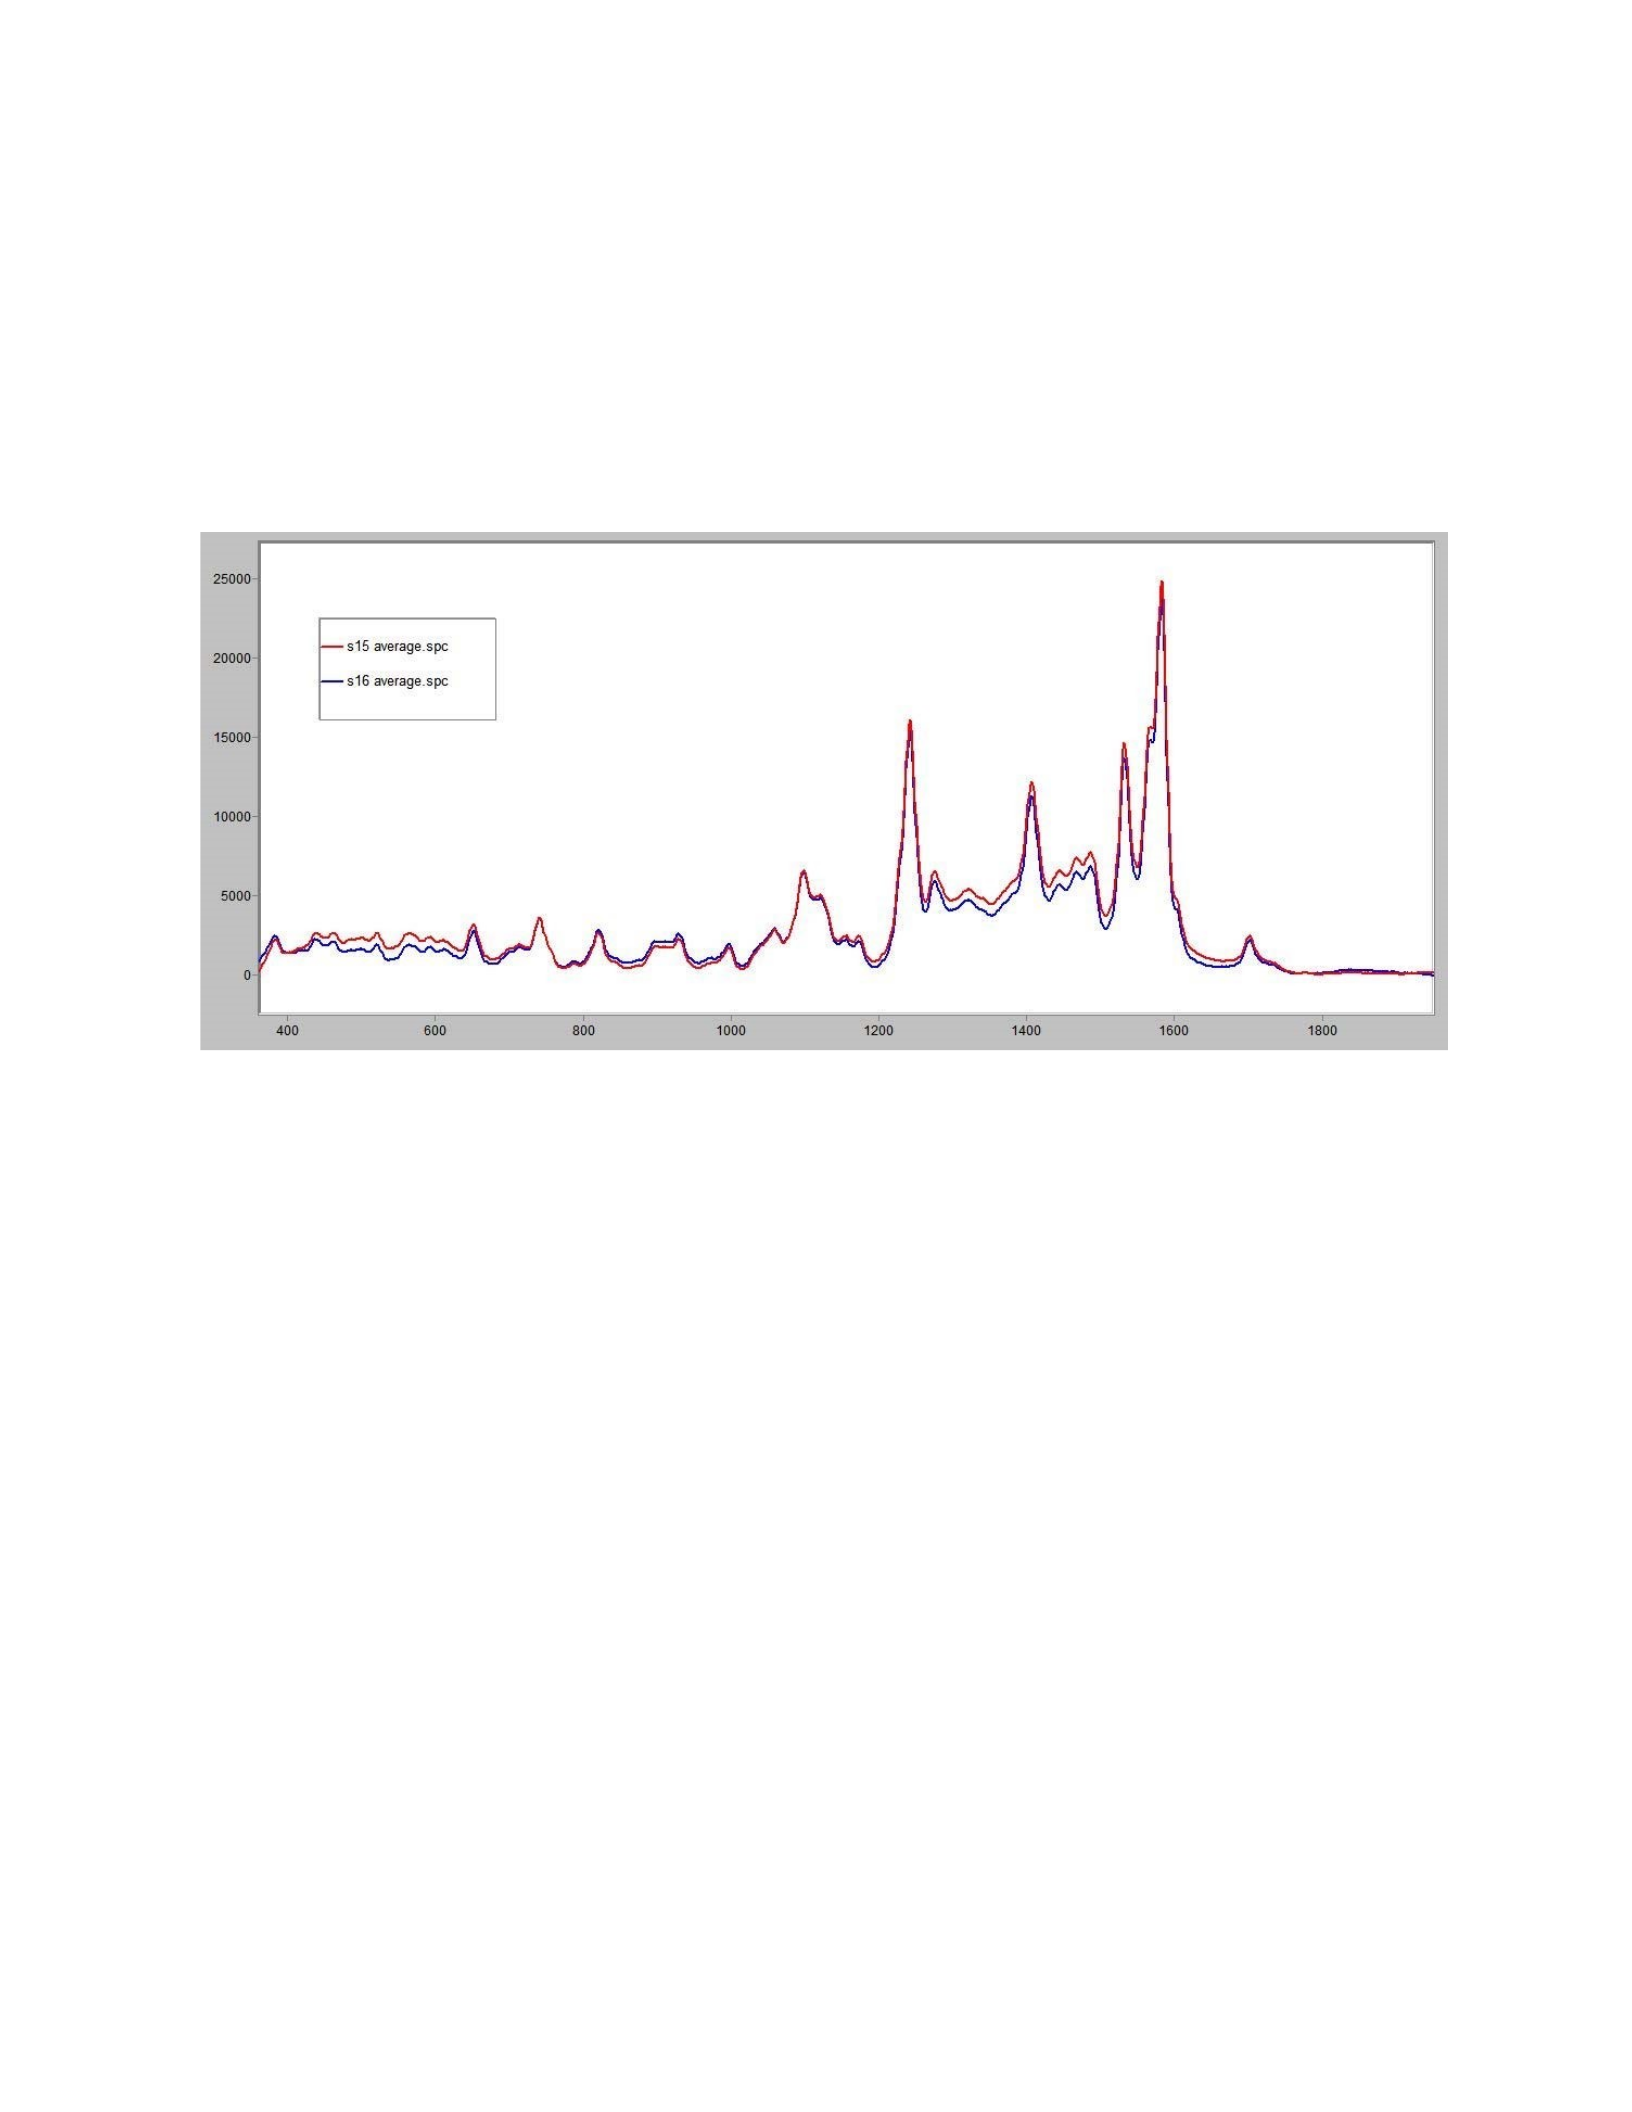
be used to differentiate authentic from non-authentic samples, but it also may be used to sort non-
authentic samples and ultimately lead to the establishment of provenance. Figure 40 shows the
average spectra collected from two different samples of authentic VIAGRA. These spectra are
consistent with each other. The same bands and were present in each sample across the
Raman spectrum with very similar band intensities.
Figure 40. Raman spectral averages of samples S15 (red) and S16
(blue). Both samples are authentic VIAGRA.
Figures 41 through 49 show the average Raman spectrum of an authentic sample
compared with the average spectrum from each of the nine non-authentic samples. Note that the
differences between the authentic and non-authentic samples were relatively small. This
indicates that the chemical composition of the non-authentic samples was similar but not
identical to the chemical composition of the authentic samples. However, the spectra collected
from different tablets of the same sample were consistent with each other. It was possible to
differentiate all non-authentic samples from the authentic samples.
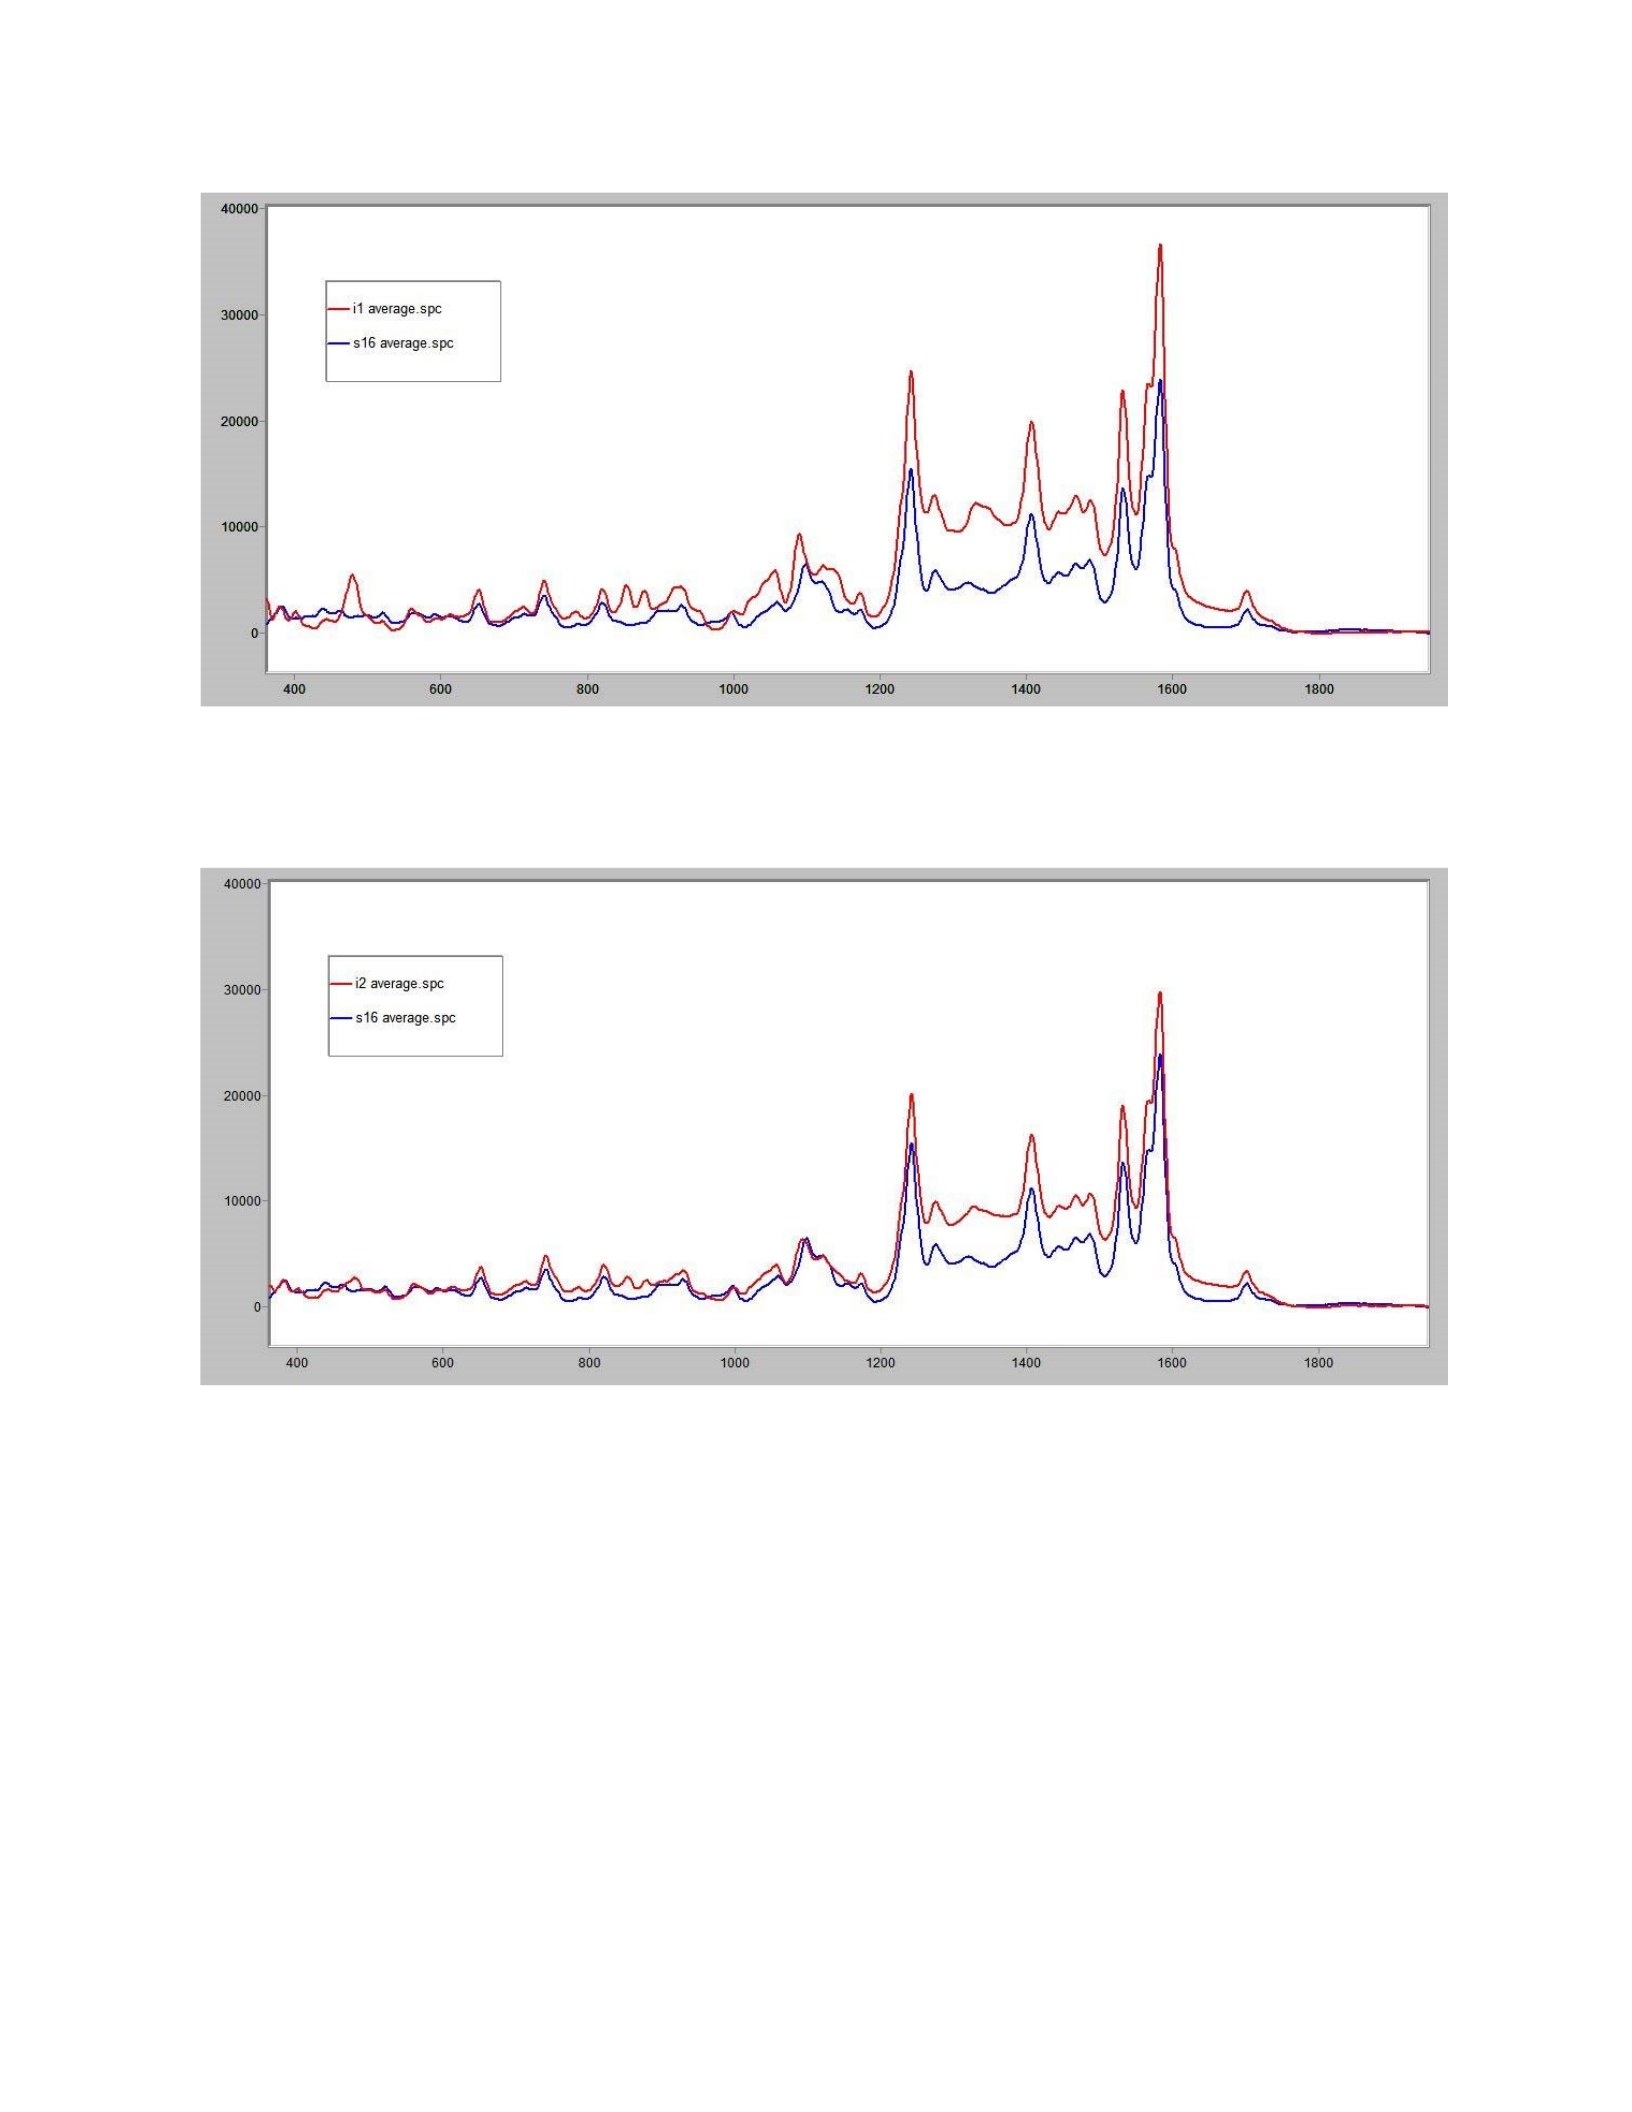
Figure 41. Raman spectrum of authentic sample (blue) compared
with non-authentic I1 (red).
Figure 42. Raman spectrum of authentic sample (blue) compared
with non-authentic I2 (red).
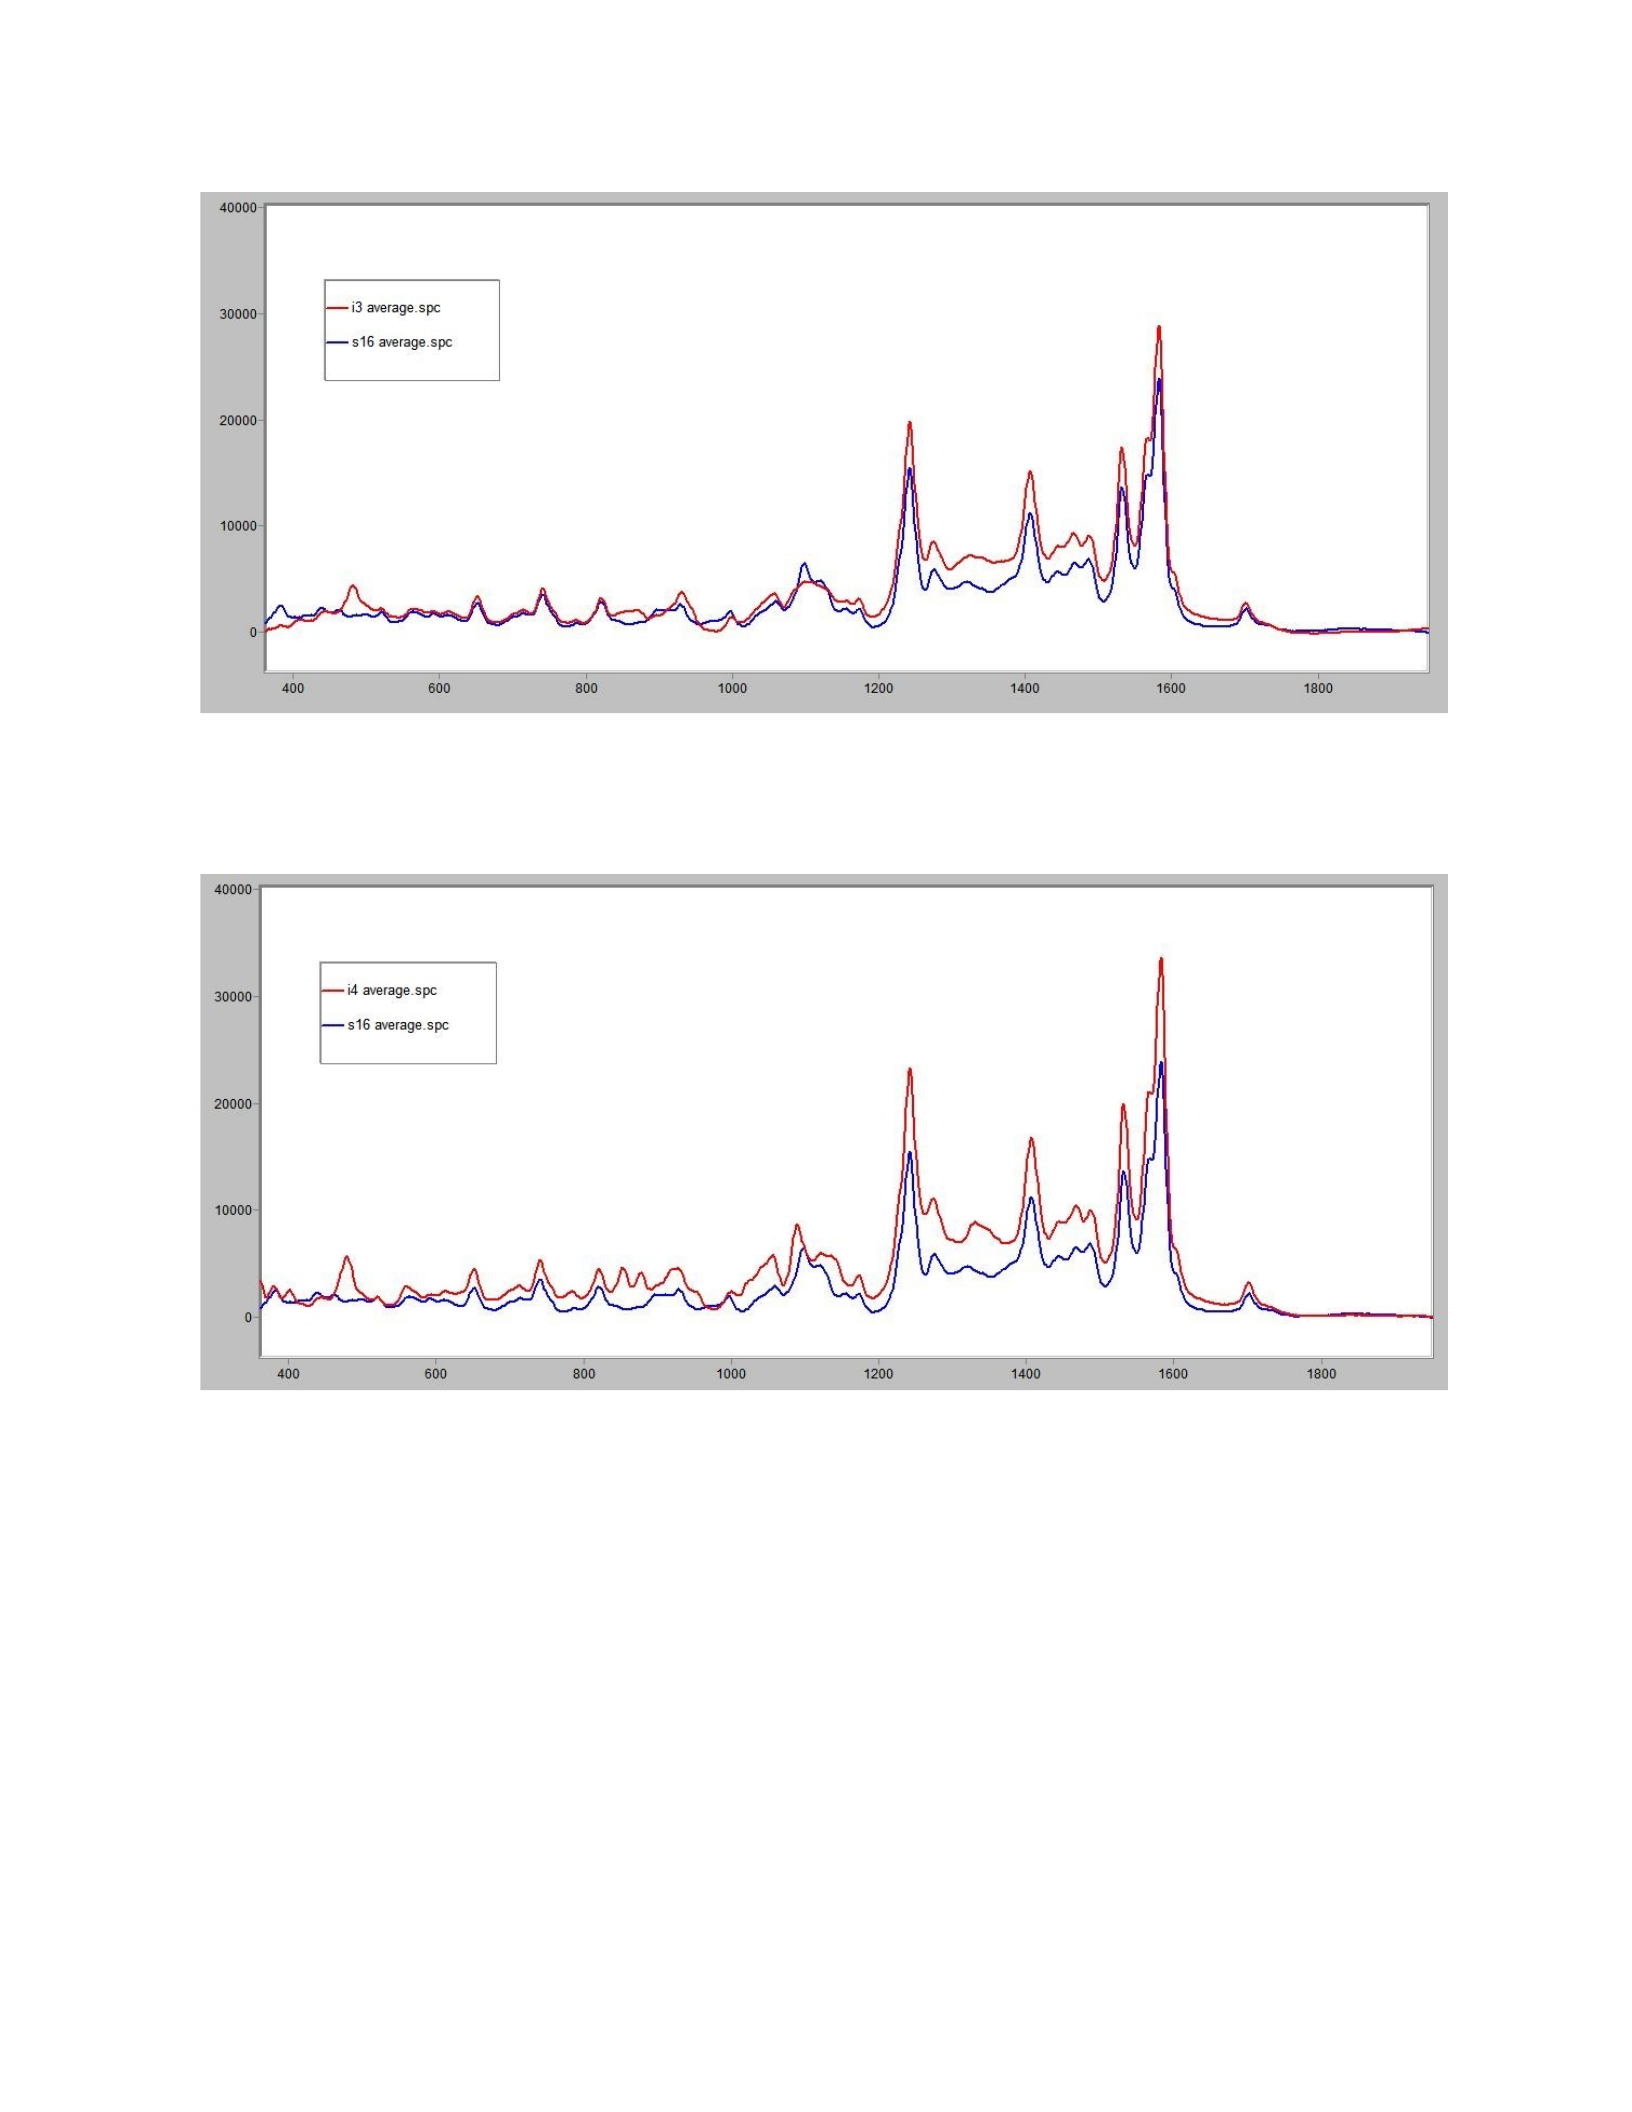
Figure 43. Raman spectrum of authentic sample (blue) compared
with non-authentic I3 (red).
Figure 44. Raman spectrum of authentic sample (blue) compared
with non-authentic I4 (red).
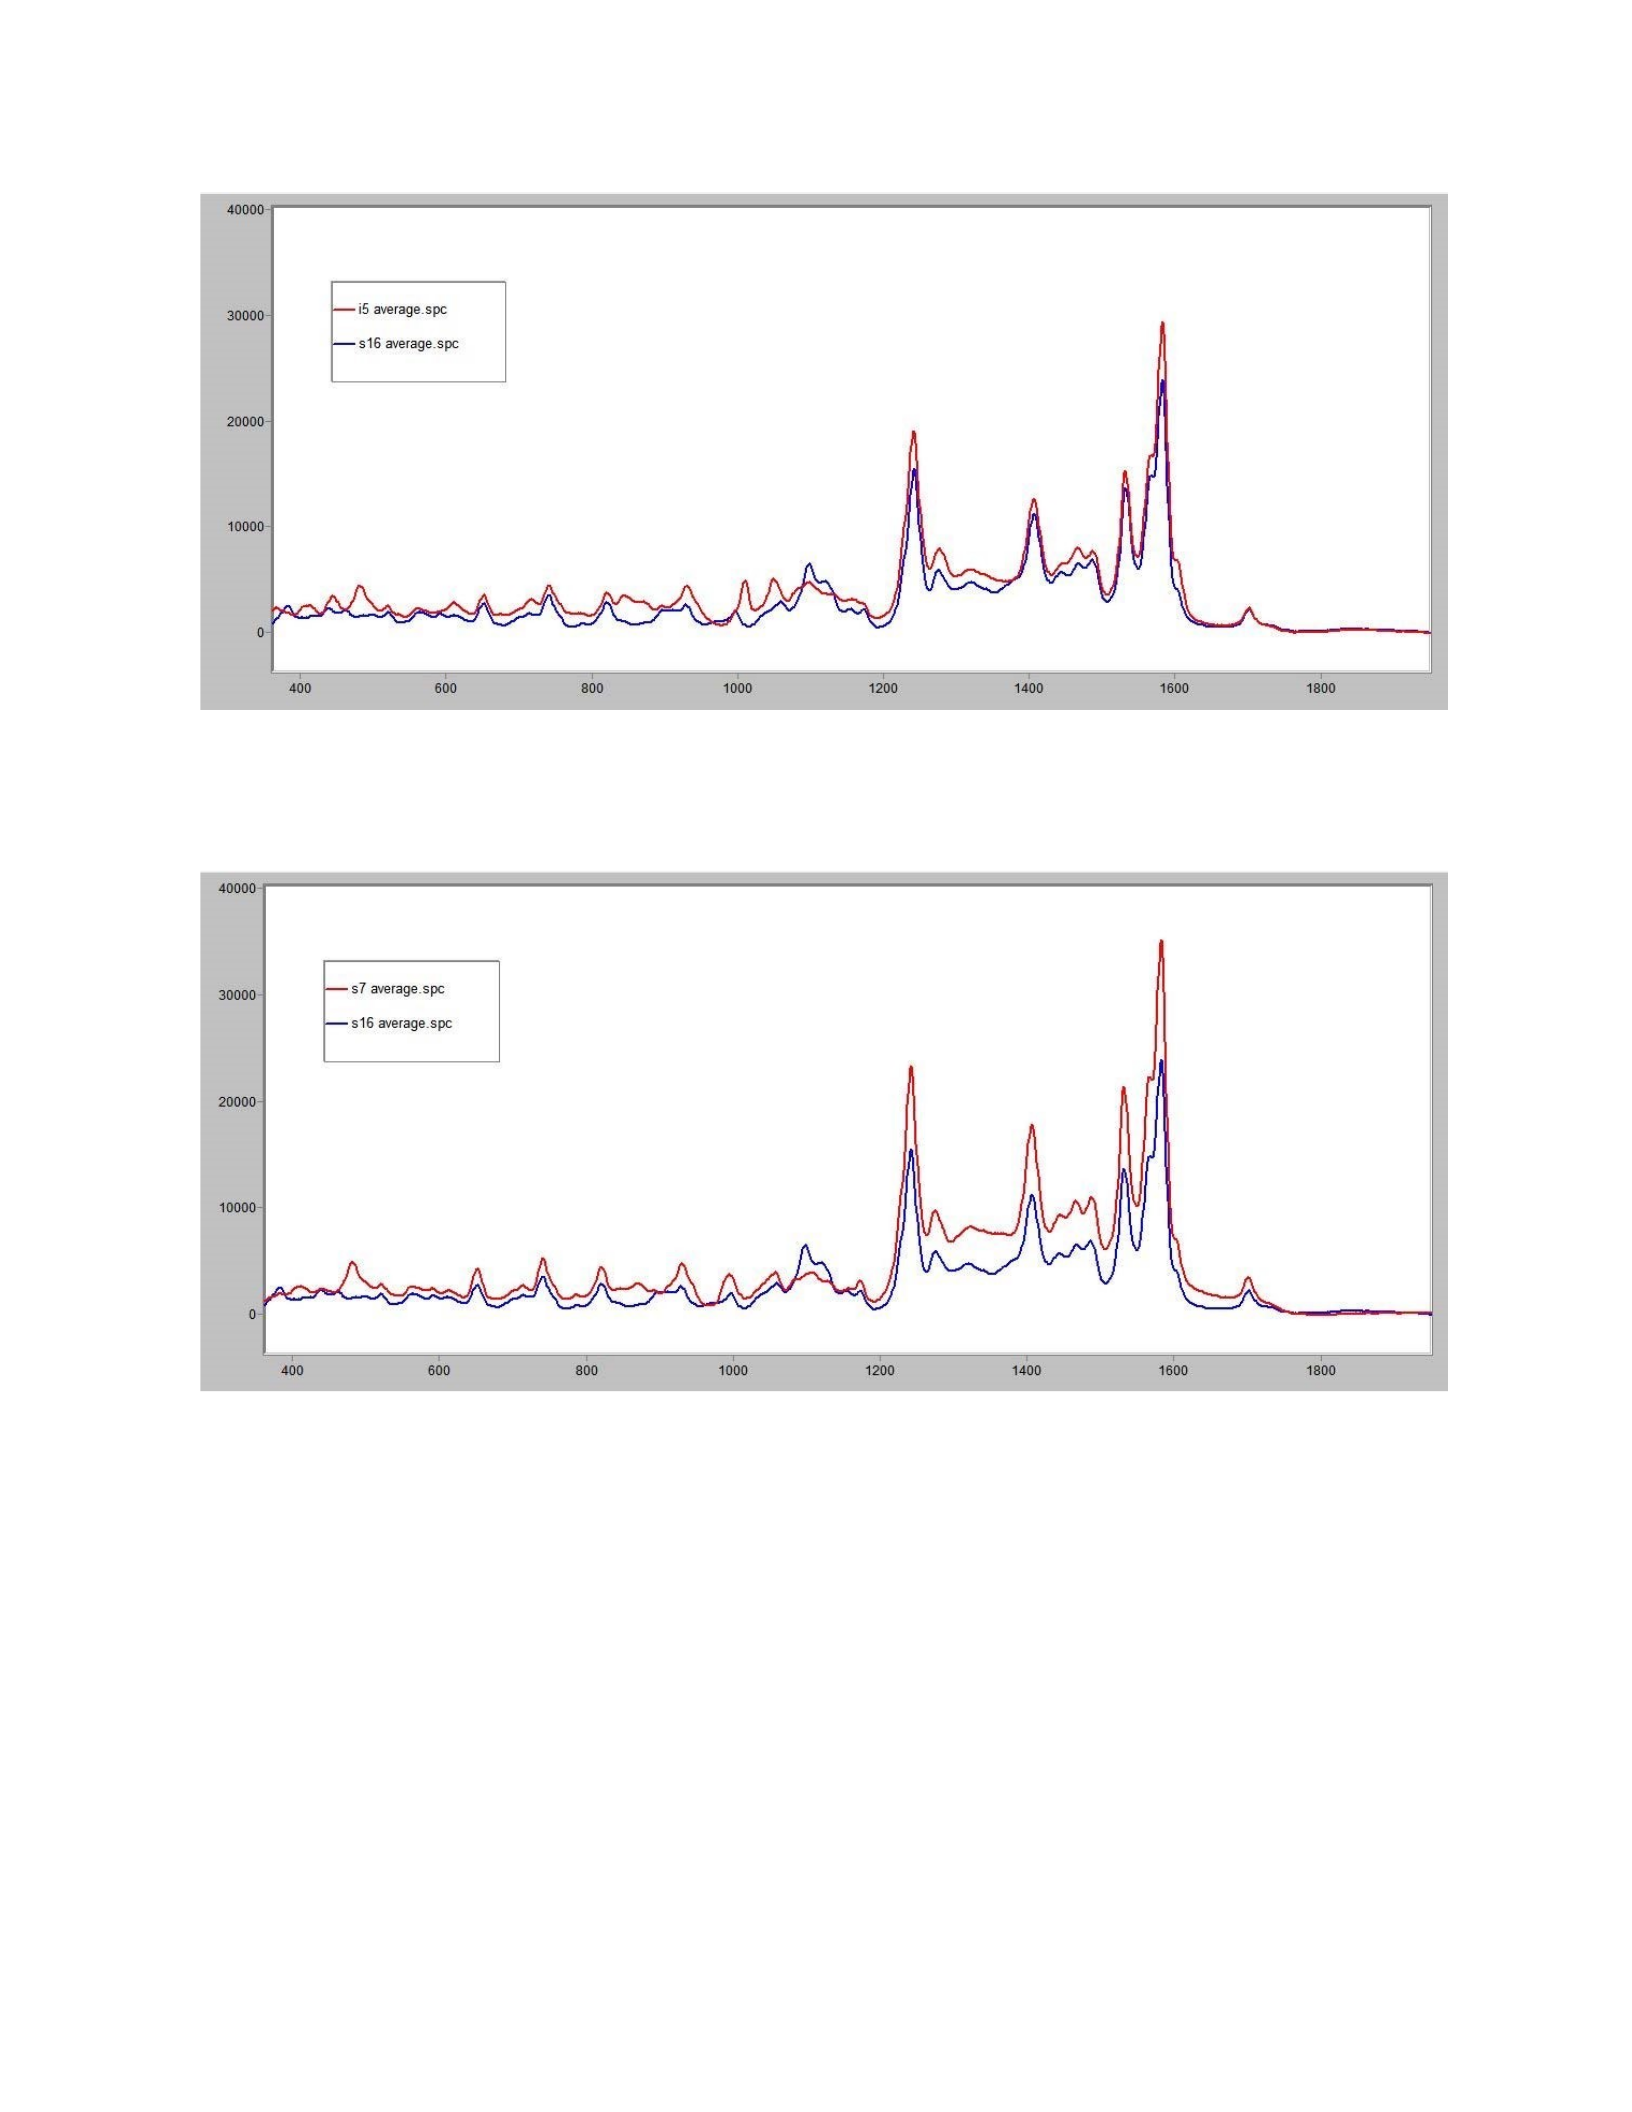
Figure 45. Raman spectrum of authentic sample (blue) compared
with non-authentic I5 (red).
Figure 46. Raman spectrum of authentic sample (blue) compared
with non-authentic S7 (red).
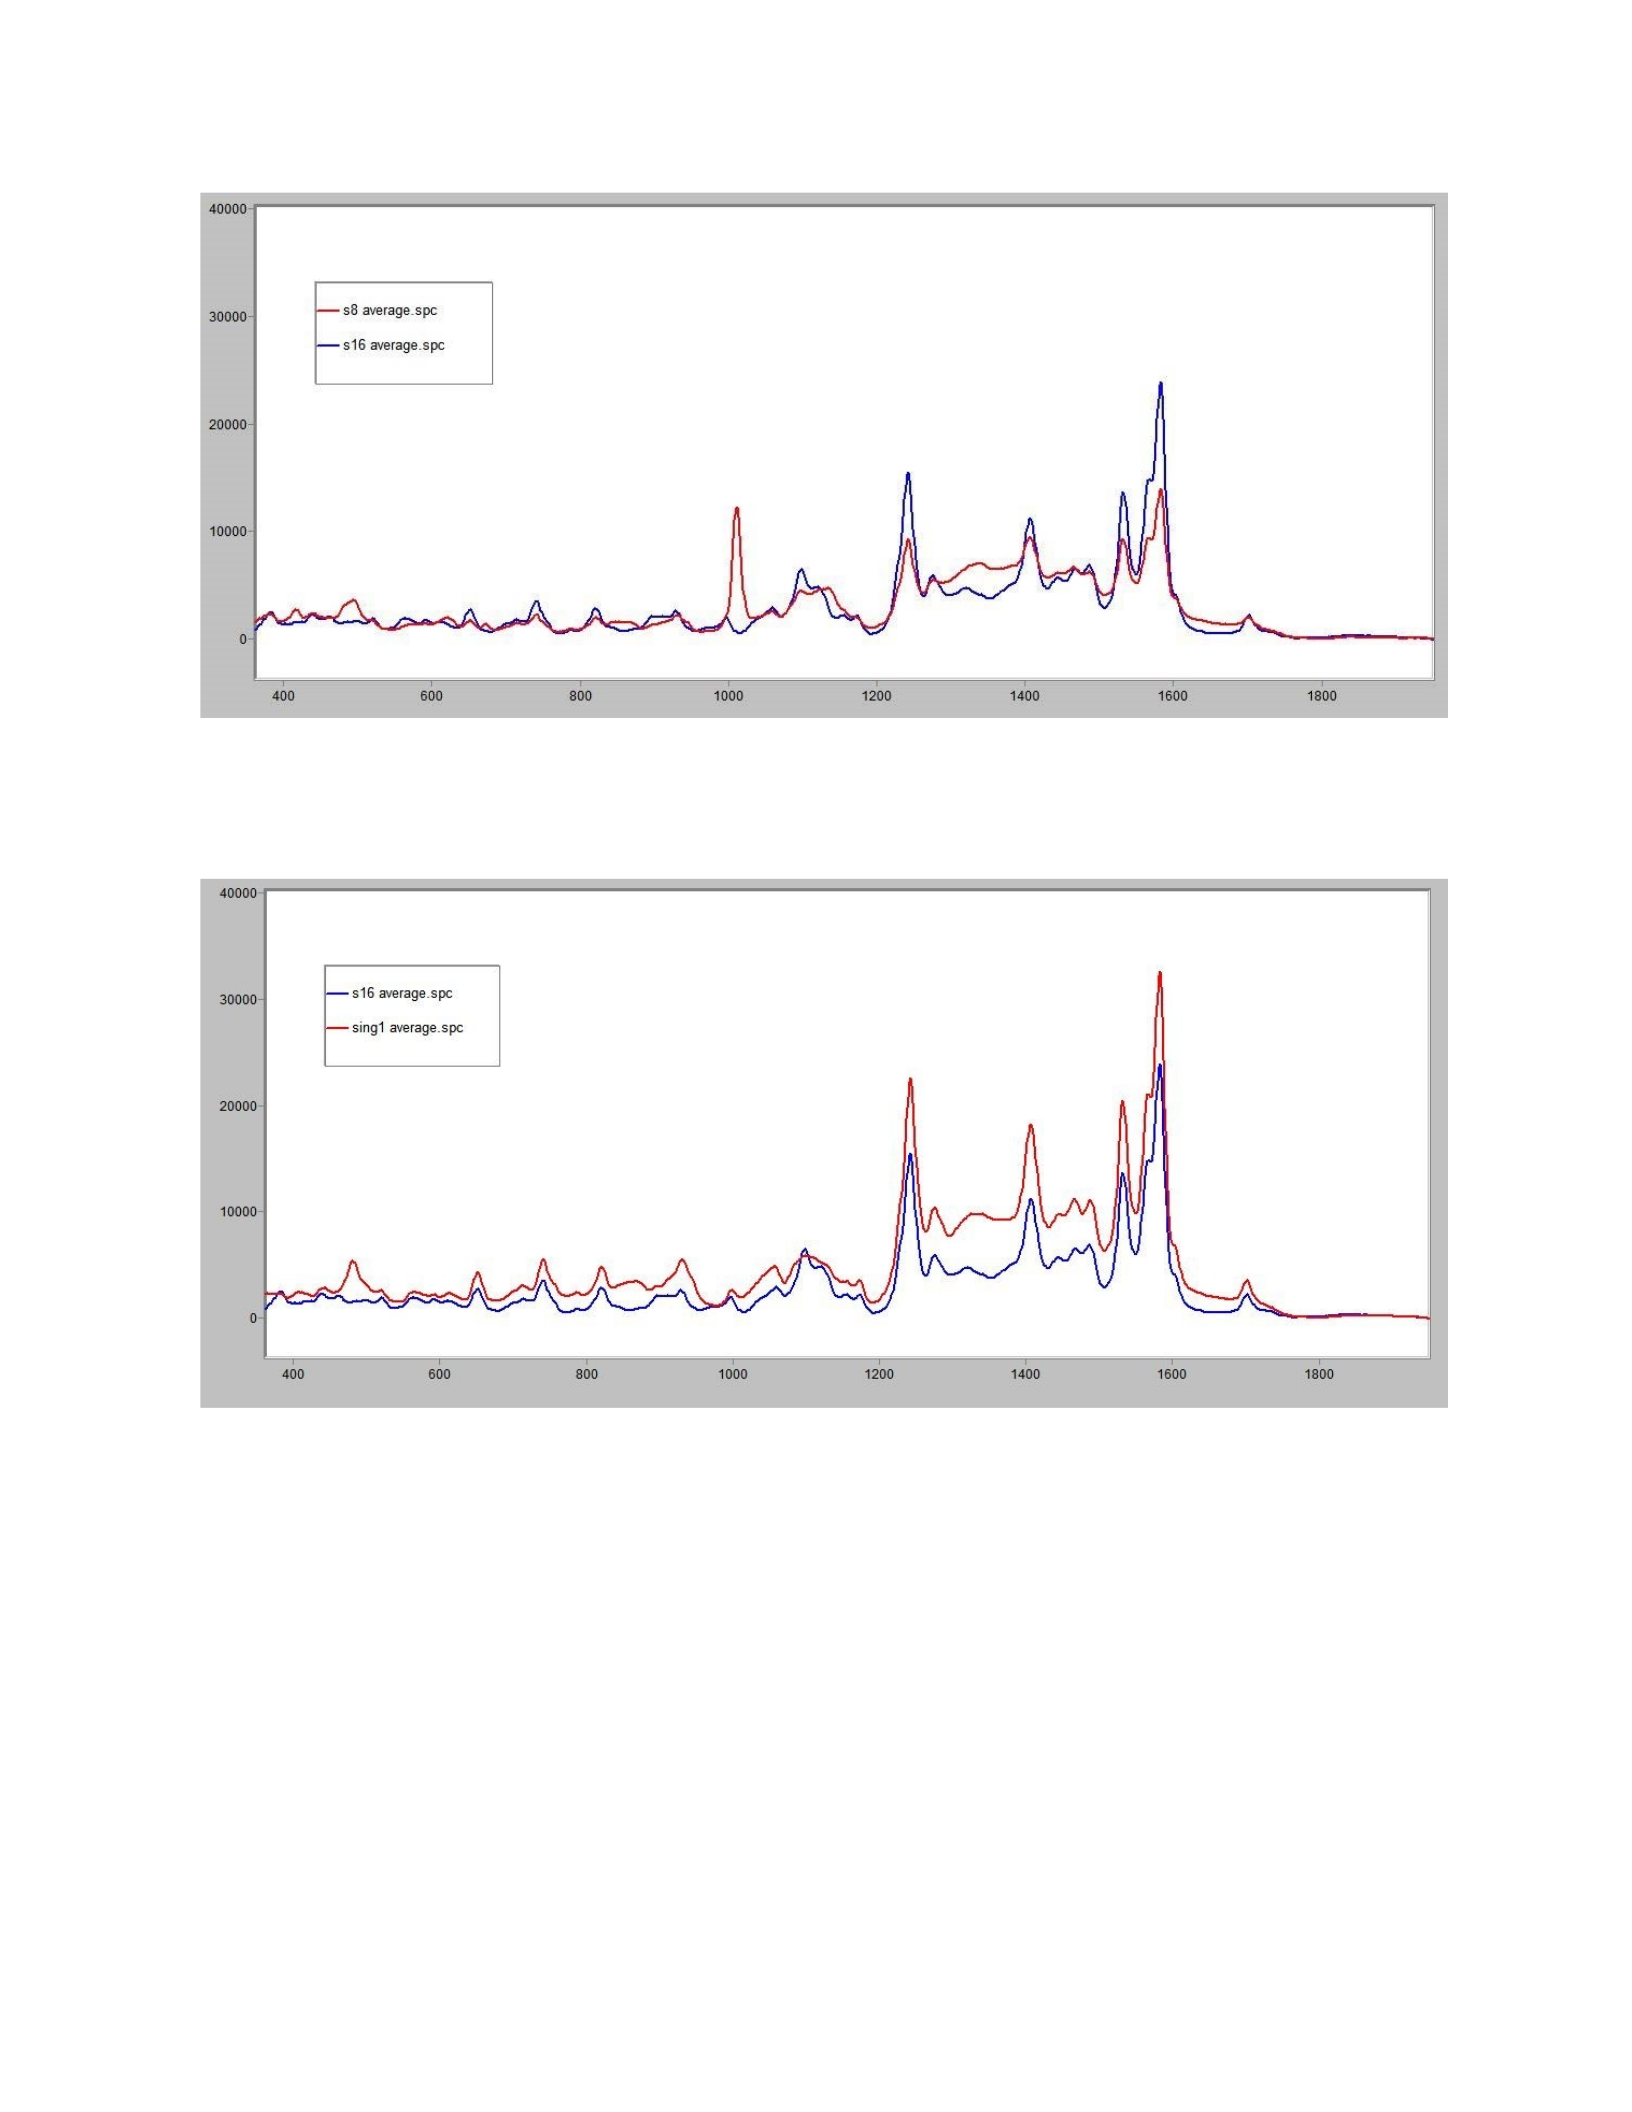
Figure 47. Raman spectrum of authentic sample (blue) compared
with non-authentic S8 (red).
Figure 48. Raman spectrum of authentic sample (blue) compared
with non-authentic SING1 (red).
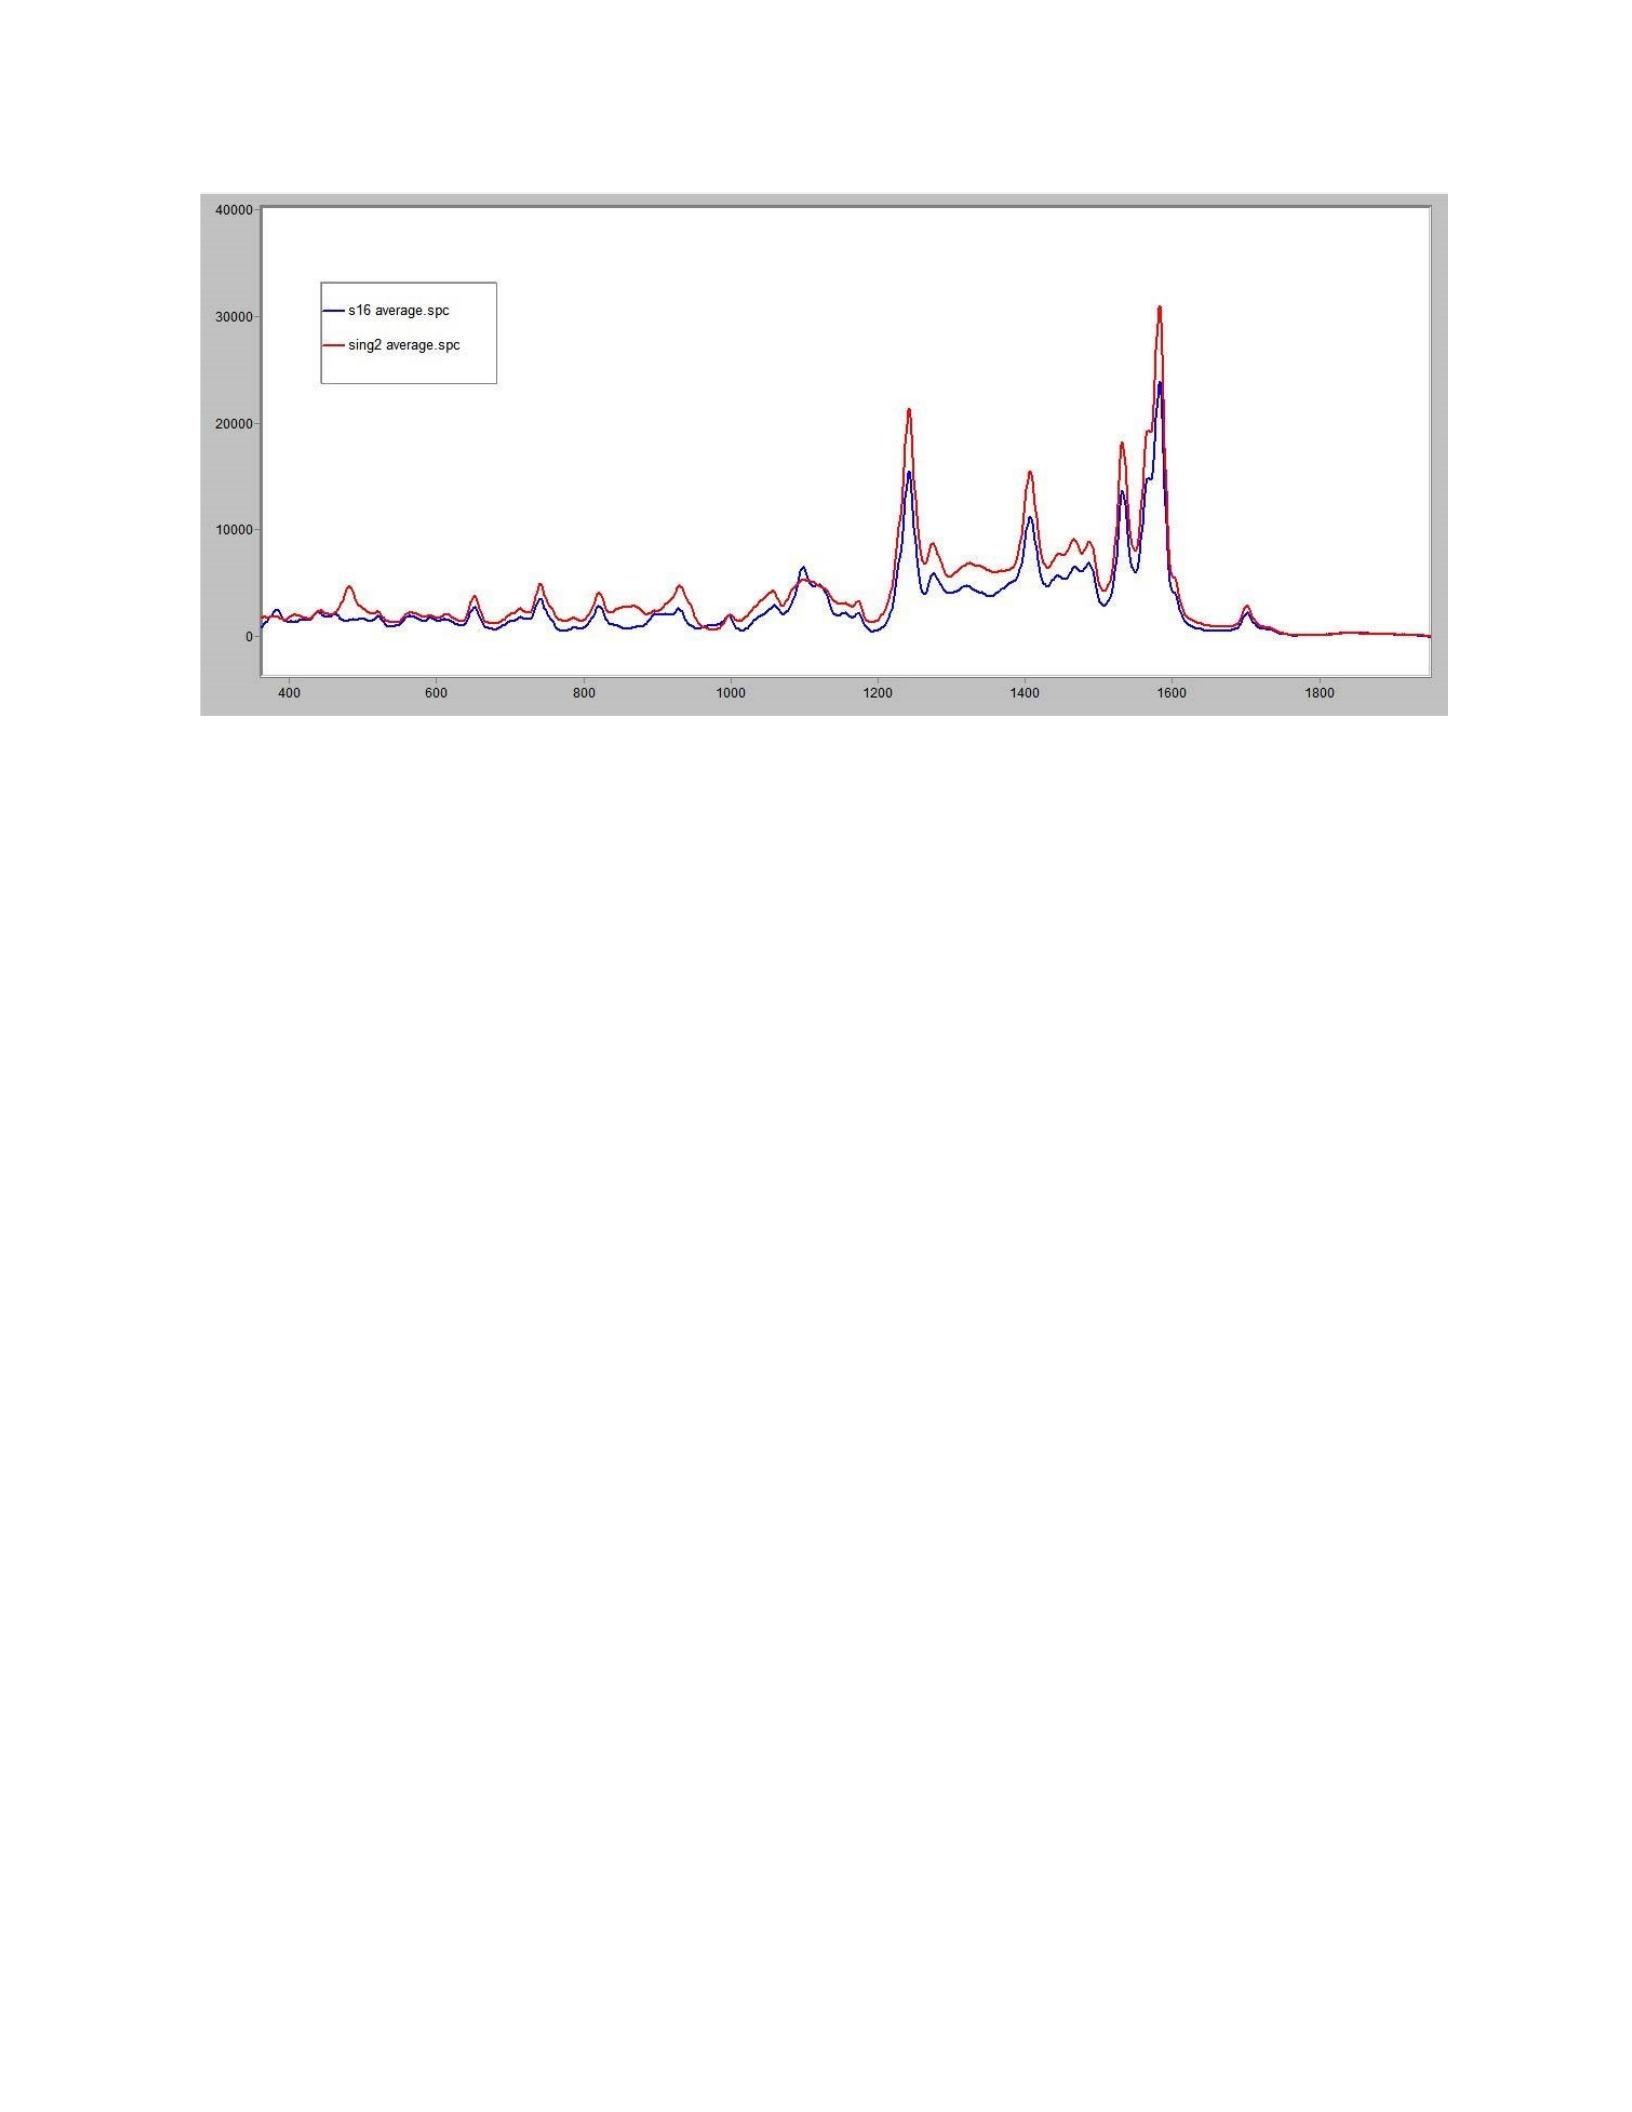
Figure 49. Raman spectrum of authentic sample (blue) compared
with non-authentic SING2.
Although only the average spectrum for each sample is shown in the preceding nine
figures, all of the spectra collected from each tablet and from the same sample were consistent
with each other. There was some variance in the intensities of some of the Raman bands within a
sample, but there wasn’t a single instance where two spectra from the same sample contained
different Raman bands.
PCA-CVA showed that it was possible to classify samples based upon their Raman
spectrum into eight groups (seven non-authentic and one authentic). HOO-CV was used to
determine an estimated error rate of 0.56% using 14 PC’s.
Figure 50 shows the 3D-scores plot from PCA-CVA. In this figure each group is colored
differently and, unlike with the infrared spectra where clustering based upon group assignment
was evident in three dimensions, no clustering is observed in the Raman data. However, figure
51 shows the estimated error rates calculated as a function of number of PC’s for the Raman, and
even though clustering is not evident in three dimensions, Raman spectroscopy was capable of
successfully classifying most samples. This difference in clustering observed in the 3D-scores
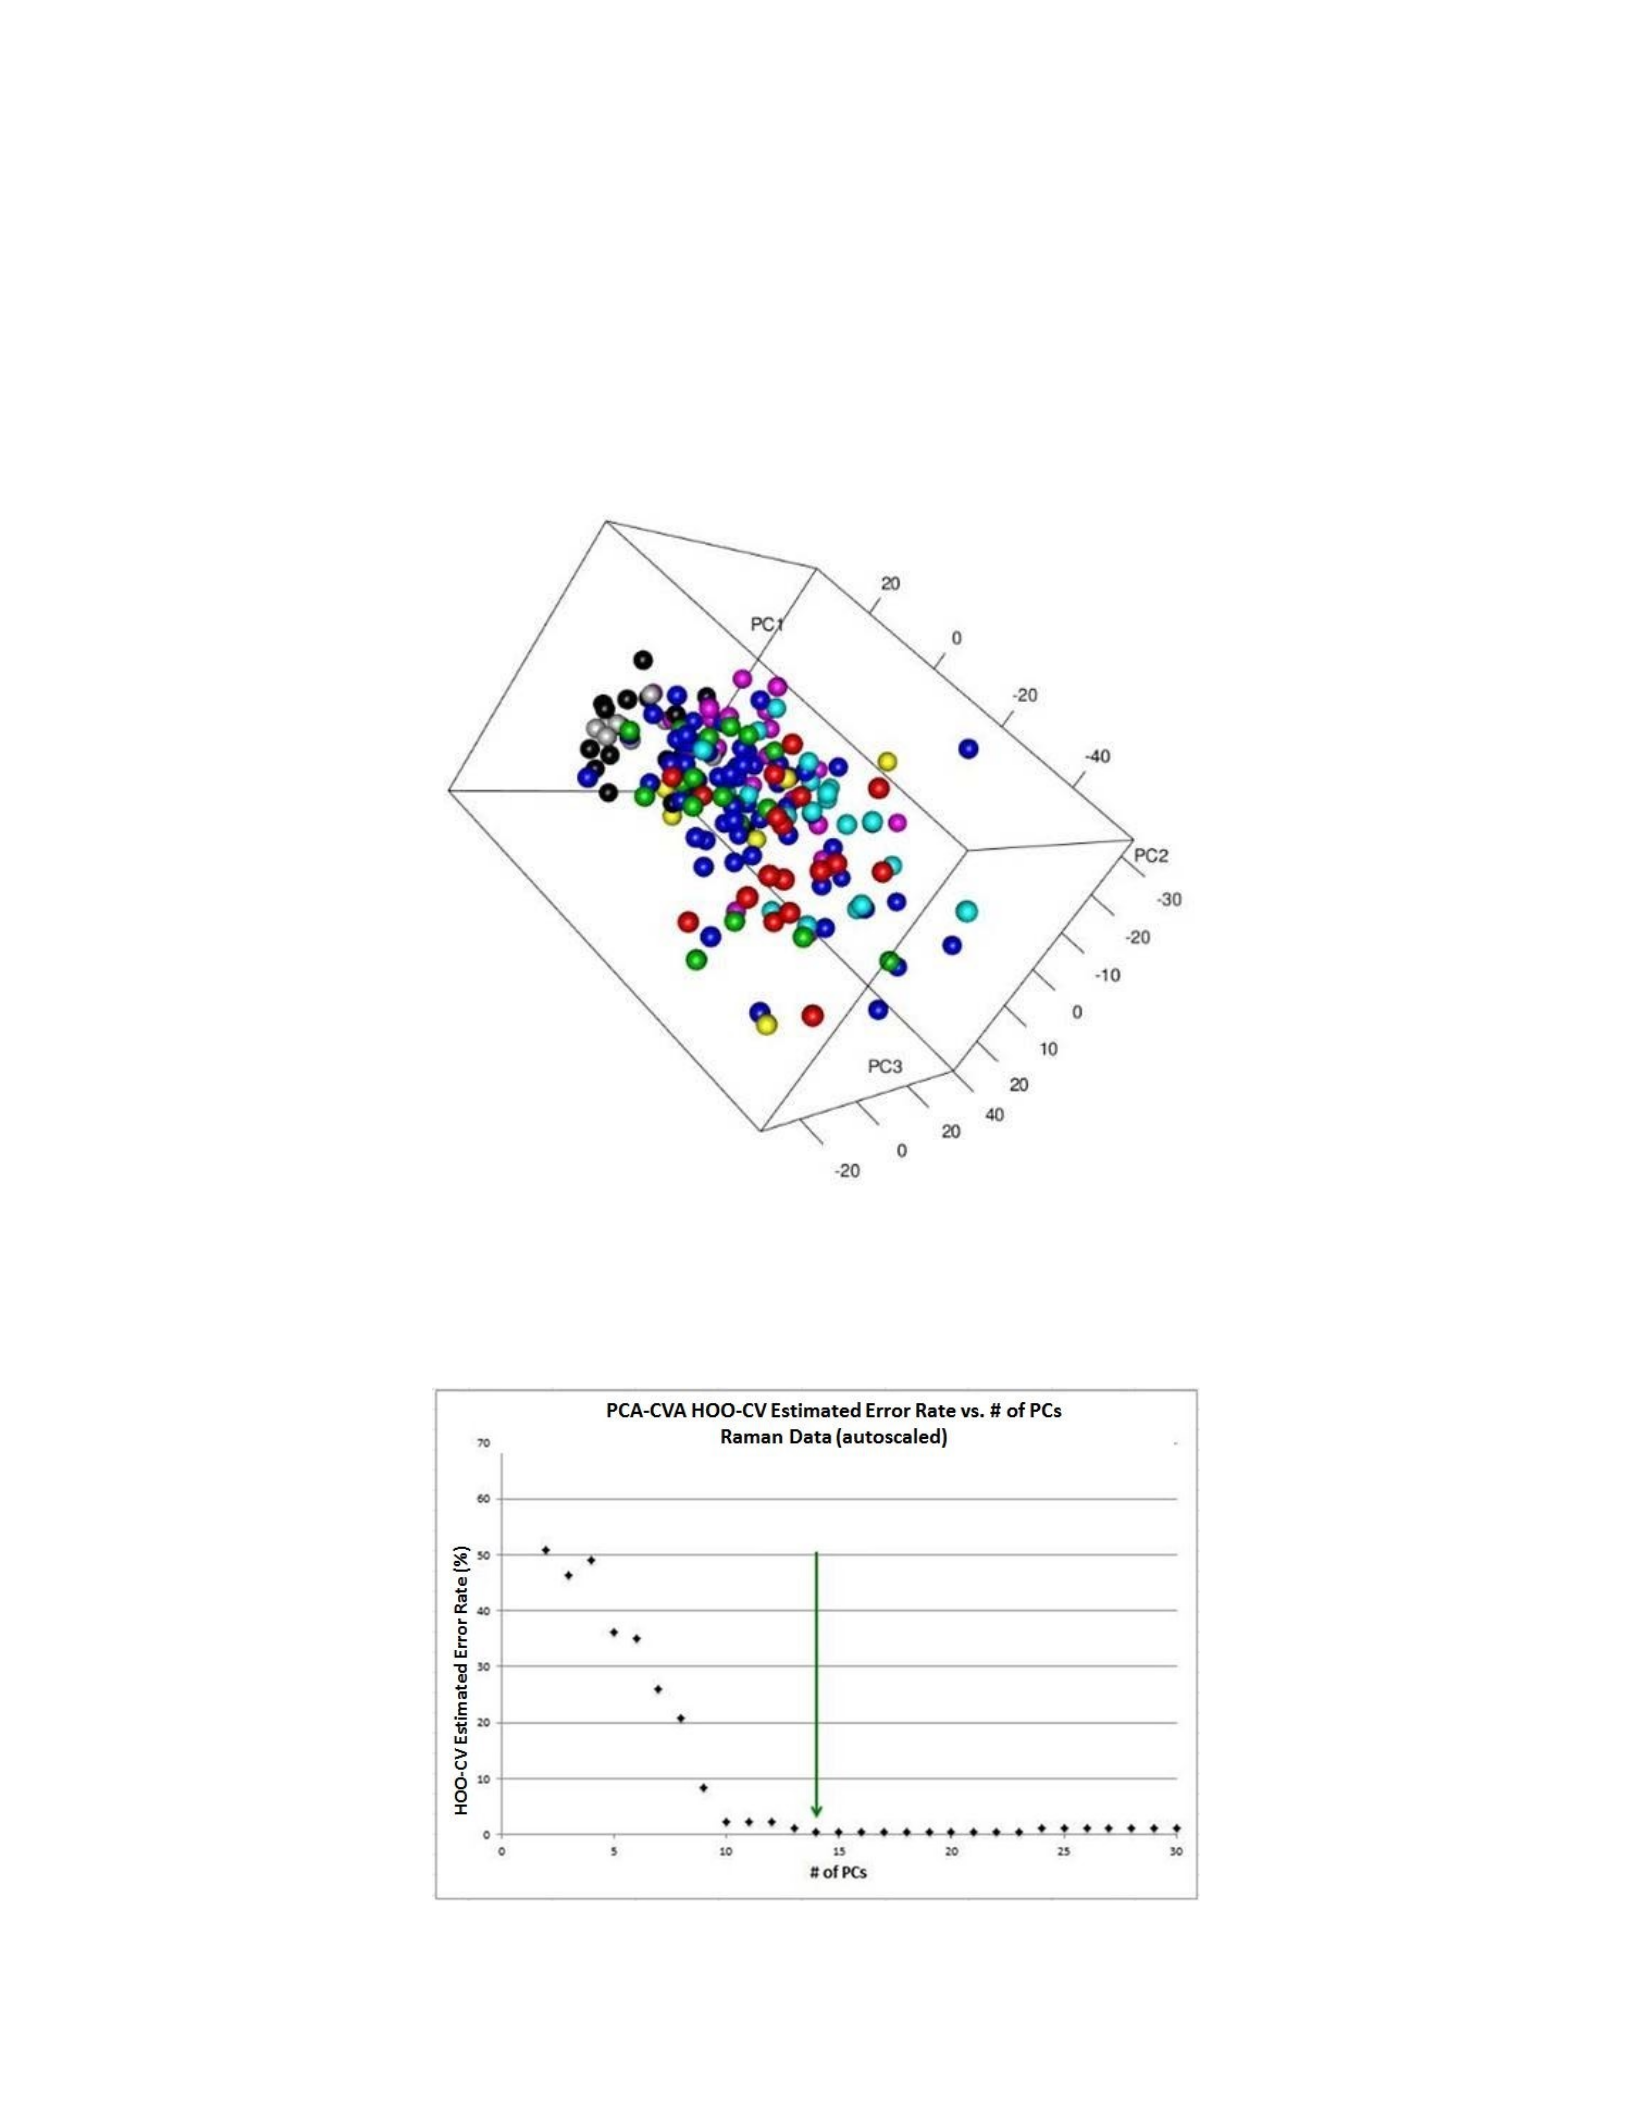
plots between the infrared (figure 36) and the Raman data (see figure 50) is interesting. It is
possible that the reason the separation was not evident in the 3D-scores plot for the Raman data
is because the Raman analysis spot size (50 μm) is significantly smaller than it is for the infrared
analysis (1.3 mm x 0.8 mm), but this hypothesis was not tested.
Figure 50. 3D-scores plot of Raman data (autoscaled).

Figure 51. Estimated error rate displayed as a function of the
number of PC’s. An estimated error rate of 0.56% was observed at
14 PC’s.
Summary and Conclusion
This research shows that detection and identification of chemical and physical differences
in drug tablets using field-portable Raman spectroscopy provides a chemical signature or profile
that may be used to authenticate product. PCA-CVA HOO-CV applied to the Raman data
showed that Raman may be used to distinguish authentic from non-authentic samples, and to
differentiate non-authentic samples from each other. This is important because it indicates that
Raman analysis performed in the field may be used to establish provenance. A visual
comparison of the Raman spectra was used to differentiate non-authentic from authentic samples.
Raman spectra were reproducible within different tablets of the same sample. These spectra may
be used to classify samples of unknown origin and provide investigators insight into the sample’s
manufacturing process and history. For example, differences in spectra may result from
differences in solid-state form of the active or inactive ingredients that are produced during
manufacture or storage, or may be have different ingredients present that could be linked to
manufacturer.
CHAPTER 10. INVESTIGATION AND COMPARISON OF ORGANIC VOLATILES
AND RESIDUAL SOLVENTS IN DRUG PRODUCTS USING GAS
CHROMATOGRAPHY-MASS SPECTROMETRY
Introduction
When pharmaceutical tablets are manufactured, chemicals are present in the finished
tablet at trace levels. These chemicals are introduced into the tablet in a variety of ways. They
may be present in the raw materials, they may be solvent residues remaining after tablet

manufacture, or they may even be degradation products that develop during manufacture or
storage. These trace chemicals generate a chemical fingerprint that is representative of the tablet.
They also provide insight about the tablet’s manufacture and storage.
When manufacturing processes are well controlled and adhere to regulatory guidelines as
is typical for an authentic product, this chemical fingerprint is well controlled and reproducible.
The ability to detect and identify these chemicals in a reliable manner is useful, providing a
chemical fingerprint that is representative of the authentic product and may be used to
differentiate authentic from non-authentic tablets. This chemical fingerprint may also be used to
classify counterfeit drugs, providing investigative leads useful in establishing source or origin.
GC-MS is useful for detecting and identifying these trace chemicals. Data generated
from this type of analysis can be evaluated many different ways when applied to the analytical
challenges associated with counterfeit drugs. In this study, total ion chromatograms were
compared to determine reproducibility amongst different tablets of the same sample. In addition,
they were used to differentiate authentic from non-authentic samples. Samples were heated and
organic volatiles and residual solvents evolved from the sample were collected and analyzed.
Experimental Details
Instrumentation
GC-MS data were collected using a GUARDION™ GC-MS (Smiths Detection, Danbury,
CT). The system uses a 5-m long MXT®-5 (Restek Corporation, Bellefonte, PA) resistively
heated capillary column (inner diameter 0.1 mm, film thickness of 0.4 μm). This column is a
crossbond® diphenyl/dimethylpolysiloxane column. The carrier gas is helium (Leland, South
Plainfield, NJ) introduced via disposable canister. Sample collection and introduction to the
GUARDION was performed via solid-phase microextraction (SPME). A CUSTODION®
(Torion Technologies, American Fork, UT) SPME holder was used with a Supelco (St. Louis,
MO) 23-gauge, 65 μm DMS/DVB fiber assembly.
Performance of the system was tested at the start and end of every work day and
periodically throughout the day. Performance testing evaluates both GC and MS performance.
To perform this testing, a standard containing 13 chemicals is analyzed. These 13 chemicals
have expected retention-time values between 0 and 90 seconds and are spaced across this range.
GC performance acceptance criteria required that the retention times for all 13 chemicals must be
the stated +/- 2 seconds. Retention-time values measured during all performance testing were
always within the accepted range. Table 9 shows the expected retention-time values for these 13
chemicals.
Chemical in Standard CAS Number Stated Retention-Time Value (s)
Acetone 67-64-1 11.34
Methylene chloride 75-09-2 13.26
Methyl-t-butyl ether 163-40-44 15.17
Heptane 142-82-5 26.25
Methylcyclohexane 108-87-2 28.92
Toluene-d8 2037-26-5 32.85
Perchloroethylene 127-18-4 37.34
Bromopentafluorobenzene 344-04-7 41.84
Bromoform 75-25-2 45.14
1,2-Dibromotetrafluorobenzene 827-08-7 65.94
Methyl salicylate 119-36-8 72.25
Tetrabromoethane 79-27-6 79.83
Tetradecane 629-59-4 85.76
Table 9. Chemicals contained within standard sample analyzed
during performance testing.
In addition to evaluating the GC, performance testing also evaluates MS performance.
MS performance includes tests for spectral quality, mass calibration, mass resolution, ion
statistics, space charge, signal-to-noise ratio, and sensitivity. The performance-test procedure
used was the procedure recommended by the instrument manufacturer. The system did not
always pass performance-test criteria. However, the reasons for all failures were due to MS
performance. Spectral data, however, were not directly interpreted and compared. Only GC
retention-time data was used for comparison.
The GUARDION is specifically intended for field use. Some of the specifications for
this system, especially those intended to support portability and ruggedization of the instrument,
are shown in Table 10. An image of the model of GC-MS instrument used is shown in figure 52.
GUARDION Specifications of Importance to this Study
Compact size 15.5 in x 15 in x 9 in excluding handle
Weight 14.6 kg
GC column MXT-5 column, 5 m, 0.1 mm inner diameter, 0.4 μm film
thickness
Carrier gas Helium carrier gas supply, internal disposable cartridge
Mass spectrometer Toroidal ion trap
Vacuum system Turbomolecular vacuum pump with diaphragm roughing pump
Sample collection Solid-phase microextraction
Hot-zone operation Sealed for operation in the hot zone
Decontamination Spray/wipe down
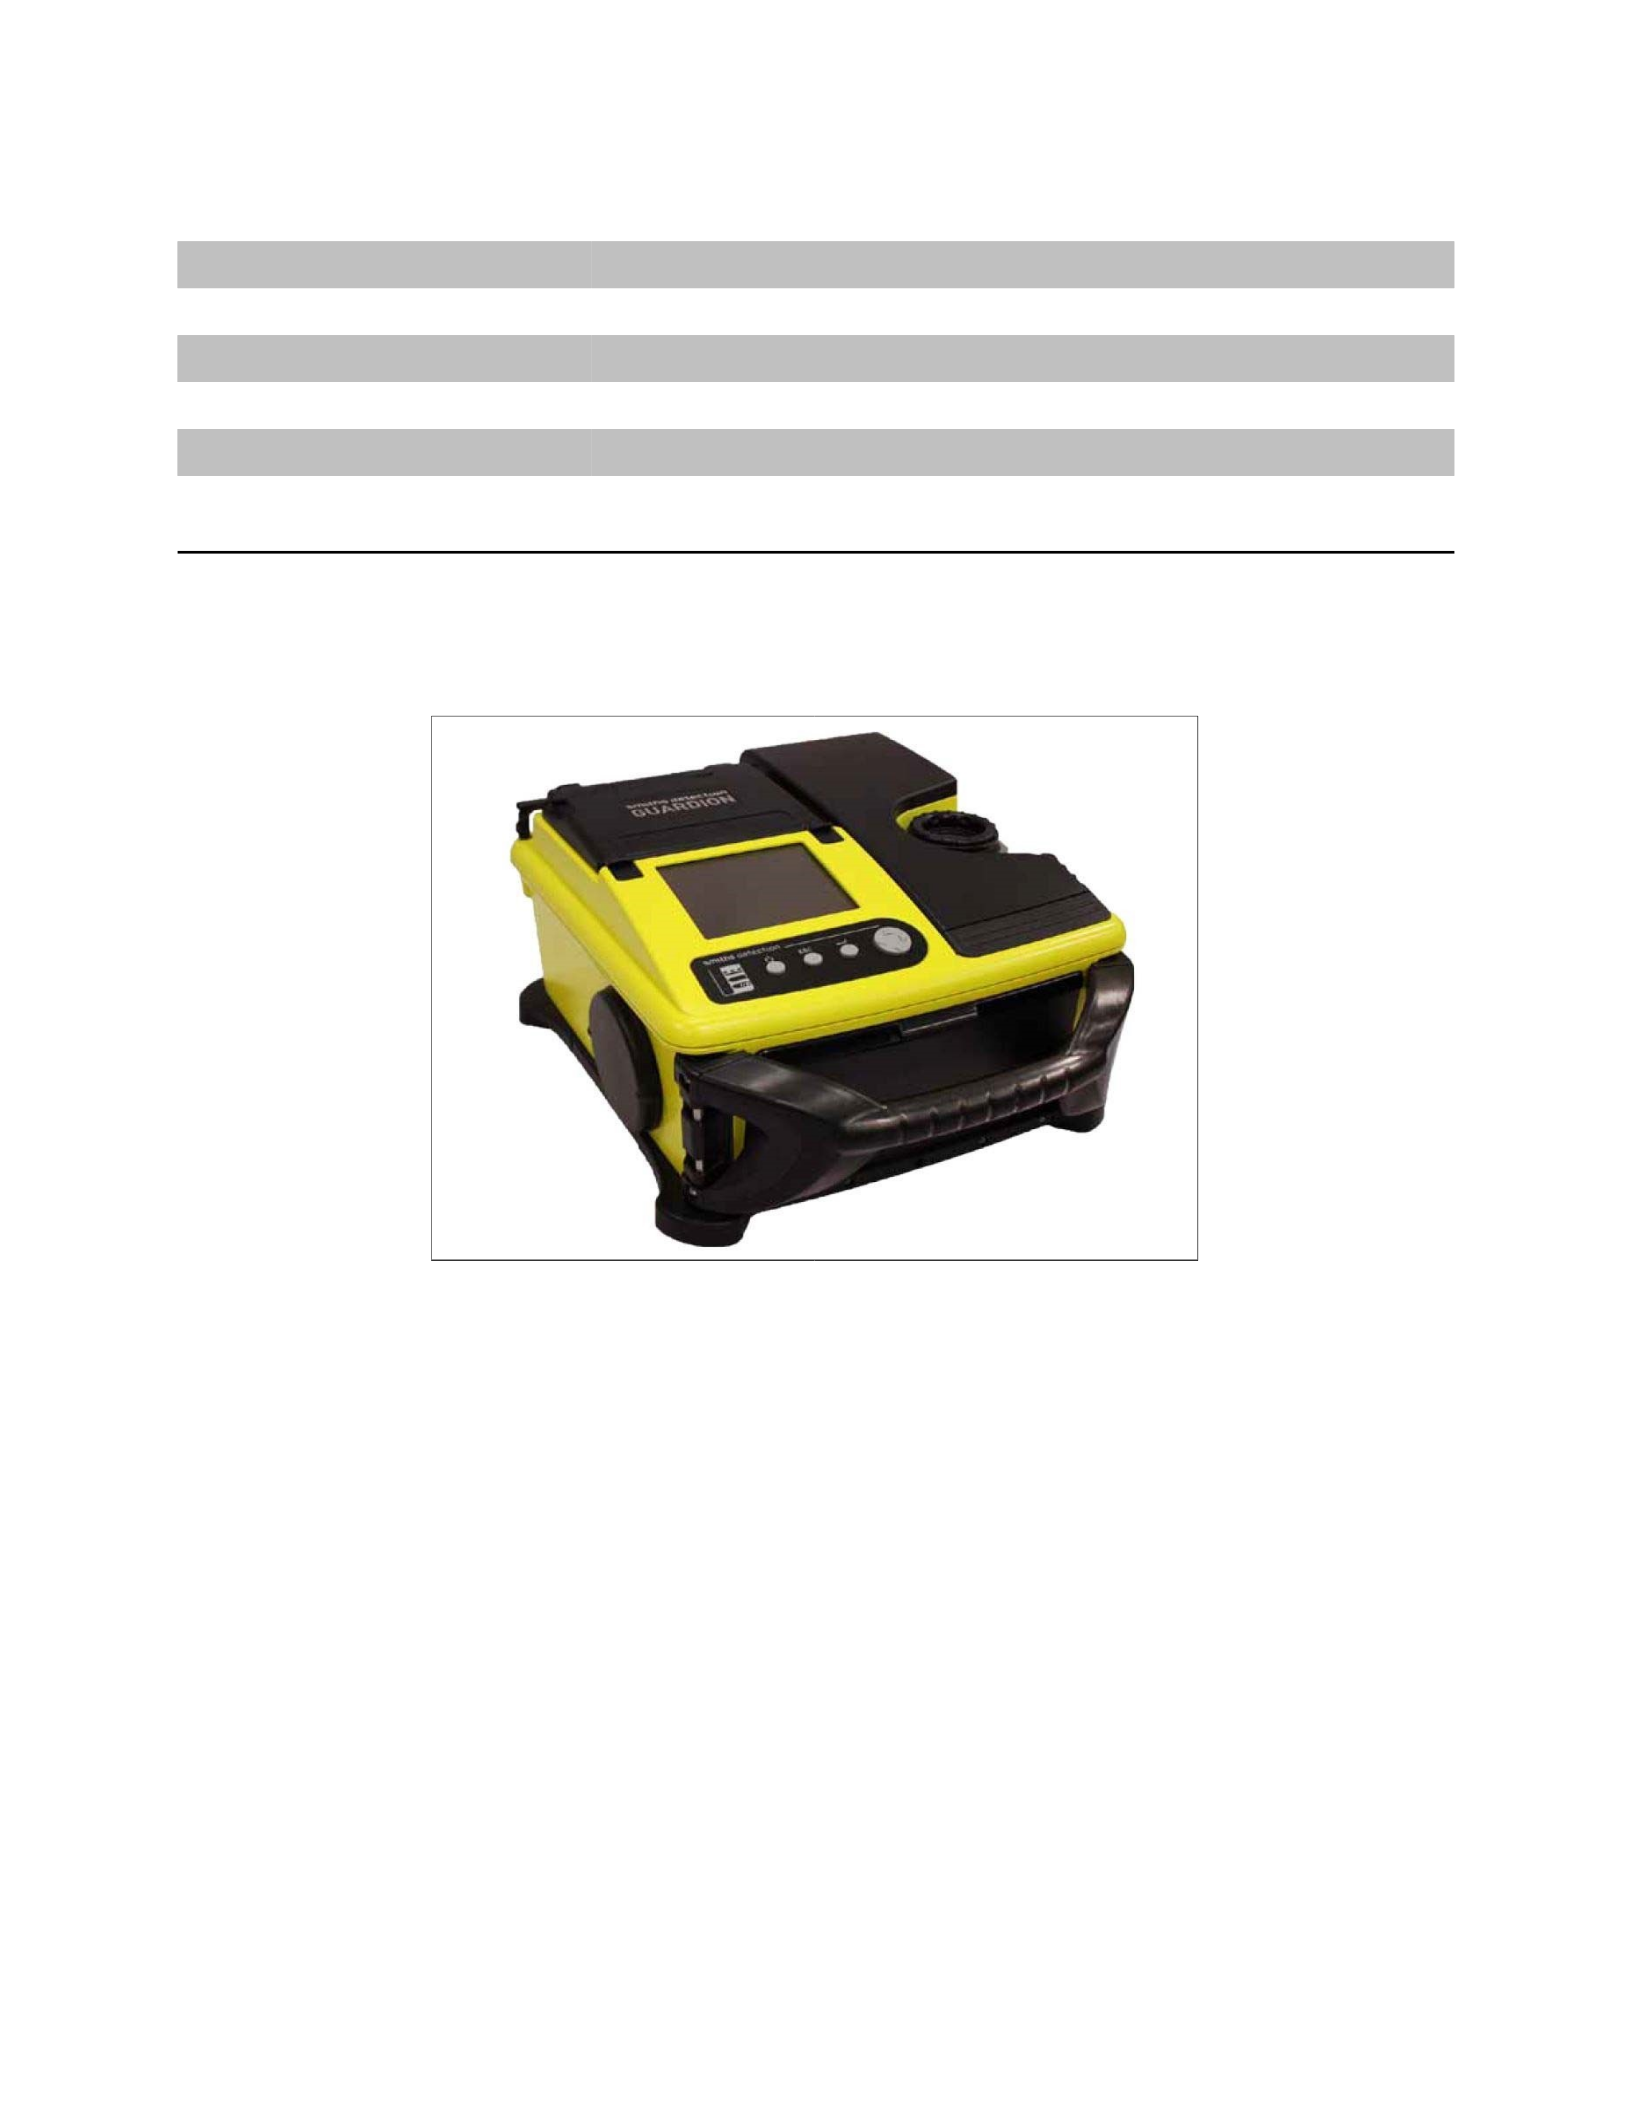
Operating temperature 0⁰C to 45⁰C
Operating humidity 0% - 95%
User interface Embedded software with touch-screen
Data export Full USB support; Ethernet capable; mouse and keyboard capable
Power Removable, rechargeable Li-ion battery; external power supply
Mass range 43 m/z - 500 m/z
Temperature programming Up to 300⁰C
Table 10. GUARDION specifications of importance for its use in
this study.
Figure 52. GUARDION GC-MS. Photograph courtesy of Smiths
Detection, Danbury, CT.
Method
Samples were analyzed using a field-portable toroidal ion-trap GC-MS. Samples were
injected directly from the SPME holder into the inlet of the GC-MS. During analysis, samples
elute directly from the GC into the ion trap of the MS. Ionization occurs via electron impact at
70 eV. The carrier gas used was helium. The injector and transfer line temperatures were
270⁰C. The column temperature was held at 50⁰C for 10 seconds and then ramped at a rate of

2⁰C per second to 296⁰C. The temperature was held at 296⁰C for 47 seconds. The voltage of the
MS was scanned continually to detect mass fragments in the range between 43 and 500 atomic
mass unit’s. Scan time was approximately 60 milliseconds and continuously performed
throughout the GC analysis.
To prepare for GC-MS analysis, each tablet was gently crushed and transferred to a GC-
MS headspace vial. Samples were heated to 60⁰C and held at this temperature for 20 minutes.
The headspace was then sampled at 60⁰C for five minutes using the SPME technique. The
sample was then injected directly from the SPME holder into the inlet of the GC-MS. Sampling
and analysis was performed two times in immediate succession. The first sampling used a
PDMS/DVB SPME sorbent; the second sampling used a carboxen SPME sorbent. Samples were
then heated to 110⁰C and held at this temperature for 20 minutes. The same SPME headspace
sampling and analysis steps performed at 60⁰C were repeated at 110⁰C. In some instances,
SPME headspace sampling was performed using only a PDMS/DVB SPME sorbent. The reason
both fibers were not always used is because, initially during data collection, the PDMS/DVB
SPME sorbent was used in duplicate to verify reproducibility. The carboxen fiber was not used
during these earlier experiments. After reproducibility was observed in the first samples
analyzed, it was deemed to be more important to evaluate the use of the carboxen sorbent during
the second injection rather than continue to verify reproducibility.
Results and Discussion
It was possible to differentiate the authentic samples from all non-authentic samples using
this method. In most cases, differences in chromatograms, i.e., chemical fingerprints, between
the authentic and the non-authentic sample were significant. An attempt was made to average

chromatograms to enable quantitation of these visual observations. Due to many instrumental
factors including the dynamic ionization that occurs in the instrument during sample analysis to
optimize MS results, retention-time values from one run to the next are unique. This makes an
averaging function impossible to perform without changing retention-time values. It was
determined that averaging chromatograms would not be performed.
Figures 53 through 61 show the chromatogram of an authentic tablet compared with a
chromatogram from each of the nine non-authentic samples. A chromatogram (in this context) is
a display showing all of the chemicals detected during GC-MS analysis. The x-axis represents
the analyte’s retention time, and the y-axis represents abundance. This study was non
quantitative, so although relative abundances within a single chromatogram may be used as a
factor to be evaluated within that single chromatogram, no quantitative assessment of each
analyte’s concentration within the tablet was or should be performed. Ideally, each peak in the
chromatogram is representative of a chemical present in the sample. In this study, the nature of
the peak was not important and overlapping peaks were not considered to be a problem. The
presence or absence of a peak at a specific retention time was the criteria used for discrimination.
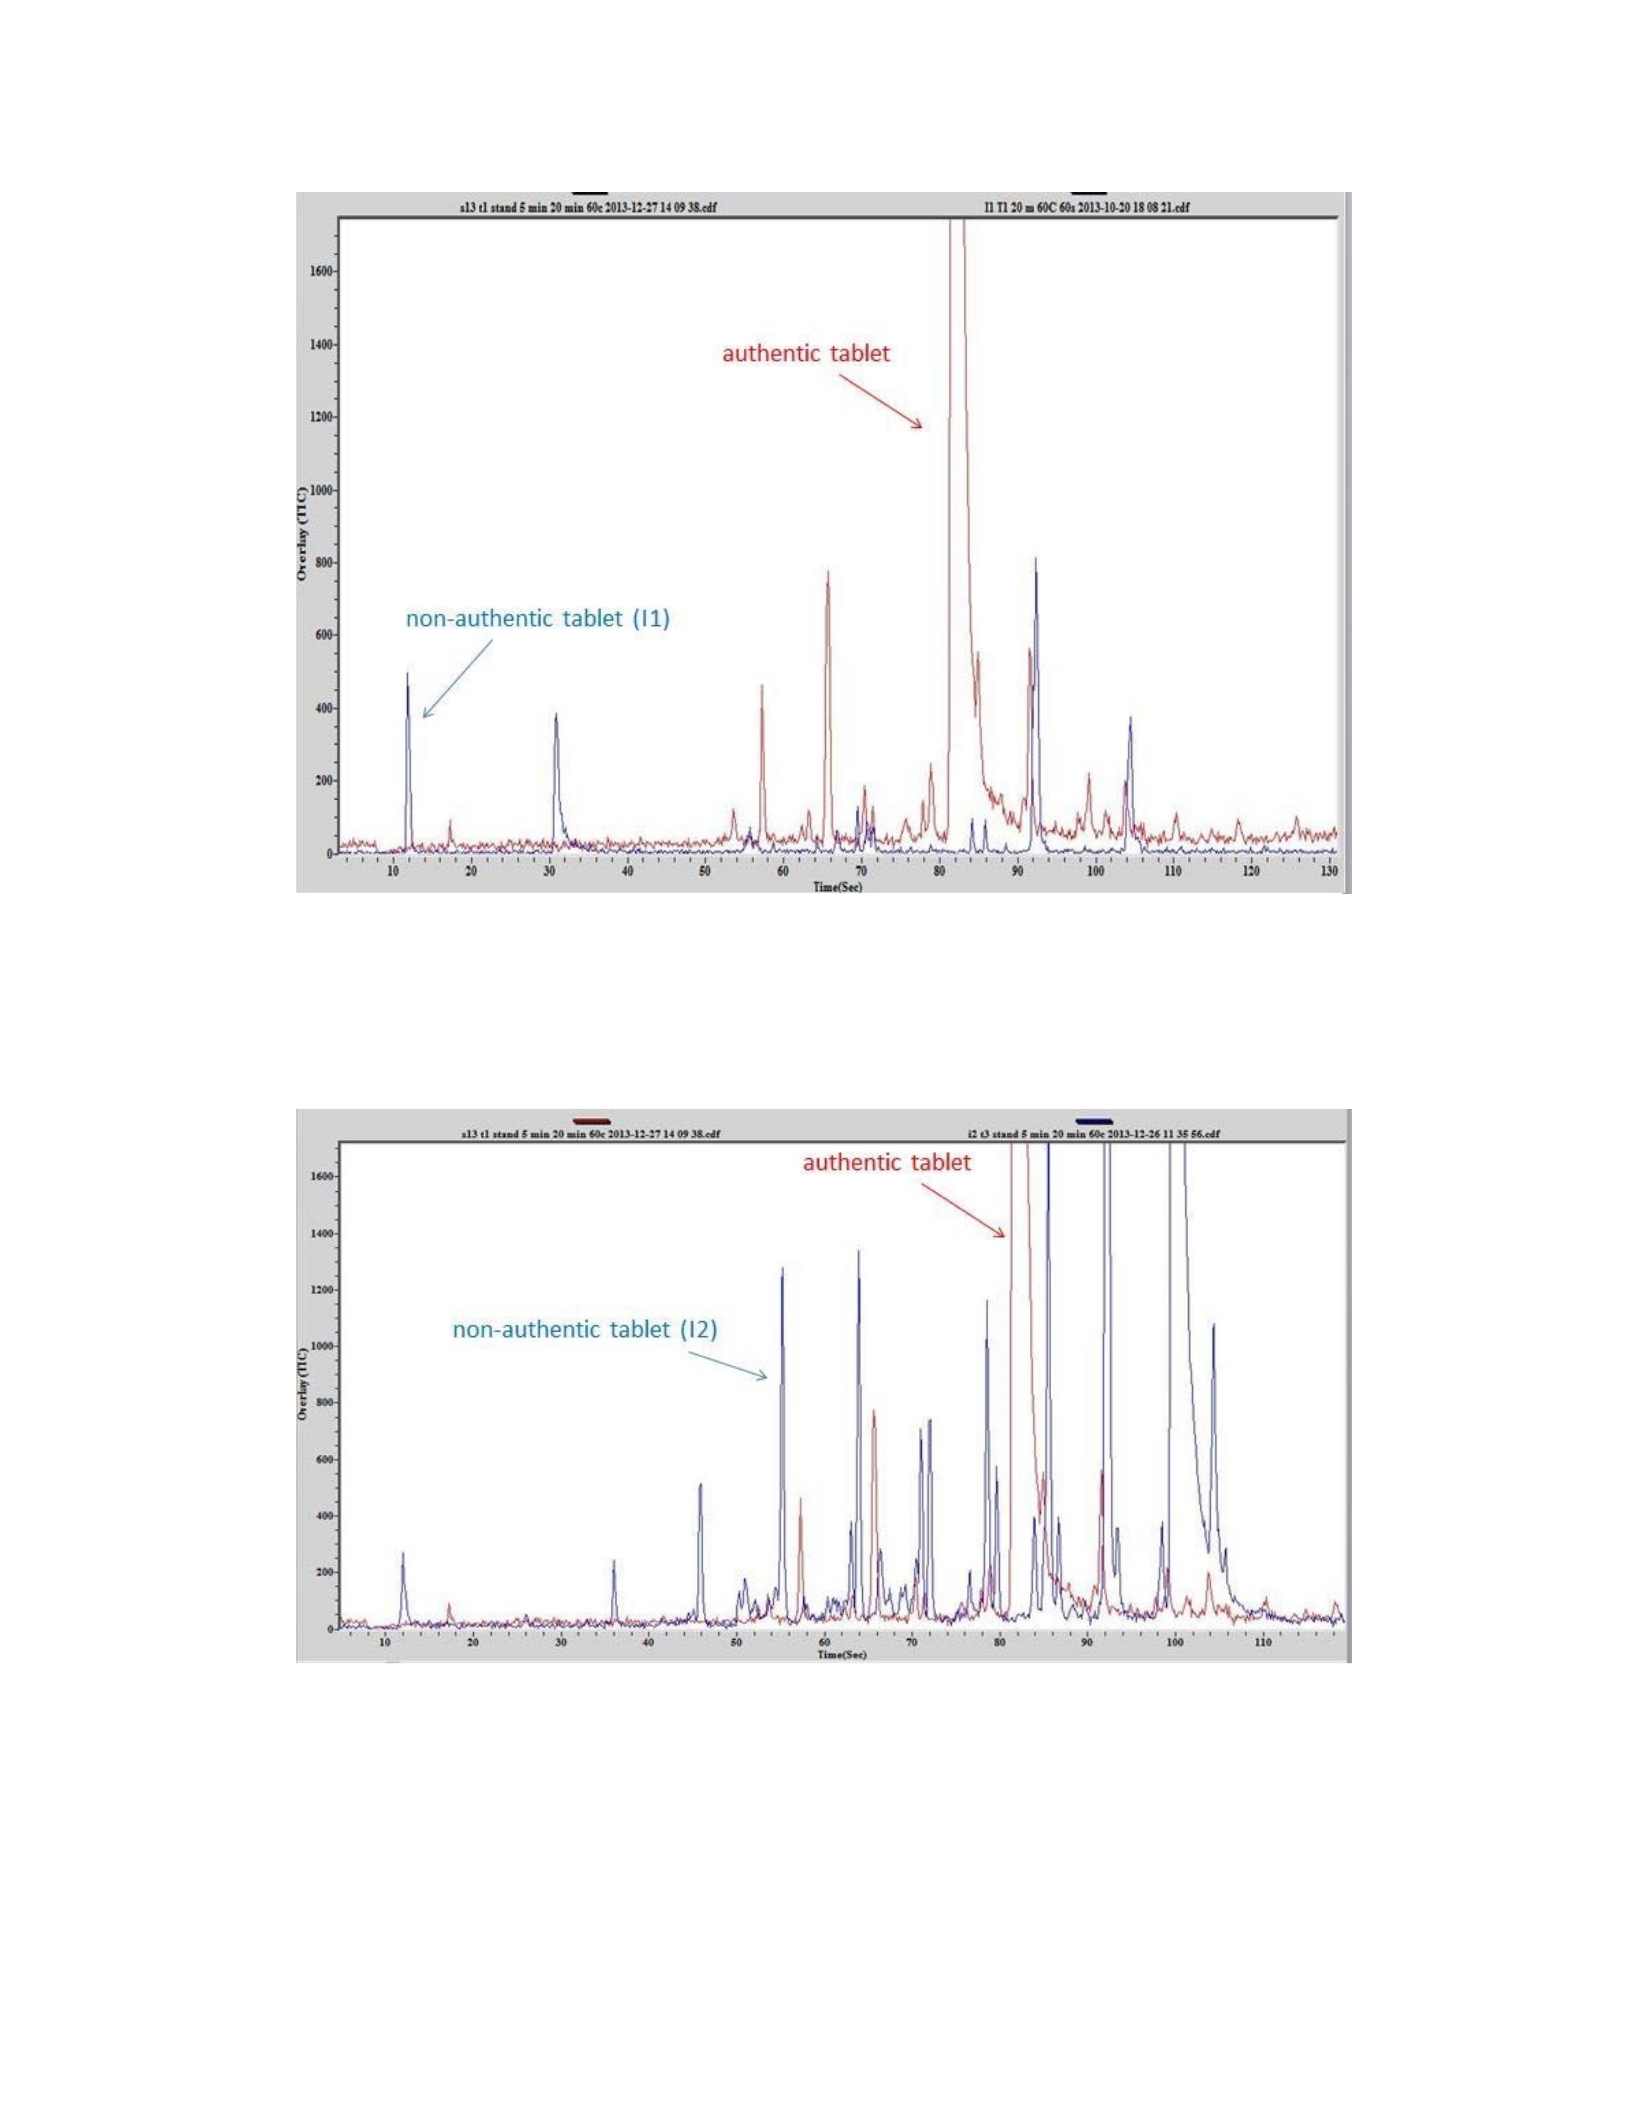
Figure 53. Chromatogram of authentic sample compared with
non-authentic I1 (PDMS/DVB SPME sorbent, 60⁰C).
Figure 54. Chromatogram of authentic sample compared with
non-authentic I2 (PDMS/DVB SPME sorbent, 60⁰C).
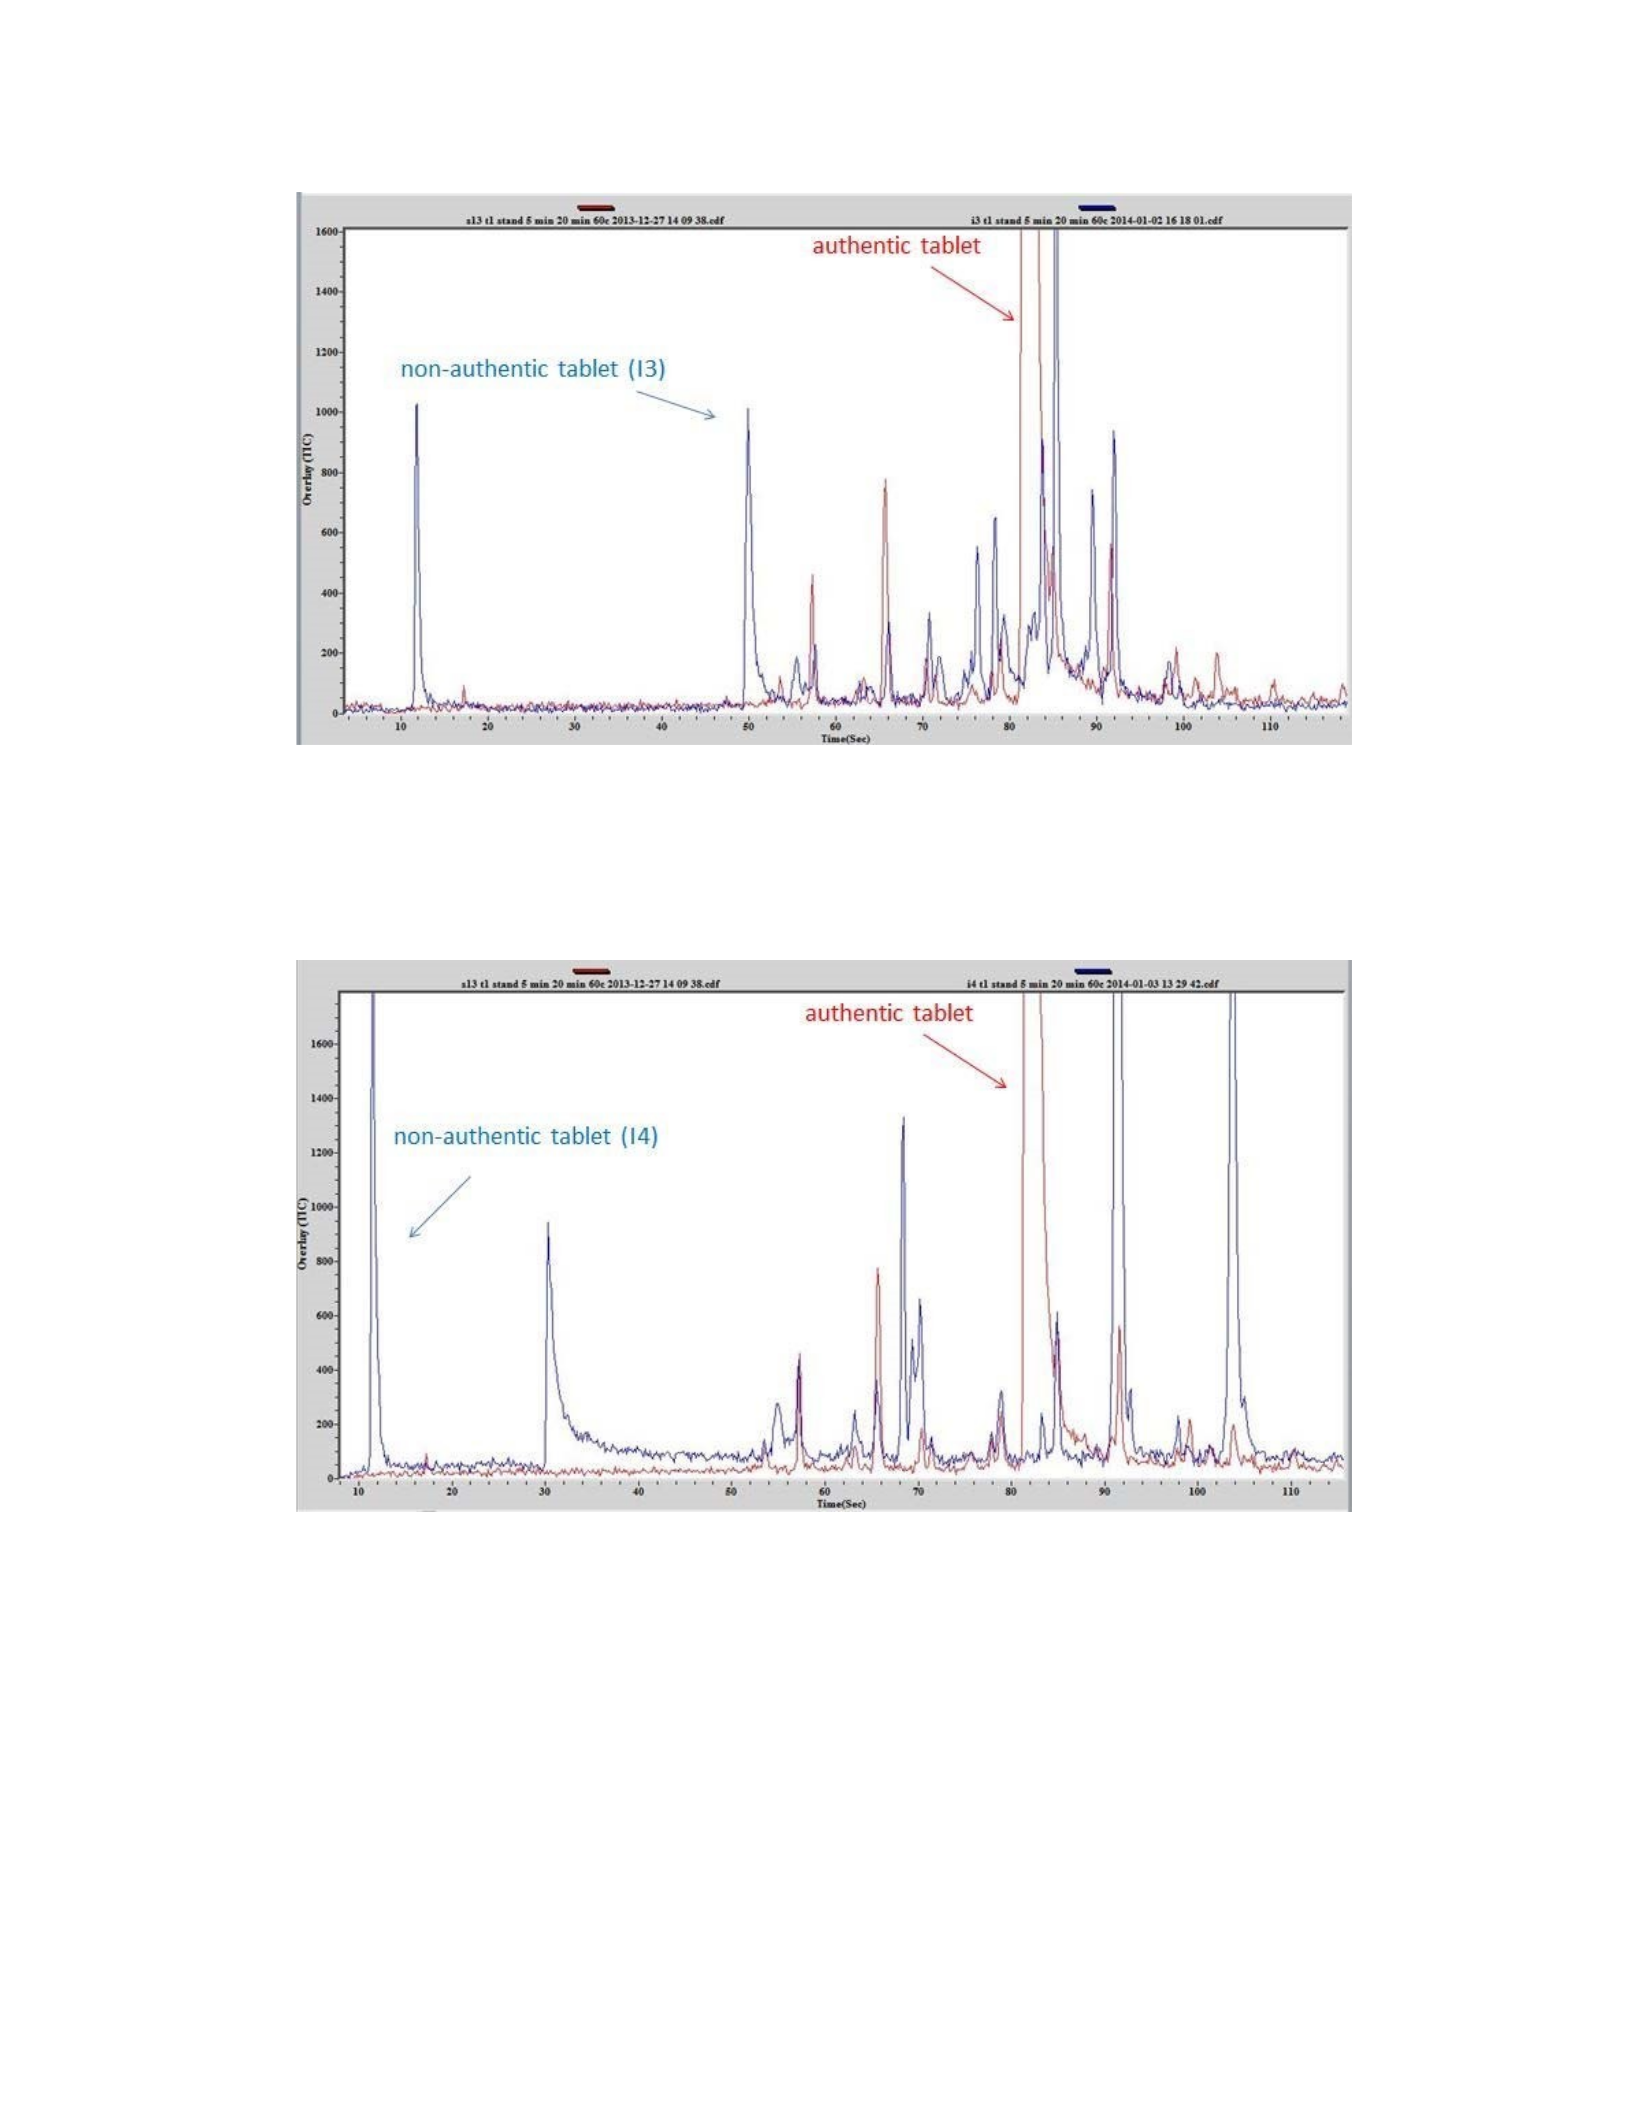
Figure 55. Chromatogram of authentic sample compared with
non-authentic I3 (PDMS/DVB SPME sorbent, 60⁰C).
Figure 56. Chromatogram of authentic sample compared with
non-authentic I4 (PDMS/DVB SPME sorbent, 60⁰C).
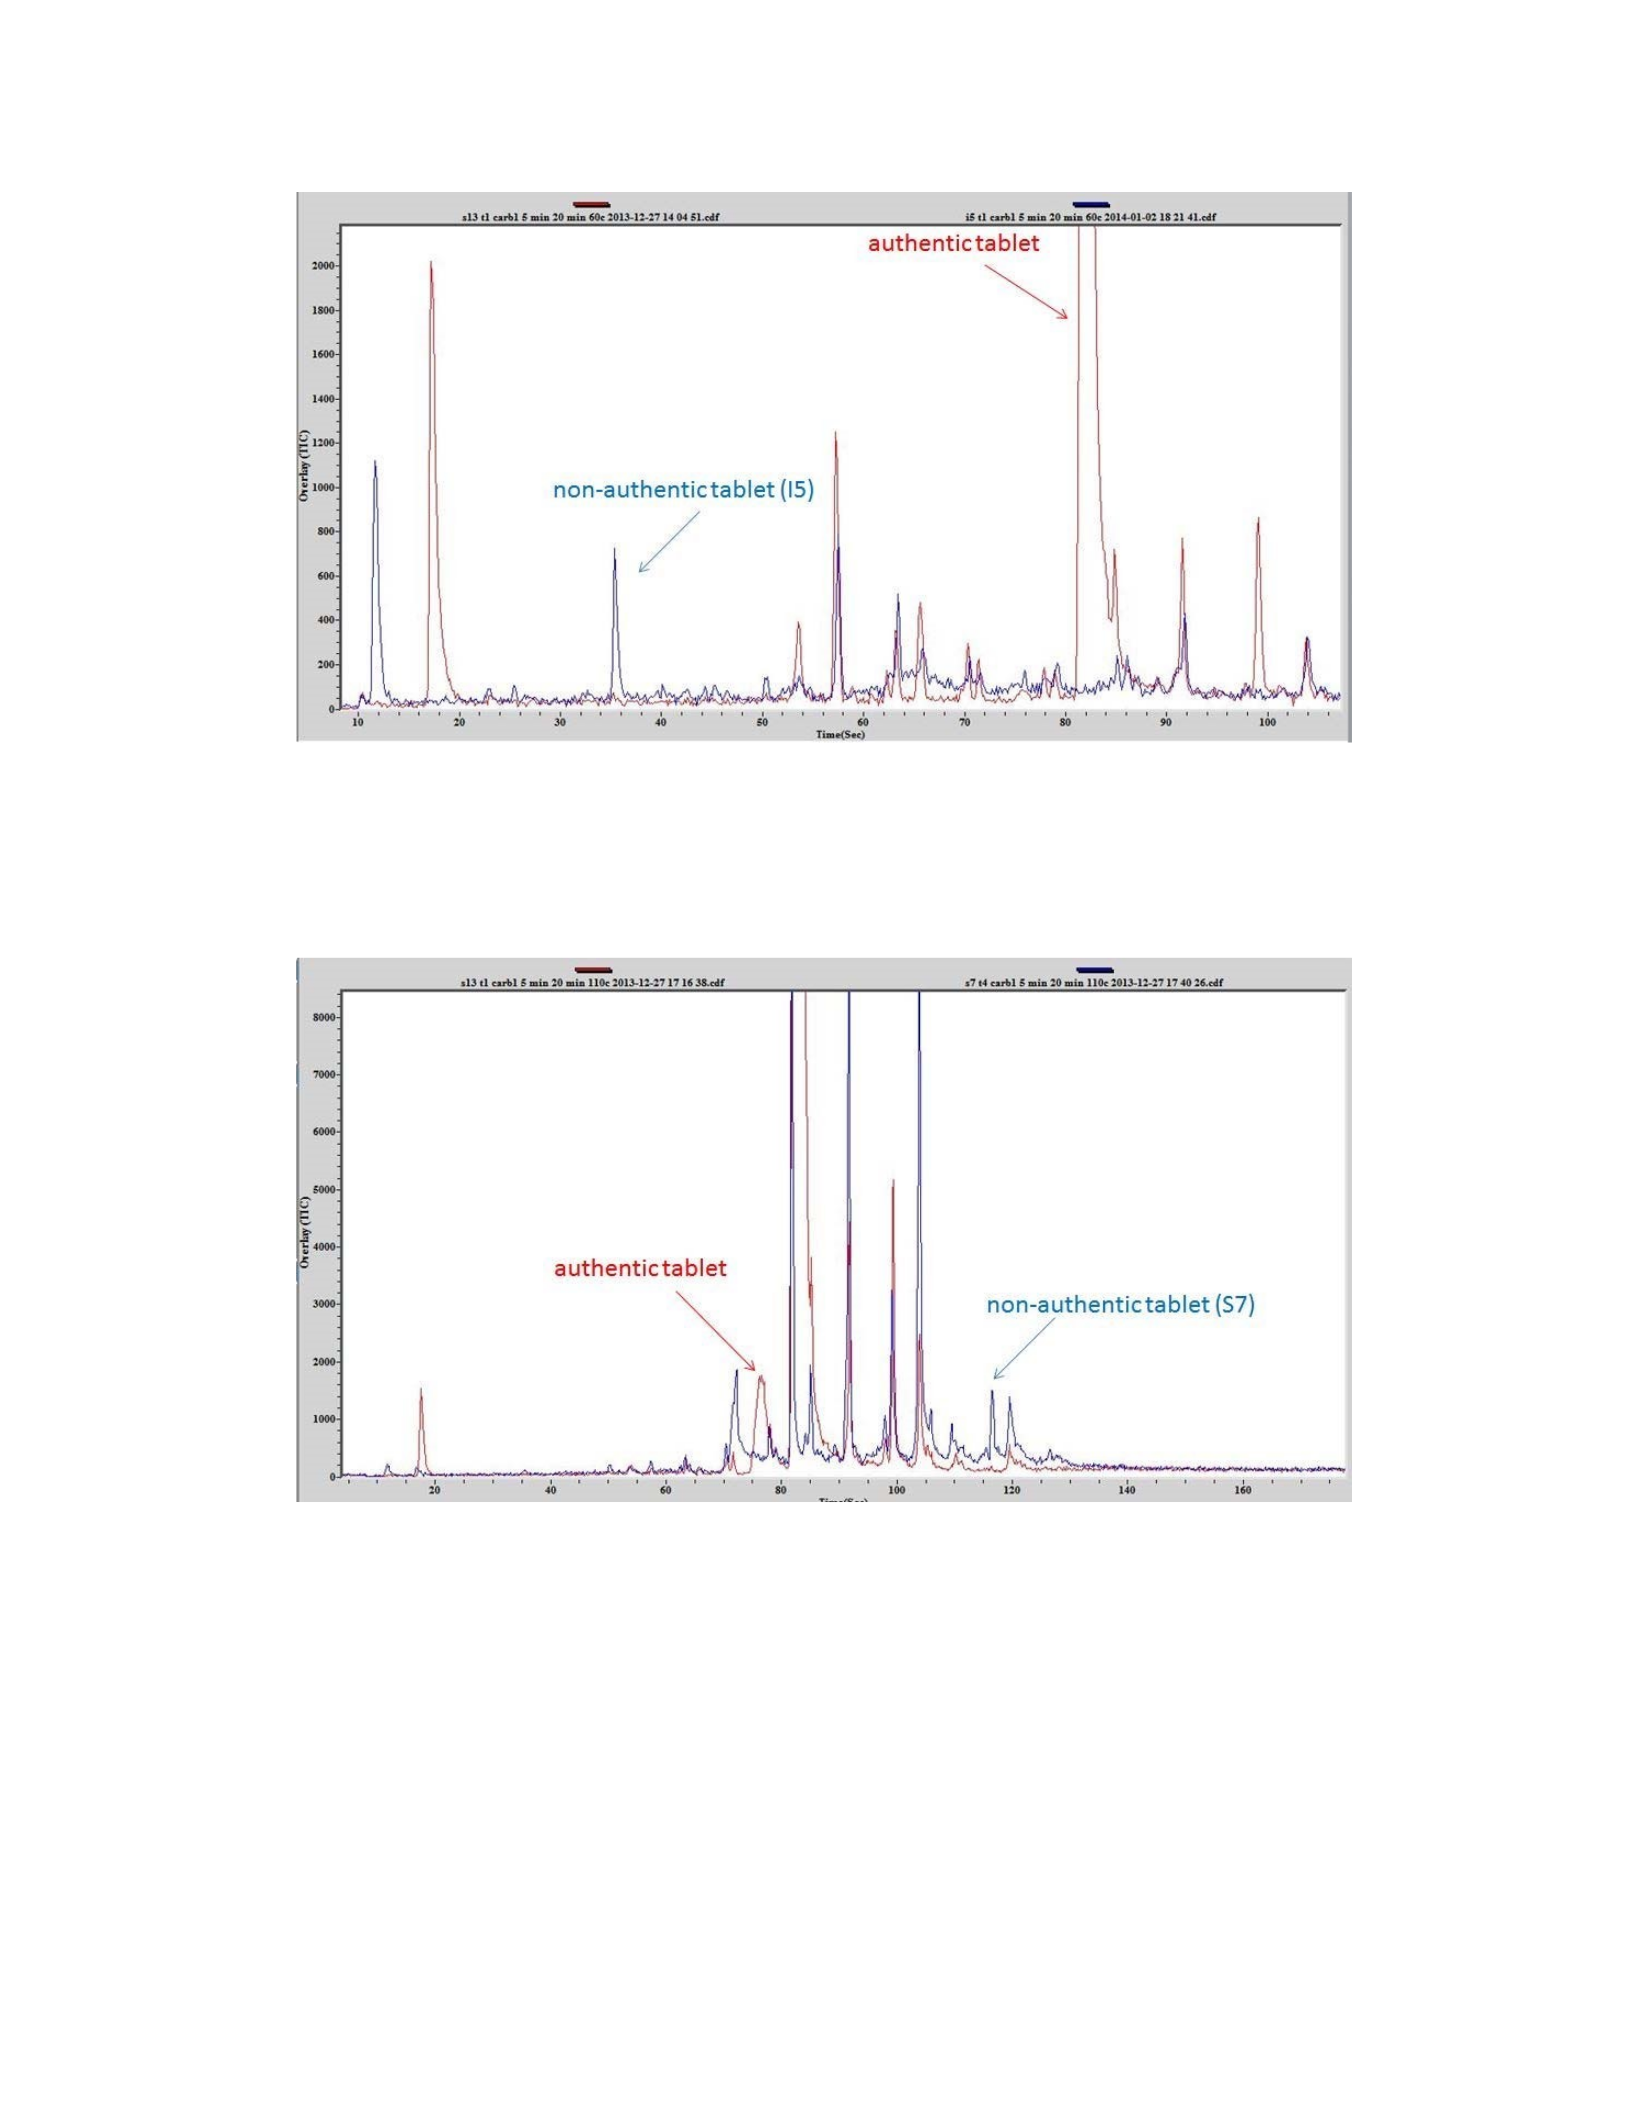
Figure 57. Chromatogram of authentic sample compared with
non-authentic I5 (PDMS/DVB SPME sorbent, 60⁰C).
Figure 58. Chromatogram of authentic sample compared with
non-authentic S7 (PDMS/DVB SPME sorbent, 60⁰C).
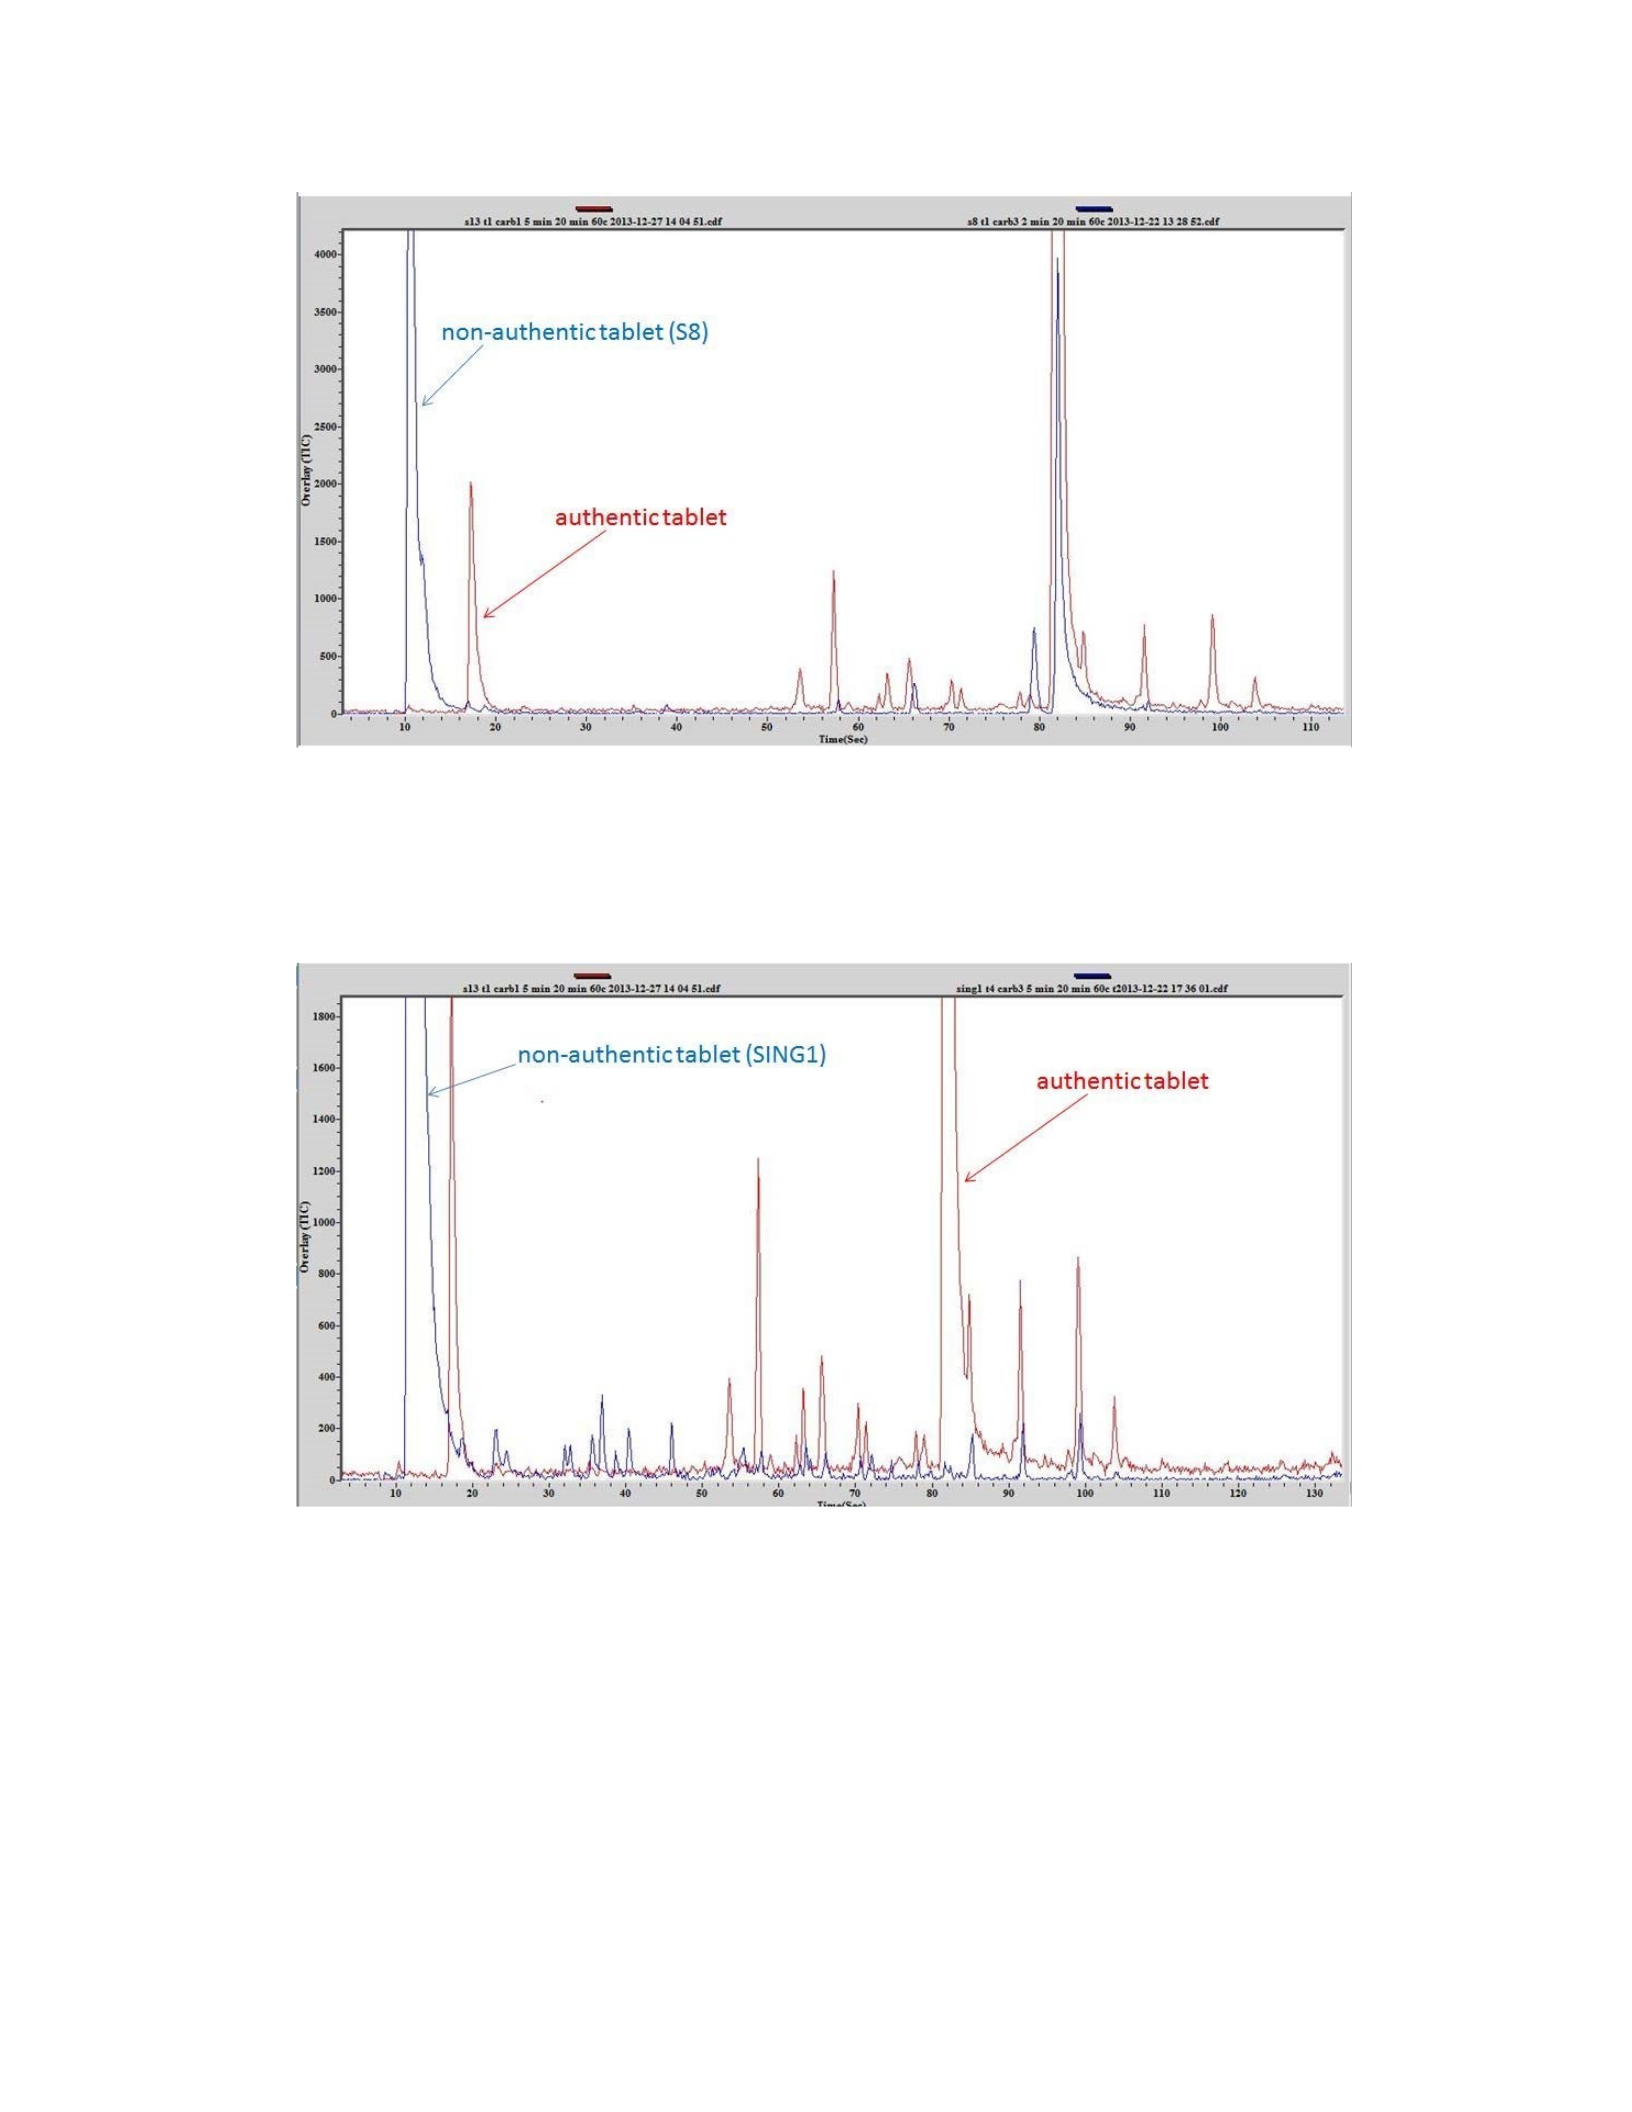
Figure 59. Chromatogram of authentic sample compared with
non-authentic S8 (PDMS/DVB SPME sorbent, 60⁰C).
Figure 60. Chromatogram of authentic sample compared with
non-authentic SING1 (PDMS/DVB SPME sorbent, 60⁰C).
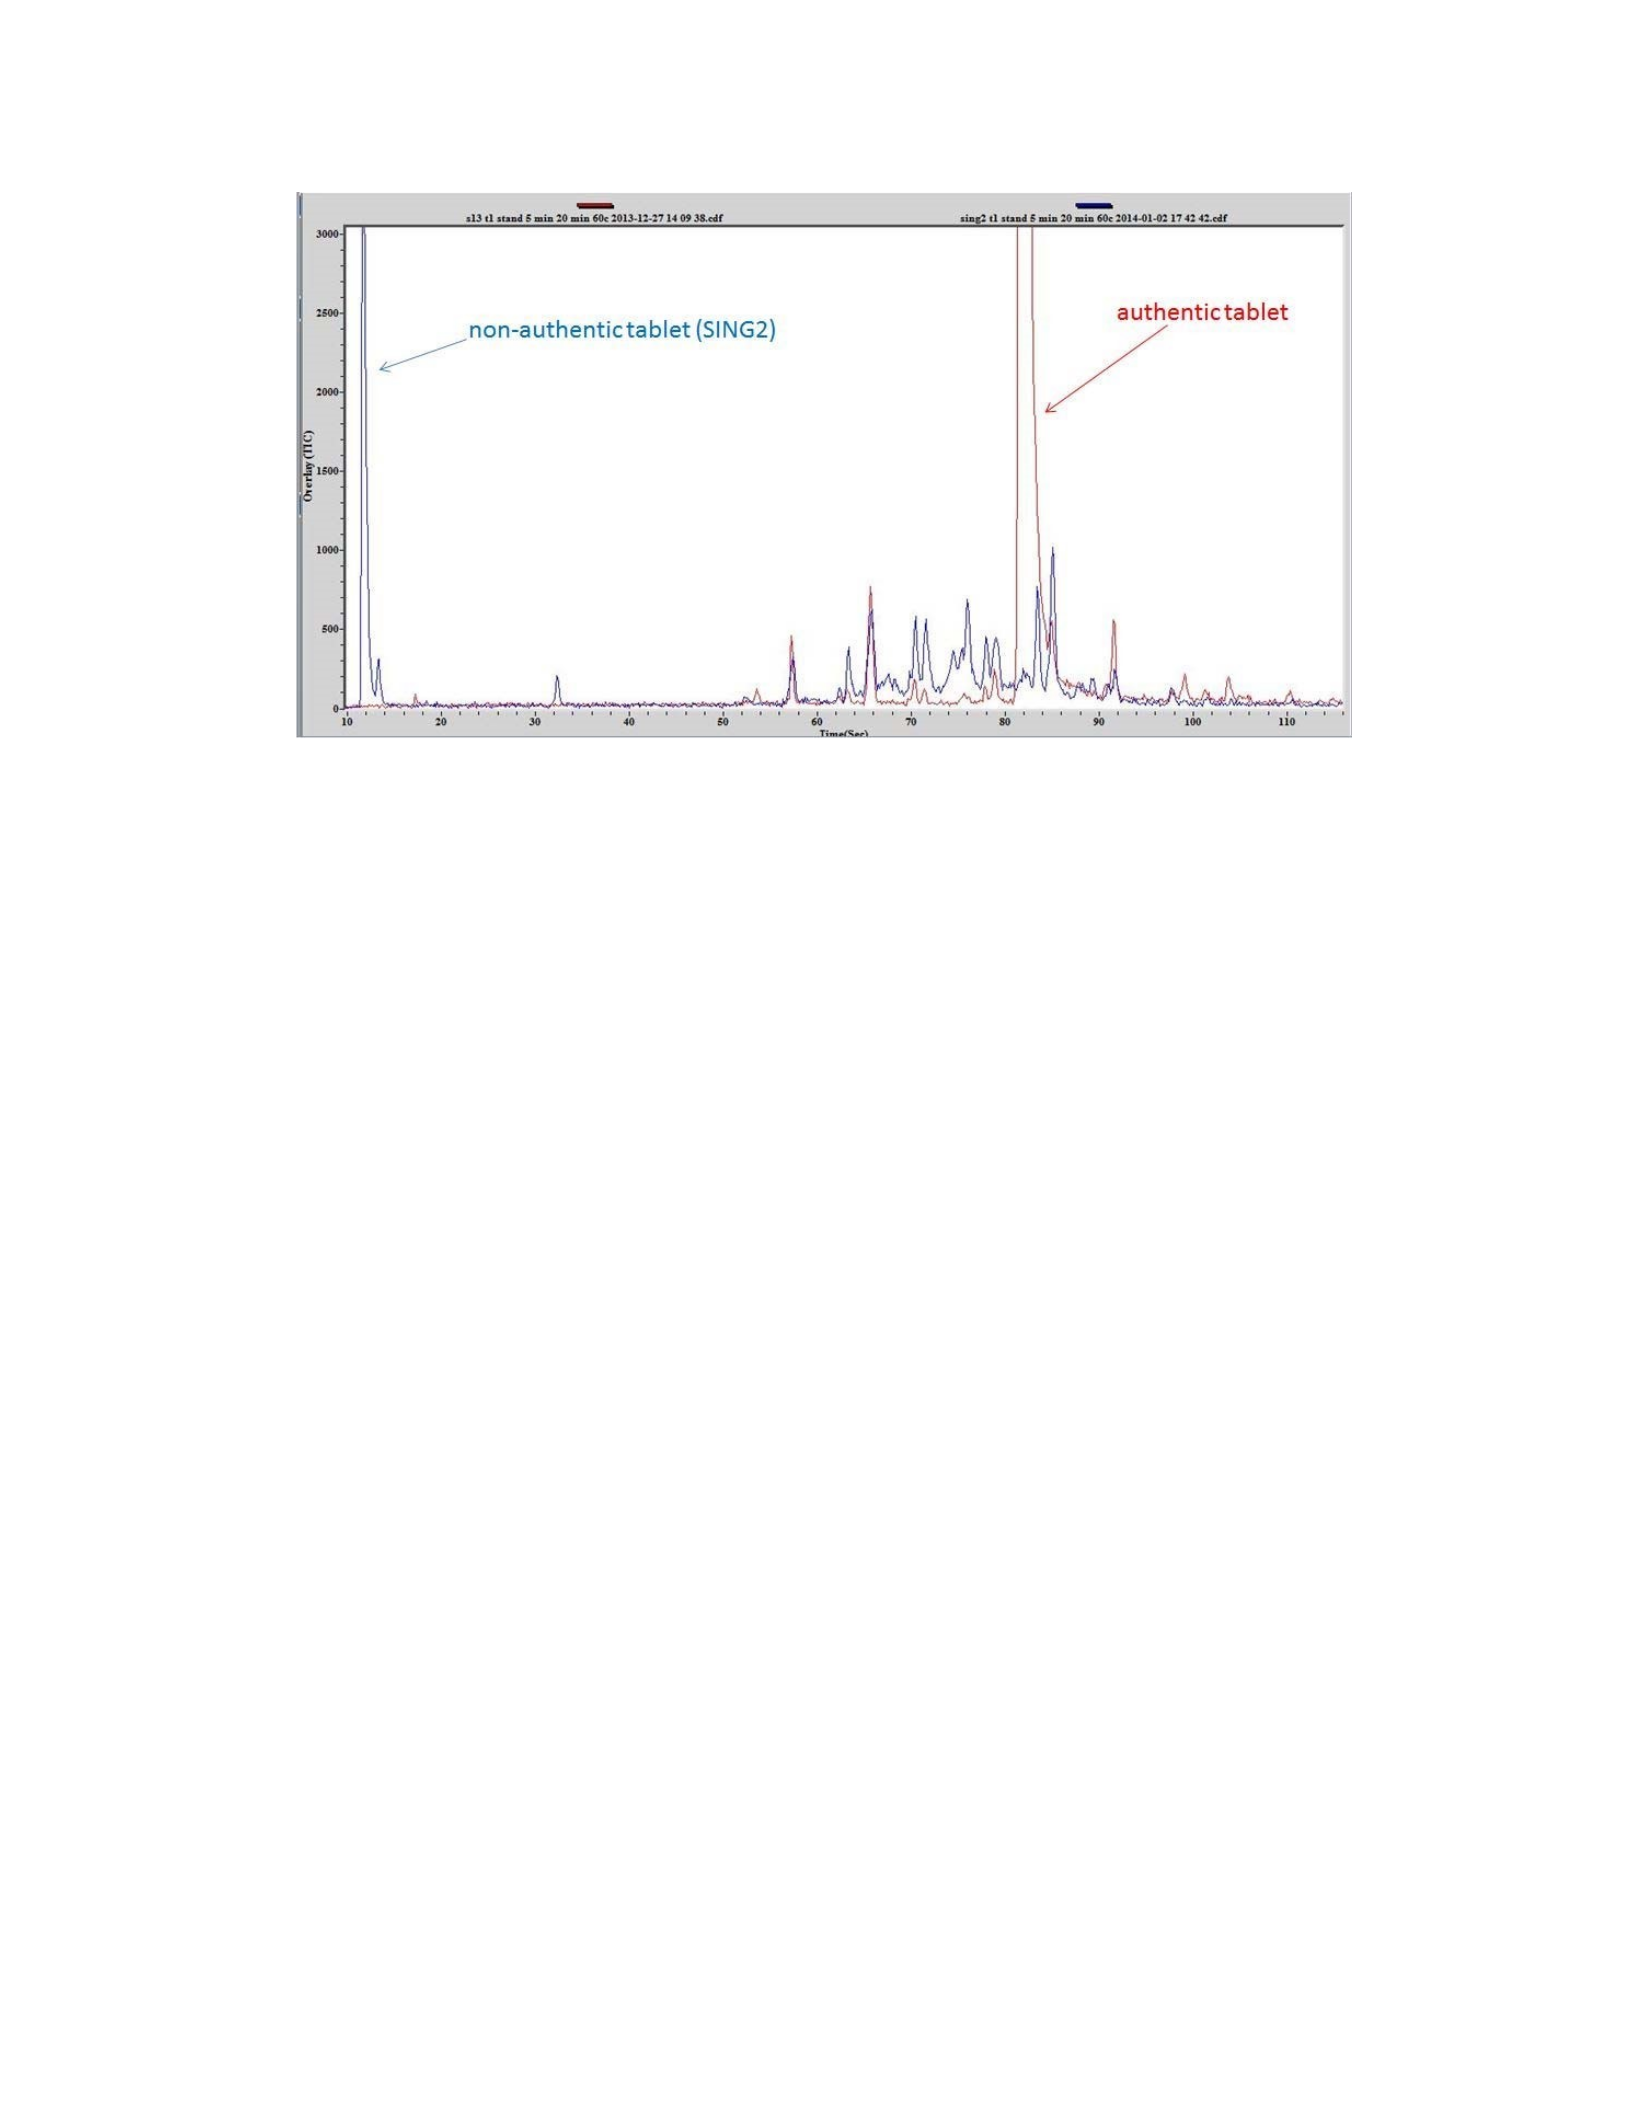
Figure 61. Chromatogram of authentic sample compared with
non-authentic SING2 (PDMS/DVB SPME sorbent, 60⁰C).
Although only a single chromatogram for each sample is shown in the preceding nine
figures, it is important to note that all chromatograms generated from each tablet within a sample
were compared to each other. In all cases, the chromatogram of each sample was generally
consistent from tablet to tablet, but was dependent upon SPME sorbent and sample temperature.
Therefore, only results generated using the same SPME sorbent and sample temperature were
used for differentiation.
Figure 62 shows an overlay of seven chromatograms from seven different tablets of the
same sample collected using the same SPME sorbent and sample temperature (PDMS/DVB
SPME sorbent, 60⁰C). Although peak heights are variable from tablet to tablet, the general
pattern is consistent across the tablets analyzed. This type of chromatogram reproducibility was
typically observed for different tablets of a given sample analyzed using the same conditions.
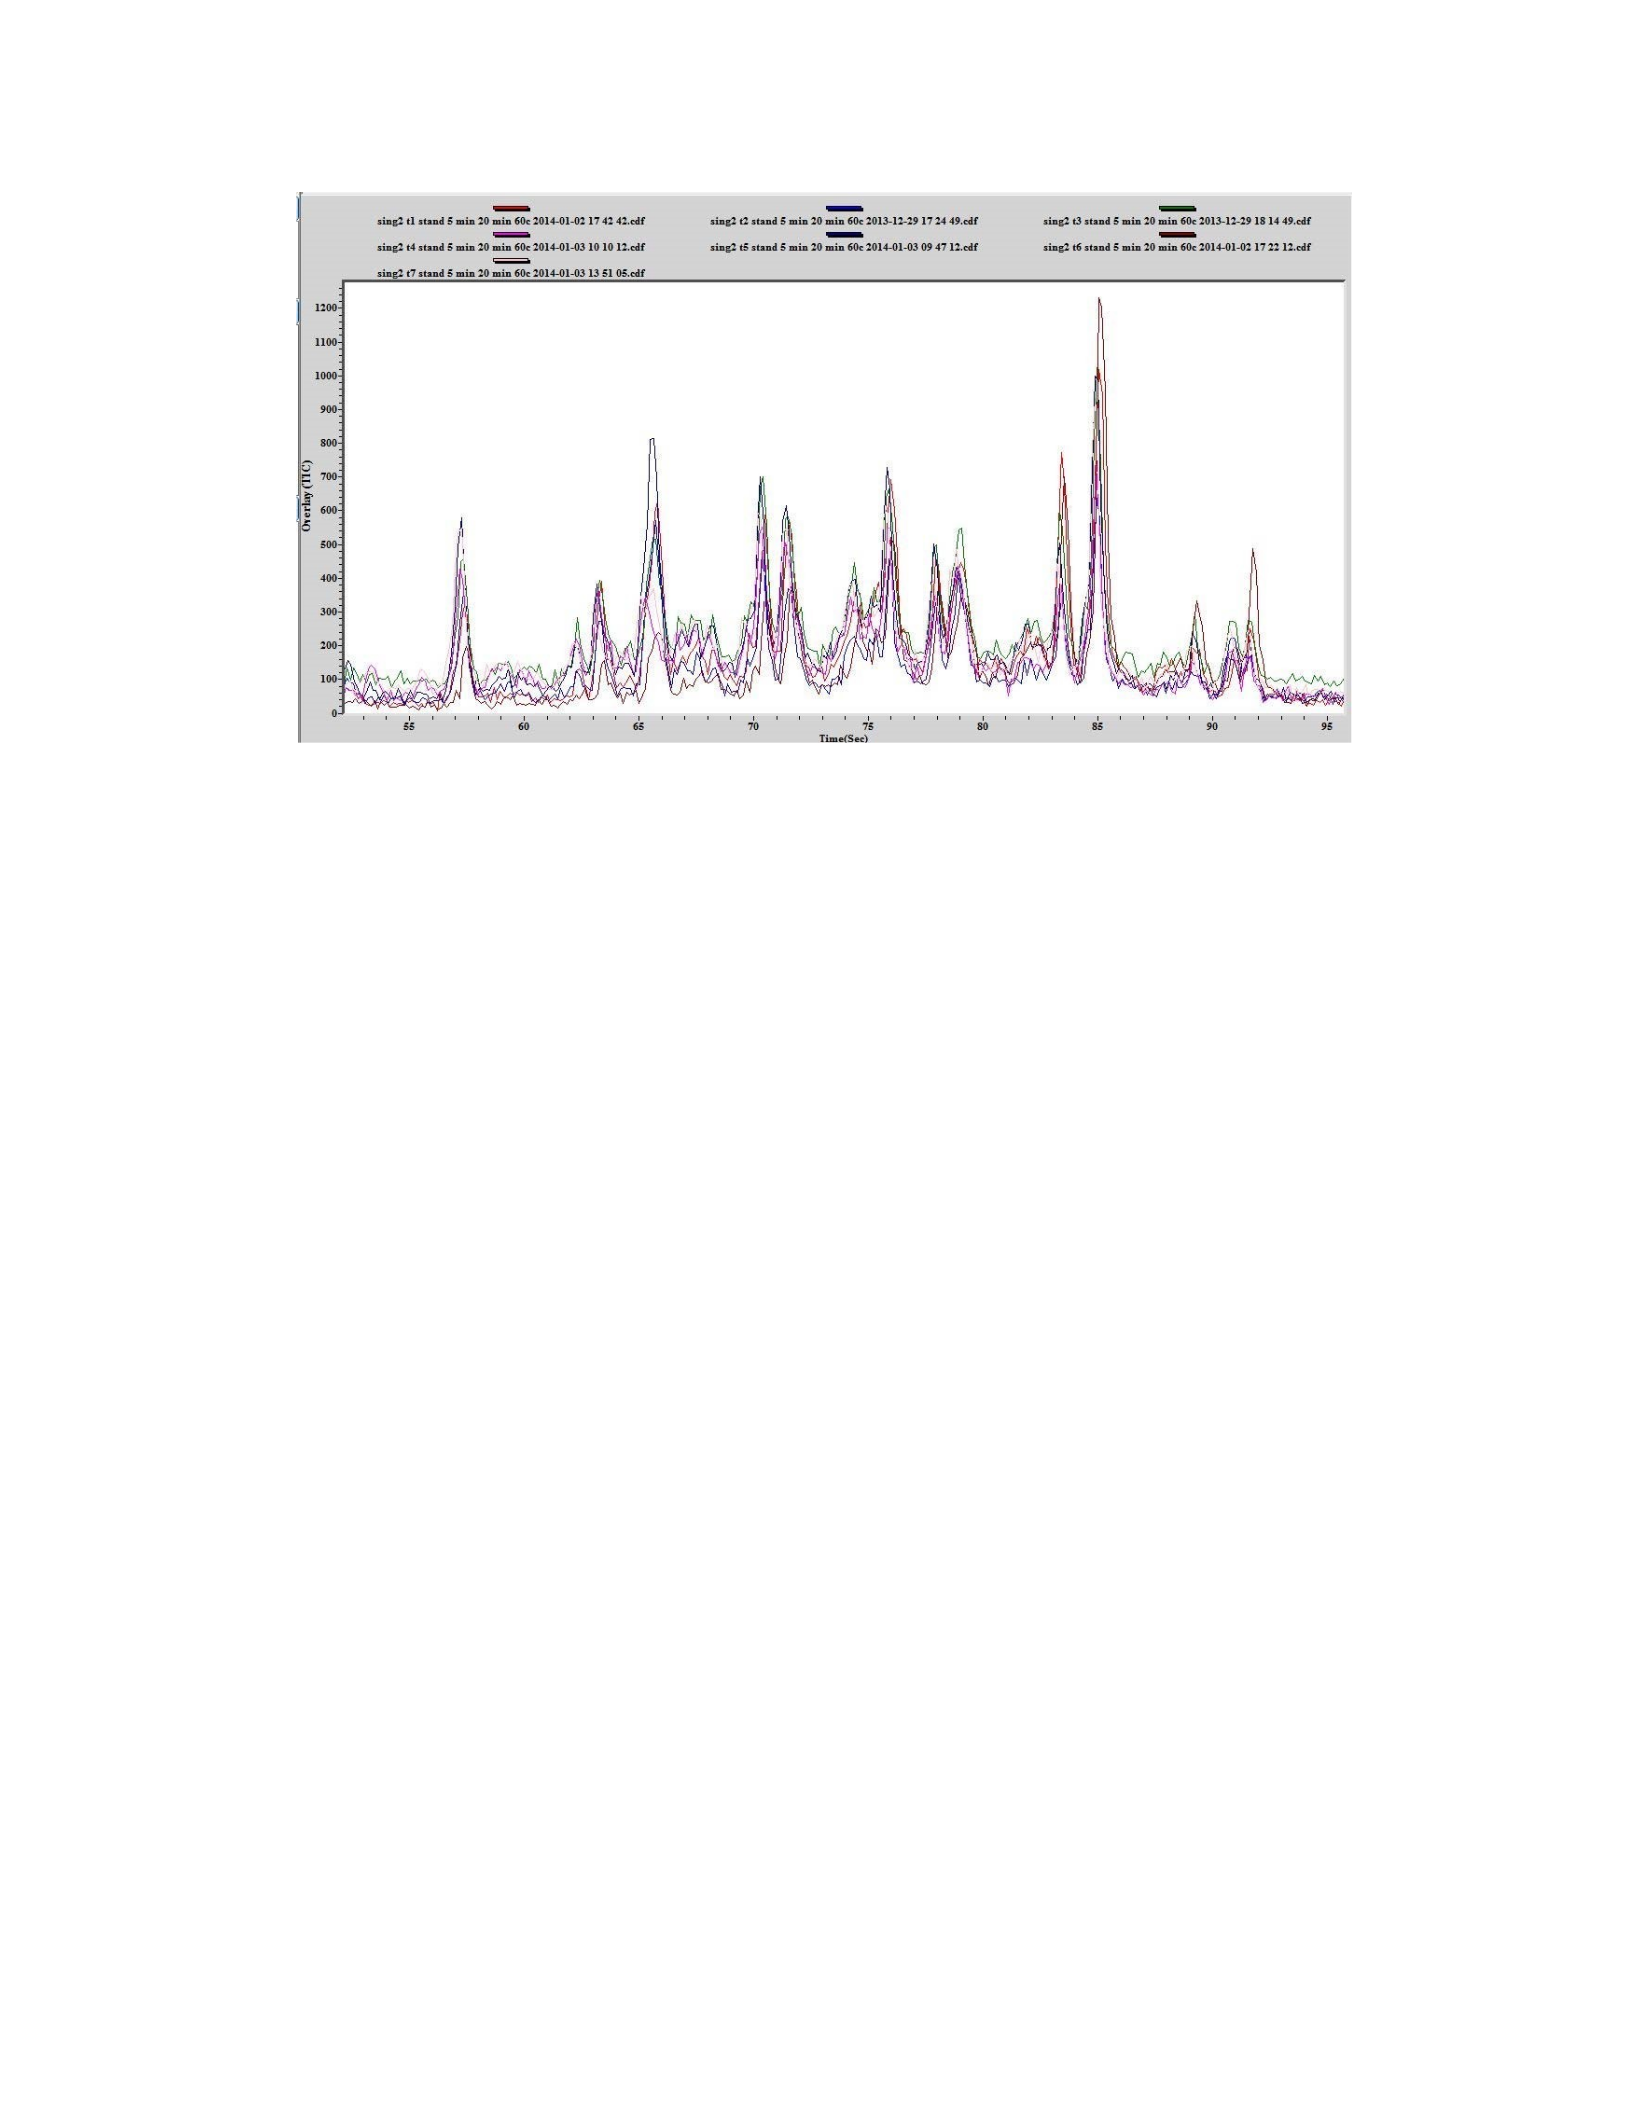
Figure 62. Seven chromatograms of seven tablets of sample
SING2. Sampling was performed using PDMS/DVC SPME
sorbent at 60⁰C.
Chromatogram differences based upon SPME sorbent are expected. Different SPME
sorbents, by design, selectively sorb different chemicals differently. This selective sorption
accounts for the differences in chromatograms from the same tablet analyzed at the same
temperature using different SPME sorbents. As an example, Figure 63 shows the chromatograms
for sample SING2 collected using two different sorbent materials analyzed at the same
temperature and under otherwise identical conditions. Note that the peak intensities for some
chemicals across the chromatogram are different. The most significant differences are observed
for chemicals with lower boiling point which elute at lower retention times on this GC column.
The carboxen fiber is a more aggressive sorbent for these types of chemicals, generating patterns
with higher concentrations of these chemicals in the chromatogram from the same sample
analyzed under otherwise identical conditions. Figure 64 shows replicate analysis from the same
tablet analyzed under the same conditions using the same SPME sorbent. Note that peak
intensities are much more consistent across the entire chromatogram when replicate analysis
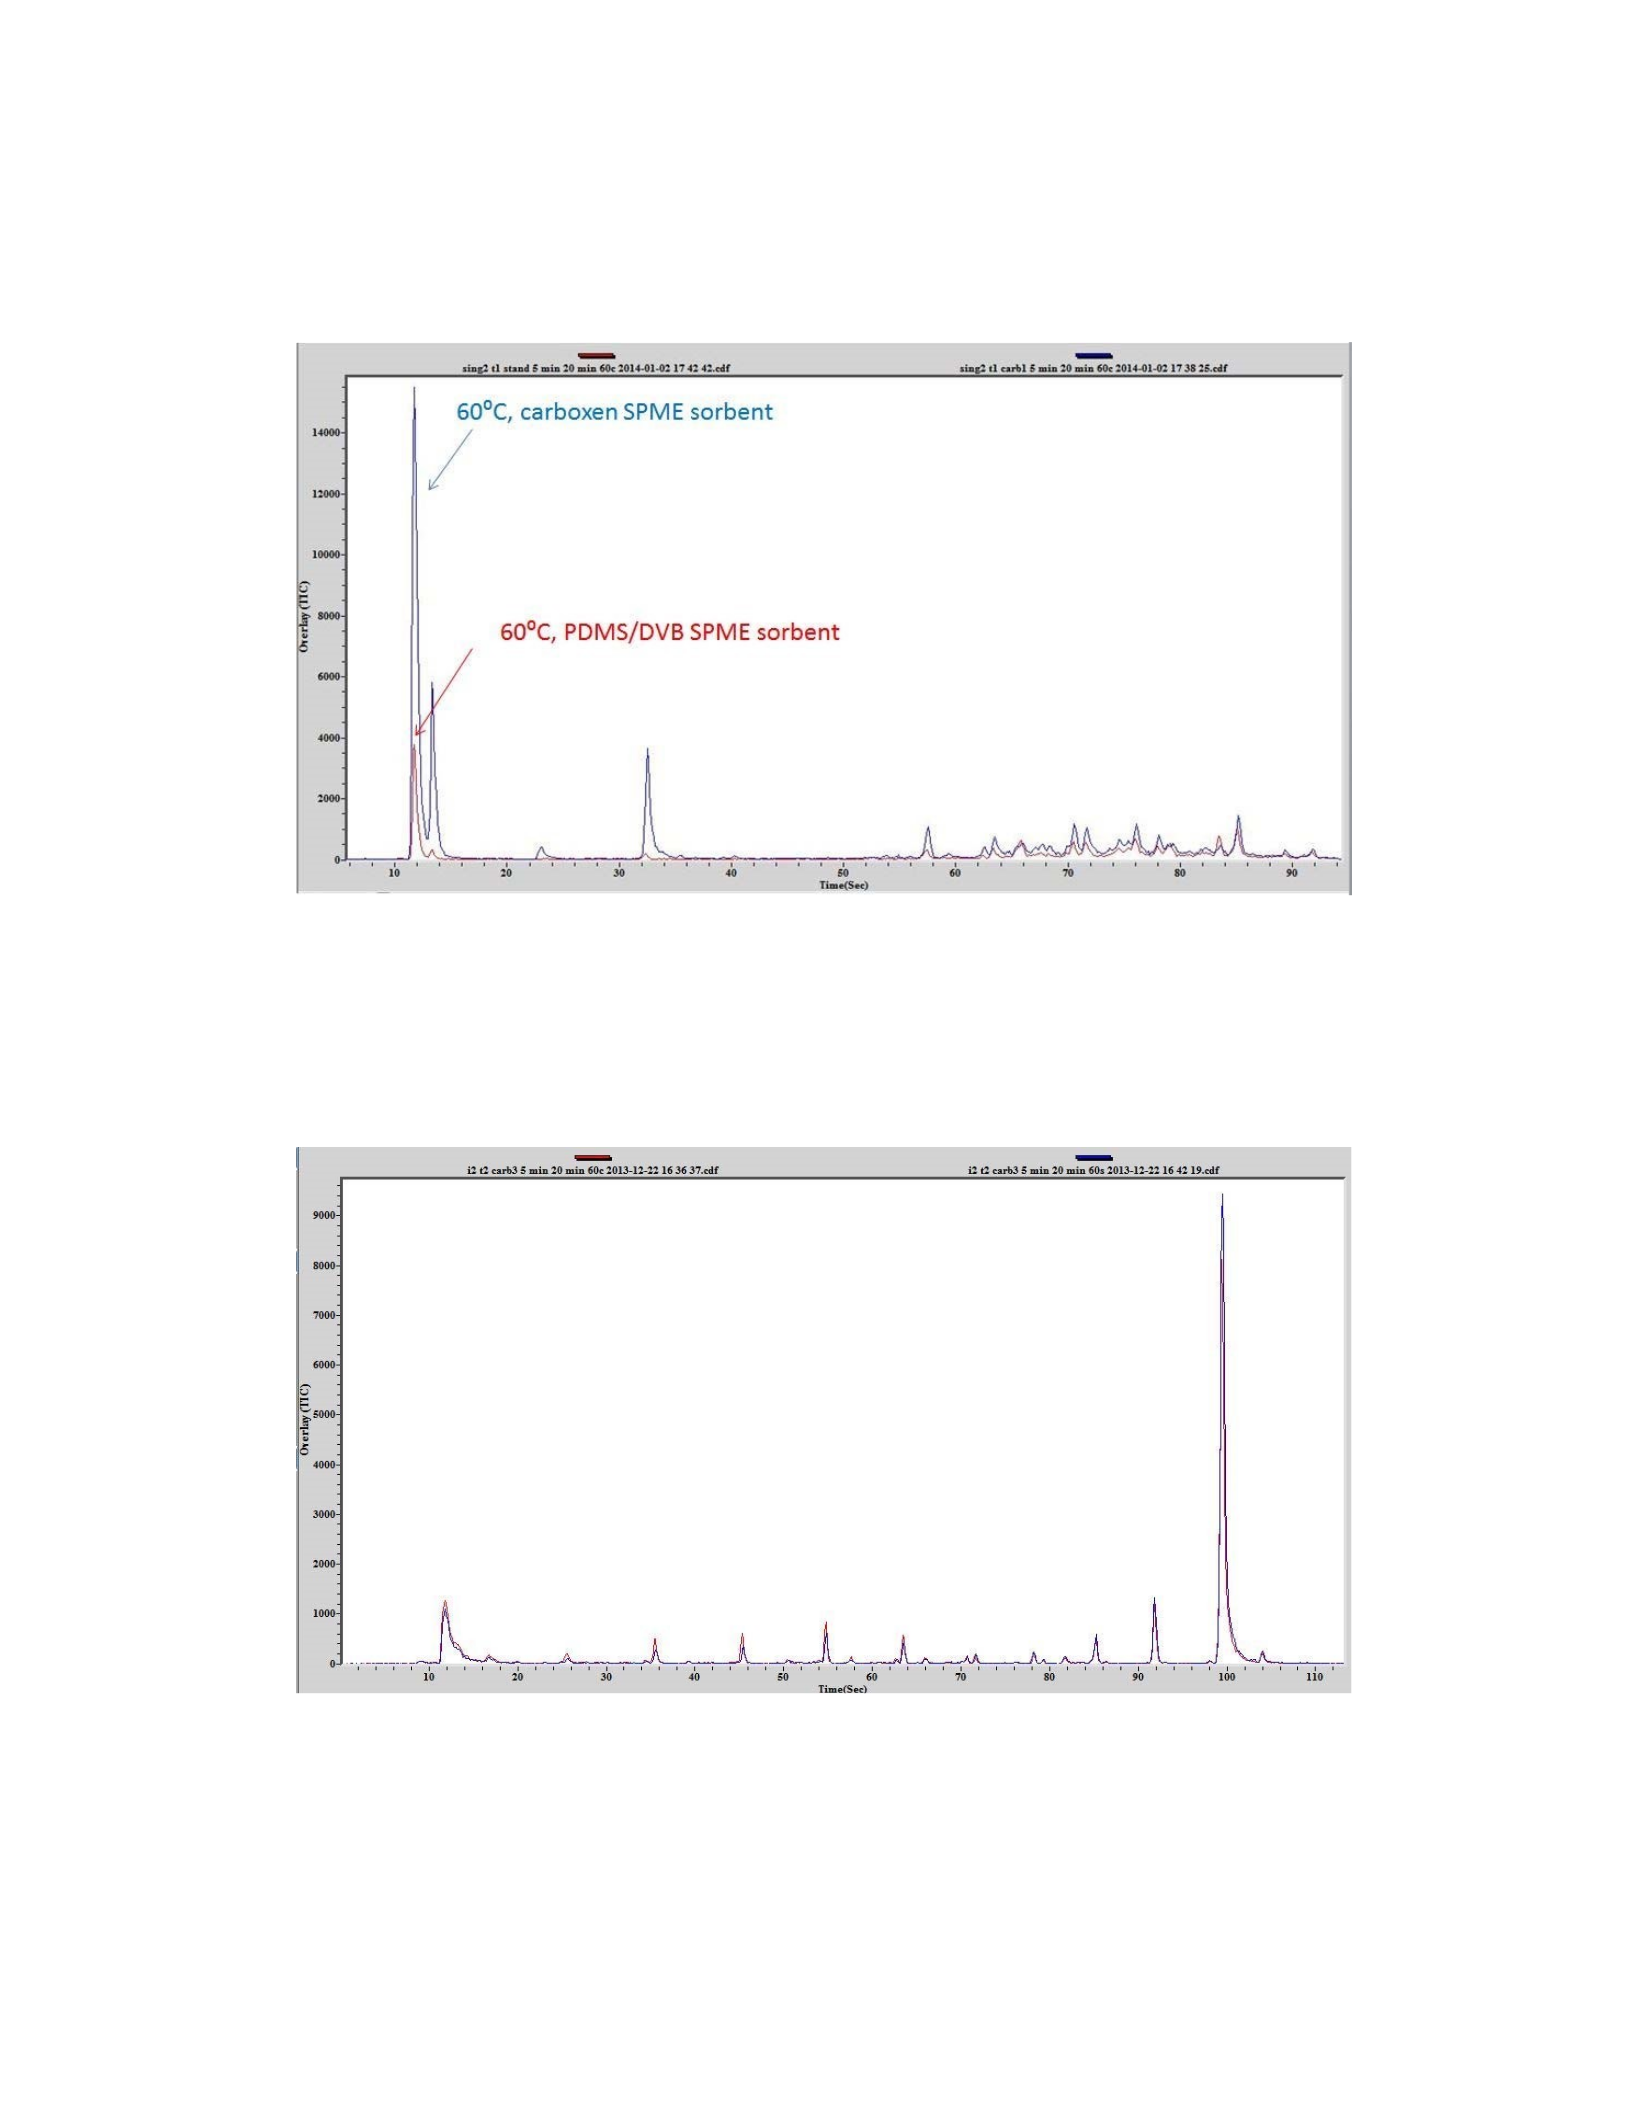
using the same sorbent materials was performed. These figures show typical behavior observed
across all samples analyzed.
Figure 63. Chromatograms of tablet 1 of sample SING2 analyzed
at the same temperature (60⁰C) using two different SPME sorbents
(PDMS/DVB and carboxen).
Figure 64. Chromatograms of tablet 2 of sample I2 analyzed at the
same temperature (60⁰C) using the same SPME sorbent
(carboxen).
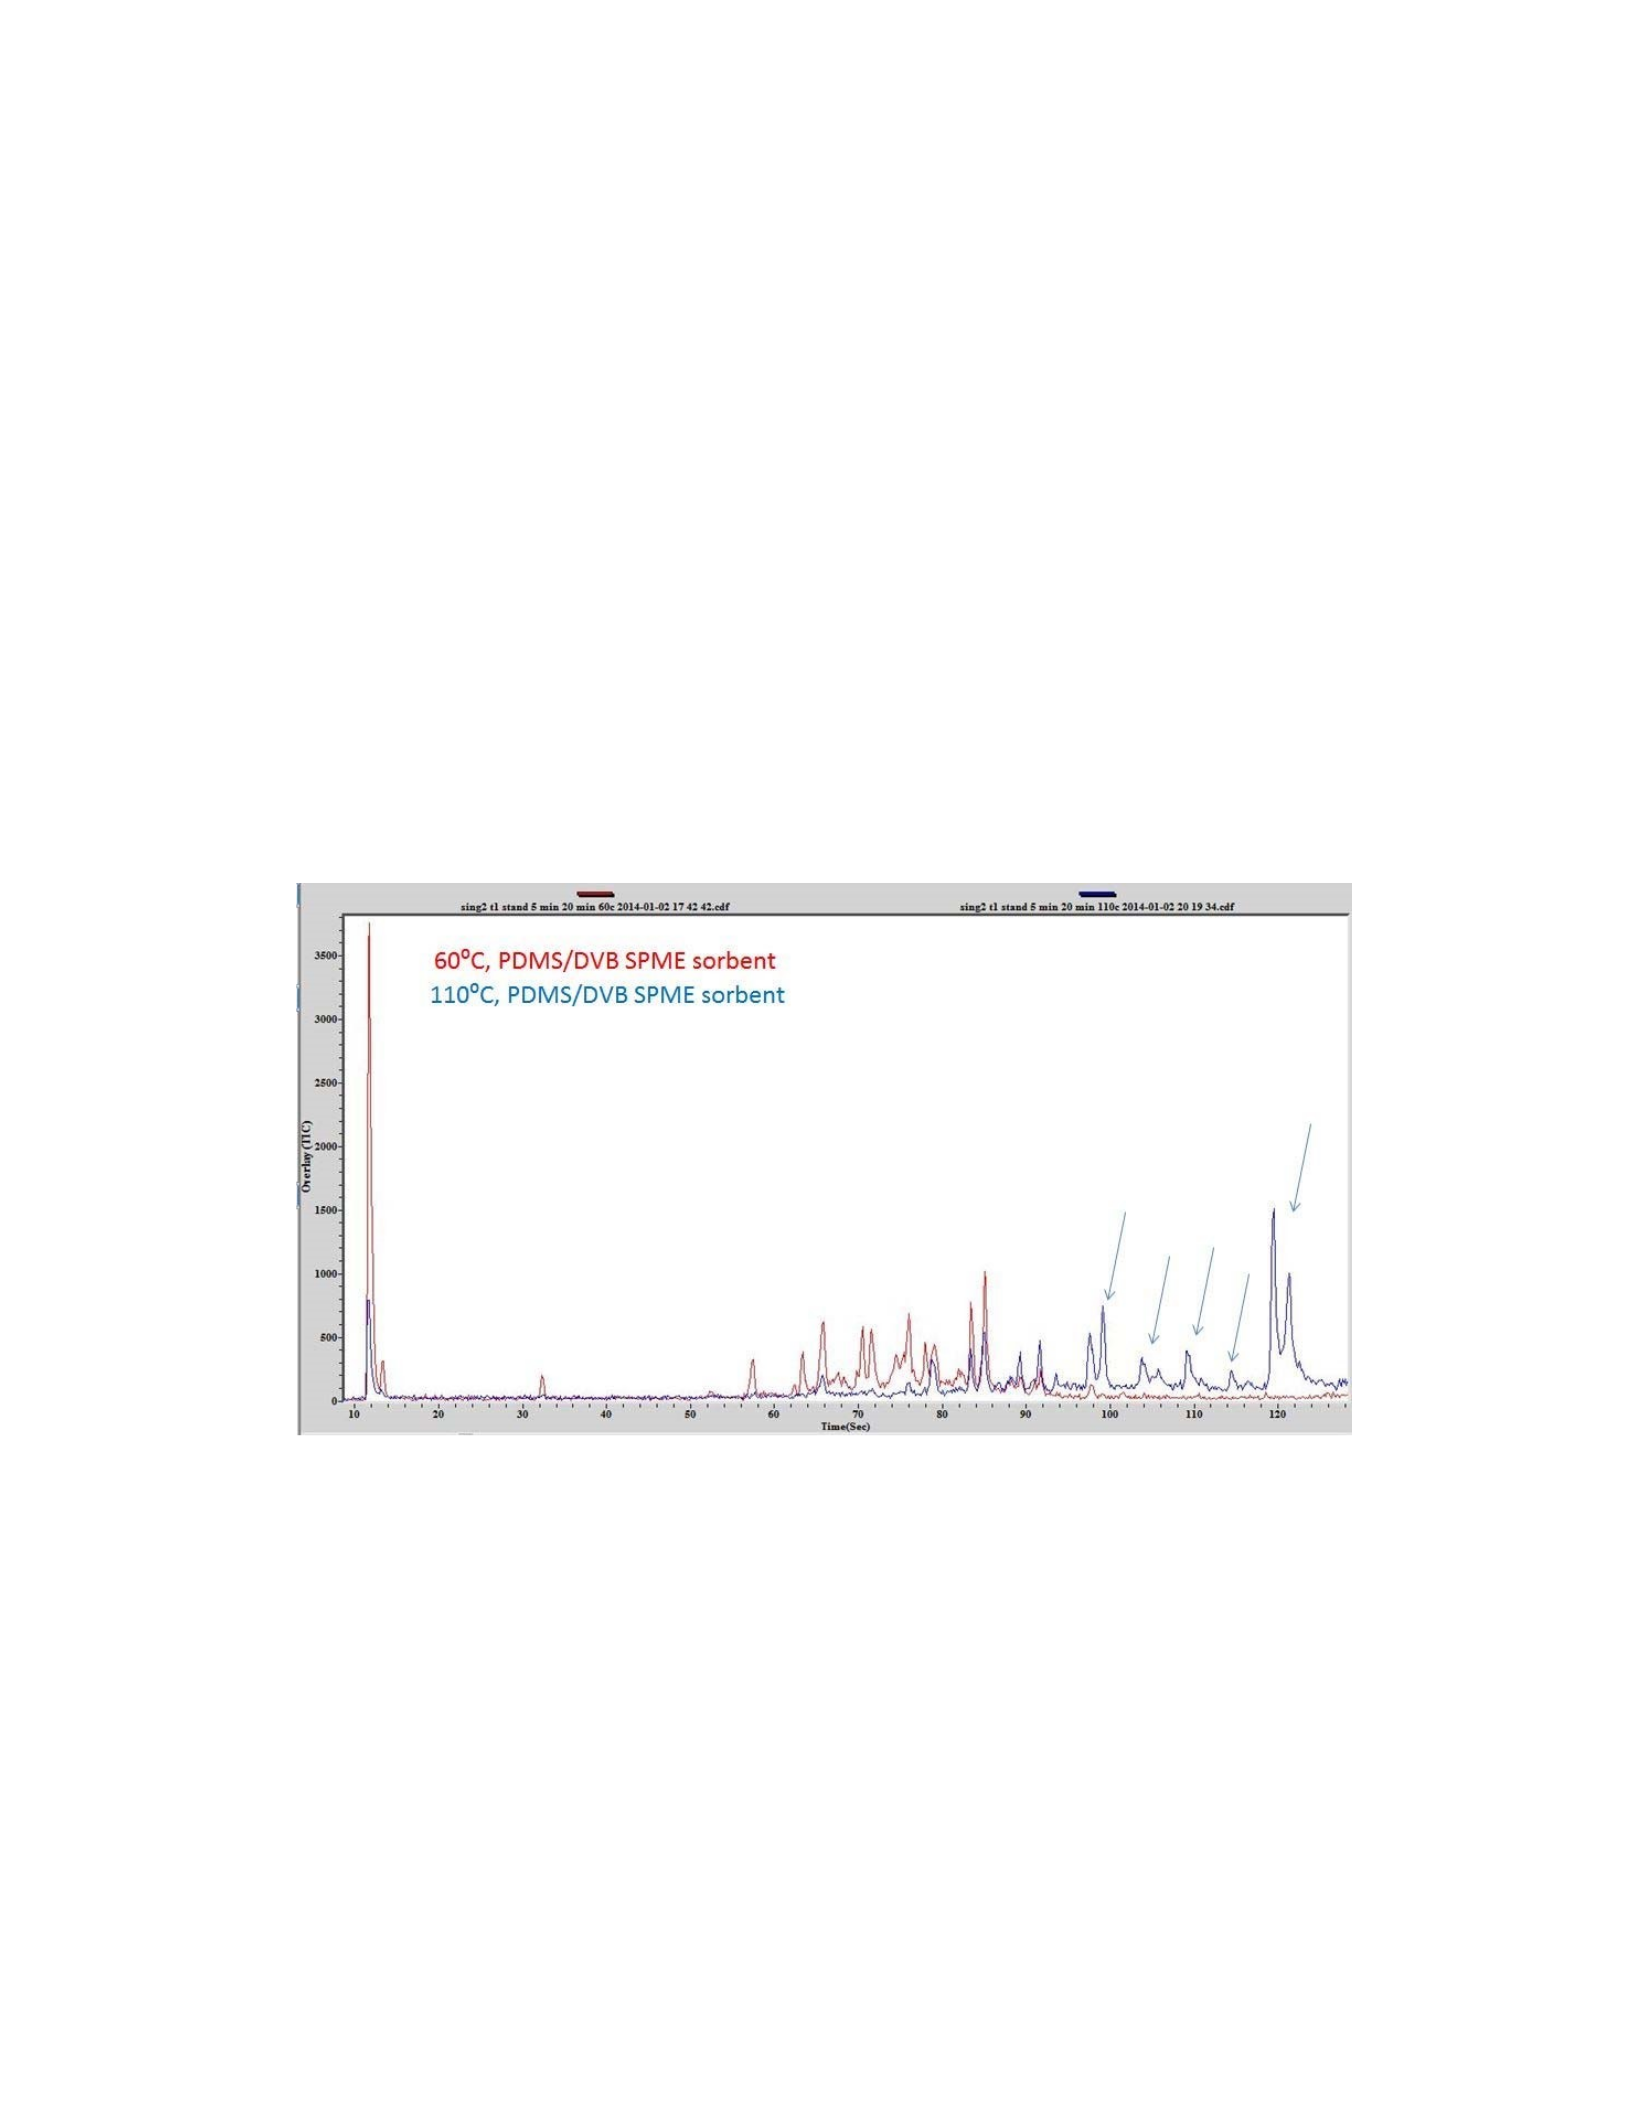
Differences in chromatograms based upon sample temperature are also expected.
Temperature will modify the chemical composition of the headspace and, therefore, the resulting
chromatogram. Figures 67 shows the types of differences that may be observed based upon
sampling temperature. Chromatograms for sample SING2 collected from the same tablet at 60⁰C
and 110⁰C using the PDMS/DVB SPME sorbent are displayed. Although there is varying peak
intensities across the chromatograms, the most significant differences lie in the region at
retention times greater than about 90 seconds, where chemicals present at 110⁰C are completely
absent in the 60⁰C sample. This type of difference based upon temperature was consistent
whether comparisons were made using PDMS/DVB SPME sorbent or carboxen sorbent.
Figure 65. Chromatograms of tablet 1 of sample SING2 analyzed
at different temperatures (60⁰C and 110⁰C) using the same SPME
sorbent (PDMS/DVB).
Summary and Conclusion
This research shows that detection and identification of organic volatiles and residual
solvents in drug tablets using field-portable GC-MS provides a chemical signature that can be
used to authenticate product. A visual comparison of the chromatograms produced by these

samples made this a relatively trivial task. Chromatograms were reproducible within different
tablets of the same sample and may be used as a chemical fingerprint of a specific tablet and
sample. These chemical fingerprints may be used to classify samples of unknown origin and
provide investigators insight into the sample’s manufacturing process and history.
CHAPTER 11. AN ANALYTICAL SCHEME FOR TESTING OF SUSPECTED
COUNTERFEIT TABLETS IN THE FIELD
The data summarized in the preceding chapters shows that differentiating authentic from
non-authentic samples using many different analytical methods in the field is possible. The
choice of method to be used is dependent upon the sample. In some instances, a visual
comparison between the authentic and non-authentic samples will suffice. In others, chemical
methods must be used to attempt differentiation. It may be prudent, though, when analyzing
suspected counterfeit drugs in the field, to measure samples using a combination of methods
regardless of whether or not a non-authentic sample is detected during the initial examination.
This is because although individual methods may be used independently to show that a suspect
sample is not authentic, the real challenge facing forensic scientists is in the sourcing of these
goods.
It seems that any attempt at sourcing requires access to data from many different samples
in order to identify patterns in the date collected from non-authentic samples. There is a need for
a database of these samples. Data collected quickly in the field at border crossings and other
law-enforcement venues can provide the content for this database and be easily saved in an
electronic format for review at a later time by members of forensic science and law-enforcement
organizations. It is important that the methods used maximize information and minimize testing
time and difficulty so that the workflow of the field analyst is not too cumbersome. The results
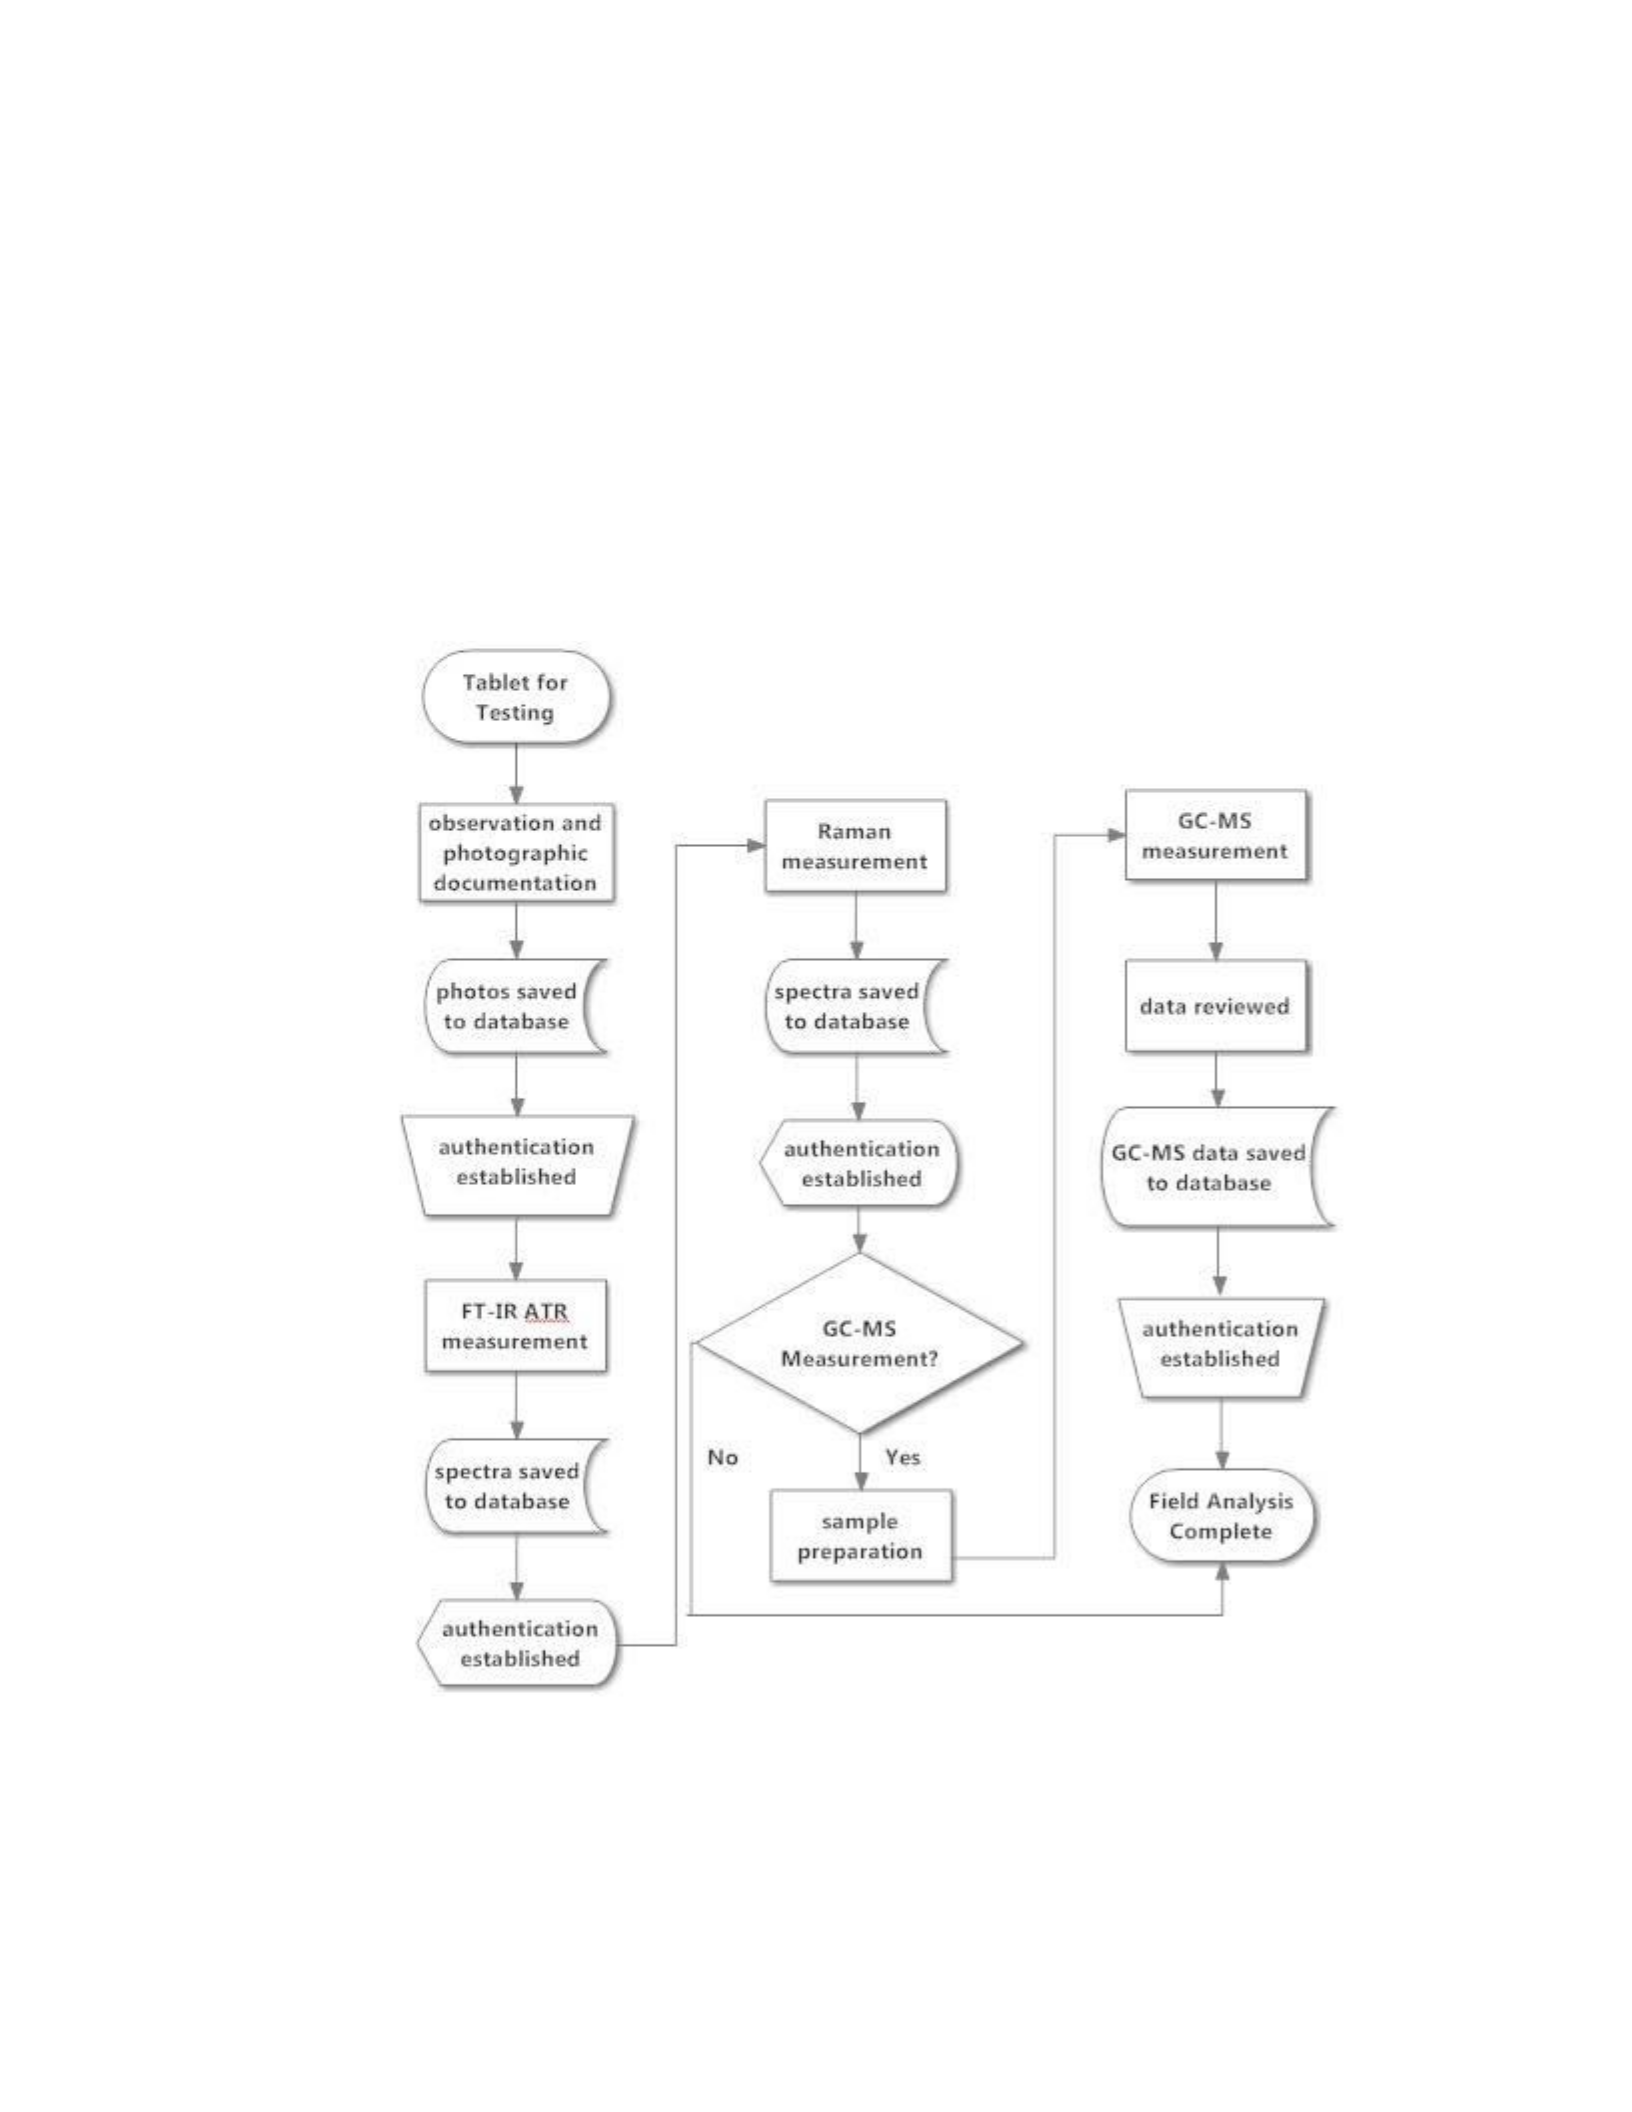
presented in the previous chapters of this dissertation show that handheld instruments can be
used to generate useful data that, if entered into a database, could be retrieved by forensic
scientists and members of the law-enforcement community to perform important review and
comparisons that may be used to source counterfeit drugs. For these reasons, a proposed
workflow for the analysis of suspected counterfeit tablets to be used in the field is shown in
Figure 66.
Figure 66. Workflow proposed for testing suspected non-authentic
tablets in the field.

It is important to highlight that this scheme is intended for field use by non-scientists, and
is not intended to replace lab-based methods that may be used to establish authenticity. The
purpose when performing this scheme in the field is to detect differences between the test sample
and authentic versions of the sample. While it is true that if differences are detected a sample can
be labeled as non-authentic, if no differences are detected, authenticity is not confirmed.
Determination of authenticity is likely best performed in a laboratory setting using methods that
are more discriminating. Frequently, conferencing between the brand owner and the forensic
scientist will prove to be the best way to determine the best methods to verify authenticity in the
laboratory. The brand owner is most familiar with their product and can provide insight as to
what methods work best. Figure 67 shows a general workflow for laboratory analysis of
methods. It’s important to note that depending upon the sample, some methods may be better
than others. It is also important to note that this scheme is not inclusive of all potential methods
of analysis.
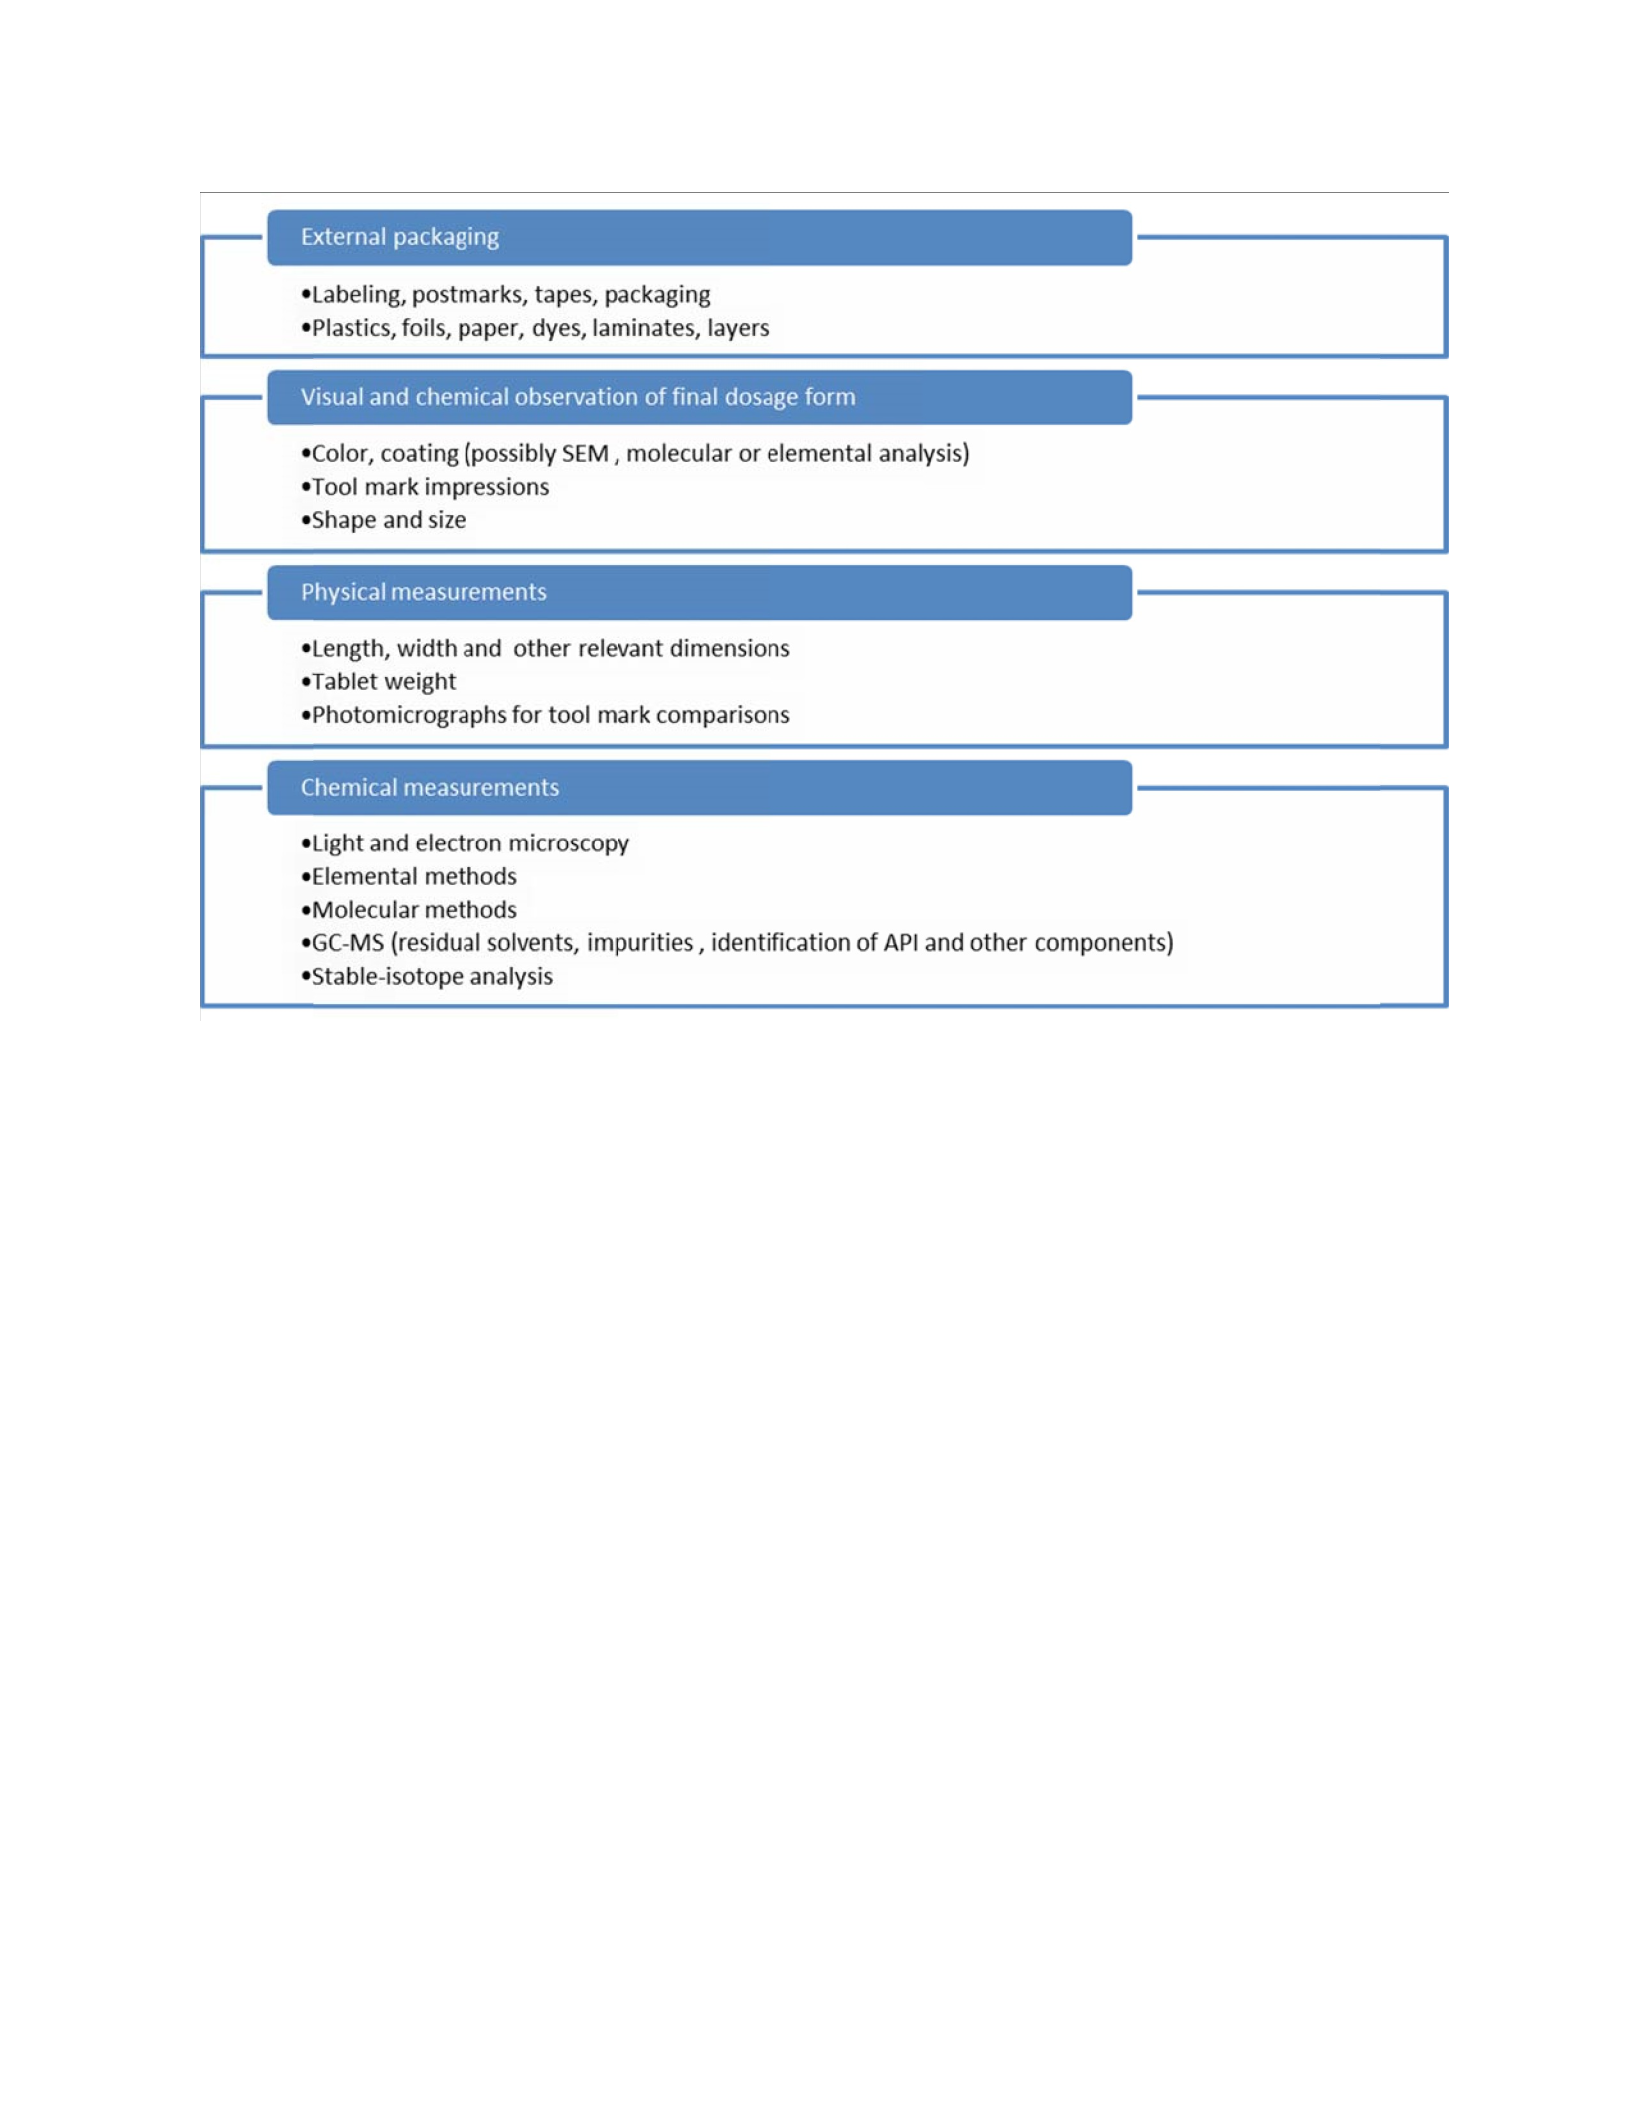
Figure 67. Workflow proposed for lab-based analysis of suspected
non-authentic samples.
Analytical Workflow
The first step of any analytical scheme to be performed in the field to test counterfeits is
observation and photographic documentation. All photographs should be taken under the same
conditions and saved to the database. Review of the data to establish authenticity from a
physical examination would likely be manual unless image-analysis methods are developed. For
this reason, it may be important to have a second analyst reviewing data in near-real time,
especially if a non-authentic sample is detected. This way any conclusions drawn by the field
analyst are supported by a second analyst. Tablet color, indicia, and dimensional measurements
will frequently enable fast and simple differentiation of authentic from non-authentic samples.

The most significant challenge to this type of analysis is that it requires that the analyst have
knowledge of the appearance of authentic versions of the drug product. This may or may not be
the case. Determination of whether or not a suspect tablet is indistinguishable from the authentic
sample is based upon physical appearance, and would be made by the analyst after direct
comparison of the physical properties of the sample. Groupings of tablets in the database by the
forensic scientist based upon visual appearance may be attempted at a later date if determination
of source or origin is desired. Measurements may be taken at the time of inspection, but if no
reference is available for comparison, this step is not necessary. Photographs captured at the
same magnification can be reviewed at a later time to determine tablet dimensions, if necessary.
The next step in the analytical process is to collect infrared spectra from the tablet using a
field portable ATR FT-IR spectrometer. A system that uses a larger analysis spot size is preferred
in order to sample the largest amount of sample during collection of a single spectrum. Infrared
spectra collected from the tablet’s coating and from its core can be automatically searched
against electronic databases of library reference spectra to establish a match factor between the
spectrum from the test sample and library spectra of authentic samples. Results are fast, and a
numerical value is assigned to establish similarity quantitatively. ATR FT-IR spectra are very
reproducible for various reasons. (Reffner & Martoglio, 1995) It would, therefore, be
appropriate to assign a minimum match factor score to be used to automatically determine that
the suspect tablet cannot be differentiated from authentic versions using this method. If the
match-factor value is met or exceeded, the tablet cannot be differentiated from the authentic
reference; if the match-factor value is not met, the sample is identified as a non-authentic. The
instrument could be programmed to generate a real-time pass/fail result to automatically appear
on the instrument’s results screen at the time of analysis dependent on the sample’s match factor.

Infrared spectra collected, regardless of whether or not they are displayed on the instrument’s
result screen, should be stored in an electronic database. Storage of spectral data in a searchable
database would allow forensic scientists and law-enforcement personnel performing
investigations to use the data to generate investigative leads. This type of review may be used to
group samples together so that provenance may be established.
The next step in the analytical process is to perform a Raman measurement of the tablet
core using a portable Raman spectrometer. Just like with infrared spectrometry, a Raman
spectrum collected from the tablet’s core can be automatically searched against electronic
databases of library reference spectra to establish a match factor between the spectrum from the
test sample and library spectra of authentic samples. Results are fast, and a numerical value is
assigned to establish similarity quantitatively. Although Raman spectra are generally
reproducible, differences in peak intensities and other small artifacts of the measurement may be
observed between spectra. A minimum match-factor score can be used to automatically
determine that the suspect tablet cannot be differentiated from authentic versions using this
method. If the match-factor value is met or exceeded, the tablet cannot be differentiated from the
authentic reference; if the match-factor value is not met, the sample is identified as a
nonauthentic. The instrument could be programmed to generate a real-time pass/fail result to
automatically appear on the instrument’s results screen at the time of analysis. Raman spectra
collected, regardless of whether or not they are displayed on the instrument’s result screen,
should be stored in an electronic database. Storage of spectral data in a searchable database
would allow forensic scientists and law-enforcement personnel performing investigations to use
the data to generate investigative leads. This type of review may be used to group samples
together so that provenance may be established.

The next and final analytical step in the process is GC-MS analysis to determine a tablet’s
residual-solvent and organic-volatiles profile. GC-MS is different from the other analytical
methods proposed as part of this analytical scheme because it requires sample preparation that is
time consuming (approximately 25 minutes). In addition, analysis time is on the order of
minutes rather than seconds. The ability to establish this profile is powerful, though, especially
in providing investigative information. Solvents and volatile compounds detected and identified
may be used as chemical markers. These markers can be used to group samples together and can
provide information about manufacturing methods and processes. This information can be very
helpful when establishing provenance.
Use of all of these methods has value. Photographs make it possible to easily observe
differences amongst samples. Both infrared and Raman methods provide chemical information
about the sample and can be used to quickly detect non-authentic samples. Each method
(infrared and Raman) has its benefits, and while some information interpreted from the data from
these two methods may be redundant, the methods complement each other well and provide a
more complete picture of the nature of the sample. Performing photography, infrared and Raman
analysis of a sample may occur in less than five minutes. It is important to recognize that the
goal of this field analysis is to detect non-authentic samples. It is not the goal of this analysis to
verify authenticity. Using these methods, the inability to detect differences should not be
confused with verification of authenticity.
GC-MS analysis is also proposed as part of this scheme because the data can be quite
informative and is not redundant to photography, infrared, or Raman analysis. However, the
challenges related to time and value to the field analyst should be considered before inclusion of
GC-MS in the field-testing scheme is finalized.

Something else to consider is that when using this analytical scheme, the tablet sample
rather than the packaging needs to be analyzed. Methods like radio frequency identification tags
are useful at tracking and tracing drugs through the supply chain (Rudolf & Bernstein, 2004), but
this technology is usually applied to packaging and is not useful in situations where packaging
has been removed or repackaging occurs. Alternative methods that require analysis of the
sample contained within the packaging are necessary.
Database Management
The methods described as part of this analytical workflow for analysis of samples in the
field provide physical and chemical information about samples analyzed. The primary goal,
especially for the field analyst, is to detect and identify non-authentic samples. However, data
collected can provide much more value if it is stored in a database for retrieval and review by
forensic scientists or other law-enforcement personnel. The methods proposed are well suited for
analysis in the field. The data is relatively easy to collect quickly. The technologies used allow
for interpretation of the physical and chemical nature of the sample. The ability to maintain this
data in a searchable database format is powerful because it enables offline processing and review
of large amounts of data from remote and potentially distant locations. This capability is well
suited to the challenge of counterfeit drugs. These goods come from different regions of the
world, and the place of manufacture is frequently geographically distant from the place of sale or
consumption. The ability to collect data from different geographical locations, combine them
into a single database, and then review and process to evaluate trends and perform other
interpretations is invaluable. There are some anticipated challenges in managing a database of
this magnitude. Creation, implementation, and maintenance of the database each offer unique
challenges that need to be addressed.

The way in which the database is formatted during its creation is important and,
ultimately, will be directly related to its success. The goal is for the forensic scientist to be able
to take data from the database and review this data in as simple and straightforward a manner as
possible. Therefore, it will be important to be able to search the database using many different
search criteria. For instance, in one situation it may be important to search based upon data
collection site, or time and date of data collection, while in another the critical search criterion
may be the purported drug product. The anticipated use and users should be considered during
database creation so that the database is as user friendly as possible. In addition, a method for
inclusion into the database of customs documents and other appropriate paperwork should be
considered.
Another important factor to consider during database creation is file formats for data
storage. It is important that if handheld devices from different instrument manufacturers of the
same technology will be contributing to the database, it should be required that files written be
converted to a general data format that is not dependent upon which manufacturer’s instrument
was used to collect the file. For instance, infrared data could be collected and reviewed in the
field using the proprietary software that generates a pass/fail response and then converted to a
general infrared data file format like *.spc or *.csv when the file is written to the database.
Determination of the best database format for each technology, as well as other data processing
requirements should be considered in detail during the creation of the database.
Other items for consideration during creation of the database include networking
capabilities and security, speed of internet, availability of internet cabling at anticipated field
sites, user interface, and other factors that impact data collection in the field like efficiency and
simplicity.

Implementation of a program to analyze and centrally store data from different locations
seems complicated, but is manageable. Practically, the field analyst’s job may be as simple as
collecting data using a handheld instrument, reading off a result and sending the file to the
database. Training is important so that useful data is collected. For instance, photographs taken
need to be in focus and at a uniform magnification. If not, their value is limited. For spectral
files, the spectrum collected needs to be of an appropriate quality so that when compared with
data collected from other sites, effective comparisons may be made.
It is also important to manage the amount of data collected and keep the data collection as
uniform as possible from site to site. Although it is nice to be able to collect every piece of data
and make available to the user every possible option for data processing and review, this would
be an overwhelming training burden and would likely not be successful. Handheld cameras and
infrared and Raman spectrometers currently available are capable of meeting the field challenges.
However, field analysis with GC-MS is more complicated and requires more time for analysis
than infrared and Raman measurements. Modifications to workflow in the field would need to
be made to streamline them into the process, specifically considering sample preparation time to
generate chemical profiles of residual-solvent and organic-volatile profiles. As previously stated,
the use of GC-MS to establish a sample’s residual-solvent profile may not be appropriate at high-
volume border crossings, but at locations where data collection of this type is possible, inclusion
in the database would provide very valuable insight as to the chemical composition of the
sample, as well as its method of manufacture. GC-MS files, although complex, can be simplified
into a common data format so transfer of the data would be possible.
Maintenance of the database would be required. Aside from maintenance issues related to
the server and other hardware, networking and service provider issues need to be addressed. A

plan for management of software changes and bug fixes need to be in place. In addition, it would
be beneficial to have routine audits of data entered into the database. This would verify that data
collected in the field meets minimum quality standards so that meaningful results are possible
when data is accessed by the forensic scientists and other users for review.
Admissibility in Court
Forensic scientists are typically considerate of whether or not the testing they perform
will be admissible in court. The question of admissibility in court is complicated to answer when
considering the testing and workflow provided in this analytical scheme. When considering
analytical testing in the field to determine authenticity, the answer to this question is not usually
dependent upon the fundamental technology of the instrument, but rather on the methods and
procedures in place for use, maintenance, and workflow when using the instrument, as well as on
the training and education of the analyst.
The methods used in this research for field analysis of suspected counterfeit drugs using
infrared, Raman and GC-MS technologies provide the end user with the ability to distinguish an
authentic from a non-authentic sample. These three technologies have all been successfully
admitted in court when tested against the standards for the detection and identification of drugs
when used in a laboratory setting. (State of Arizona, Appellee, v. Ronald Michael Lucero,
Appellant, 2004; The People of the State of New York, Plaintiff, v. Donald Roraback, Also
Known as Donald Reed, Also Known as Tom Reed, Defendant, 1997; Warner Chilcott
Laboratories Ireland Limited, et al., Plaintiffs, v. Impax Laboratories, Inc., et al., Defendants.
Warner Chilcott Laboratories Irealnd Limited, et al., Plaintiffs, v. Mylan Pharmaceuticals Inc., et
al., Defendants., 2012) For all three technologies, instrument design and the type of data
generated between lab-based and field-portable instruments is consistent. This would lend

support for the admissibility of these field-portable technologies in court. However, there are
many factors that need to be considered if results collected from a field user were to be
admissible in court including maintenance and calibration, instrument performance testing, user
training and experience, standards for comparison, and purpose of testing.
Proper maintenance and calibration allows the end user to be confident the instrument is
performing in accordance with manufacturer specifications. It is critical that these functions be
documented, and that a record of all maintenance performed is available for review. Procedures
should be in place to verify that this is the case. Procedures should also be in place to verify that
as soon as an instrument fails performance testing, these issues are addressed. It is reasonable to
expect that these criteria could be met when deploying these instruments in the field. Infrared,
Raman, and GC-MS portable instruments are quite robust, and it is reasonable to assume that
when using commercially available instruments, maintenance and calibration requirements in the
field would not be too great a burden to meet.
Instrument performance testing is also very important for field-portable instruments.
When using infrared and Raman systems, this type of testing is fast and easy and could easily be
incorporated into the workflow for these instruments in the field. For GC-MS systems, the
methods used to verify performance are a little bit more complicated and may require more time
for this aspect of the workflow. This requirement of time would further burden an already
timeconsuming field method. However, the value of this data collected, as previously described,
is high and likely worth the investment of time required in situations where time for analysis is
available.
User training and experience is a big factor impacting admissibility of scientific evidence
because users of the technology may not have a strong background in a scientific discipline.

Typically, end users are not scientists, but are expected to be able to interpret scientific evidence.
Field-portable instruments are intended to be easy to use and designed to generate results rather
than data. In fact, most field users rely on the match factor generated by the handheld instrument
rather than concern themselves with interpreting a spectrum. However, extensive training on the
theory and design of the technology and the instrument or on the process used to generate match
factors is not usually extensive. Therefore, the field analyst is not acting as a scientist in their
approach to the evidence, but rather as a technician. This does not mean that the evidence is
inherently inadmissible (Kumho Tire Company, Ltd., et al., Petitioners v. Patrick Carmichael,
etc., et al., 1999). However, it does present challenges in court when considering the individual
who performs the test and his or her suitability to testify in court. It would be prudent to
establish procedures that verify that the person determining that a sample is non-authentic
understands how the instrument works, what aspects of the sample are being tested, what the data
means, and other things an expert witness should understand about the method and technology.
There are ways to address this. For instance, a scientist could be assigned to supervise all non-
authentication determinations. This person could be responsible to review the data in real time
and verify the data and results used to determine authenticity are accurate. Field-portable
instruments and the workflow described in this dissertation are not used to verify authenticity, but
rather to detect non-authentic samples. The supervisory scientist could represent the data when
presented in court.
It is not reasonable to expect that standards for comparison would be available for testing
during field analysis, or that time would be available in the field to perform such testing. When
performing analysis in the field, end users are forced to rely on electronic libraries of data
available on the handheld unit. This is not necessarily a problem because the quality of the

library used can be secured and verified prior to deployment of the instrument. However, it is
typical for an analyst working in the lab to not make analysis determinations based upon a
library-search result, but rather to use these electronic databases to identify unknowns. The lab
analyst identifies an unknown by comparison with a library spectrum. They then will typically
prepare a standard of the unknown and analyze the sample using the same methods they use to
analyze the unknown sample for direct comparison of the standard with their unknown, and to
evaluate whether or standard spectrum is incongruous with the unknown spectrum. This
paradigm would likely extend to field applications and would be a problem for admissibility of
field-testing results. It is important that these points be considered if results for field tests are to
be admissible in court.
One of the most important items to consider when determining whether data from
fieldportable instruments is admissible in court is the purpose of the field test. In this analytical
scheme presented, data collected in the field would be used to demonstrate that a sample is not
authentic. It is reasonable to expect that infrared, Raman, and GC-MS methods would all be
capable of performing this function effectively. However, it is important that the analyst
understands the value and the limitations of the instrument, the technology, and the test. It is also
important that the testing be performed accurately, and in accordance with established methods
and procedures.
When meeting the standards for admissibility of evidence in court in the United States, it
is likely, as previously referenced, that field-portable infrared, Raman, and GC-MS methods
would meet the standards for admissibility. However, challenges to admissibility would be faced
based upon the systems and procedures in place to describe the workflow, the standards used for
comparison, and the training and experience of the analyst. These points are important to

consider, but it is reasonable to expect that when these issues are considered and addressed,
court-admissible field tests for the detection of non-authentic samples is possible.
Summary
Analysis of pharmaceutical tablets using this analytical scheme would enable the field
analyst to identify non-authentic samples rapidly and reliably. The availability of this data
collected in the field in a searchable database format would be very valuable to forensic
scientists. A significant challenge to forensic science is the determination of source or origin of
these illicit goods. It seems that any attempt at sourcing requires access to data from many
different samples in order to identify patterns in the samples and the data to ultimately link
nonauthentic samples to each other. Collection of such data is time consuming and challenging
for various reasons. Data compiled from field analysis using appropriate methods as detailed in
this dissertation would be very beneficial. This research shows the methods proposed in this
analytical scheme for testing in the field may be used to determine source or origin of
nonauthentic samples. An accessible database would enable forensic scientists to apply these
methods to large numbers of samples and generate meaningful investigative information about
the source and origin of non-authentic samples.

CHAPTER 12. PHARMACEUTICAL DEVELOPMENT IN THE UNITED STATES
Drug development in the United States is regulated by the United States FDA. The
mission of FDA's Center for Drug Evaluation and Research (CDER) is to ensure that drugs
marketed in the United States are safe and effective. (Food and Drug Administration, 2014)
Although the mission of CDER is safety, it is important to recognize that drug development (as
regulated by FDA) is inseparably intertwined with the patenting system and protection of IP
rights. Pharmaceutical innovation and the patenting of new drugs and inventions are part of the
business strategy of pharmaceutical development (Thakur & Ramacha, 2012), and mechanisms
are in place throughout the regulated drug-development life cycle to allow IP owners to protect
their inventions.
Patent protection is the primary incentive to pharmaceutical innovators to develop new
medicines. Patent protection ensures market exclusivity for a specified period of time. This
market exclusivity is guaranteed under FDA regulation for drugs sold in the United States upon
FDA approval. This period of market exclusivity is when most of the financial investment in
development is recouped. (Melethil, 2005) For instance, Lipitor® total sales were well over
$100 billion before its patent expired. (Phelps, 2012) Failure to protect IP rights by allowing the
infiltration of counterfeit and substandard drugs that violate IP rights threatens the future of
pharmaceutical innovation because it minimizes the incentives to invest in the development of
new drugs. In terms of profitability, backing blockbuster drugs over many decades made big
pharma what it is today, but as time goes on, it has become evident that the risks of this strategy
are increasing and the rewards decreasing. (Phelps, 2012) Lack of incentive to innovate could
have devastating consequences on public health because new drugs are needed to address the
continually changing needs of public health. A review of some of the aspects of FDA regulation

of the drug-development process, especially related to the granting and protection of IP rights, is
presented for consideration.
Drug Applications
Approval to sell pharmaceuticals in the United States is usually granted through one of a
few types of applications. The first is a new drug application (NDA); the second is an
abbreviated new drug application (ANDA). Pharmaceutical innovators that bring a new drug to
the market will typically submit their request for approval using an NDA. Generic
manufacturers requesting approval to market generic versions of approved drugs will submit
their request for approval using an ANDA. A third type of application, sometimes referred to as
a section 505(b)(2) application, is different from both the NDA and ANDA. This type of
application has significant IP considerations. Section 505(b)(2) applications are specific for
applications that contains full reports of investigations of safety and effectiveness, but where at
least some of the information required for approval comes from studies not conducted by or for
the applicant, and for which the applicant has not obtained a right of reference (IP claim). In
comparison, NDA filings contain full reports of investigations of safety and effectiveness
conducted by or for the applicant; ANDA filings do not contain full reports of investigations of
safety and effectiveness, but rather contain information to show that the proposed product is,
among other things, identical to a previously approved product in active ingredient, dosage form,
strength, route of administration, labeling, quality, performance characteristics, and intended use.
The requirements for these three different applications are dramatically different and are briefly
reviewed.

New Drug Application
The NDA is the formal step a drug sponsor takes to ask that the FDA consider approving
a new drug for marketing in the United States. An NDA includes all animal and human data and
analyses of the data, as well as information about how the drug is manufactured and behaves in
the body. Once an NDA is received by FDA, the FDA has 60 days to decide whether to file it so
that it can be reviewed. The FDA can refuse to file an application that is incomplete. For
example, some required studies may be missing. (Food and Drug Administration, 2014)
Once an NDA is filed, an FDA review team of medical doctors, chemists, statisticians,
microbiologists, pharmacologists, and other experts evaluate whether the studies the sponsor
submitted show that the drug is safe and effective for its proposed use. "Safe" in this sense
means that the benefits of the drug appear to outweigh the known risks. The review team
analyzes study results and looks for possible issues with the application such as weaknesses of
the study design or analyses. Reviewers determine whether they agree with the sponsor's results
and conclusions, or whether they need any additional information to make a decision. Approval
to market is granted after the NDA and labeling are approved, and facility inspections are
completed. (Food and Drug Administration, 2014)
Although the NDA is the formal request a drug sponsor makes to FDA for approval to
market a drug in the United States, the process of drug development and the sponsors interactions
with FDA begin years before an NDA is filed. Initially, researchers work to either synthesize or
identify new drugs viable for development. Many drugs may be synthesized, but only a small
number are deemed viable for testing in animals. Most drugs that undergo testing in animals
never make it to human testing and review by the FDA. Further to this, hundreds more fail in
clinical trials. (Phelps, 2012)

Permission to test drugs in humans (which is a required step before submitting an NDA)
is only granted after an investigational new drug (IND) application has received FDA approval.
Drug sponsors submit an IND to the FDA to request permission to test the drug in humans.
During the IND approval process, the FDA scrutinizes everything about the drug from the design
of clinical trials, to the severity of side effects, to the conditions under which the drug is
manufactured. Sponsors, which may be companies, research institutions, or other organizations
taking responsibility for developing a drug, must show the FDA results of preclinical testing in
laboratory animals, as well as what they propose to do for clinical testing. At this stage, the FDA
decides whether it is reasonably safe for the drug sponsor to move forward with clinical testing.
Once an IND application is approved, the drug sponsor spends years in development to
pass through the various phases of clinical trials to ultimately submit an NDA. Clinical trials can
begin only after an IND is reviewed by the FDA and a local institutional review board (IRB). The
board is a panel of scientists and non-scientists in hospitals and research institutions that oversee
clinical research. IRB’s approve the clinical-trial protocols, which describe the type of people
who may participate in the clinical trial, the schedule of tests and procedures, the medications
and dosages to be studied, the length of the study, the study's objectives, and other details. IRB’s
make sure the study is acceptable, that participants have given consent and are fully informed of
their risks, and that researchers take appropriate steps to protect patients from harm.
The process of NDA approval is time consuming and expensive. DiMasi and colleagues
estimated average out-of-pocket direct cost per new drug is $403 million United States dollars
(2000 dollars). Capitalizing out-of-pocket costs to the point of marketing approval at a real
discount rate of 11% yielded a total pre-approval cost estimate of $802 million United States
dollars (2000 dollars). (DiMasi, Hansen, & Grabowski, 2003) There are some that have argued
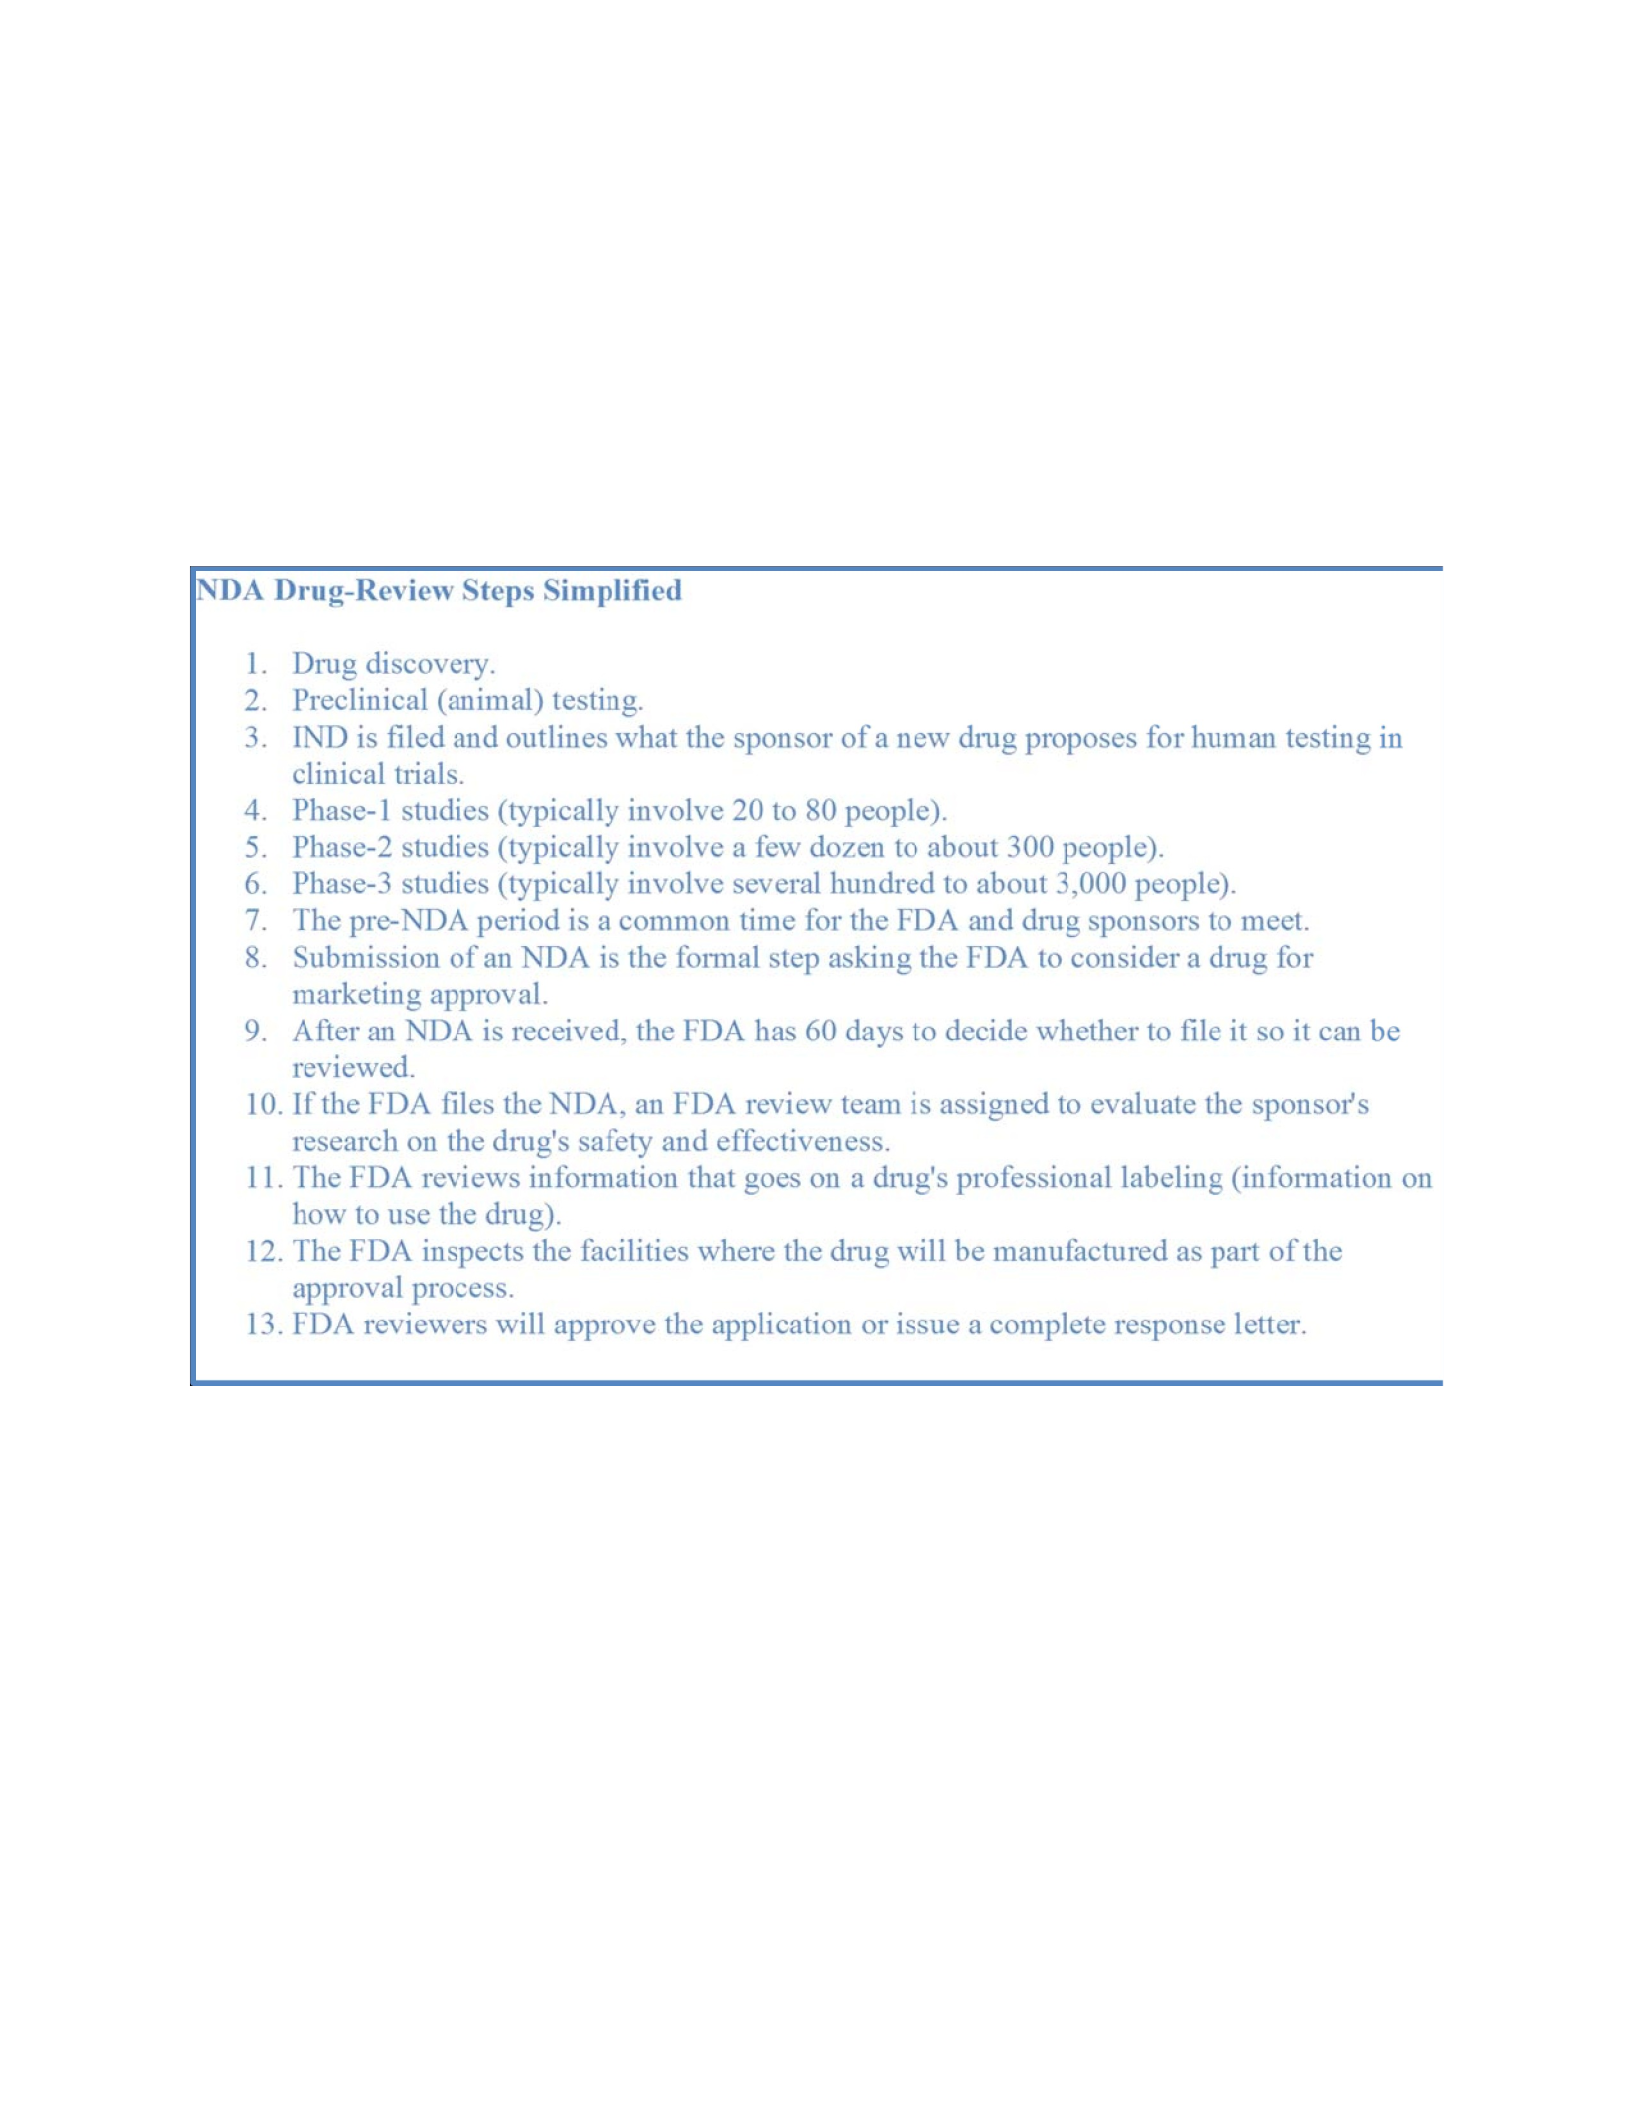
these cost estimates, but most agree that the cost to develop new drugs is in the hundreds of
millions of dollars, but can be quite variable. Variability in cost estimates depends upon many
factors including the type of drug developed. In addition to the high costs, the process of new
drug approval is long and complicated. Figure 68 summarizes the drug-development process for
drugs approved through the NDA process.
Figure 68. Summary of the drug-development process for drugs
approved for innovative drugs.
Ultimately, the result of a successful NDA filing for the drug sponsor is a new or
innovative drug for sale in the United States. Some of these drugs have been shown to control or
even eradicate sickness and disease. (International Task Force for Disease Eradication, 1993)
The role these drugs play in developed societies is critical.

Abbreviated New Drug Application
An ANDA contains data which, when submitted to FDA's CDER, Office of Generic
Drugs (OGD), provides for the review and ultimate approval of a generic drug product. The
OGD is responsible for providing regulatory oversight to expedite the availability of safe,
effective, high-quality generic drugs to patients. The OGD also provides guidance to the
regulated industry on a wide variety of clinical, scientific, and regulatory matters relating to
generic drugs. Part of the mission of the OGD is to maintain the public’s confidence in an FDA
as it continues to meet the ever-changing needs of public health.
Once an ANDA is approved, an applicant may manufacture and market the generic drug
product in the United States to provide a safe, effective, low-cost alternative to the American
public. A generic drug product is defined as one that is comparable to an innovator drug product
in dosage form, strength, route of administration, quality, performance characteristics, and
intended use.
Generic drug applications are termed "abbreviated" because drug sponsors are generally
not required to include animal and human data to establish safety and effectiveness. Instead,
generic applicants must scientifically demonstrate that their product is bioequivalent (i.e.,
performs in the same manner as the innovator drug). One way scientists demonstrate
bioequivalence is to measure the time it takes the generic drug to reach the bloodstream in 24 to
36 healthy volunteers. This gives them the rate of absorption, or bioavailability, of the generic
drug, which they can then compare to that of the innovator drug. The generic version must
deliver the same amount of active ingredients into a patient's bloodstream in the same amount of
time as the innovator drug. ANDA reviewers focus on bioequivalence data, chemistry and
microbiology data, requests for plant inspection, and drug labeling information.

The costs to bring generic drugs to the market are significantly lower than they are for
new drugs. A primary reason for this is because many of the expensive and risky aspects
required of an NDA filing are not a factor when seeking approval to market generics. Drug
discovery and proof of safety and efficacy are major components of the NDA life cycle that are
risky and expensive. They are not required steps for a generic drug sponsor. In addition, much
of the marketing and advertising costs for generics are lower than they are for new drugs. (Food
and Drug Administration, 2014)
505(b)(2) Application
A 505(b)(2) application may be submitted for a new chemical entity (NCE) when some
part of the data necessary for approval is derived from studies not conducted by or for the
applicant and to which the applicant has not obtained a right of reference. For an NCE, this data
is likely to be derived from published studies, rather than FDA's previous finding of safety and
effectiveness of a drug. It is important to note that if the applicant had a right of reference to all
of the information necessary for approval, even if the applicant had not conducted the studies, the
application would be a considered an NDA.
A 505(b)(2) application may also be submitted for applications based upon changes to a
previously approved drug product. The application may rely on FDA’s finding of safety and
effectiveness of the previously approved product, coupled with the information needed to support
the change from the approved product. The additional information could be new studies
conducted by the applicant or published data. This use of section 505(b)(2), described in the
regulations at 21 CFR 314.54, was intended to encourage innovation without creating duplicate
work and reflects the same principle as the ANDA, i.e., it is wasteful and unnecessary to carry
out studies to demonstrate what is already known about a drug.

An applicant should file a 505(b)(2) application if it is seeking approval of a change to an
approved drug that would not be permitted using an ANDA, because approval will require the
review of clinical data. However, section 505(b)(2) applications should not be submitted for
duplicates of approved products that are eligible for approval using an ANDA. In addition, an
applicant may submit a 505(b)(2) application for a change in a drug product that is eligible for
consideration pursuant to a suitability petition under Section 505(j)(2)(c) of the Act.
Although a 505(b)(2) applications is not applicable to all drug applications, there are
situations when this option is useful. Some examples include (1) applications for pediatric drugs
where results of studies performed on adults may be extrapolated to apply to pediatric patients,
(2) instances of bioequivalence where alternative formulations and new dosage strengths may be
assessed on the basis of evidence of bioequivalence, instances where modified release dosage
forms may be approved on the basis of pharmacokinetic data linking the new dosage form to an
approved immediate-release dosage form, and (3) in situations of different doses, regimens, or
dosage forms where blood levels and exposure are not very different. In these instances, it may
be possible to conclude that a new dose, regimen, or dosage form is effective on the basis of
pharmacokinetic data alone.
Drug Development and Patent Protection
The research and development required to gain approval to market a drug in the United
States is dramatically different depending upon whether the drug is innovative or generic.
Regardless of which application and subsequent development process is followed, however, FDA
regulation holds the value of IP rights to be fundamental. Mechanisms for protection of these
rights through enforcement and legal action are integrated throughout all application processes.

Pharmaceutical innovators seeking approval to market drugs in the United States using
the NDA typically are performing development for drugs they’ve either patented or licensed
from a patent holder. Patents are granted and protected for a specified amount of time. FDA
regulations require drugs be approved before they are sold in the United States. Therefore, the
earlier in the development process the inventions are patented, the shorter the amount of market
exclusivity time (because the longer the patented invention will be in the approval process.) Loss
of market-exclusivity time due to time spent gaining approval to market is an inherent
consideration of the drug-development process in the United States. The Hatch-Waxman Act
addresses these details and will be considered in chapter 14. At this point, though, it is important
to understand that it is typical that drugs approved through the NDA process are patented, and
patenting strategies attempt to reconcile time lost in development with protection of
marketexclusivity time.
The approach to patents and IP rights when seeking approval to market generic drugs
using the ANDA process is extremely different than it is for the NDA process. As part of the
ANDA process, generic manufacturers attempt to demonstrate bioequivalence of innovative
drugs that are patent protected so that these patented inventions become available to the public on
the date the patent expires. Using bioequivalence as the basis for approving generic copies of
drug products was established as part of the Hatch-Waxman Act. Typically, it is not legal to
practice patented inventions until after patents expire. In the case of pharmaceutical patents, this
is not the case. Giving generic drug sponsors rights to practice patented inventions are also an
inherent consideration of the Hatch-Waxman Act. Fundamentally, though, generic drug sponsors
are required to adhere to laws governing patented inventions. The ANDA process requires that
patent holders are advised of potential infringements of their inventions.

To begin the FDA approval process, the generic applicant must: 1) certify in its ANDA
that the patent in question is invalid or is not infringed by the generic product (known as
"paragraph IV certification"); and 2) notify the patent holder of the submission of the ANDA. If
the patent holder files an infringement suit against the generic applicant within 45 days of the
ANDA notification, FDA approval to market the generic drug is automatically postponed for 30
months, unless, before that time, the patent expires or is judged to be invalid or not infringed.
This 30-month postponement allows the patent holder time to assert its patent rights in court
before a generic competitor is permitted to enter.
Patents and IP rights are also integrated with section 505(b)(2) applications. The filing or
approval of a 505(b)(2) application may be delayed due to patent or exclusivity protections
covering an approved product. Section 505(b)(2) applications must include patent certifications
described at 21 CFR 314.50(i) and must provide notice of certain patent certifications to the
NDA holder and patent owner under 21 CFR 314.52. The certifications are required and force
the 505(b)(2) applicant to directly address their impact on the IP of others.
The drug-development process in the United States is complicated, but it is important to
recognize the value placed upon IP rights and their protection during this process. These rights
are fundamental in developed societies, and violation of them when considering pharmaceutical
innovation is a significant threat to public health. Sale of counterfeit drugs violates these IP
rights. Sale of drugs in the United States without FDA approval is illegal and poses a significant
threat to public health. Marketing of substandard drugs throughout the world undermines the
system in place in the United States for pharmaceutical innovation and poses long-term threats to
public health.

CHAPTER 13. PATENTS AND SOCIETY
As described in chapter 12, IP and IP-rights protection are fully integrated in the process
of gaining approval to market a drug for sale in the United States, regardless of which application
route the drug sponsor pursues. This type of integration demonstrates the value developed
societies like the United States place upon IP. IP fosters innovation and promotes society. The
patenting system is an important part of the drug-development process. A brief review of
patenting in the United States and patenting related to drug applications to FDA is considered.
The first issued patent in the American colonies was granted in 1641. The first United
States patent was granted in 1790. A patent in the United States is an IP right granted by the
government of the United States to an inventor “to exclude others from making, using, offering
for sale, or selling the invention throughout the United States, or importing the invention into the
United States” for a limited time in exchange for public disclosure of the invention when the
patent is granted. (The United States Patent and Trademark Office, 2014)
Patents are granted to inventions that are novel and nonobvious. They are a trade
between society and the invention owner that provides benefit to both parties. The inventor
benefits because they enjoy market exclusivity for a specified period of time. This market
exclusivity provides financial incentive to the IP owner, and may be considered reward for the
contribution their invention will make to society. In the pharmaceutical industry, the potential to
recoup the initial investment required to develop the invention during this time of market
exclusivity is a significant incentive. This is because the process to achieve FDA approval to sell
a pharmaceutical in the United States is long, risky, and expensive. Society benefits from the
patented invention because details of the best practice of the invention are available for all
members of society to use upon patent-term expiration.

Disclosure and details of the best practice of the invention is required as part of the
patenting process. However, even though the details of the invention are available for review at
the time the patent is granted, patent law forbids the practice of the invention until the patent
term expires. This is fundamental to patent law in the United States, and the concept dates back
in patent history as far as 1474 to the Venetian Senate Act.
The Venetian Act lays out all the essential features of a modern patent statute including an
established procedure to determine infringement, as well as a remedy for infringement. It covers
“devices”; states that they must be registered with a specific administrative agency; it says that
they must be “new and useful”, “reduced to perfection”, and “not previously made in this
Commonwealth”. The Venetian Act also provides a fixed patent term of ten years. Interestingly,
the Venetian Act reserved to the Republic the right to use any invention without compensating
the inventor. This is an early attempt to reconcile individual interest with the good of the
community. It implies that the inventor’s protection, provided by the grace of the state, ought
naturally to be subject to the needs of the state. (Merger, 1997)
Patents, patent terms, and patent law for drug-related and medical-device inventions in
the United States are different than they are for all other types of patents. The reason for this
difference is because society in the United States recognizes the balance between pharmaceutical
innovation and providing access to affordable medication. To account for this balance,
pharmaceutically relevant patents allow for time spent gaining FDA approval. Patent law for
pharmaceutically relevant patents also allow for legal infringement of patented inventions when
the purpose of such infringement is to provide generic alternatives upon patent-term expiration.

These differences in patent law are a result of the Hatch-Waxman Act. Patent terms and legal
infringement of patent inventions related to the pharmaceutical industry are described in this
chapter. The impact of the Hatch-Waxman Act will be discussed in chapter 14.
Patent Terms
Normally, patent terms are for 20 years. It is expected that for a normal patent, the
patent-application process may take up to three years. Since the patent term begins on the initial
date of the patent application, the effective term of a normal patent for market exclusivity is
usually about 17 years. (Pensabene & Gregory, 2013) Prior to the Hatch-Waxman Act, patent
terms were the same in the drug-development world, but the effective time for market exclusivity
was usually shorter due to the inability of the patent owner to market until FDA approval.
Gaining this approval would usually take years. Patents for inventions related to drug
development now compensate for market exclusivity time lost gaining regulatory approval.
Patent-term extension is provided for patents covering certain products and methods, including
human drug products, that are subject to FDA approval. (35 U.S.C. § 156; Eli Lilly and
Company, Petition v. Medtronic, Inc., 1990) The patent’s term can be extended by a maximum
of five years or 14 years of effective patent life, whichever is less. Specifically, the patentee is
entitled to a credit for the time the FDA was reviewing the first drug application. (Pensabene &
Gregory, 2013)
Legal Infringement
To balance the extension in patent term for pharmaceutically relevant compounds,
practice of the patented invention is allowed in some circumstances. Specifically, practice of the
invention is allowed when the goal of the practice is to develop a generic alternative that can be

provided to the consumer on the date of patent-term expiration. Prior to this change in the patent
system, generic manufacturers were required to wait until patent-term expiration before they
could begin the process of developing generic alternatives. This led to an effective extension of
market exclusivity prior to the Hatch-Waxman Act and, therefore, generic (more affordable)
alternatives took longer to reach the public.
It is clear the benefit to society of pharmaceutical innovation resulting from IP protection
cannot be overlooked. Innovative drugs have eradicated many diseases and improved quality-
oflife standards throughout the world. (International Task Force for Disease Eradication, 1993)
The patenting system is an integral part of the drug-development process. This system promotes
pharmaceutical innovation and offers long-term benefits to society.

CHAPTER 14. THE DRUG PRICE COMPETITION AND PATENT TERM
RESTORATION ACT OF 1984
The patenting of pharmaceuticals in the United States went through a significant change
as a result of the Hatch-Waxman Act. This Act establishes certain rights and procedures in
situations where a company seeks FDA approval to market a generic product prior to the
expiration of a patent or patents relating to a brand-name drug upon which the generic is based.
Congress enacted this law to facilitate the entry of lower-priced generic drugs, while maintaining
incentives for pharmaceutical companies to invest in developing new drugs.
There were two related issues that were addressed by the Hatch-Waxman Act. First,
when new drugs were discovered and patents applied for, the patent was issued before the FDA
regulatory approval process for the new drug was complete. Thus, the patent term would begin
to run, but the patent owner could not market the product until the FDA approval process was
completed. As a result, patent owners seeking FDA approval would lose time in market
exclusivity due to regulatory requirements. Second, generic drugs could not begin development
until patent terms expired. Under Roche v. Bolar (Roche Products, Inc. Appellant, v. Bolar
Pharmaceutical Co., Inc., Appellee, 1984), the testing of a generic equivalent to a patented drug
for the purpose of obtaining the FDA’s approval was deemed infringement. Accordingly, a
competitor hoping to produce a generic product had to wait until the patent expired before it
could begin the often-lengthy process of obtaining FDA approval. Thus, from the time the patent
expired until a competitor obtained FDA approval, the patent owner enjoyed an effective
extension of market exclusivity. The Hatch-Waxman Act reconciled these two related issues and
fundamentally changed the patenting system in the United States.

With regard to the first issue, patent terms under the Hatch-Waxman Act for
pharmaceutically relevant inventions were extended. Most United States patents have a 20-year
term measured from the original application’s filing date. Since the application process for most
patents (non-drug) typically takes about three years, a patent’s effective life is usually about 17
years from its issuance. This term is subject to patent-term adjustment for USPTO delays during
patent prosecution that shrink the effective life below 17 years. (35 U.S.C. § 154(b))
Virtually all drugs approved through the NDA process are covered by one or more
patents. The effective patent term for patents covering a new drug is typically much shorter than
17 years because the time-consuming FDA approval process typically occurs during part of the
patent term. As a result, branded drug companies do not enjoy the full benefit of the patent until
the FDA approves the drug and sales can begin. (Pensabene & Gregory, 2013) As a partial
remedy, the Hatch-Waxman Act provides a patent-term extension for patents covering certain
products and methods, including human drug products, which are subject to FDA approval. (35
U.S.C. § 156; Eli Lilly and Company, Petitioner v. Medtronic, Inc., 1990) The patent’s term can
be extended by a maximum of five or 14 years of effective patent life, whichever is less.
Specifically, the patentee is entitled to a credit for the time the FDA was reviewing the first drug
application. (35 U.S.C. § 156(a) and 35 U.S.C. § 156(f)(1)(A) and (2)(A))
With regard to the second issue, the Hatch-Waxman Act allowed provisions to benefit
generic drug sponsors. The first provision was that the generic drug sponsor did not have to file
an NDA to be granted approval to market the generic drug; they could file an ANDA. The
importance of this was that requirements for an ANDA filing were different than for the NDA
filing, and it was expected that the approval time and cost to gain approval via the ANDA process
would be significantly less. Under an ANDA, a generic drug company must establish that the

generic drug is effectively a duplicate of the branded, NDA drug, which is referred to as the
Reference Listed Drug (RLD). Generic manufacturers were no longer required to perform
clinical testing to prove safety and efficacy. Specifically, the generic drug company must show
that the proposed generic drug:
1. Has the same active ingredient, route of administration, dosage form, strength,
and intended use as the RLD. It also must have the same labelling, except that the generic drug
company sometimes may remove information related to a patented method or use subject to
exclusivity from its label. However, the generic drug is not required to have the same inactive
ingredients as the RLD (21 U.S.C. § 355(j)(2)(A)).
2. Is bioequivalent with the RLD so that it performs in the same manner as the RLD
in the body. Generally, a drug is bioequivalent when it delivers the same amount of active
ingredient in a patient’s bloodstream over the same amount of time as the RLD. Different but
analogous rules apply to drugs that are not delivered in the bloodstream, for example, by topical
application (21 U.S.C. § 355(j)(8)).
Additionally, the generic manufacturer must file a certification regarding patents listed in
the Orange Book (also known as Approved Drug Products with Therapeutic Equivalence
Evaluations). A Paragraph-IV Certification states that the patent is invalid or will not be
infringed and begins a process by which that question may be answered by the courts prior to
expiration of the patent. Under the Hatch-Waxman Act, FDA approval of an ANDA is
automatically stayed for 30 months when a patent owner files a patent infringement lawsuit
within 45 days of receiving a Paragraph-IV notification. During the stay, the FDA is prohibited
from approving another ANDA. Additionally, the first ANDA is granted a 180-day exclusivity
period, as an incentive whereby the generic company does not have competition from other

generic companies and can both establish market share and charge a higher price. This higher
price is quite an incentive because the prices charged are usually only slightly below the branded
drug and are a time of high profit for the generic manufacturer.
The second provision of the Hatch-Waxman Act to the benefit of generic manufacturers is
with regard to practice of a patented invention. Typically, the patented invention may not be
practiced until patent-term expiration. In these cases, though, the Hatch-Waxman Act allows
generic drug sponsors to practice the invention if the intention is to bring a generic drug to the
market. What this practically means is that the generic manufacturer no longer has to wait until
patent-term expiration to begin their development and, therefore, could be ready to market their
generic product on the date of patent-term expiration. It is important to recognize that this
privilege that is extended to generic drug sponsors to legally infringe unexpired patents is unique
to specific types of patents.
There are other details of the Hatch-Waxman Act that are important and have changed the
business of pharmaceutical development. It is clear that the intention of the Act was to balance
pharmaceutical innovation with access to affordable medication. There have, however, been
many unexpected consequences. For instance, especially in more recent years, there have been
more patent applications by generic companies and increased generic research and development
for branded products. Although clearly not a goal of Hatch-Waxman, generics innovate, often
obtaining “design-around” patents or a more efficient manufacturing process, new formulations,
or new forms of the API. (Rumore, 2009) Conversely, although there are more ANDAs filed
than ever, there are an increased number of “me-too” ANDAs and ANDAs for products which
already have a generic version. (Silver, 2007)

Although the net effect of Hatch-Waxman on pharmaceutical innovation is ambiguous, its
effect on generic drug development has been explicit, and the effect on consumers has been
beneficial. It will be argued in later chapters that the long-term benefits to the consumer are not
as clearly beneficial. One thing is for sure. The Act has instigated a flurry of litigation this is
costly. The Hatch-Waxman Act resulted in increased ANDA applications and Paragraph-IV
challenges, especially since 1998. There has also been a high success rate for patent invalidation,
particularly formulation and polymorph patents. Since the Hatch-Waxman Act, virtually all top-
selling drugs not covered by patent face generic competition; whereas pre– Hatch-Waxman, only
35% had generics available. Similarly, today more than 70% of prescriptions are for generics,
whereas prior to the Act generic prescriptions numbered 15%. (Rumore, 2009) In addition to
substantial generic penetration, Saha and colleagues also reported on the generic-to-brand price
ratio. Their analysis indicated that the average generic-to-brand price ratio for the drugs in their
sample set falls continuously following generic entry. Furthermore, as the entry rate slows down
after the first year, so does the decline in the price ratio. A month after the first generic entry, the
average price of generics for all 40 drugs evaluated was 76% of the brand price; by the end of the
first year it was 54%; by the end of the second year it was 41%. (Saha, Grabowski, Birnbaum,
Greenberg, & Bizan, 2006) Under the Hatch- Waxman Act, the average length of patent
extension is three years. Overall, there have been some reduced returns on new drugs, but
product life cycles have not changed significantly.
(Rumore, 2009)
The impact of the Hatch-Waxman Act upon pharmaceutical development has been
significant. Not only was the drug-development process impacted, but also the fundamental
rights of IP owners were changed to accommodate the balance between innovation and

accessibility of the public to needed drugs. There is no question it has increased the availability
of generic drugs to that market. Whether it will continue to foster innovation is still to be seen.
CHAPTER 15. ENFORCEMENT OF INTELLECTUAL PROPERTY RIGHTS
Enforcement of IP rights in the pharmaceutical industry, especially as related to the
availability of counterfeit, substandard and illegal-imitation drugs, is an important issue that is
complex. Even in the United States enforcement can be challenging. In other regions of the
world where societies are less developed and IP is not valued, enforcement can be daunting. It
seems appropriate, therefore, to discuss the challenges of enforcement in developed societies like
the United States separate from enforcement in underdeveloped societies.
Enforcement of Intellectual Property Rights in the United States
In the United States, IP rights are granted and infringement criteria are established. This
is especially true during drug development and approval where patent litigation amongst
innovative and generic drug companies is common. In addition, the United States has a highly
regulated drug-supply chain which minimizes the introduction of counterfeit and other illegal
pharmaceuticals into the legitimate marketplace. Finally, the functional legal system of the
United States provides a mechanism to enforce instances of IP-rights violation. This is not to say
they are always enforced; just that the framework within which they can be enforced is
established.
FDA and other regulations make it very difficult, although not impossible, for counterfeit
and other illegal pharmaceuticals to infiltrate the legitimate supply chain within the United
States. It is reasonable for a consumer in the United States to expect that, assuming they
purchase within authorized pharmacies, the pharmaceuticals they receive are what they are

purported to be. This is not to say that counterfeit and other illegal pharmaceuticals are not
available in the United States, but rather that within the regulated supply chain, they are not
encountered with great frequency.
The internet in recent years has made it significantly easier for the American consumer to
encounter counterfeit and substandard drugs. Unregulated website pharmacies have added a new
dimension to the trade of these goods. Online purchasing has become more common in the
United States. However, not all people understand that purchasing medication online is different
from purchasing other consumer goods online. Online purchases of pharmaceuticals are very
risky, especially if the consumer is unaware of what to look for to detect fraud or other illegal
activities when making these purchases. Purchases of pharmaceuticals online may result in the
receipt of drugs that are counterfeit, the wrong dose, not approved by FDA, not stored properly
during shipment, may contain unsafe ingredients, or have expired. (AWARxE Consumer
Protection Program, 2014)
During drug development, the Hatch-Waxman Act works reasonably well to enforce IP
rights. The primary legal actions taken during this time include litigation triggered by a
Paragraph-IV Certification, where a patent holder believes their patent rights have been violated
by an ANDA filing. The patent holder has the right to file a patent infringement suit against the
generic applicant and, essentially, will halt the approval of the ANDA for a period of at least 30
months. This 30-month stay allows time for the patent-infringement claims to be reviewed in a
court of law. This mechanism for enforcement is quite powerful, and enforcement of patent
protection in these cases is possible.
For the patentee, the primary reward for winning a Hatch-Waxman Act patent litigation is
continued enjoyment of any unexpired exclusivity. The typical remedies for a prevailing patentee

include (1) an order directing the FDA not to approve the ANDA before the expiration of the
relevant patents and any other exclusivity, or (2) an injunction against future infringement.
Money damages also may be available if the ANDA filer made actual sales. (Pensabene &
Gregory, 2013) In exceptional cases, attorney fees also are available to a prevailing patentee. (35
U.S.C. § 271(e)(4))
For a generic drug company, the primary reward for victory is the ability to enter the
market before patent expiration without patent-infringement liability. A prevailing generic drug
company that has tentative approval usually can immediately gain final approval and enter the
market. If not, the prevailing generic drug company can enter the market once any other
exclusivity has expired. This is typically the 180 days of exclusivity held by a first filer. A
prevailing generic drug company also may obtain attorney fees in an exceptional case.
(Pensabene & Gregory, 2013)
There are some reported abuses of the Hatch-Waxman Act and loopholes that have made
for some interesting legal actions. Some of these loopholes have been closed. One such example
of a closed loophole is with regard to the 30-month-stay provision related to ParagraphIV
Certification. Once pharmaceutical companies began to realize the advantageous nature of the
30-month stays, they also began looking for ways to extend them beyond the 30-month period.
(Eurek, 2003) According to a study conducted by the Federal Trade Commission (FTC) in 2002,
one of the most common ways that patent-holding companies were able to further delay the
market entry of generic drugs was through multiple patent listings in the Orange Book, which is
the FDA’s official list of all approved products. The FTC study identified several instances in
which brand-name companies listed patents in the Orange Book after an ANDA had already been
filed by a generic manufacturer. The effect of these later listings is that the generic applicant was

then required to recertify that the later-listed patent is also invalid or not infringed, and notify the
patent holder of the recertification. In essence, the generic applicant was required to repeat the
ANDA process for the later-listed patent. It was estimated that abuses of this kind resulted in
billions of dollars in additional sales to the patent holders. (Federal Trade
Commission, 2002)
The Hatch-Waxman Act was amended several times to close this and other loopholes, as
well as to decrease generic drug approval times. (Medicare Prescription Drug, Improvement, and
Modernization Act, 2003) Additionally, over the years, the FDA issued many guidance
documents clarifying the Hatch-Waxman Act with the goals of reducing generic approval time,
improving ANDA application quality to avoid multiple review cycles, and avoiding
timeconsuming legal delays which delayed competition. (Rumore, 2009)
There are still many legal strategies that are used by brand owners and generic sponsors
to gain or maintain market exclusivity. One such strategy is the “at-risk” product launch by
generic companies. This strategy involves launching the generic product prior to a district court
decision and injunction. In Sanofi- Synthelabo and Bristol-Myers Squibb v. Apotex
(SanofiSynthelabo, Inc.; and Bristol-Myers Squibb Sanofi Pharmaceuticals Holding Partnership,
Plaintiffs, -against- Apotex Inc.; and Apotex Corp., Defendants, 2006), three months prior to the
FDA’s approving the ANDA, the generic company notified Sanofi that it would launch the
generic Plavix® product “at risk.” The generic was launched stocking pharmacy shelves over
the course of approximately three weeks, while Sanofi filed for a preliminary injunction. Three
weeks later, an injunction was granted with Apotex being “at risk” that it would lose some of the
180-day exclusivity period and the market share associated with it. Even though the court ruled
in favor of Sanofi, it denied Sanofi’s request to recall the product that had already been

distributed prior to the injunction. During the three-week distribution that occurred as a result of
the at-risk launch, Apotex supplied enough of the generic drug pharmacy’s to maintain
domination over Plavix for five months. Apotex received hundreds of millions of dollars in
revenue for this at-risk launch. (Pechersky, 2007)
In spite of this and other “legal strategies” that have changed the business of drug
development in the United States as a result of the Hatch-Waxman Act, it is important to
recognize that this law is intended to balance innovation with access to affordable medicines.
Fundamentally, the United States legal and regulatory system affords patent owners with the
tools they need to enforce IP rights. This is clearly the case when it comes to patent litigation
surrounding drug approvals.
Patent infringement related to other kinds of sales of pharmaceutical is a little bit more
difficult to manage than it is for infringement related to legal sales of pharmaceutical within the
United States. For example, generic versions of VIAGRA ordered online that are shipped to the
consumer located in the United States from overseas are illegal. In this case, the generic
VIAGRA is illegal because it is sold in the United States without FDA approval. It is also illegal
because it violates the IP rights of Pfizer, the brand owner. These are related, but fundamentally
different reasons they are illegal, and each violation has different implications. With regard to
lack of FDA approval, the primary implication of this problem is that the safety and efficacy of
the drug is truly unknown. With regard to the IP-rights violations of Pfizer, this is a problem that
is monetary and social. However, it is difficult to prosecute against the infringer primarily due to
jurisdictional issues.

As an example, sample SING1 analysed as part of this research was purportedly (by
package labelling) manufactured in India, shipped from Singapore (by postmarking), and
received in the United States. What is the appropriate jurisdiction within which to file action?
Jurisdiction issues challenges not only IP rights in the pharmaceutical industry, but also other
industries as well. (Dean, 2013)
It seems one of the best ways, at least in theory, to enforce IP rights protection for
violations against United States patents is by way of trade agreements. In the 1980’s the United
States was undergoing a transformation from an industrial to an information economy. Waning
industrial competitiveness hurt United States companies and international trade. The United
States began searching for new areas of commerce which would help it maintain
competitiveness. IP emerged as "a new basis of comparative advantage." (Subramanian, 1991)
There were several IP-related industries, namely the entertainment industry (records and
movies), pharmaceutical industry, and the computer industry who were becoming extremely
important contributors to the United States economy. Computer technology, for example, was a
huge growth industry during the mid-1980s. During this time, personal computers hit the
market, computer programs became hot items, and computer software emerged as a commercial
product with a high level of economic return. (Halbert, 1997) The United States worked to
protect its IP interests and as a result has become the international advocate for strong IP-rights
protection. This advocacy has been the motivating force behind the inclusion of IP rights in the
general agreement on tariffs and trade (GATT), the United States-Canada free-trade agreement,
and the North American free-trade agreement. (Halbert, 1997)
In 2009, the Commissioner of the USPTO stated that his office works closely with the
Office of the United States Trade Representative (USTR) and other agencies to establish, on a

bilateral and multilateral basis, workable treaty commitments and trade agreements. For
example, Commerce worked with the USTR on matters concerning the IP chapter for several free
trade agreements (FTAs) during FY 2008, most notably negotiation of the IP chapter of the
United States-Malaysia FTA, Costa Rica’s implementation of the Dominican Republic-Central
America-United States FTA, Peru’s implementation of the United States-Peru Trade Promotion
Agreement, as well as implementation of the United States-Chile FTA. Commerce also
contributed to the development of the United States’ World Trade Organization (WTO) dispute
settlement case against China relating to deficiencies in its IP rights enforcement regime. In
addition, the USPTO co-chair’s the IP Rights Working Group in the United States-China Joint
Commission on Commerce and Trade (JCCT), the United States’ ongoing trade dialogue with
China. (Stoll, 2009)
It can be argued that unless trade agreements mandate the consideration of IP rights by
foreign countries, there is virtually no recognition of such rights. (Bird, 2006) For instance, in
China and India, coercion attempts by the United States centered on trade agreements were used
to get these countries to recognize IP rights. While there is genuine concern about whether or not
coercion attempts garner real benefit in the long term, infringement of IP rights in these countries
is a significant problem for brand owners. (Bird, 2006) Brazil faced overwhelming
pharmaceutical industry pressure and governmental threats of sanctions from the United States to
improve its patent protection for drug products and processes. Brazil finally gave up its
resistance by discussing IP rights at the GATT, joined the trade-related aspects of IP rights
(TRIPS) agreement, and now possesses a functioning patent approval system that is fairly
consistent with the minimum standards of protection required by TRIPS. (Schulz & Wu, 2004)

The TRIPS agreement is quite comprehensive in its understanding of the complexity of
the IP-protection problem, especially within the pharmaceutical industry. The agreement
requires all WTO Member States to grant patents for pharmaceutical products or process
inventions for a minimum of 20 years. (Velásquez & Boulet, 1999) Prior to the agreement, many
Member States granted no rights at all, or had terms as short as five years. It also provides time
for developing countries to modify and enact laws that made them compliant with the agreement.
The agreement also allows the use of compulsory licenses, a provision that demonstrates an
understanding of the need for some consideration of public-health crises (even if at the expense
of an individual’s right to own property). Compulsory licensing enables a competent government
authority to license the use of a patented invention to a third party or government agency without
the consent of the patent holder. Article 31 of the Agreement sets forth a number of conditions
for the granting of compulsory licenses. These include a case-bycase determination of
compulsory-license applications, the need to demonstrate prior (unsuccessful) negotiations with
the patent owner for a voluntary license, and the payment of adequate remuneration to the patent
holder. Where compulsory licenses are granted to address a national emergency or other
circumstances of extreme urgency, certain requirements are waived in order to hasten the
process, such as that for the need to have had prior negotiations to obtain a voluntary license
from the patent holder. (World Health Organization, 2014)
Enforcement of Intellectual Property Rights in Underdeveloped Societies
Even in the best of cases where IP-rights protections are in place like for Member States
of the TRIPS agreement, enforcement of these rights is not common. (Bird, 2006) Interestingly,
it has been proposed that although social benefits may arise from patent protection through the
discovery of new drugs, the TRIPS agreement standards derive from those of industrialized

countries and are not necessarily appropriate for the development level of all countries.
(Velásquez & Boulet, 1999) As previously mentioned, IP rights are fundamental in a developed
society like the United States, but their value is not always appreciated.
It is likely the best chance for success of IP-rights enforcement on a global scale is by
way of trade agreements. However, there are other efforts taking place that may lead to the
improvement of IP-rights enforcement. The USPTOs Attaché Program was formally instituted in
2006 for the benefit of United States economic and political interests abroad to promote the value
and importance of strong IP protection and enforcement in selected, high-profile countries where
United States IP challenges are greatest. In partnership with Commerce’s more broadly scoped
Foreign Commercial Service and the Department of State, the IP Attachés are sent out to
strengthen global IP protection and enforcement overseas. The IP-rights experts support United
States embassies and consulates on IP-rights issues, including devising strategies to stop
counterfeiting and piracy, and supporting United States government efforts to improve the
protection and enforcement of IP rights. The Attachés also advocate United States IP policies,
coordinate training on IP rights matters, and assist United States businesses that rely on IP-rights
protection abroad. These Attachés serve at posts in Brazil, Russia, India, China, Thailand, and the
United States Missions in Geneva. (Stoll, 2009)
Although hopeful, it seems lofty to expect that IP-rights enforcement related to
pharmaceuticals in countries of extreme poverty will improve in the short term. The unfortunate
aspect of this realization is that these environments are the ones where counterfeit drugs pose the
greatest short- and long-term threat to public health. (USP Council of the Convention Section on
Global Public Health, USP Council of Experts International Health Expert Committee, USP
Regionalization Team, Heyman, & Williams, 2011) The administration of counterfeit or

substandard drugs is the norm in these regions of the world. In fact, without the availability of
counterfeit or substandard drugs, no drugs at all would be available for these citizens, which can
be considered a problem if government has a moral responsibility to guarantee its citizens a right
to a healthy life. (Gewertz & Amado, 2004) Some researchers have proposed charging
counterfeiters with manslaughter when a patient’s death can be directly linked to performance of
a counterfeit medication. (Newton, et al., 2006) This approach may help curb the influx of
counterfeit goods because it would increase the risk counterfeiters take when supplying these
harmful drugs. Currently, the risk/reward ratio for counterfeiters in these regions of the world is
so low that there is no incentive to deter these criminals from engaging in these activities.
(Thomas, 2011) There have been instances where safe medications have been delivered to these
regions of the world, but abuses in these situations are also a big problem that frequently lead to
the diversion of these goods from their intended recipients. (Médecins Sans Frontières Australia,
2014)
It seems fair to say that public health is the most significant factor when considering the
counterfeit-drug problem, but coupled with this public-health aspect is the need for developed
societies to support innovation by enforcing IP rights. Extending this to underdeveloped and
extremely poor countries is important. Enforcement is challenging, and in many regions of the
world mechanisms are in place to support this enforcement.
CHAPTER 16. FUNDING OF PHARMACEUTICAL RESEARCH
Funding of pharmaceutical research comes from a variety of different sources including
private industry, government, not-for-profit organizations, and individual donors. The cost of
research and development, by all accounts, is high. In 2003, DiMasi and colleagues reported in

the Journal of Health Economics that the pre-approval cost estimate for development of a drug in
the United States is $802 million (year 2000 United States dollars). (DiMasi, Hansen, &
Grabowski, 2003) Since this time, this number has been cited frequently.
In addition to the cost estimate, these authors determined that the average successfully
developed new molecular entity (NME) in the study’s sampled required 4.3 years for discovery
and preclinical development, and another 7.5 years for clinical trials and FDA approval.
Approval itself took an average of 1.5 years. Thus, developing an NME and bringing it to
market required 11.8 years, on average. (DiMasi, Hansen, & Grabowski, 2003) Table 11
summarizes estimated costs calculated in the study, as well as average research length for both
preclinical and clinical phases of development. The study surveyed drugs from a representative
set of therapeutic classes, but it excluded some types of new drugs that have lower average
research and development costs. In addition, the estimate may not be representative of research
and development costs for smaller pharmaceutical firms which did not participate in the survey
on which the study was based. However, by focusing on NME’s, the study did base its cost
estimate on the types of drugs that have been the source of most pharmaceutical breakthroughs.
(Congressional Budget Office, 2006)
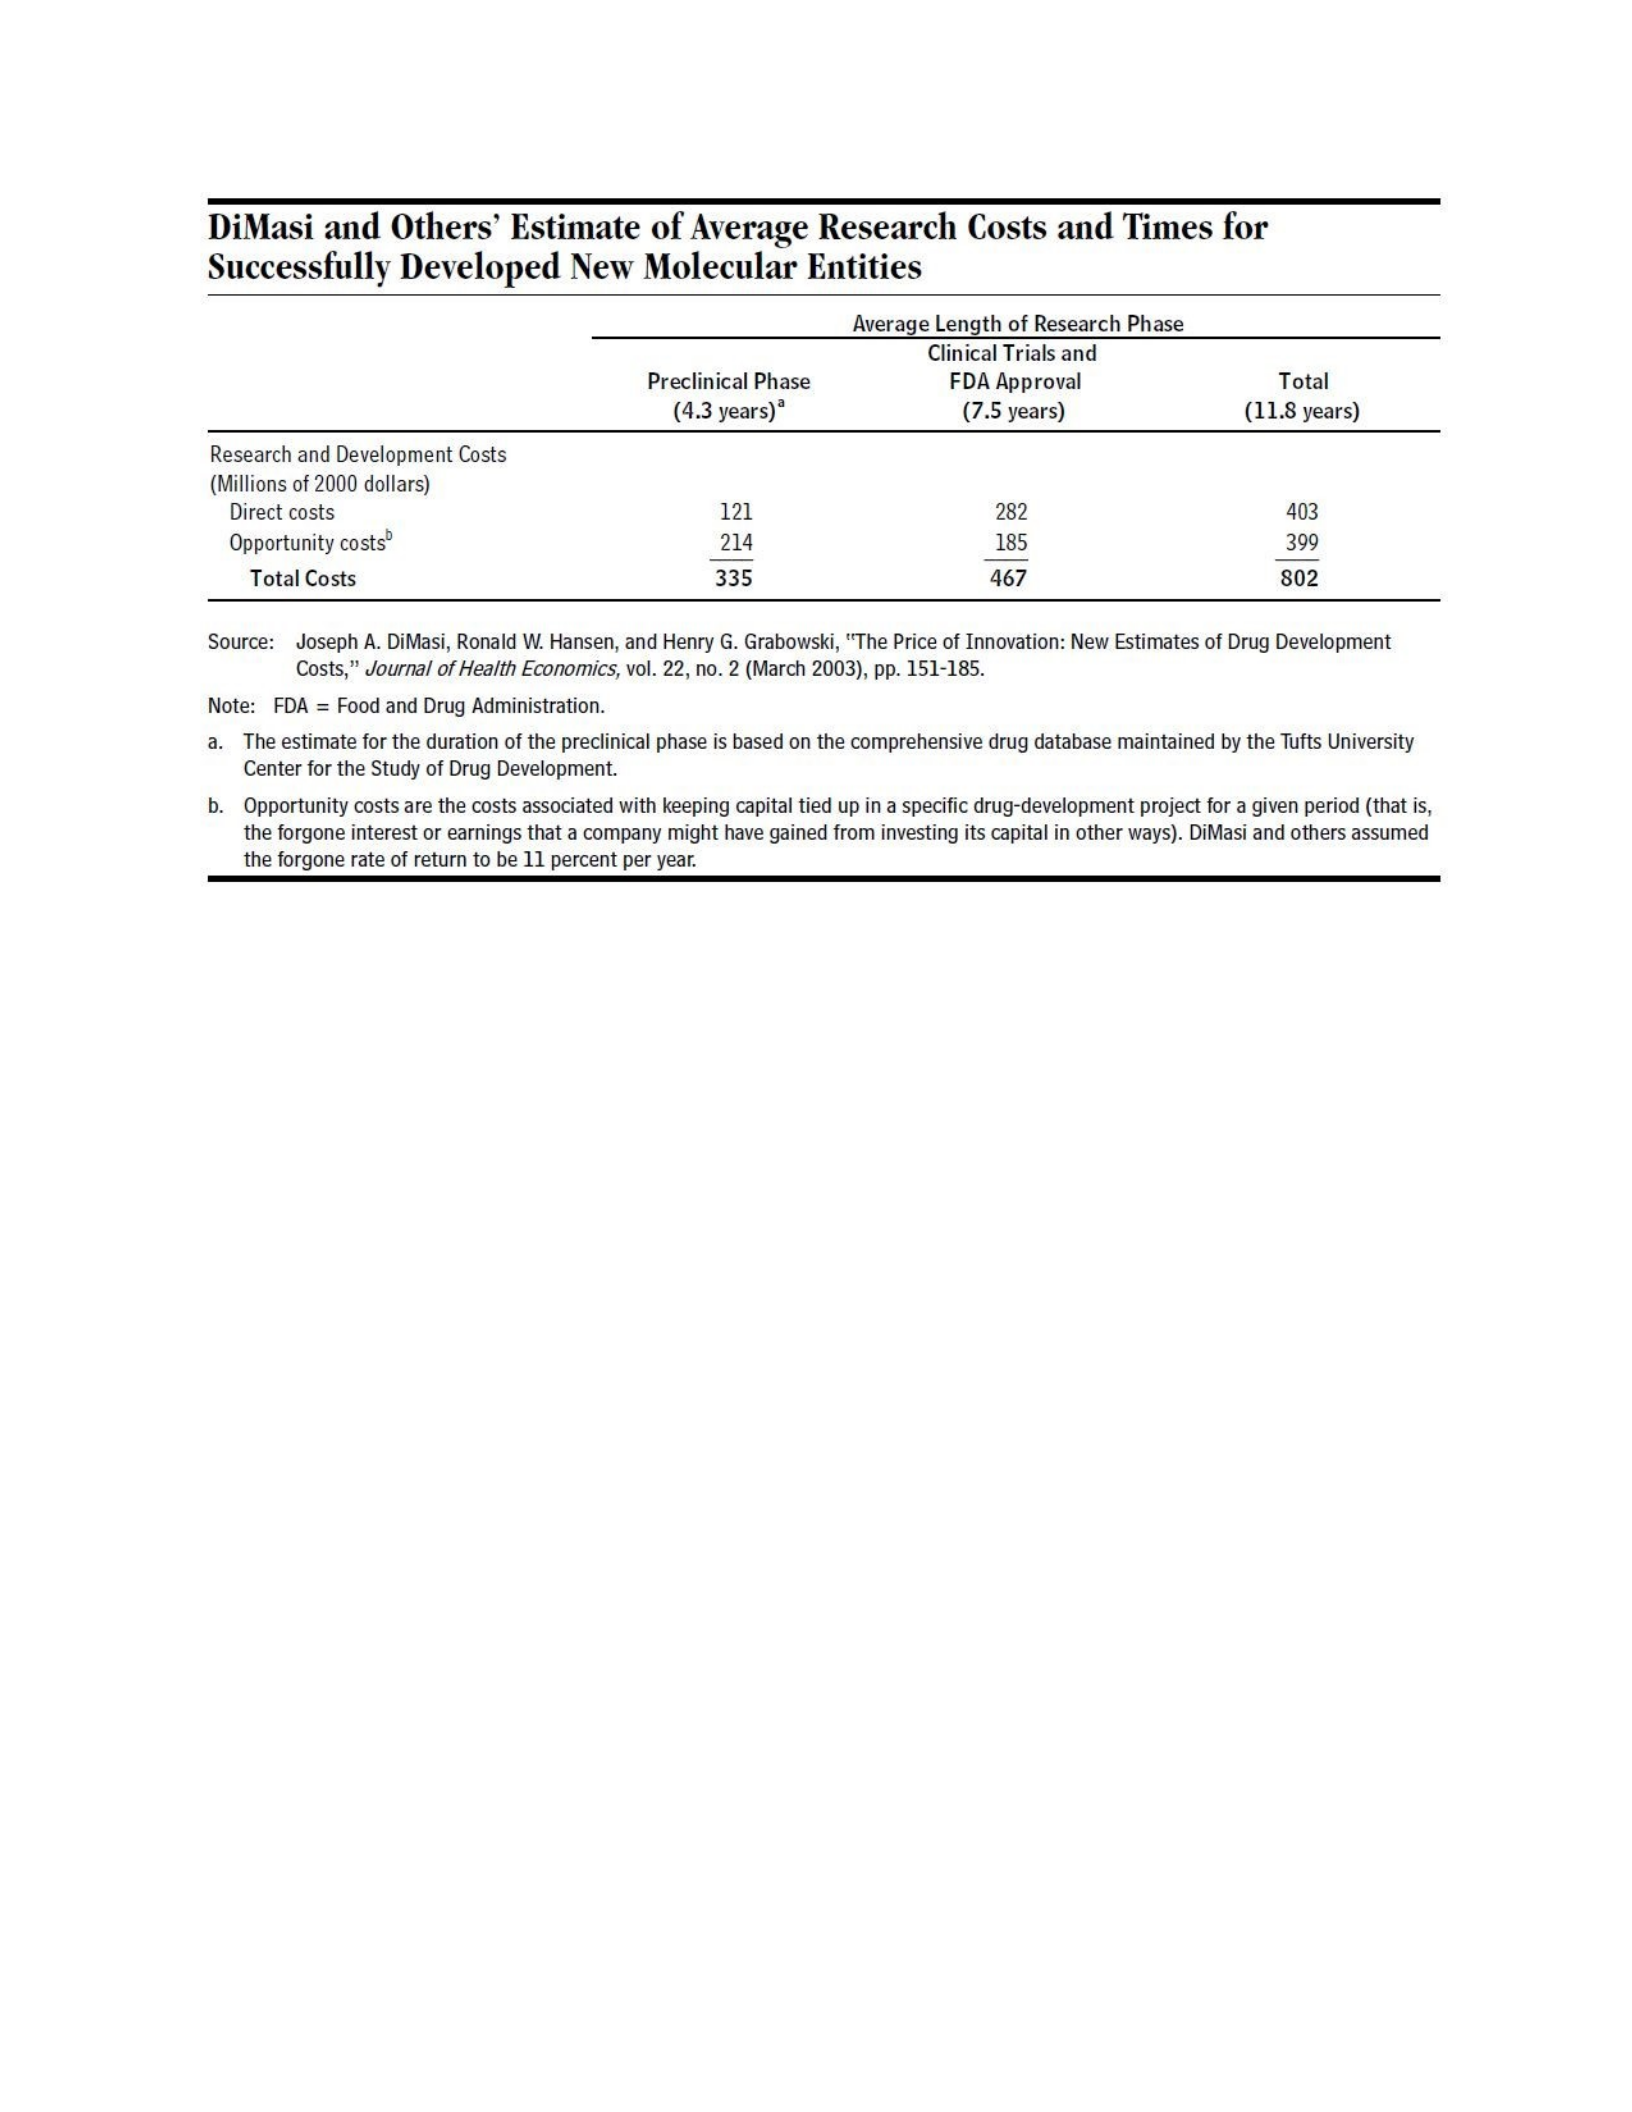
Table 11. Estimated costs calculated by DiMasi et al. Reprinted
from Congressional Budget Office Report, Research and
Development in the Pharmaceutical Industry, 2006.
Although this study is widely cited, it is not without its critics. Light and Warburton
discuss various reasons for the problems with the calculation of this $802-million value. They
state, among other things, that this cost figure, like the ones that preceded it, is based on
confidential, unsystematic data, and has dubious scientific validity. (Light & Warburton, 2005)
They state that given the many independent sources of variability, any point estimate is
misleading. Reporting that research and development costs of major new drugs range from $300
million to $1.3 billion rather than a single average point estimate, therefore, is more informative.
(Light & Warburton, Setting the record straight in the reply by DiMasi, Hansen and Grabowski,
2005)
While the exact cost of pharmaceutical research and development is an elusive number to
determine, it is clear that regardless of how costs are calculated, the numbers are high and usually

in the hundreds of millions of dollars. (Adams & Brantner, 2006; Light & Warburton, 2011) For
every successful new drug, a firm will have had many failed drug projects that did not survive
clinical trials or that never won approval from the FDA. Estimates of average research and
development costs per drug include the costs of those failures. (Congressional Budget Office,
2006)
These high costs have been used by the pharmaceutical industry for decades to justify
high sale prices. (Melethil, 2005) When you follow the money, it seems that the largest
percentage of research and development costs is, indeed, paid by industry. Government makes a
significant contribution, though. Health-related research receives the second largest amount of
federal support for research and development (behind only defense-related research). That
support has been steadily growing for several decades. Spending on research by the National
Institutes of Health (NIH)—by far the primary recipient of government funding for health-related
basic research—was $5.8 billion (in 2005 dollars) in 1970. This figure more than doubled to
$12.3 billion by 1990. By 2004, it reached $28.5 billion. In comparison, spending reported on
research and development by the members of the Pharmaceutical Research and Manufacturers of
America (PhRMA) was just two-thirds the size of NIH’s spending in 1980. PhRMA’s research
and development spending surpassed NIH’s in 1987 and has remained higher since then,
although both grew at similar rates in the late 1990s and early 2000s. Figure 69 shows the
spending data from 1970 to 2004. (Congressional Budget Office, 2006)
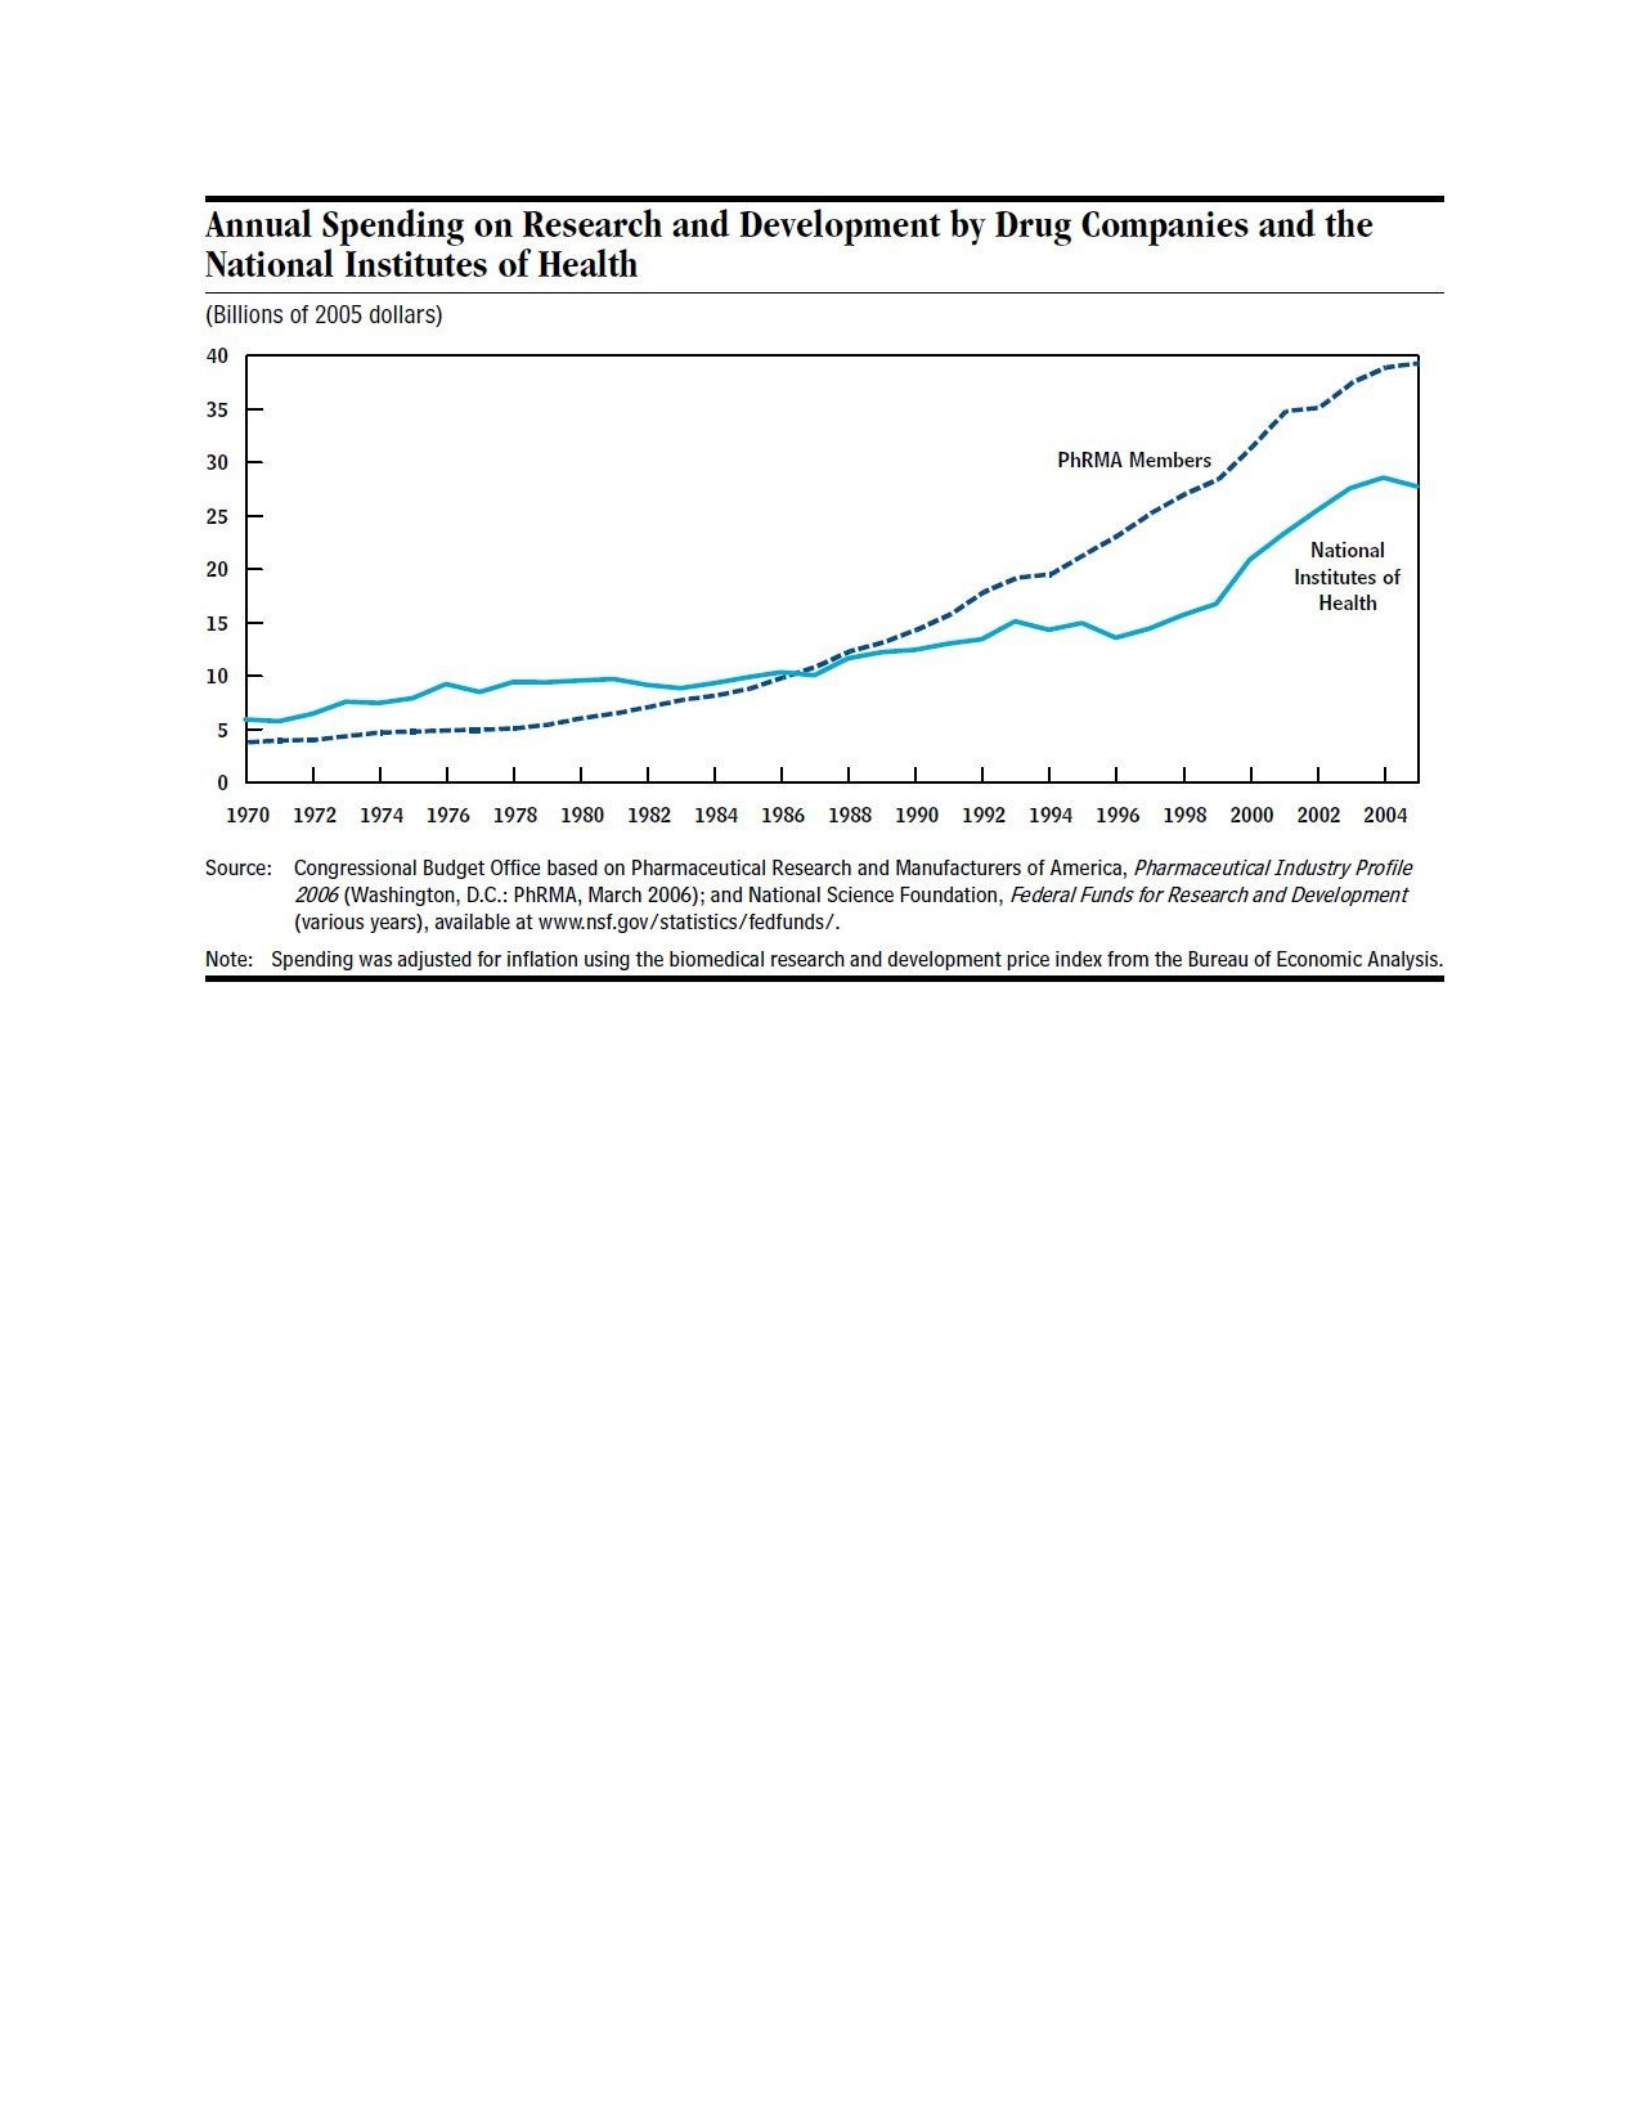
Figure 69. Research and development spending between 1970 and
2004 for PhRMA members and NIH.
Government funding has a very strong role in pharmaceutical research and development,
but it tends to have a different role than funding from private industry. Much of the government
funding is for the basic research on disease mechanisms that underlie the search for new drugs.
Although the distinction between basic and applied research and development is not always clear,
it is useful to think of basic research as generating information or knowledge that is not readily
embodied in physical products. Federally supported basic research in genomics, molecular
biology, and other life sciences has greatly expanded the drug industry’s technological
opportunities, stimulating private investment in pharmaceutical research and development.
(Congressional Budget Office, 2006)
The rationale for government funding of basic scientific research is simple; if such
research was left solely to the private sector, too little of it would be done. The benefits to

society from doing additional basic research and development (beyond what firms alone would
conduct) far outweighs the costs. A company’s incentive to invest in research and development
is limited to its own expected returns. In the case of basic research and development, those
returns can be particularly low compared with the social benefits. It can be difficult for private
companies to capture more than a small fraction of the total social value of their basic research.
A basic research discovery cannot be patented unless the inventor can credibly describe the
discovery’s “specific and substantial” utility. (Congressional Budget Office, 2006) It seems the
role government funding plays is to create new knowledge and new tools and produce large
numbers of highly trained researchers, all of which are a direct and important input to
privatesector research. (Cockburn & Henderson, 2001)
Some argue that since government funding can be linked to drug products that provide
profit to private industry, government should share in the profits. This approach has historically
not been met with success. Prior to 1980, invention rights supported by government agencies
belonged strictly to the federal government. Nobody could exploit such research without tedious
negotiations with the federal agency concerned. Worse, companies found it near impossible to
acquire exclusive rights to a government-owned patent. Without these rights, few firms were
willing to invest millions more of their own money to turn a raw research idea into a marketable
product. The result was that inventions and discoveries made in American universities, teaching
hospitals, national laboratories, and non-profit institutions sat in warehouses gathering dust. Of
the 28,000 patents that the American government owned in 1980, fewer than 5% had been
licensed to industry. Although taxpayers were footing the bill for 60% of all academic research,
they were getting hardly anything in return. (Innovation's golden goose, 2002)

The transfer of rights from the government to the inventor and university came about in
1980 with the Bayh-Dole Act. The philosophy behind the Act was the belief that the solution lay
with the individual and that the best thing government could do to provide incentives for success
was get out of the way of these individuals. (Stevens, 2004) Government would then benefit in
two ways. First, the therapeutic value of new medications would be considered a primary benefit
to government. (Congressional Budget Office, 2006) Second, even though rights were
transferred from government to individual researchers and universities, some financial benefits to
government were written into the law to help the government get better results from their funding
than had historically been the case. Even so, the government has not had much success
collecting on these financial incentives primarily due to its own administrative issues. (Gerth &
Stolberg, 2000)
The Bayh-Dole Act was designed to push federally financed research from the university
laboratory into the marketplace. Scientists who made discoveries using taxpayer money were
required to file invention reports with the government. Universities were directed to license
patented inventions to companies that would commercialize them. The law was originally
passed to aid small businesses, but later it was modified so that even big companies could
benefit. If a company did not develop a product quickly enough, the government could revoke
the company's license and hand the job over to a competitor. It could also take control of an
invention to alleviate "health or safety needs," the law said. (Gerth & Stolberg, 2000)
Once an invention is on the market, the law grants the government the right to buy it
without paying customary royalties. At the same time, other laws enable federal agencies to put
taxpayer-financed inventions out to competitive bidding. For example, the government could
give other companies (besides the developer) the opportunity to manufacture and sell at a lower

price, but only to the government. However, in the first 20 years after the Bayh-Dole bill became
law, the government had not taken advantage of these provisions. One reason is because the
government already buys drugs cheaply by purchasing them in bulk. More importantly, even if
federal officials wanted to use the Bayh-Dole Act to get medicines at still cheaper prices, they
could not because they do not keep track of products, including drugs that are invented with
taxpayer money. In addition, a preliminary report by the inspector general's office of the
Department of Health and Human Services found that as many as 22 percent of discoveries
financed by the health institutes were not reported by universities, as is required. More than
2,000 inventions developed with government money were reported to the health institutes in
1999, but agency officials said in interviews that they had no idea which, if any, companies had
licensed those inventions, or how they were being used. (Gerth & Stolberg, 2000)
In spite of the government’s inability to fully collect financially from the drug products it
funds, the Bayh-Dole Act did invigorate industry. Coming out of World War II, the United States
was unchallenged in its political and economic leadership of the free world. However, by the end
of the 1970s it was clear that United States industry had lost its international competitiveness to
Europe and, particularly, to Japan. This process had started with the success of the United States
programs to rebuild its Allies and former enemies and was completed by the impact of the oil
shocks of the 1960s and 1970s on an economy dependent on cheap domestic energy. Examples
of the loss of competitiveness abounded. The United States lost leadership roles in both mature
industries, such as automobiles and televisions, and emerging industries, such as memory chips.
In addition, Japanese companies dominated in industries based on American and European
innovations, such as video cassette recorders and compact discs. The Bayh-Dole Act was

intended to reverse this trend toward non-competitiveness. It is unlikely that anyone in the
technology-transfer community would dispute its importance. (Stevens, 2004)
Although initially the concern with this Act was that the government would provide
investment and not reap reward, it seems the most important criticism is the impact it has had on
academic institutions and their role in society. Many scientists, economists, and lawyers believe
the Act distorts the mission of universities, diverting them from the pursuit of basic knowledge,
which is freely disseminated, to a focused search for results that have practical and industrial
purposes. Whether that is a bad thing is a matter of debate. (Bayhing for blood or Doling out
cash? Intellectual property, 2005)
What is not in dispute is that it makes American academic institutions behave more like
businesses than neutral arbiters of truth. For example, a study published in 2003 by Jerry and
Marie Thursby of Emory University and the Georgia Institute of Technology, respectively,
reported that a survey of industry licensing executives shows 27% of their university licenses
include clauses that allow deletion of information from papers before submission, and 44% ask
for publication delay (3.9 months on average) (Thursby & Thursby, 2003)
Moreover, there is ample evidence that scientific research is being delayed, deterred, or
abandoned due to the presence of patents and proprietary technologies. Researchers (and
particularly their minders in university patent-licensing offices) are increasingly reluctant to
share materials and knowledge with others unless such sharing is accompanied by legal
agreements about "reach-through" royalties on potential findings and the right to restrict
publication of results. A study released by the American Association for the Advancement of
Science noted that 35% of academic biotechnology researchers experience difficulties getting
hold of patented technologies that they need for their work, even though non-commercial

research is supposed to be exempt from the normal restrictions of patents. The question is just
how "non-commercial" is such research? Lawsuits between universities and researchers over
patents and royalties are now common. (Bayhing for blood or Doling out cash? Intellectual
property, 2005)
Even industry is starting to complain about a gold-digger mentality among academic
administrators. The most notorious example is Columbia University, which tested the boundaries
of the law by seeking to re-patent a technique whose patent had already expired. The patent was
for a technology called co-transformation that is used to place external DNA into cells, and is
important in making certain drugs. Columbia eventually backed down, but only in the face of
both public criticism and a series of writs from biotechnology companies. (Bayhing for blood or
Doling out cash? Intellectual property, 2005)
Another case ensnared the University of Utah, which licensed its patent on a gene
underlying hereditary breast cancer exclusively to one company, Myriad Genetics. That gave
Myriad Genetics a monopoly on diagnostic testing for the disease, which was controversial
enough. Then the firm started suing universities that were using its technology in follow-up
research, bringing the non-commercial research exemption still further into question. (Bayhing
for blood or Doling out cash? Intellectual property, 2005)
Funding has changed in recent years. Neither federal nor private support for research and
development is likely to return to the trend that produced a tripling in support between 1995 and
2005. As a portion of gross domestic product, research and development expenditures are
declining in the United States and Europe, whereas they are increasing at an accelerated rate in
China, South Korea, and India. (Moses III & Dorsey, 2012) In 2009, privately funded

